Related Experiment Video
Updated: Mar 28, 2026

Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Critical Assessment of TD-DFT for Excited States of Open-Shell Systems: I. Doublet-Doublet Transitions
1Beijing National Laboratory for Molecular Sciences, Institute of Theoretical and Computational Chemistry, State Key Laboratory of Rare Earth Materials Chemistry and Applications, College of Chemistry and Molecular Engineering, and Center for Computational Science and Engineering, Peking University , Beijing 100871, People's Republic of China.

Abstract:
A benchmark set of 11 small radicals is set up to assess the performance of time-dependent density functional theory (TD-DFT) for the excited states of open-shell systems. Both the unrestricted (U-TD-DFT) and spin-adapted (X-TD-DFT) formulations of TD-DFT are considered. For comparison, the well-established EOM-CCSD (equation-of-motion coupled-cluster with singles and doubles) is also used. In total, 111 low-lying singly excited doublet states are accessed by all the three approaches. Taking the MRCISD+Q (multireference configuration interaction with singles and doubles plus the Davidson correction) results as the benchmark, it is found that both U-TD-DFT and EOM-CCSD perform well for those states dominated by singlet-coupled single excitations (SCSE) from closed-shell to open-shell, open-shell to vacant-shell, or closed-shell to vacant-shell orbitals. However, for those states dominated by triplet-coupled single excitations (TCSE) from closed-shell to vacant-shell orbitals, both U-TD-DFT and EOM-CCSD fail miserably due to severe spin contaminations. In contrast, X-TD-DFT provides balanced descriptions of both SCSE and TCSE. As far as the functional dependence is concerned, it is found that, when the Hartree-Fock ground state does not suffer from the instability problem, both global hybrid (GH) and range-separated hybrid (RSH) functionals perform grossly better than pure density functionals, especially for Rydberg and charge-transfer excitations. However, if the Hartree-Fock ground state is instable or nearly instable, GH and RSH tend to underestimate severely the excitation energies. The SAOP (statistically averaging of model orbital potentials) performs more uniformly than any other density functionals, although it generally overestimates the excitation energies of valence excitations. Not surprisingly, both EOM-CCSD and adiabatic TD-DFT are incapable of describing excited states with substantial double excitation characters.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:40
MO Theory and Covalent Bonding
 02:45
The Quantum-Mechanical Model of an Atom
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 03:21
The Energies of Atomic Orbitals
 02:35
Molecular Orbital Theory I
 02:38
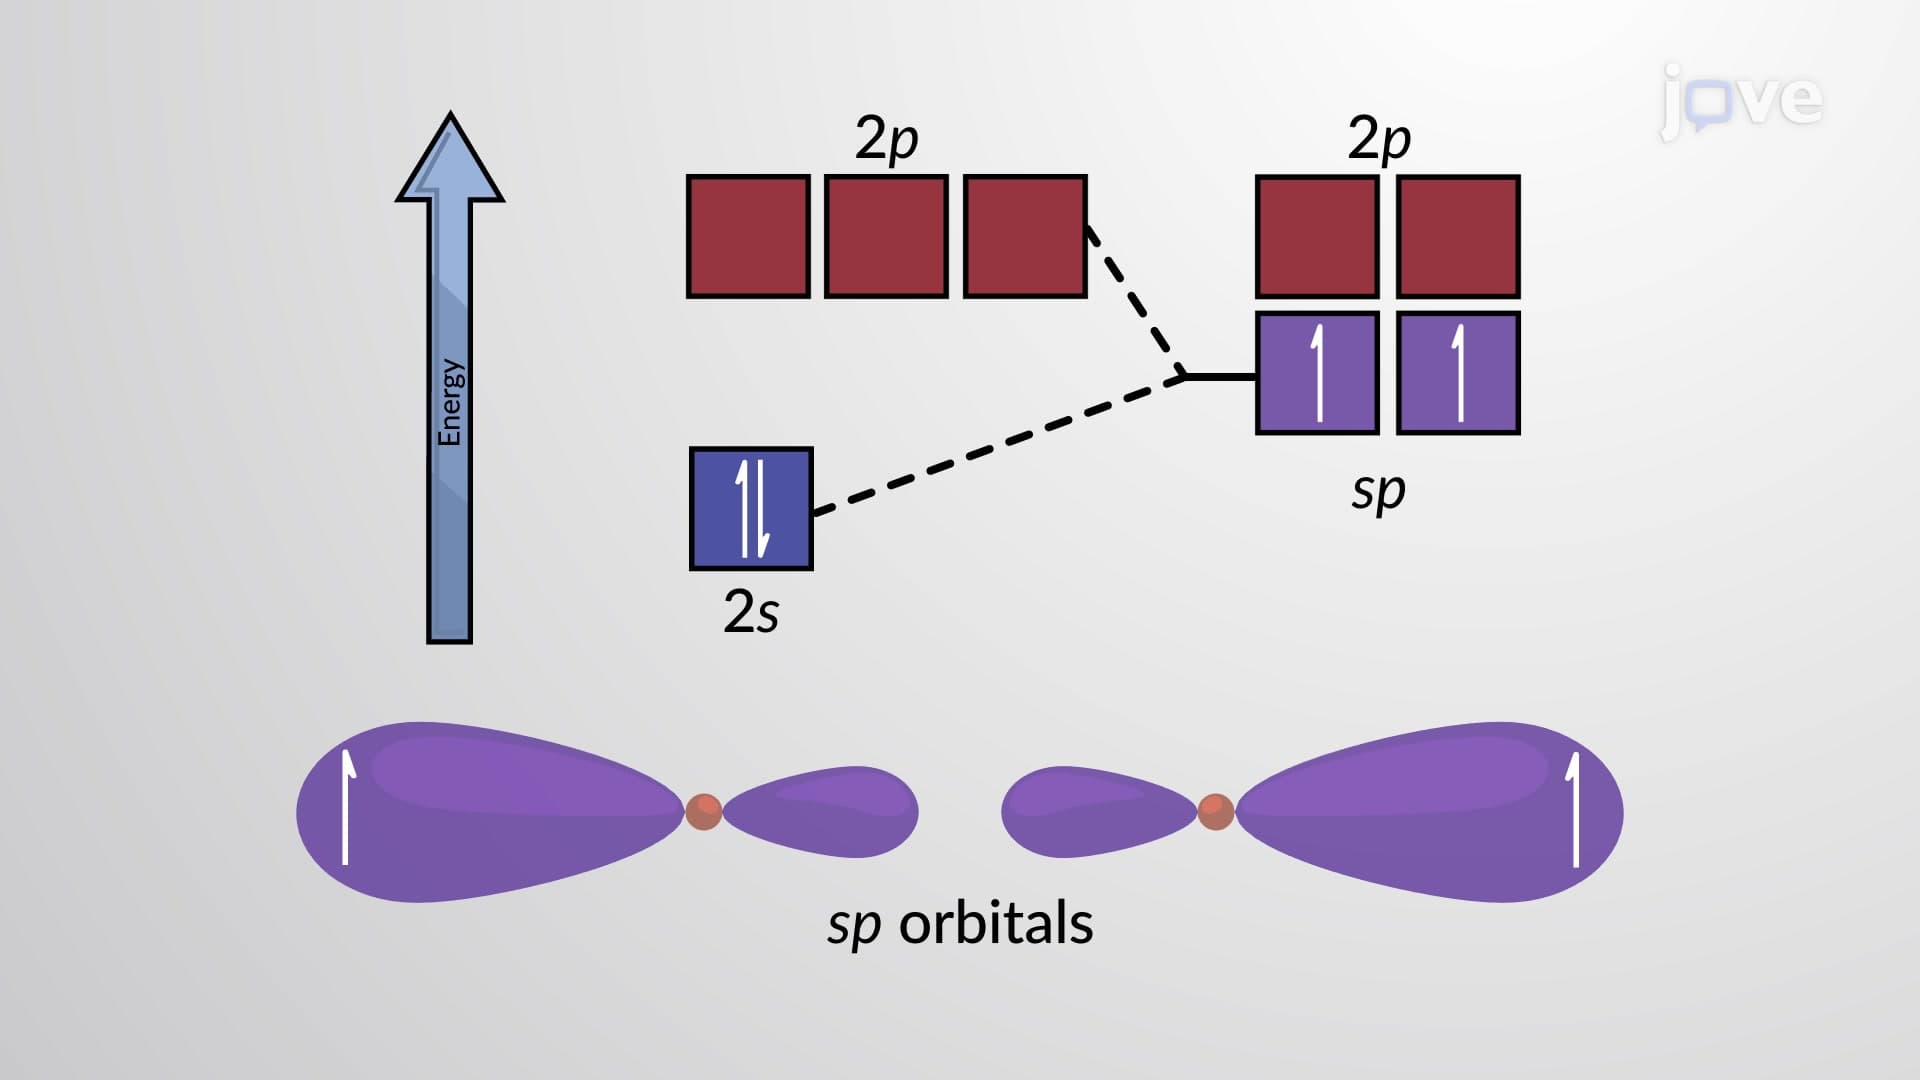
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...