Related Experiment Video
Updated: Mar 28, 2026

Synthesis and Characterization of Functionalized Metal-organic Frameworks
Published on: September 5, 2014
Exceptional H2 sorption characteristics in a Mg(2+)-based metal-organic framework with small pores: insights from
Tony Pham1, Katherine A Forrest, Eduardo H L Falcão
1Department of Chemistry, University of South Florida, 4202 East Fowler Avenue, CHE205, Tampa, FL 33620-5250, USA. juergen@usf.edu brian.b.space@gmail.com.

Abstract:
Experimental sorption measurements, inelastic neutron scattering (INS), and theoretical studies of H2 sorption were performed in α-[Mg3(O2CH)6], a metal-organic framework (MOF) that consists of a network of Mg(2+) ions coordinated to formate ligands. The experimental H2 uptake at 77 K and 1.0 atm was observed to be 0.96 wt%, which is quite impressive for a Mg(2+)-based MOF that has a BET surface area of only 150 m(2) g(-1). Due to the presence of small pore sizes in the MOF, the isosteric heat of adsorption (Qst) value was observed to be reasonably high for a material with no open-metal sites (ca. 7.0 kJ mol(-1)). The INS spectra for H2 in α-[Mg3(O2CH)6] is very unusual for a porous material, as there exist several different peaks that occur below 10 meV. Simulations of H2 sorption in α-[Mg3(O2CH)6] revealed that the H2 molecules sorbed at three principal locations within the small pores of the framework. It was discovered through the simulations and two-dimensional quantum rotation calculations that different groups of peaks correspond to particular sorption sites in the material. However, for H2 sorbed at a specific site, it was observed that differences in the positions and angular orientations led to distinctions in the rotational tunnelling transitions; this led to a total of eight identified sites. An extremely high rotational barrier was calculated for H2 sorbed at the most favorable site in α-[Mg3(O2CH)6] (81.59 meV); this value is in close agreement to that determined using an empirical phenomenological model (75.71 meV). This rotational barrier for H2 exceeds those for various MOFs that contain open-metal sites and is currently the highest yet for a neutral MOF. This study highlights the synergy between experiment and theory to extract useful and important atomic level details on the remarkable sorption mechanism for H2 in a MOF with small pore sizes.
More Related Videos
 10:27
Preparation of Highly Porous Coordination Polymer Coatings on Macroporous Polymer Monoliths for Enhanced Enrichment of Phosphopeptides
Published on: July 14, 2015
 07:14
Author Spotlight: Experimental Approaches for the Synthesis of Low-Valent Metal-Organic Frameworks from Multitopic Phosphine Linkers
Published on: May 12, 2023
Related Concept Videos
 00:56
Extraction: Advanced Methods
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 01:23
Properties of Organometallic Compounds
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:09
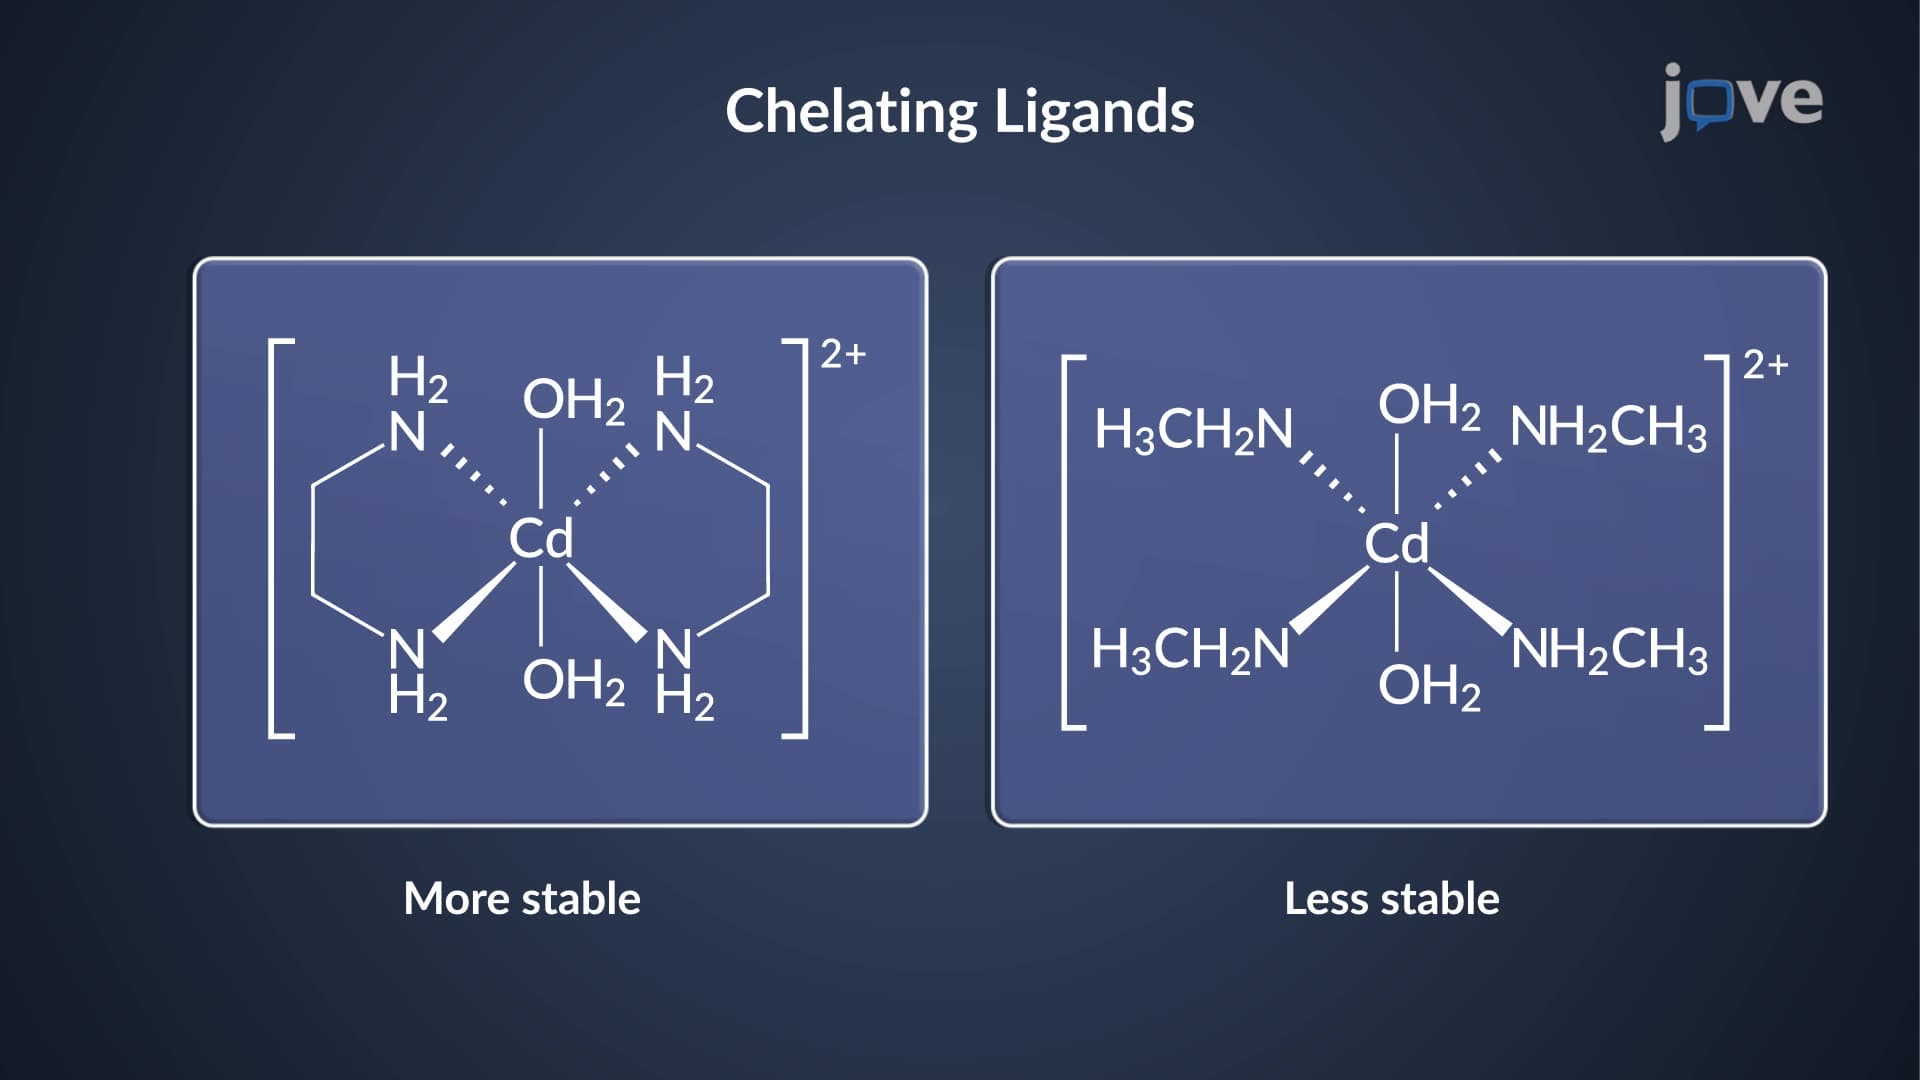
Complexation Equilibria: Factors Influencing Stability of Complexes
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...