Related Experiment Video
Updated: Mar 27, 2026
Interactive Molecular Model Assembly with 3D Printing
Published on: August 13, 2020
Factor Analysis of Conformations and NMR Signals of Rotaxanes: AIMD and Polarizable MD Simulations
Pingying Liu1,2, Wei Li1, Zigui Kan1
1School of Chemistry and Chemical Engineering, Key Laboratory of Mesoscopic Chemistry of MOE, Nanjing University , Nanjing 210093, People's Republic of China.

Abstract:
The interlocked ⟨rod | ring⟩ structures of pseudorotaxanes and [2]rotaxanes are usually maintained by the complex hydrogen-bonding (H-bonding) network between the rod and ring. Ab initio molecular dynamics (AIMD) using generalized energy-based fragmentation approach and polarizable force field (polar FF)-based molecular dynamics (MD) simulations were performed to investigate the conformational changes of mechanically interlocked systems and to obtain the ensemble-averaged NMR chemical shifts. Factor analysis (FA) demonstrates that the ring H-donor (2,6 pyridinedicarboxamide group) plays an important role in the ring-rod recognition. In comparison to the conventional fixed-charge force field, the polarization effect is crucial to account for the H-bonding interactions in supramolecular systems. In the hybrid scheme, the polar FF-based MD simulations are used to generate different initial states for the AIMD simulations, which are able to give better prediction of ensemble-averaged NMR signals for chemically equivalent amide protons. The magnitude of the deshielding shift of NMR signal is correlated with the length of hydrogen bond. The polar FF model with variable charges shows that the dipole-dipole interactions between the flexible diethylene glycol chain of ring and polar solvents induce the upfield shifts of NMR signals of rod H-donors and the directional distribution of the neighboring CH3CN solvents.
More Related Videos
 14:44
Structure and Coordination Determination of Peptide-metal Complexes Using 1D and 2D 1H NMR
Published on: December 16, 2013
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...
 02:18
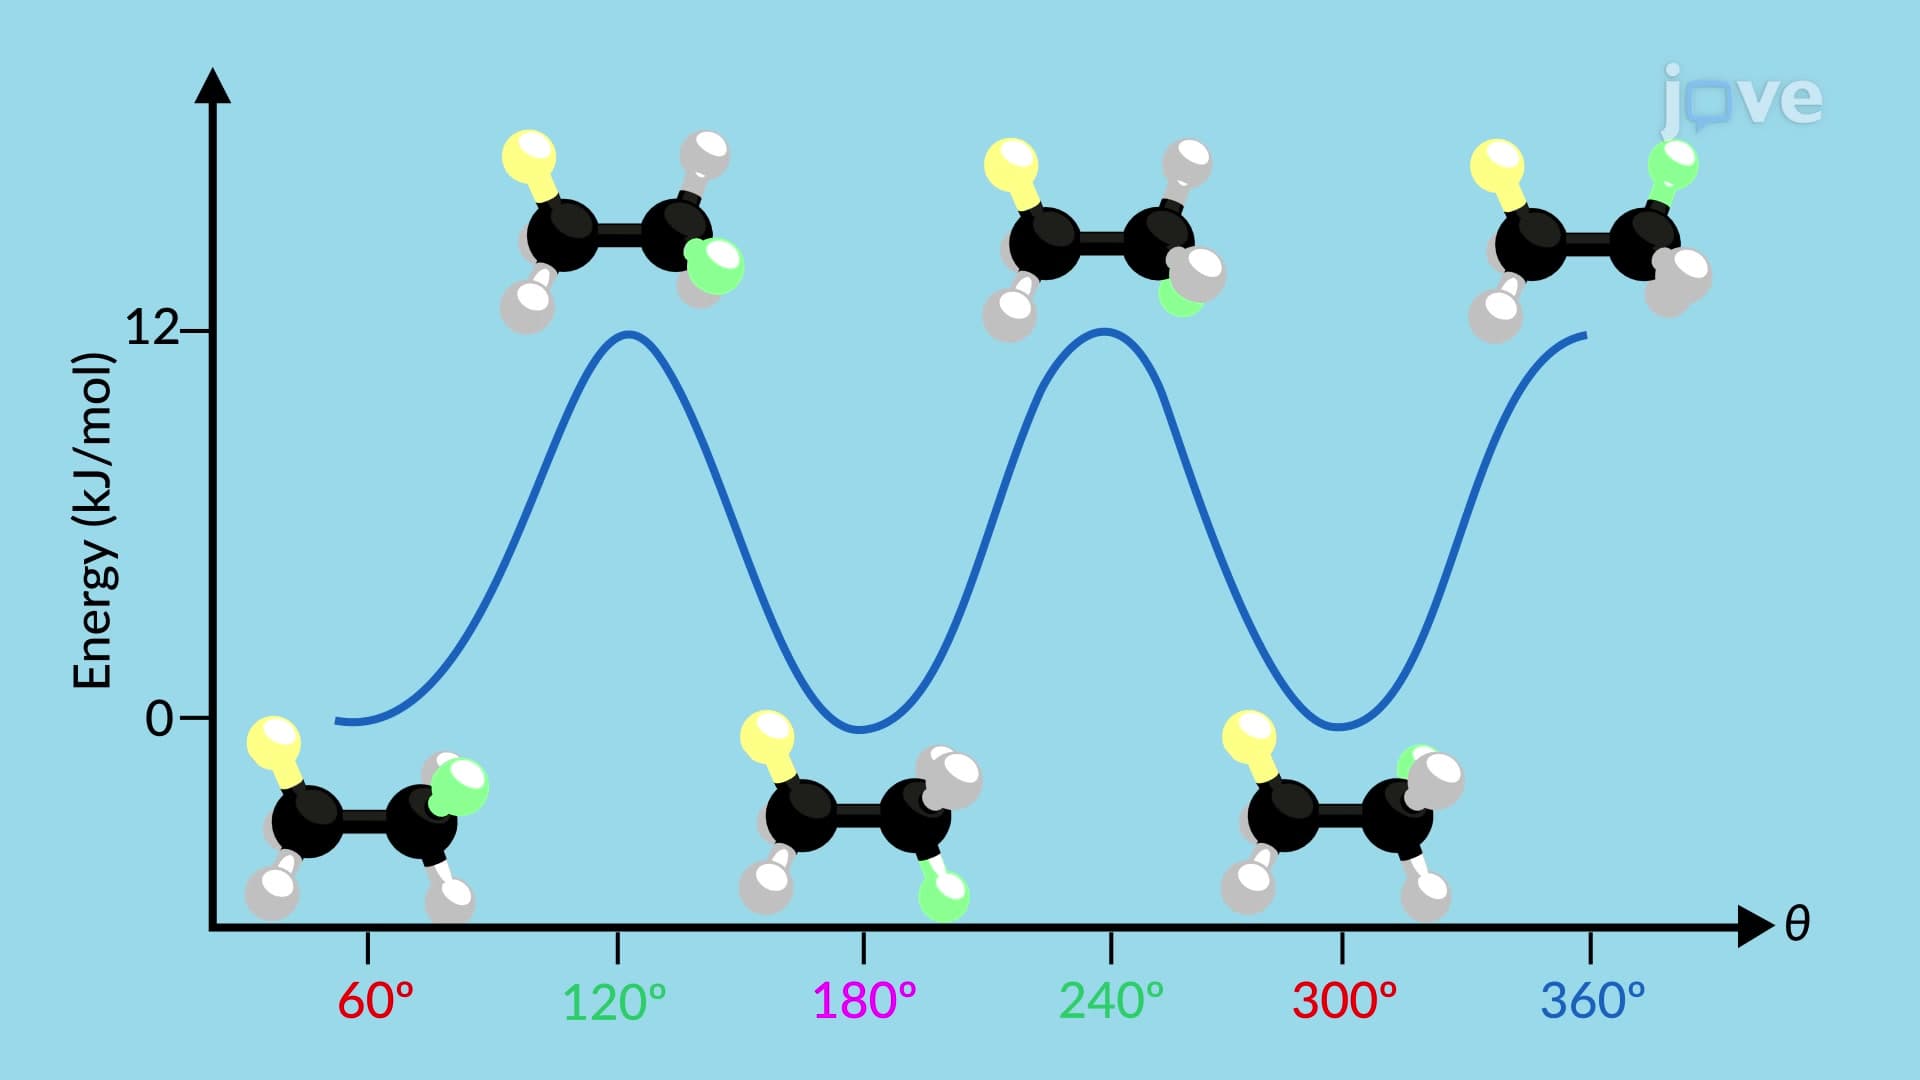
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered...
 02:02
Chair Conformation of Cyclohexane
The hydrogen atoms linked to carbons are arranged in two different axial and equatorial orientations to achieve this...
 02:06
Newman Projections
The organic molecules rotate across the single bonds leading to numerous temporary three-dimensional structures of varying energy known as...