Related Experiment Video
Updated: Mar 25, 2026

Workflow and Tools for Crystallographic Fragment Screening at the Helmholtz-Zentrum Berlin
Published on: March 3, 2021
CCSD(T)/CBS fragment-based calculations of lattice energy of molecular crystals
Ctirad Červinka1, Michal Fulem1, Květoslav Růžička1
1Department of Physical Chemistry, University of Chemistry and Technology, Prague, Technická 5, CZ-166 28 Prague 6, Czech Republic.

Abstract:
A comparative study of the lattice energy calculations for a data set of 25 molecular crystals is performed using an additive scheme based on the individual energies of up to four-body interactions calculated using the coupled clusters with iterative treatment of single and double excitations and perturbative triples correction (CCSD(T)) with an estimated complete basis set (CBS) description. The CCSD(T)/CBS values on lattice energies are used to estimate sublimation enthalpies which are compared with critically assessed and thermodynamically consistent experimental values. The average absolute percentage deviation of calculated sublimation enthalpies from experimental values amounts to 13% (corresponding to 4.8 kJ mol(-1) on absolute scale) with unbiased distribution of positive to negative deviations. As pair interaction energies present a dominant contribution to the lattice energy and CCSD(T)/CBS calculations still remain computationally costly, benchmark calculations of pair interaction energies defined by crystal parameters involving 17 levels of theory, including recently developed methods with local and explicit treatment of electronic correlation, such as LCC and LCC-F12, are also presented. Locally and explicitly correlated methods are found to be computationally effective and reliable methods enabling the application of fragment-based methods for larger systems.
Related Concept Videos
 02:54
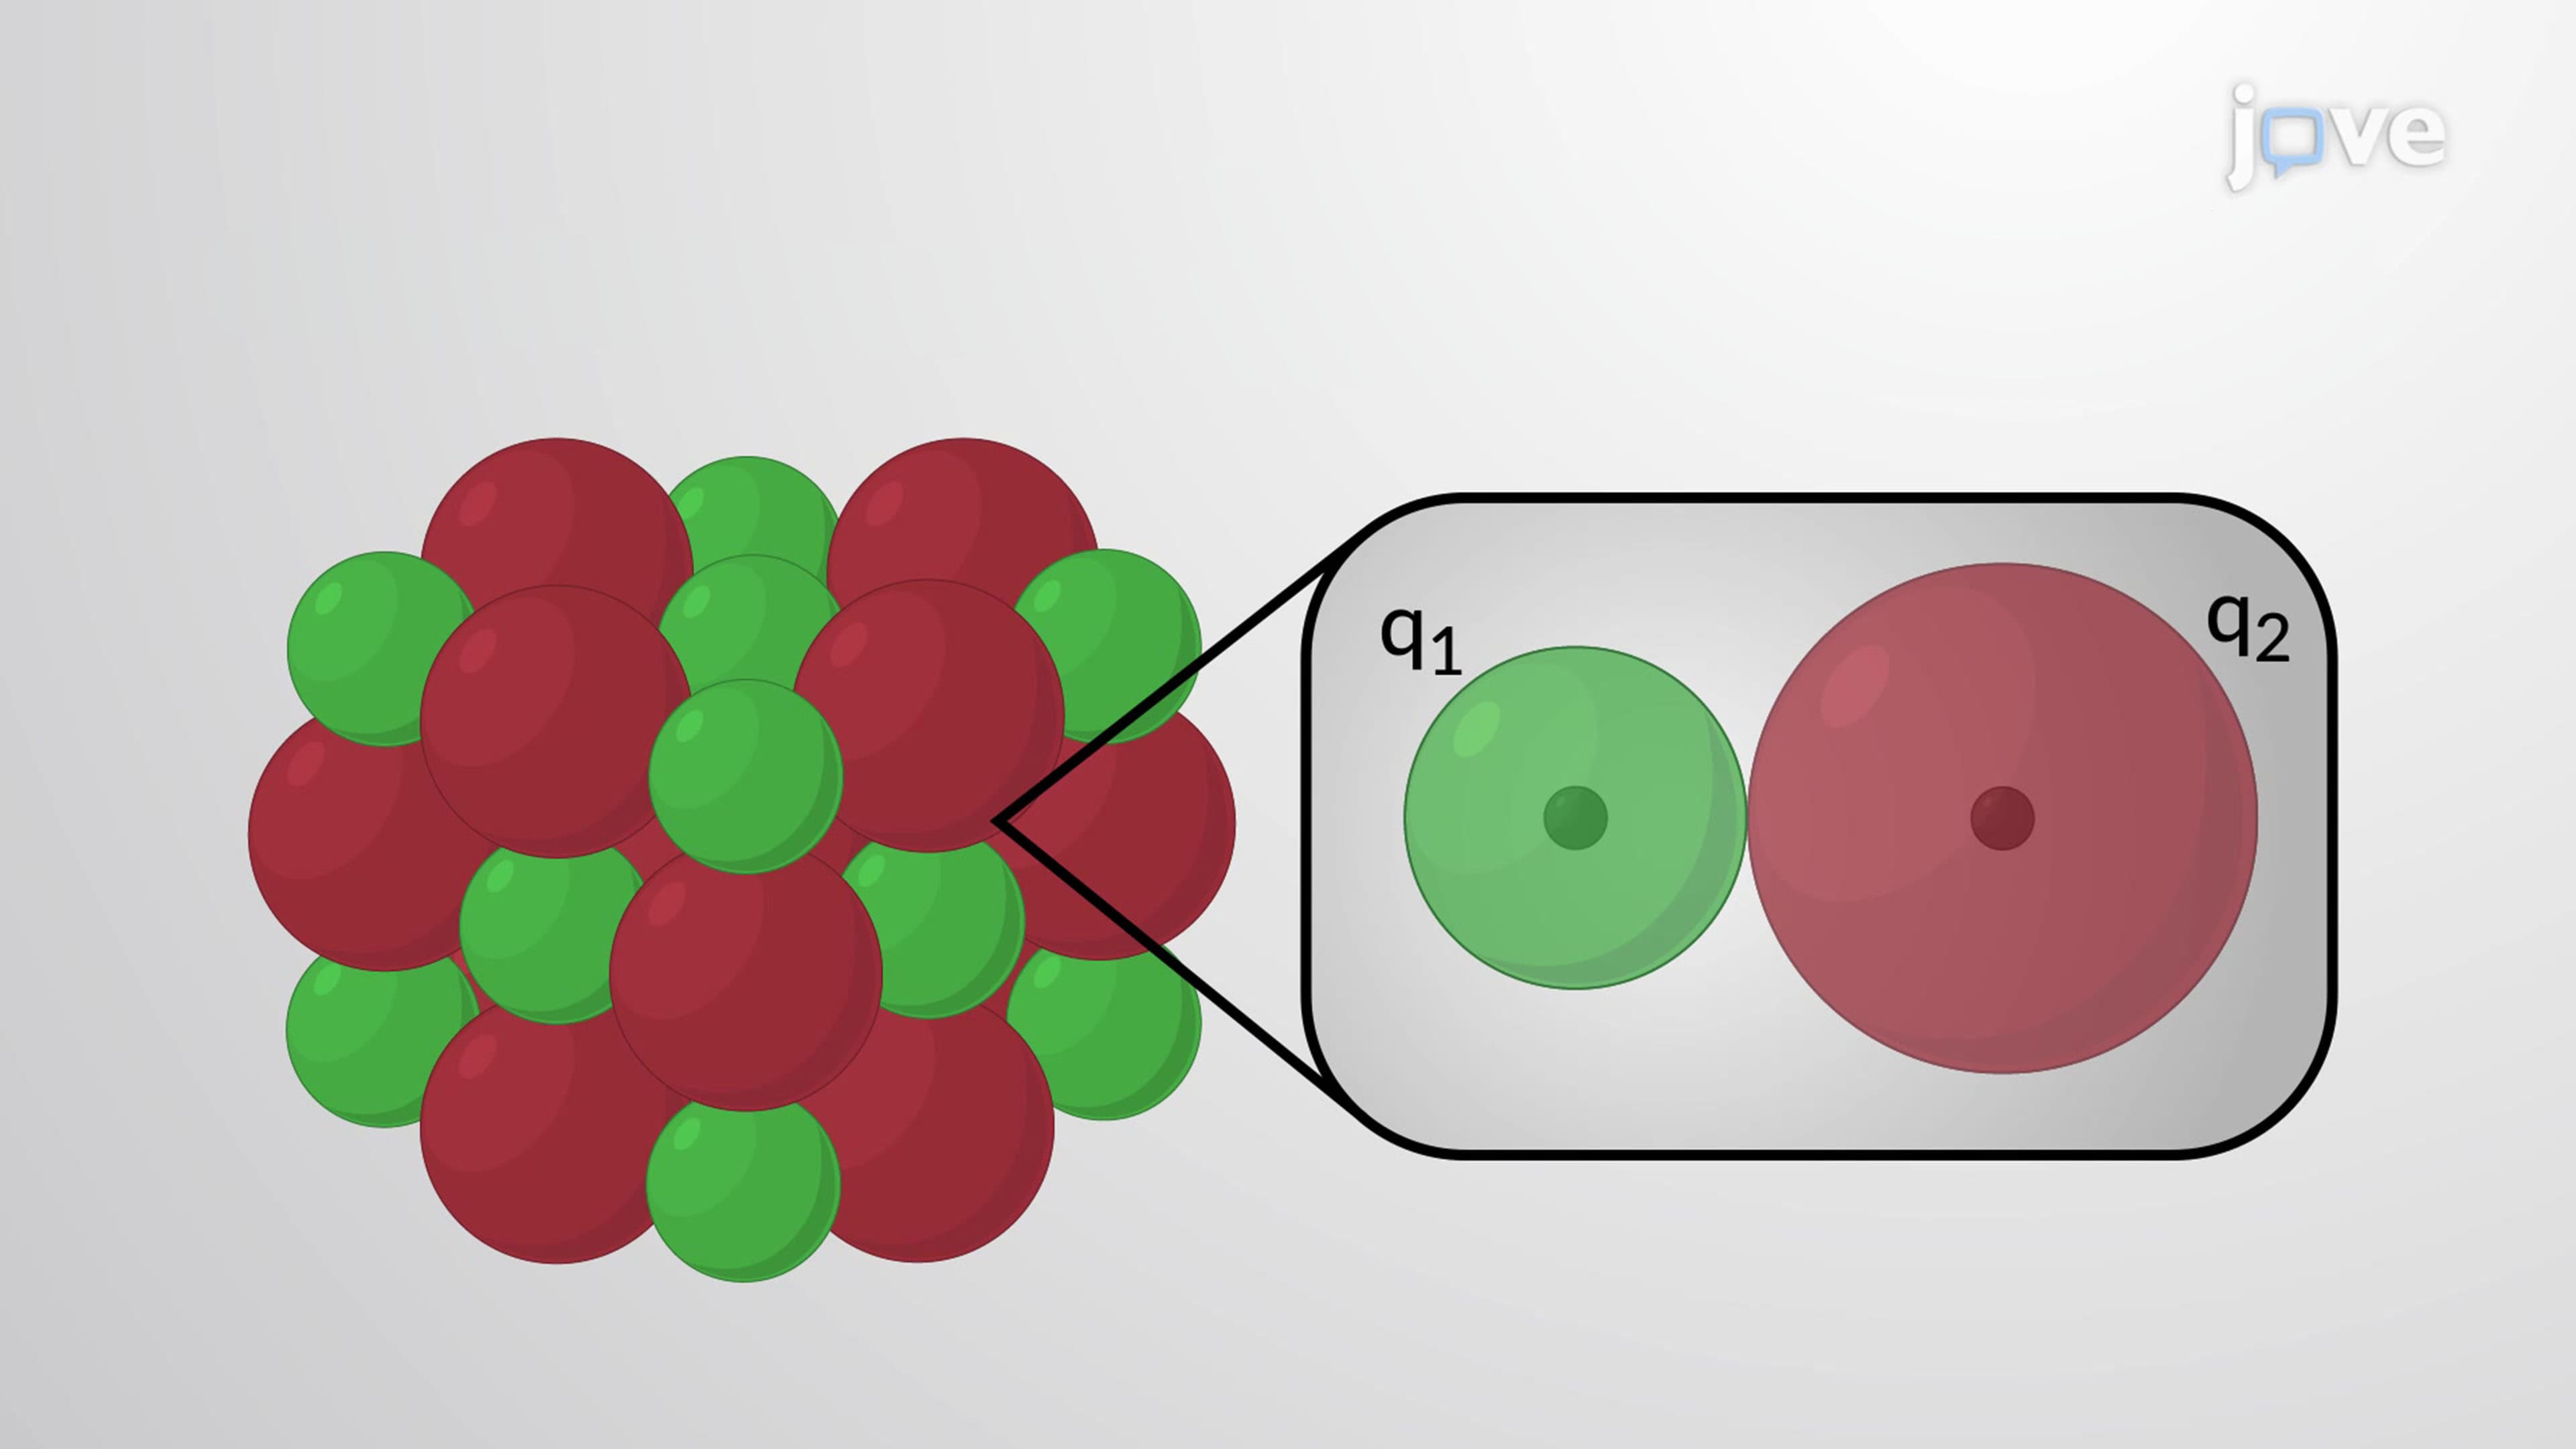
Trends in Lattice Energy: Ion Size and Charge
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:44
The Born-Haber Cycle
 02:18
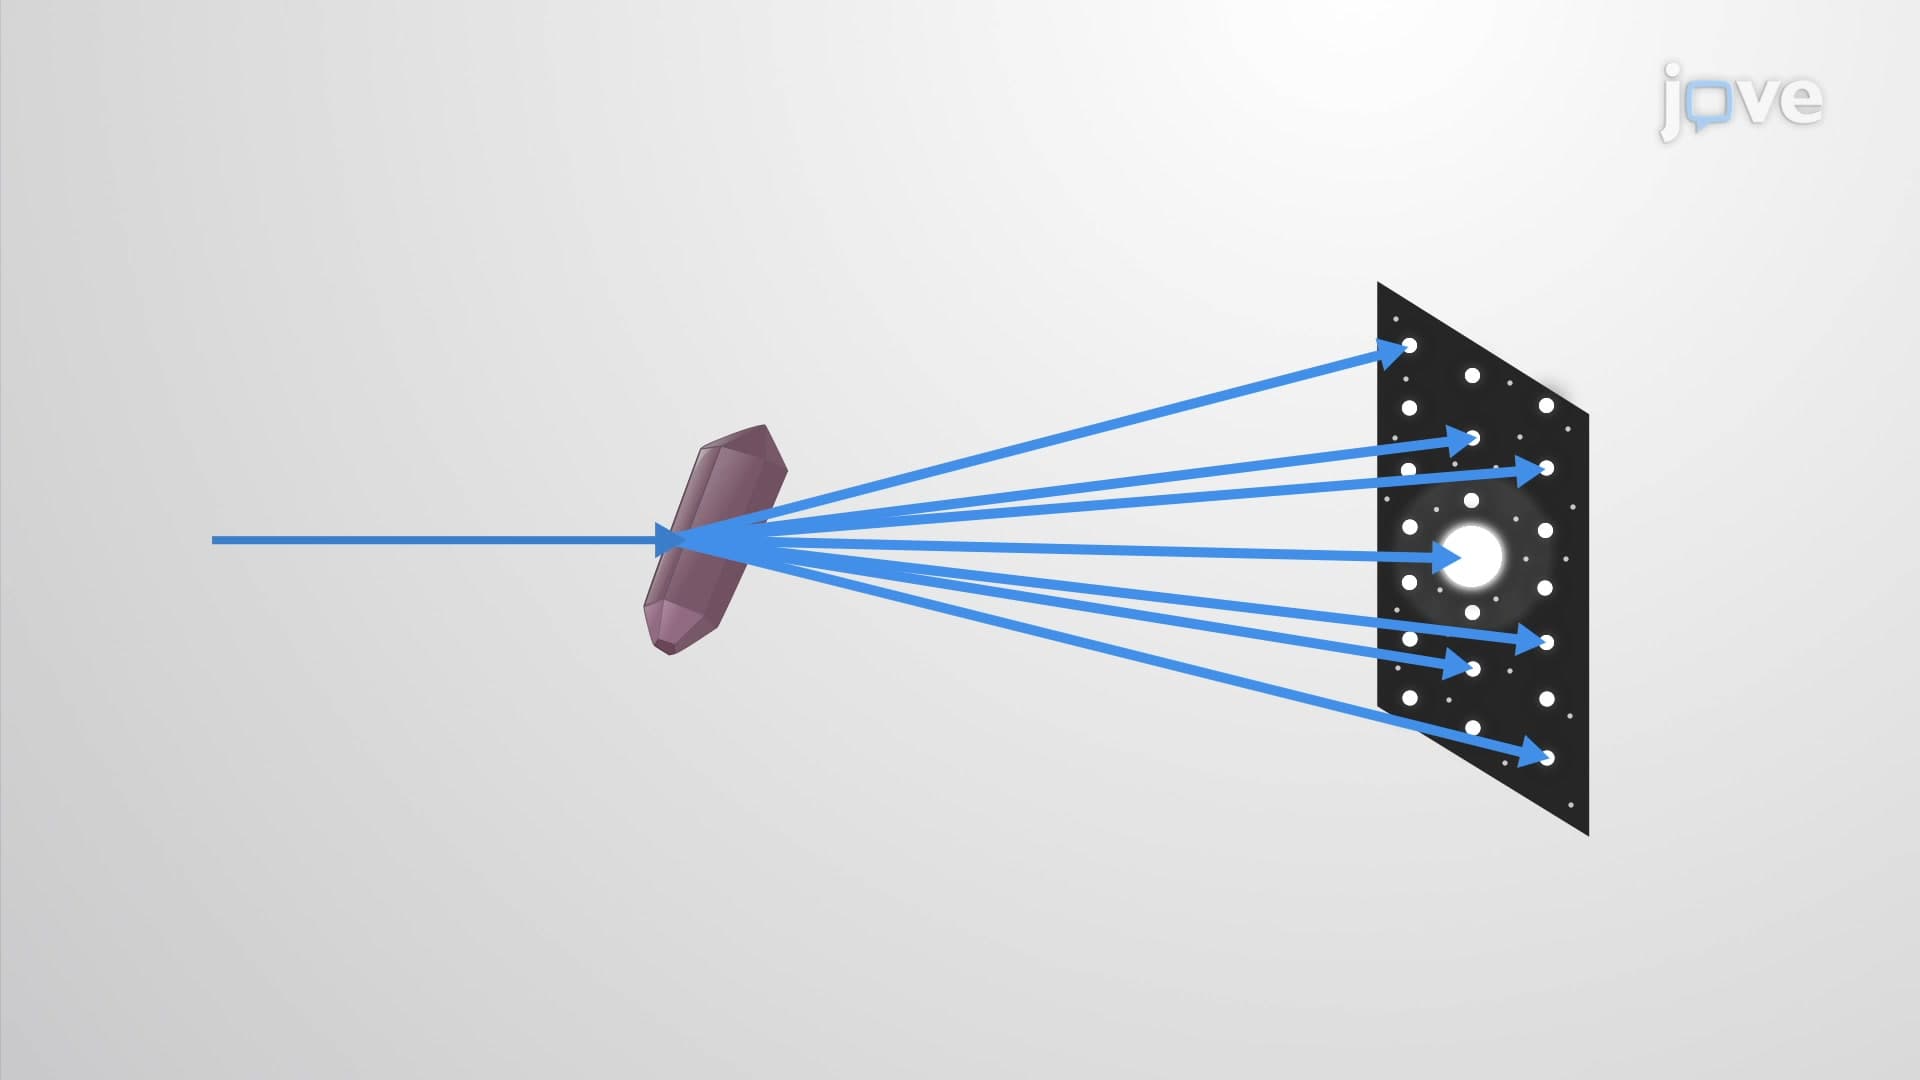
X-ray Crystallography
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 01:29
Determination of Crystal Structures