Related Experiment Video
Updated: Mar 25, 2026


In Vivo Functional Study of Disease-associated Rare Human Variants Using Drosophila
Published on: August 20, 2019
A homozygous nonsense variant in IFT52 is associated with a human skeletal ciliopathy
K M Girisha1, A Shukla1, D Trujillano2
1Department of Medical Genetics, Kasturba Medical College, Manipal University, Manipal, India.

Insights
Intraflagellar transport (IFT) defects cause ciliopathies. A novel IFT52 gene variant was identified in a child with Sensenbrenner syndrome, linking IFT52 to human disease for the first time.
Area of Science:
- Genetics
- Molecular Biology
- Developmental Biology
Background:
- Primary cilia are crucial for cellular function, and defects in intraflagellar transport (IFT) complexes lead to ciliopathies.
- Ciliopathies manifest with diverse symptoms affecting skeletal, neurological, ocular, ectodermal, and renal systems.
Observation:
- A child presented with a complex phenotype including short stature, skeletal dysplasia, polydactyly, and pigmentary retinopathy.
- The child's symptoms shared significant overlap with cranioectodermal dysplasia type I (Sensenbrenner syndrome).
Findings:
- Whole-exome sequencing identified a homozygous nonsense variant (p.R142*) in the IFT52 gene.
- This variant in IFT52, encoding an IFT-B core complex protein, is the likely cause of the observed condition.
Implications:
- This study reports the first human disease associated with mutations in IFT52.
- Understanding IFT52's role expands knowledge of ciliopathies and their genetic underpinnings.
Abstract:
Intraflagellar transport (IFT) is vital for the functioning of primary cilia. Defects in several components of IFT complexes cause a spectrum of ciliopathies with variable involvement of skeleton, brain, eyes, ectoderm and kidneys. We examined a child from a consanguineous family who had short stature, narrow thorax, short hands and feet, postaxial polydactyly of hands, pigmentary retinopathy, small teeth and skeletal dysplasia. The clinical phenotype of the child shows significant overlap with cranioectodermal dysplasia type I (Sensenbrenner syndrome). Whole-exome sequencing revealed a homozygous nonsense variant p.R142* in IFT52 encoding an IFT-B core complex protein as the probable cause of her condition. This is the first report of a human disease associated with IFT52.
More Related Videos
 09:37
Navigating MARRVEL, a Web-Based Tool that Integrates Human Genomics and Model Organism Genetics Information
Published on: August 15, 2019
 09:34
Targeted Next-generation Sequencing and Bioinformatics Pipeline to Evaluate Genetic Determinants of Constitutional Disease
Published on: April 4, 2018
Related Concept Videos
 01:31
Translation
Translation Produces the Building Blocks of Life
Proteins are...
01:31Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of...
 02:59
Animal Mitochondrial Genetics
 01:33
Pleiotropy
 02:27
Nonsense-mediated mRNA Decay
Usually, Upf3 binds to an Exon Junction Complex (EJC) at mRNA splice sites. If a ribosome fully translates the mRNA,...
 01:31
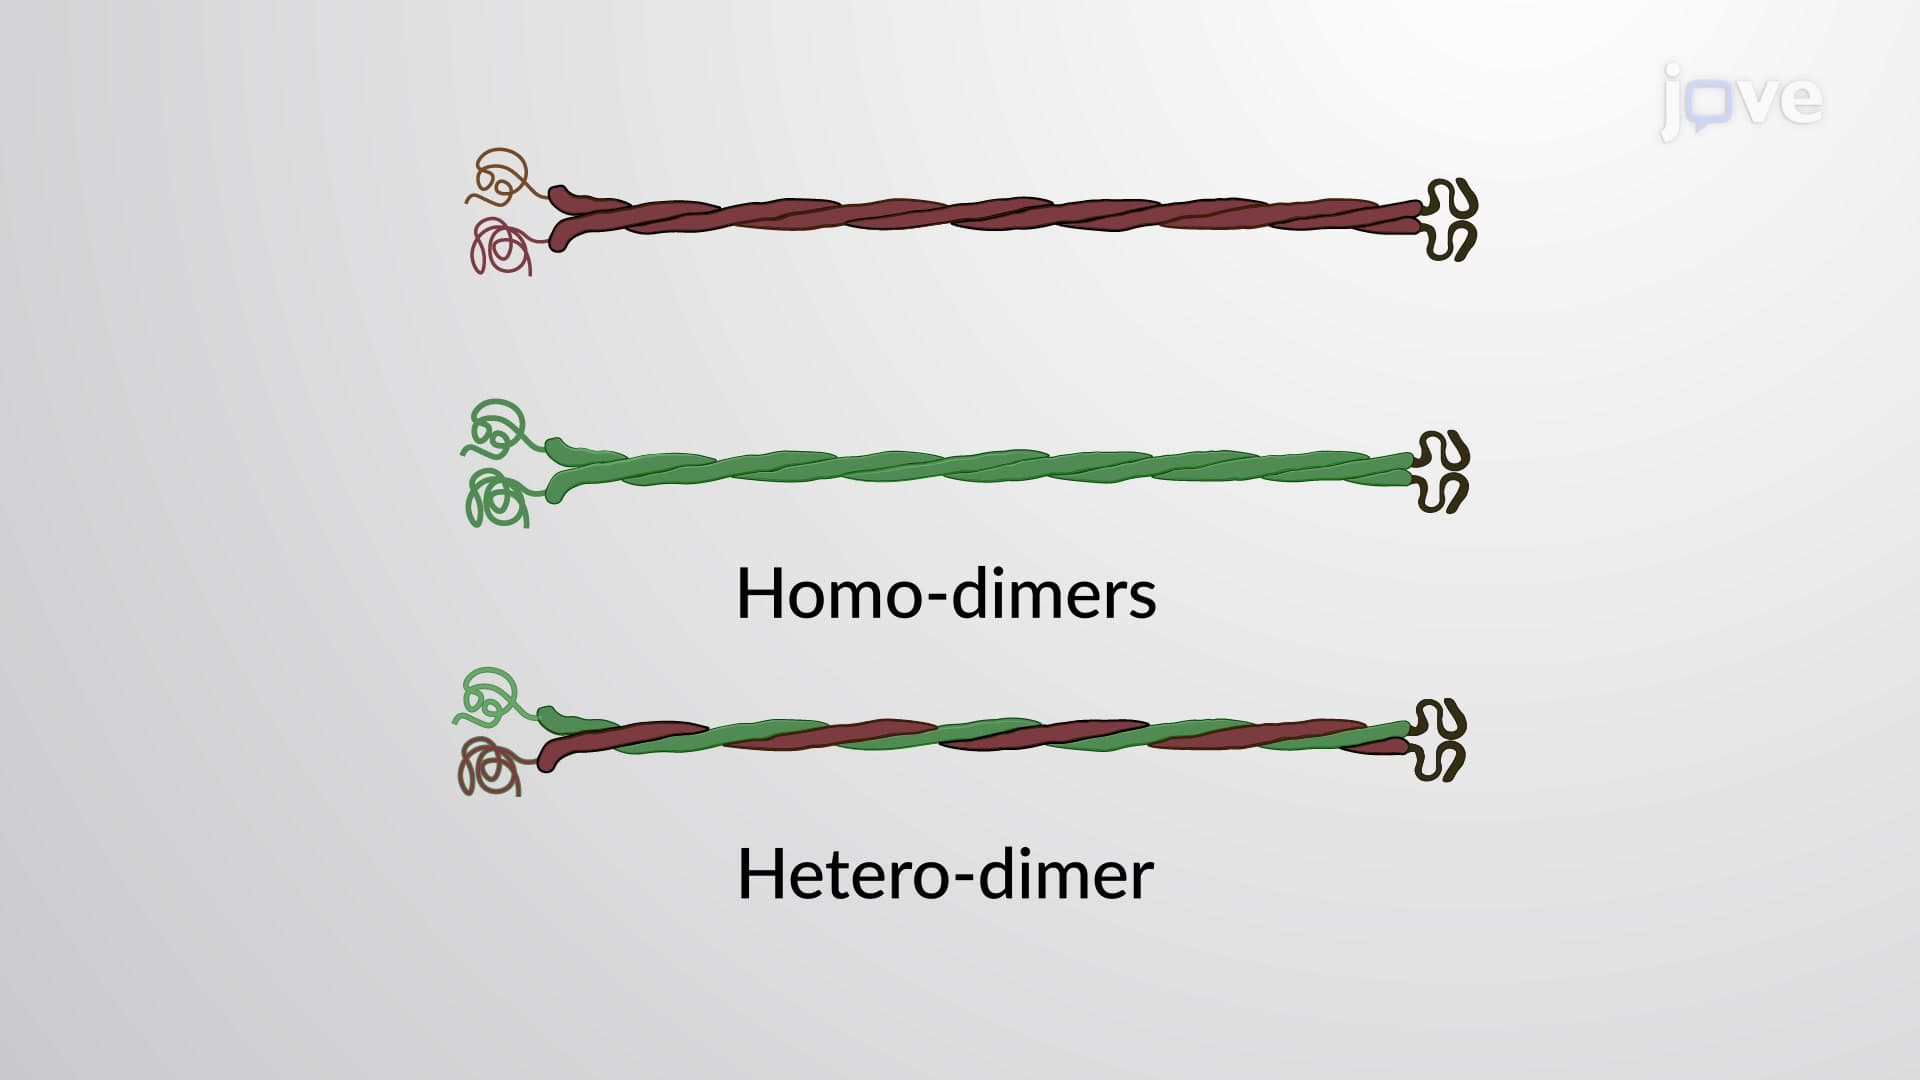
Types of Intermediate Filaments