Related Experiment Video
Updated: Mar 25, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Parameterization of the DFTB3 method for Br, Ca, Cl, F, I, K, and Na in organic and biological systems
Maximilian Kubillus1, Tomáš Kubař, Michael Gaus
1Institute of Physical Chemistry, Karlsruhe Institute of Technology, Kaiserstr. 12, 76021 Karlsruhe, Germany

Abstract:
We present an extension to the recent 3OB parametrization of the Density Functional Tight Binding Model DFTB31,2 for biological and organic systems. Parameters for the halogens F, Cl, Br, and I have been developed for use in covalently bound systems and benchmarked on a test set of 106 molecules (the ‘OrgX’ set), using bonding distances, bonding angles, atomization energies, and vibrational frequencies to assess the performance of the parameters. Additional testing has been done with the X40 set of 40 supramolecular systems containing halogens,3 adding a simple correction for the halogen bonds that are strongly overbound in DFTB3. Furthermore, parameters for Ca, K, and Na as counterions in biological systems have been created. To benchmark geometries as well as ligand binding energies a test set ‘BioMe’ of 210 molecules has been created that cover coordination to various functional groups frequently occurring in biological systems. The new DFTB3/3OB parameter set outperforms DFT calculations with a double-ζ basis set in terms of energies and can reproduce DFT geometries, with some minor deviations in bond distances and angles due to the use of a minimal basis set.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 05:57
Author Spotlight: In Silico Creation and Impact of Carbonylated Amino Acids on Protein Structure and Function
Published on: April 26, 2024
Related Concept Videos
 01:29
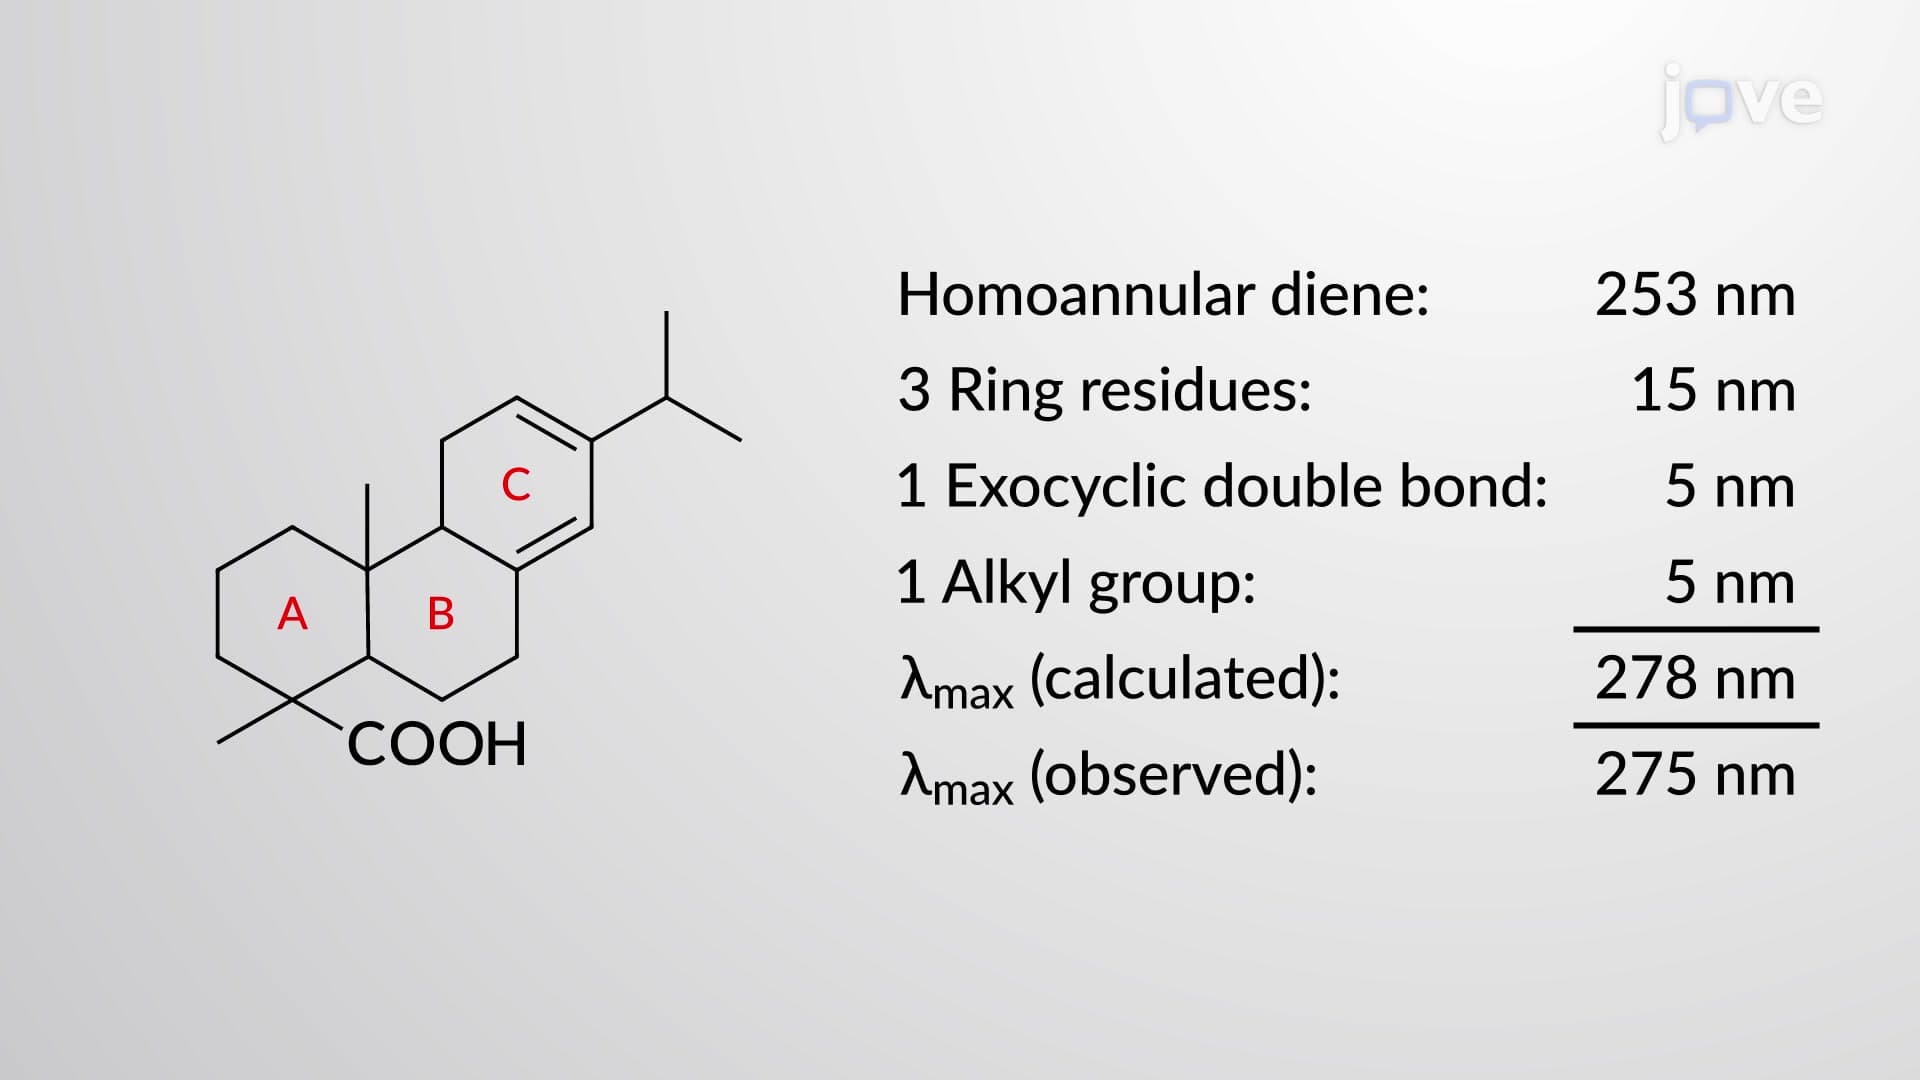
UV–Vis Spectroscopy: Woodward–Fieser Rules
 03:35
Hybridization of Atomic Orbitals II
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:20
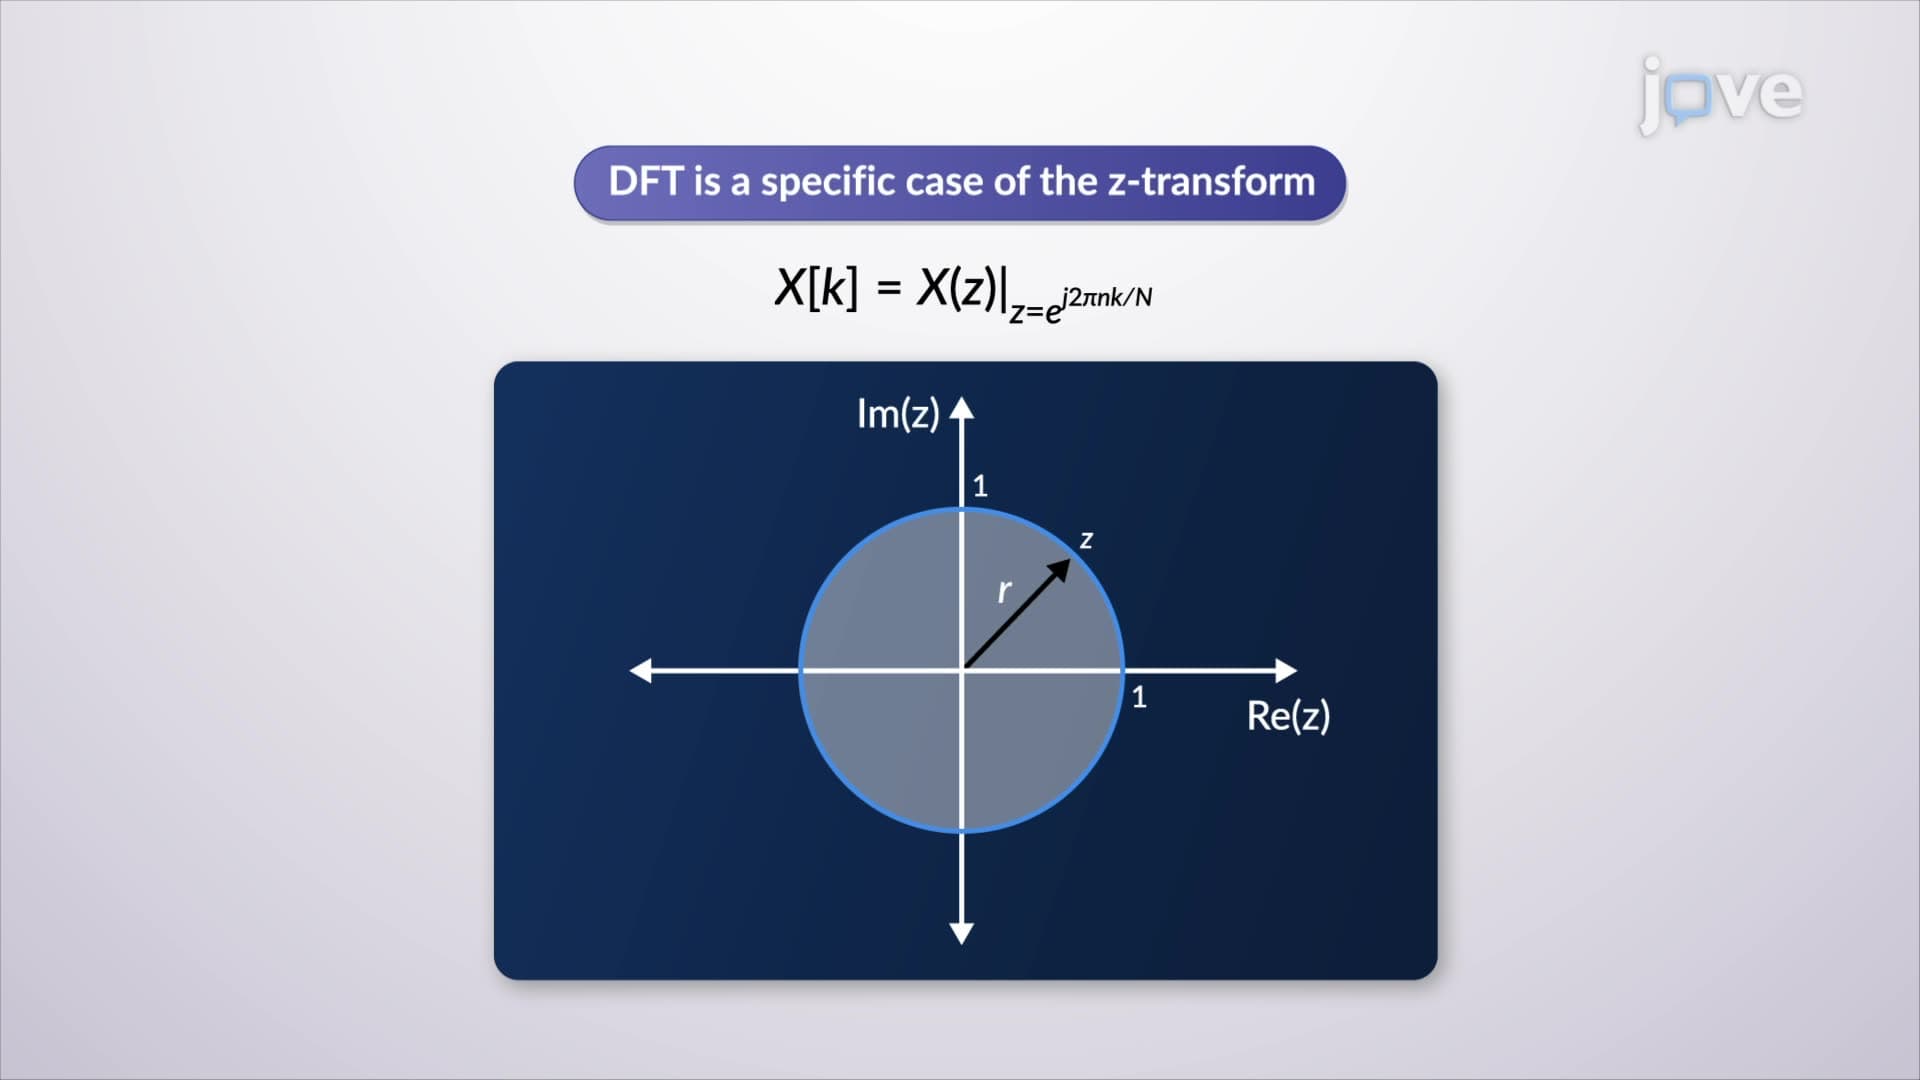
Relation of DFT to z-Transform
To understand how the DFT works, it's helpful to consider the z-transform, which is a method for representing discrete sequences in the complex frequency domain. The z-transform involves summing the...
 01:24
Properties of DTFT I
The linearity property of DTFTs is fundamental. If two discrete-time signals are multiplied by constants a and b respectively, and then combined to...
 02:42
Valence Bond Theory