Related Experiment Video
Updated: Mar 25, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Validation of local hybrid functionals for TDDFT calculations of electronic excitation energies
Toni M Maier1, Hilke Bahmann1, Alexei V Arbuznikov1
1Technische Universität Berlin, Institut für Chemie, Theoretische Chemie/Quantenchemie, Sekr. C7, Straße des 17. Juni 135, D-10623 Berlin, Germany.

Abstract:
The first systematic evaluation of local hybrid functionals for the calculation of electronic excitation energies within linear-response time-dependent density functional theory (TDDFT) is reported. Using our recent efficient semi-numerical TDDFT implementation [T. M. Maier et al., J. Chem. Theory Comput. 11, 4226 (2015)], four simple, thermochemically optimized one-parameter local hybrid functionals based on local spin-density exchange are evaluated against a database of singlet and triplet valence excitations of organic molecules, and against a mixed database including also Rydberg, intramolecular charge-transfer (CT) and core excitations. The four local hybrids exhibit comparable performance to standard global or range-separated hybrid functionals for common singlet valence excitations, but several local hybrids outperform all other functionals tested for the triplet excitations of the first test set, as well as for relative energies of excited states. Evaluation for the combined second test set shows that local hybrids can also provide excellent Rydberg and core excitations, in the latter case rivaling specialized functionals optimized specifically for such excitations. This good performance of local hybrids for different excitation types could be traced to relatively large exact-exchange (EXX) admixtures in a spatial region intermediate between valence and asymptotics, as well as close to the nucleus, and lower EXX admixtures in the valence region. In contrast, the tested local hybrids cannot compete with the best range-separated hybrids for intra- and intermolecular CT excitation energies. Possible directions for improvement in the latter category are discussed. As the used efficient TDDFT implementation requires essentially the same computational effort for global and local hybrids, applications of local hybrid functionals to excited-state problems appear promising in a wide range of fields. Influences of current-density dependence of local kinetic-energy dependent local hybrids, differences between spin-resolved and "common" local mixing functions in local hybrids, and the effects of the Tamm-Dancoff approximation on the excitation energies are also discussed.
More Related Videos


Related Concept Videos
 02:38
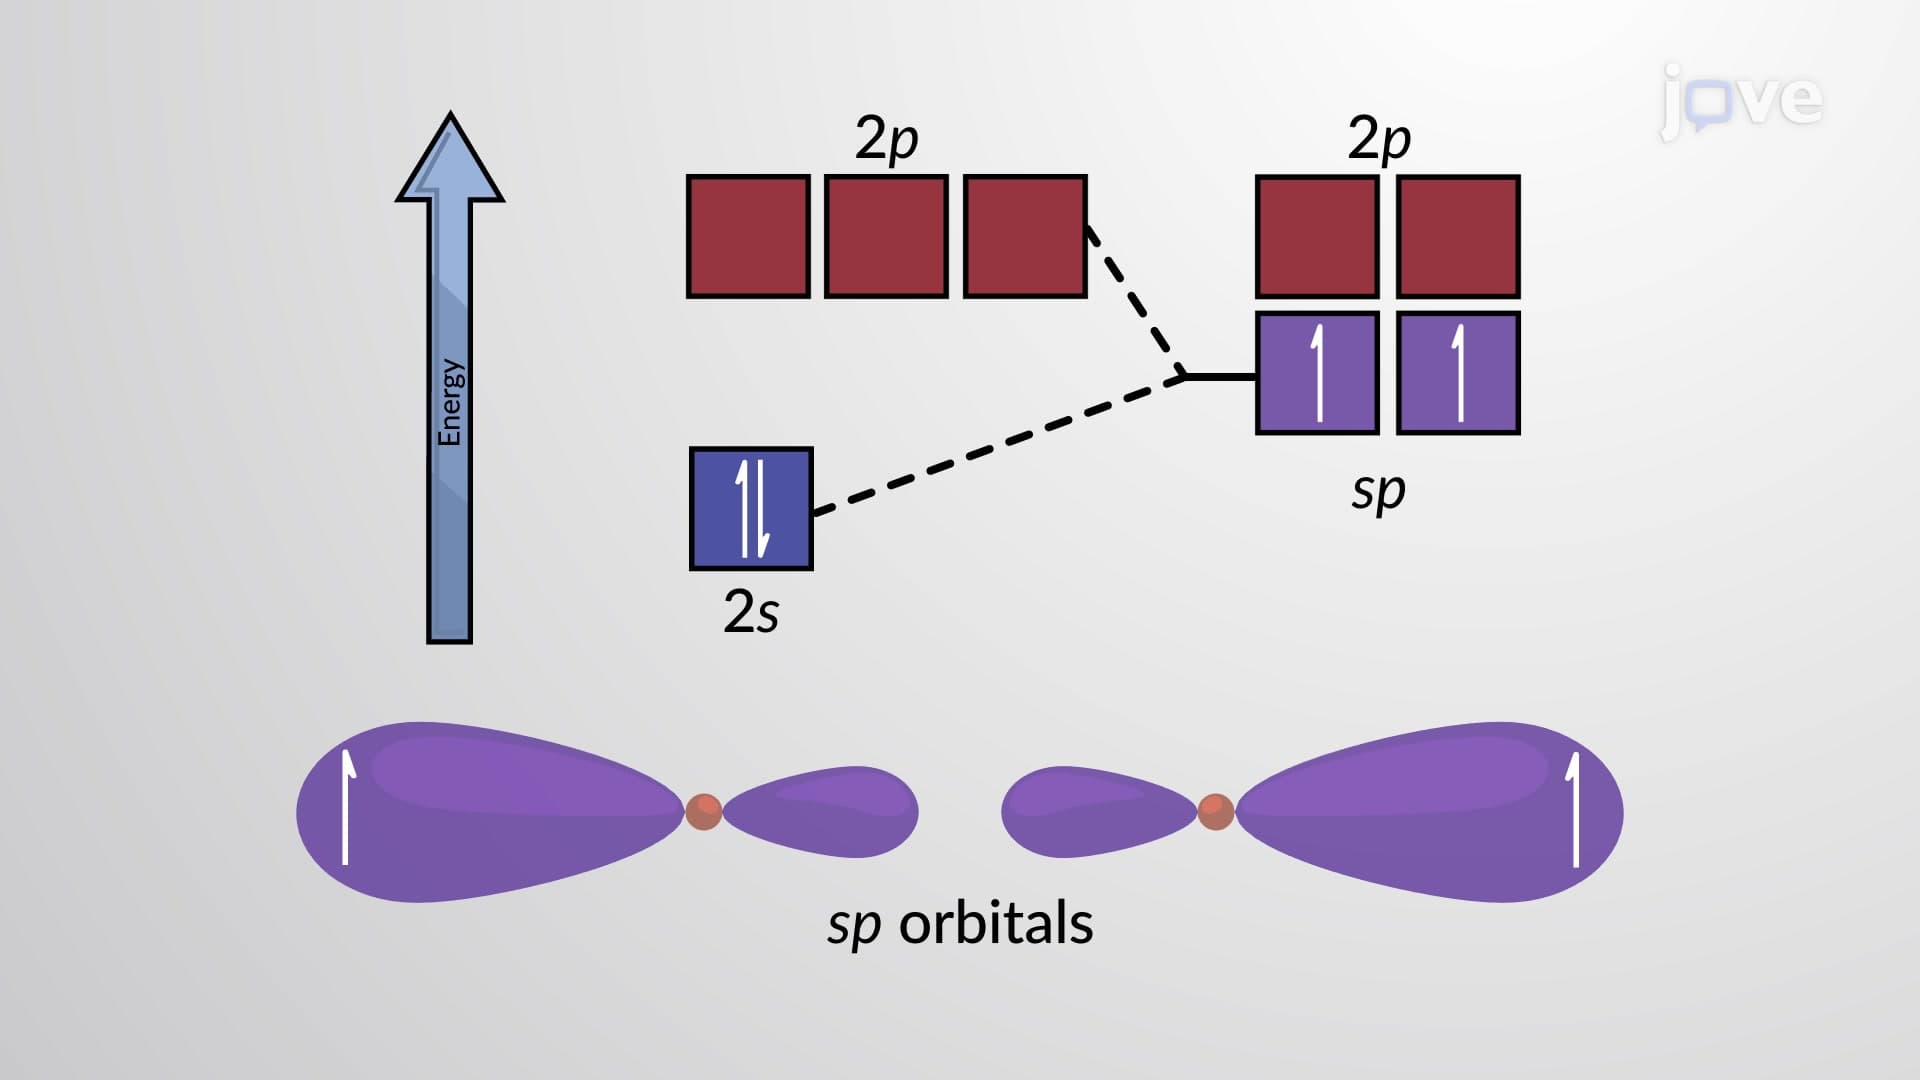
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:16
Resonance and Hybrid Structures
Resonance Structures and Resonance Hybrids
The Lewis structure of a nitrite anion (NO2−) may actually be drawn in two different ways, distinguished by the locations of the N–O and N=O bonds.