Related Experiment Video
Updated: Mar 25, 2026
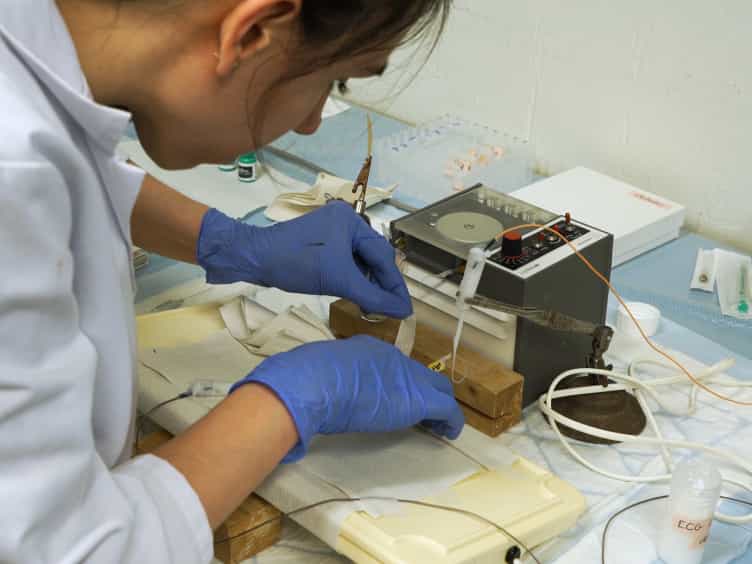
Nasal Potential Difference to Quantify Trans-epithelial Ion Transport in Mice
Published on: July 4, 2018
Loss of carbonic anhydrase XII function in individuals with elevated sweat chloride concentration and pulmonary
Melissa Lee1, Briana Vecchio-Pagán1, Neeraj Sharma1
1McKusick-Nathans Institute of Genetic Medicine, Johns Hopkins University School of Medicine, Baltimore, MD, USA.

Abstract:
Elevated sweat chloride levels, failure to thrive (FTT), and lung disease are characteristic features of cystic fibrosis (CF, OMIM #219700). Here we describe variants in CA12 encoding carbonic anhydrase XII in two pedigrees exhibiting CF-like phenotypes. Exome sequencing of a white American adult diagnosed with CF due to elevated sweat chloride, recurrent hyponatremia, infantile FTT and lung disease identified deleterious variants in each CA12 gene: c.908-1 G>A in a splice acceptor and a novel frameshift insertion c.859_860insACCT. In an unrelated consanguineous Omani family, two children with elevated sweat chloride, infantile FTT, and recurrent hyponatremia were homozygous for a novel missense variant (p.His121Gln). Deleterious CFTR variants were absent in both pedigrees. CA XII protein was localized apically in human bronchiolar epithelia and basolaterally in the reabsorptive duct of human sweat glands. Respiratory epithelial cell RNA from the adult proband revealed only aberrant CA12 transcripts and in vitro analysis showed greatly reduced CA XII protein. Studies of ion transport across respiratory epithelial cells in vivo and in culture revealed intact CFTR-mediated chloride transport in the adult proband. CA XII protein bearing either p.His121Gln or a previously identified p.Glu143Lys missense variant localized to the basolateral membranes of polarized Madin-Darby canine kidney (MDCK) cells, but enzyme activity was severely diminished when assayed at physiologic concentrations of extracellular chloride. Our findings indicate that loss of CA XII function should be considered in individuals without CFTR mutations who exhibit CF-like features in the sweat gland and lung.
More Related Videos
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Renal Tubule and Collecting Duct
Proximal Convoluted Tubule (PCT):
The PCT is the initial segment of the renal tubule, extending from the Bowman's capsule that encloses the glomerulus. Its convoluted structure and microvilli-lined cells increase the surface area for reabsorption. The PCT reabsorbs glucose, amino acids, sodium, and water from the filtrate, ensuring essential...
 01:30
Acute Respiratory Failure-III
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:29
Roles of Electrolytes: Chloride and Bicarbonate
Conditions such as hypochloremia can arise from insufficient chloride reabsorption by the kidneys, often compounded by extended bouts of diarrhea, vomiting,...
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation