Related Experiment Video
Updated: Mar 24, 2026
Metabolic Profiling to Determine Bactericidal or Bacteriostatic Effects of New Natural Products using Isothermal Microcalorimetry
Published on: October 29, 2020
Application of a Physiologically Based Pharmacokinetic Model to Study Theophylline Metabolism and Its Interactions
1Biosciences & Biotechnology Division, Physics & Life Sciences Directorate, Lawrence Livermore National Laboratory Livermore California USA.

Abstract:
Theophylline is a commonly used bronchodilator. However, due to its narrow therapeutic range, moderate elevation of serum concentration can result in adverse drug reactions (ADRs). ADRs occur because of interhuman pharmacokinetic variability and interactions with coprescribed medicines. We developed a physiologically based pharmacokinetic (PBPK) model of theophylline, caffeine, and ciprofloxacin metabolisms to: examine theophylline pharmacokinetic variability, and predict population-level outcomes of drug-drug interactions (DDIs). A simulation-based equation for personalized dosing of theophylline was derived. Simulations of DDI show that calculated personalized doses are safe even after cotreatment with large doses of strong inhibitors. Simulations of adult populations indicate that the elderly are most susceptible to ADRs stemming from theophylline-ciprofloxacin and theophylline-caffeine interactions. Females, especially Asians, due to their smaller average size, are more susceptible to DDI-induced ADRs following typical dosing practices. Our simulations also show that the higher adipose and lower muscle fractions in females significantly alter the pharmacokinetics of theophylline or ciprofloxacin.
More Related Videos
Related Concept Videos
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model
 01:26
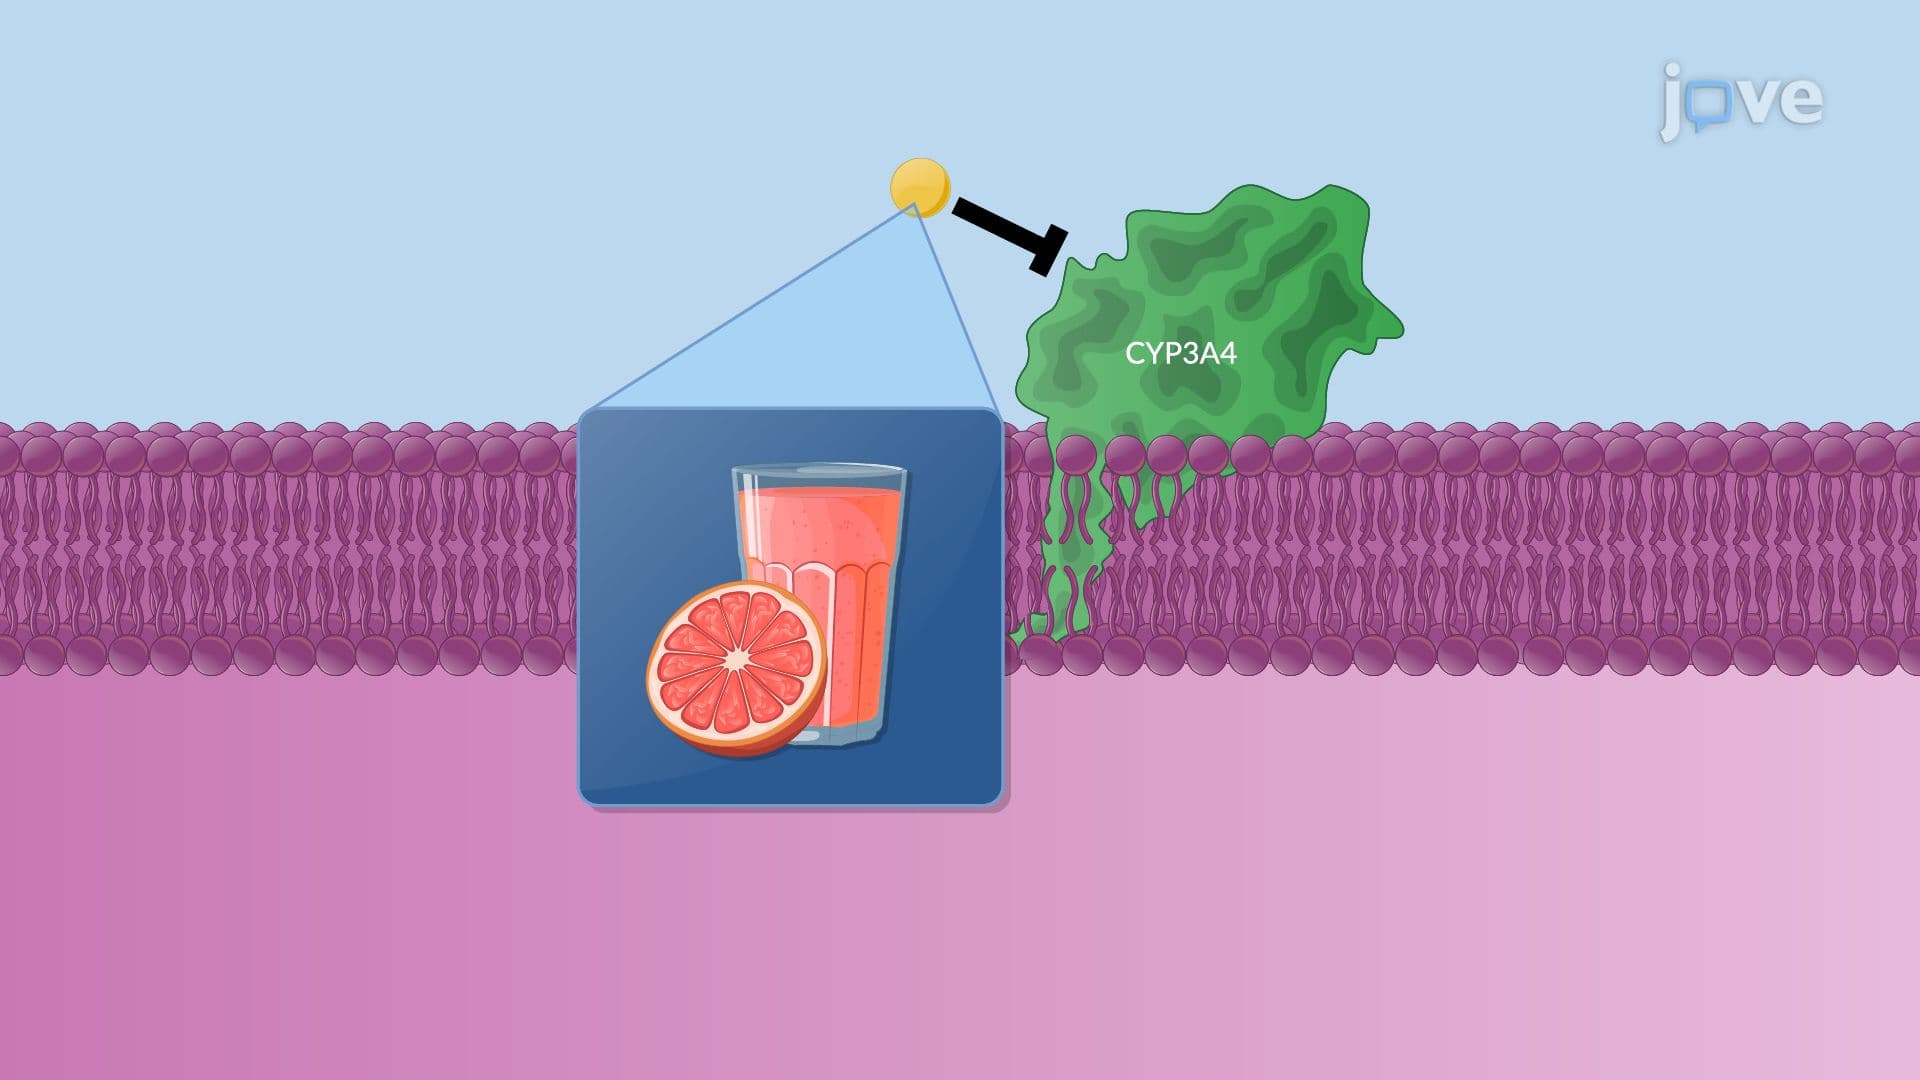
Pharmacokinetics: Drug–Food and Drug–Viral Interactions
 01:15
Model Approaches for Pharmacokinetic Data: Physiological Models
 01:25
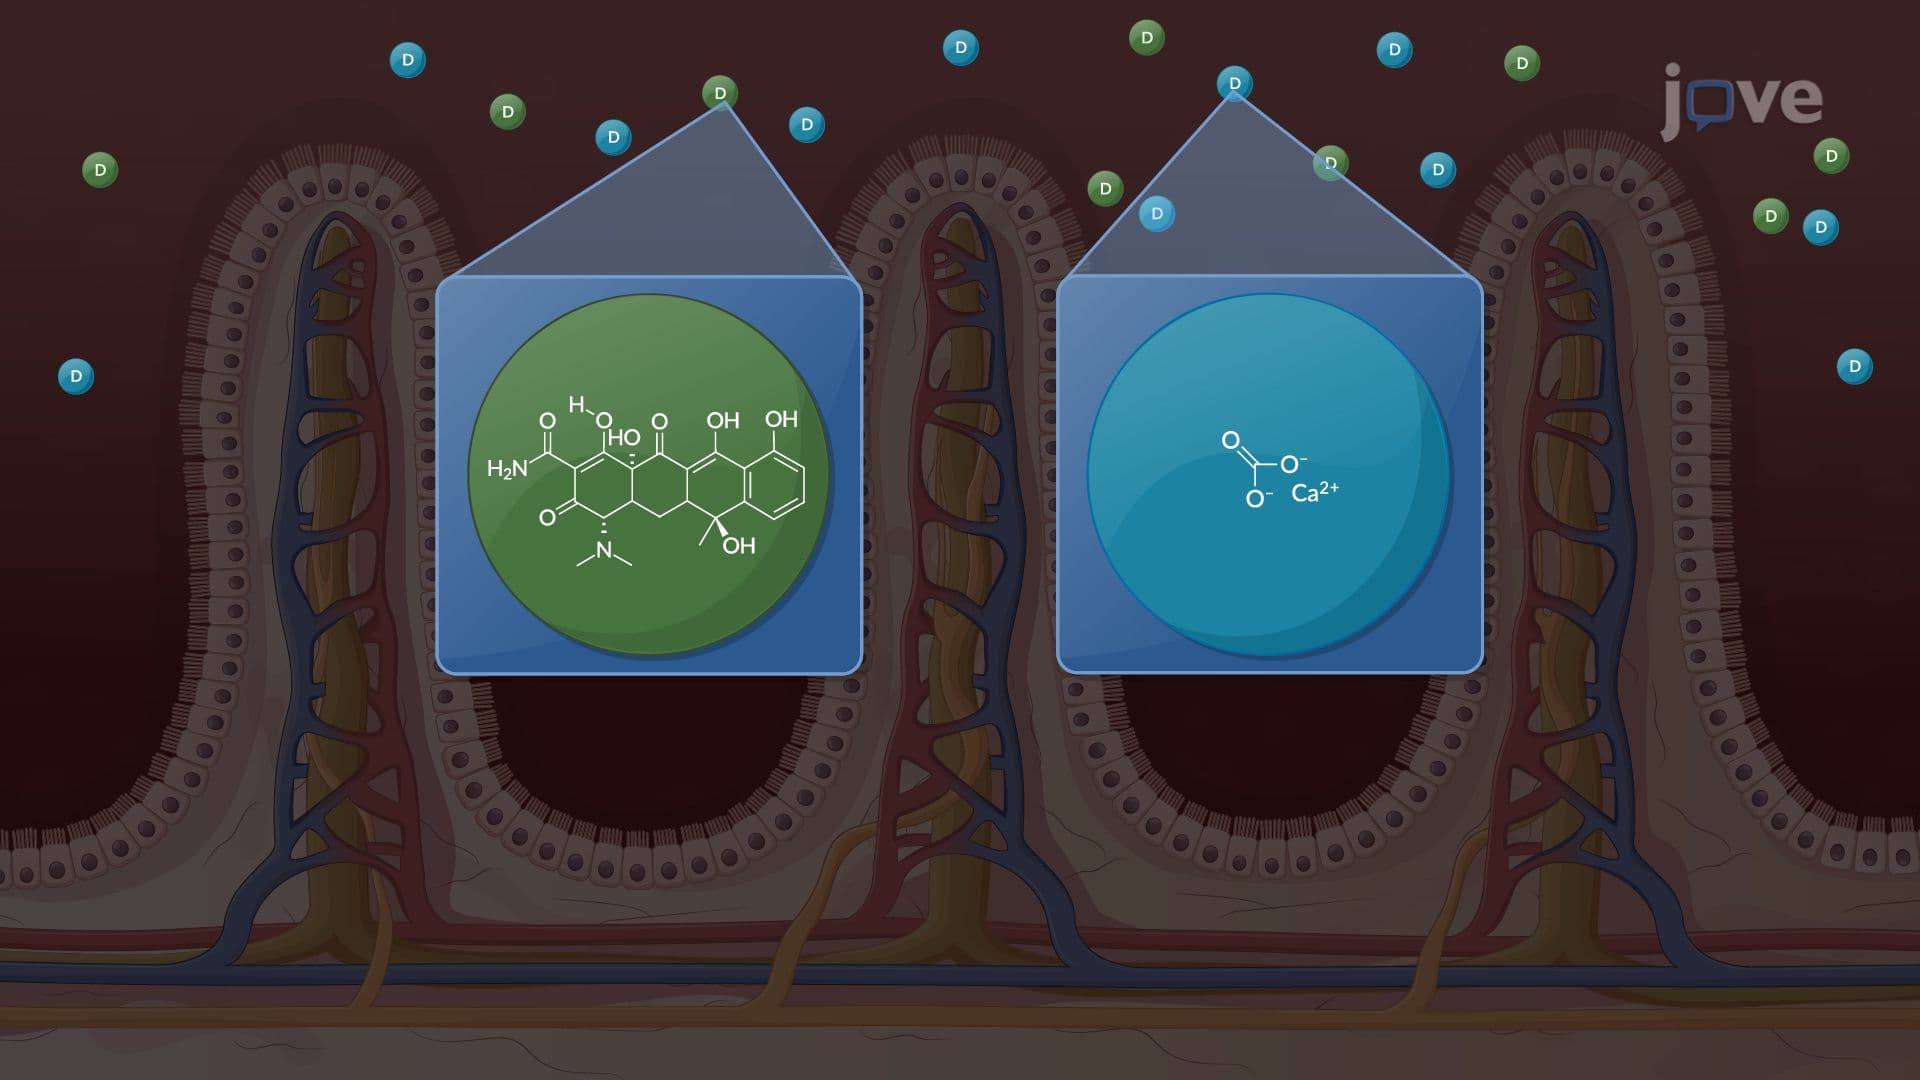
Pharmacokinetics: Drug–Drug Interactions