Single concentration loss of activity assay provides an improved assessment of drug-drug interaction risk compared to IC50-shift
Related Experiment Video
Updated: Mar 24, 2026
Diagonal Method to Measure Synergy Among Any Number of Drugs
Published on: June 21, 2018
Simon G Wong1, Mey Lee1, Bradley K Wong1
1a Department of Pharmacokinetics and Drug Metabolism , Amgen , South San Francisco , CA , USA.

Abstract:
1. The utility of two abbreviated, higher-throughput assays [IC50-shift and the loss of activity (LOA) assay] to evaluate time-dependent inhibition (TDI) of 24 structurally related compounds was compared. 2. Good correlation (R(2) = 0.90) between % inhibition and kinact/KI suggested that the LOA assay has utility as an indicator of TDI potential. Weaker correlation was observed for the shifted IC50 (IC50(T = 30)) (R(2) = 0.61) and the fold-shift in IC50 (R(2) = 0.17). 3. Primary mechanism for poor correlation was depletion of active enzyme at concentrations > 1 μM leading to greater than predicted inhibition in the IC50-shift assay. 4. Previously reported strong correlations between IC50(T = 30) and kinact/KI were found to be dependent on potent TDI compounds with kinact/KI > 30; correlation was reduced for moderate inhibitors (kinact/KI < 30). LOA assay maintained good correlation even when strong TDI compounds were excluded. 5. LOA assay (% Inhibition at 30 min, 10 μM) was a good predictor of in vivo DDI (AUCr), providing a graded response with low potential for false negatives or positives. IC50-shift assay had bias for over-predicting in vivo DDI and was more likely to identify false positives.
More Related Videos
 10:28
A Semi-High-Throughput Adaptation of the NADH-Coupled ATPase Assay for Screening Small Molecule Inhibitors
Published on: August 17, 2019
 09:09
High Throughput, Real-time, Dual-readout Testing of Intracellular Antimicrobial Activity and Eukaryotic Cell Cytotoxicity
Published on: November 16, 2016
Related Concept Videos
 01:14
Time Course of Drug Effect
 01:30
Drug toxicity: Drug–Drug Interaction
 01:14
Effect of Hepatic Disease on Pharmacokinetics: Active Drug, Metabolite and Fraction of Metabolized Drug
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:29
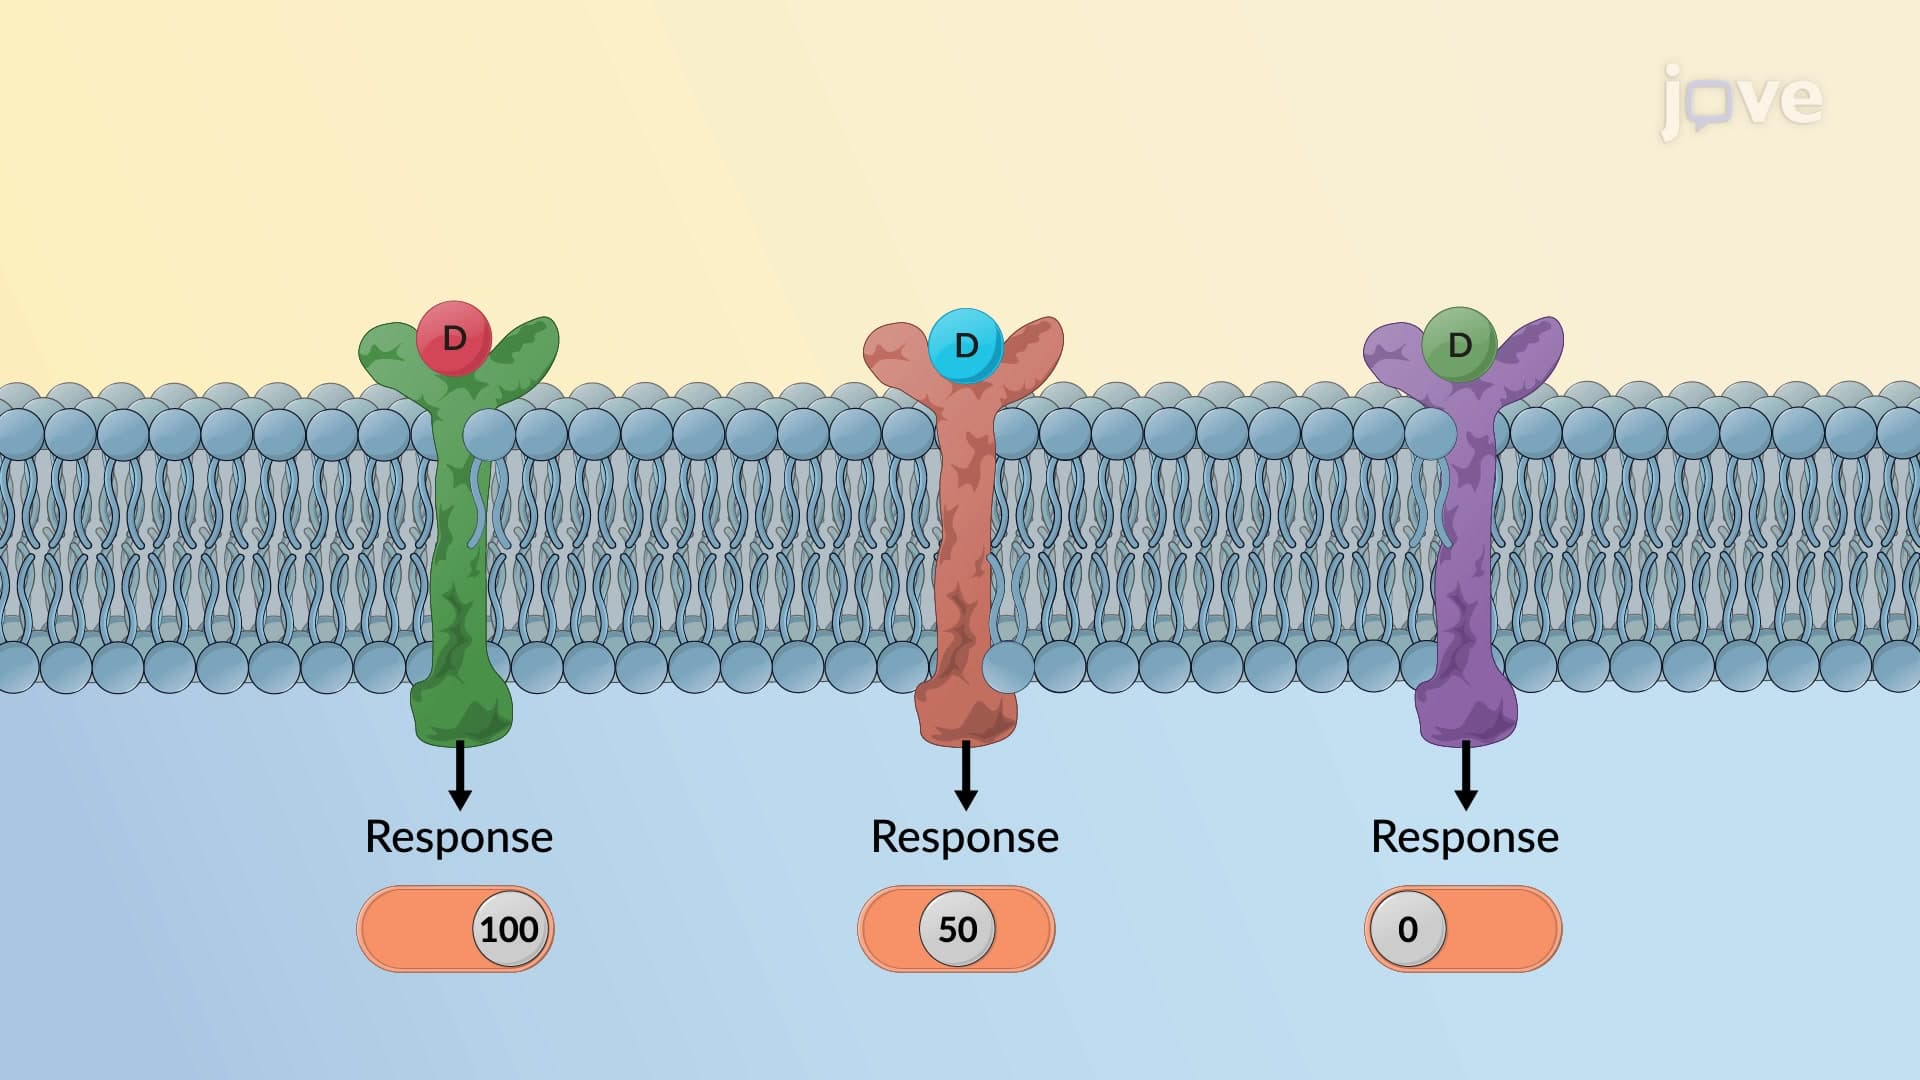
Drug-Receptor Interactions
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue....