Related Experiment Video
Updated: Mar 24, 2026
The WinCF Model - An Inexpensive and Tractable Microcosm of a Mucus Plugged Bronchiole to Study the Microbiology of Lung Infections
Published on: May 8, 2017
Neonatal Gastrointestinal and Respiratory Microbiome in Cystic Fibrosis: Potential Interactions and Implications for
1Division of Neonatology, Department of Pediatrics, Children's Hospital at Dartmouth, Geisel School of Medicine at Dartmouth, Hanover, New Hampshire.

Insights
The gut and respiratory microbiome are crucial in cystic fibrosis (CF). Interventions like probiotics may improve health outcomes and reduce illness in children with CF by modulating the microbiome.
Area of Science:
- Microbiology
- Pediatric Health
- Gastroenterology
Background:
- The gastrointestinal microbiome is vital for infant nutrition, metabolism, and immunity, impacting long-term health.
- Cystic fibrosis (CF) transmembrane conductance regulator (CFTR) mutations lead to altered gut and respiratory microbiomes (dysbiosis), causing inflammation, infection, and impaired growth in children.
- Dysbiosis in CF contributes to undernutrition, growth failure, and chronic respiratory and systemic diseases.
Purpose of the Study:
- To explore the role of the gut and respiratory microbiome in health and disease progression in cystic fibrosis (CF).
- To identify potential interventions targeting the microbiome to improve clinical outcomes in CF patients.
Main Methods:
- A literature review was conducted on studies concerning CF, the microbiome, and probiotics.
- Pertinent literature was summarized to understand current knowledge and research gaps.
Main Results:
- Studies reveal bacteria typically found in the gut are present in the CF respiratory tract.
- Enteral exposures, including breast milk and probiotics, correlate with extended periods of respiratory stability in CF patients.
Conclusions:
- Understanding the interplay between CFTR mutations, microbial colonization, and immunity is key for developing novel therapeutic strategies.
- Interventions aimed at restoring a healthy microbiome, such as probiotics or dietary changes, hold promise for improving nutritional status, immune function, and reducing morbidity and mortality in CF.
Purpose:
The gastrointestinal microbiome plays a critical role in nutrition and metabolic and immune functions in infants and young children and has implications for lifelong health. Cystic fibrosis (CF) transmembrane conductance regulator (CFTR) mutations in CF result in viscous mucous production, frequent exposure to antibiotics, and atypical colonization patterns, resulting in an evolving dysbiosis of the gastrointestinal and respiratory microsystems; dysbiosis in CF results in systemic inflammation, chronic infection, and dysregulation of immune function. Dysbiosis in both the respiratory system and gut contributes to undernutrition, growth failure, and long-term respiratory and systemic morbidity in infants and children with CF. Understanding the role that the gut and respiratory microbiome plays in health or disease progression in CF will afford opportunities to better identify interventions to affect clinical changes.
Methods:
Summary was done of the pertinent literature in CF and the study of the microbiome and probiotics.
Findings:
New studies have identified bacteria in the respiratory tract in CF that are typically members of the intestinal microbiota, and enteral exposures to breast milk and probiotics are associated with prolonged periods of respiratory stability in CF.
Implications:
Understanding the complex interactions between the CFTR mutations, microbial colonization, and mucosal and systemic immunity is of major importance to inform new treatment strategies (such as restoring a healthier microbiome with probiotics or dietary interventions) to improve nutritional status and immune competence and to decrease morbidity and mortality in CF.
Related Concept Videos
 01:29
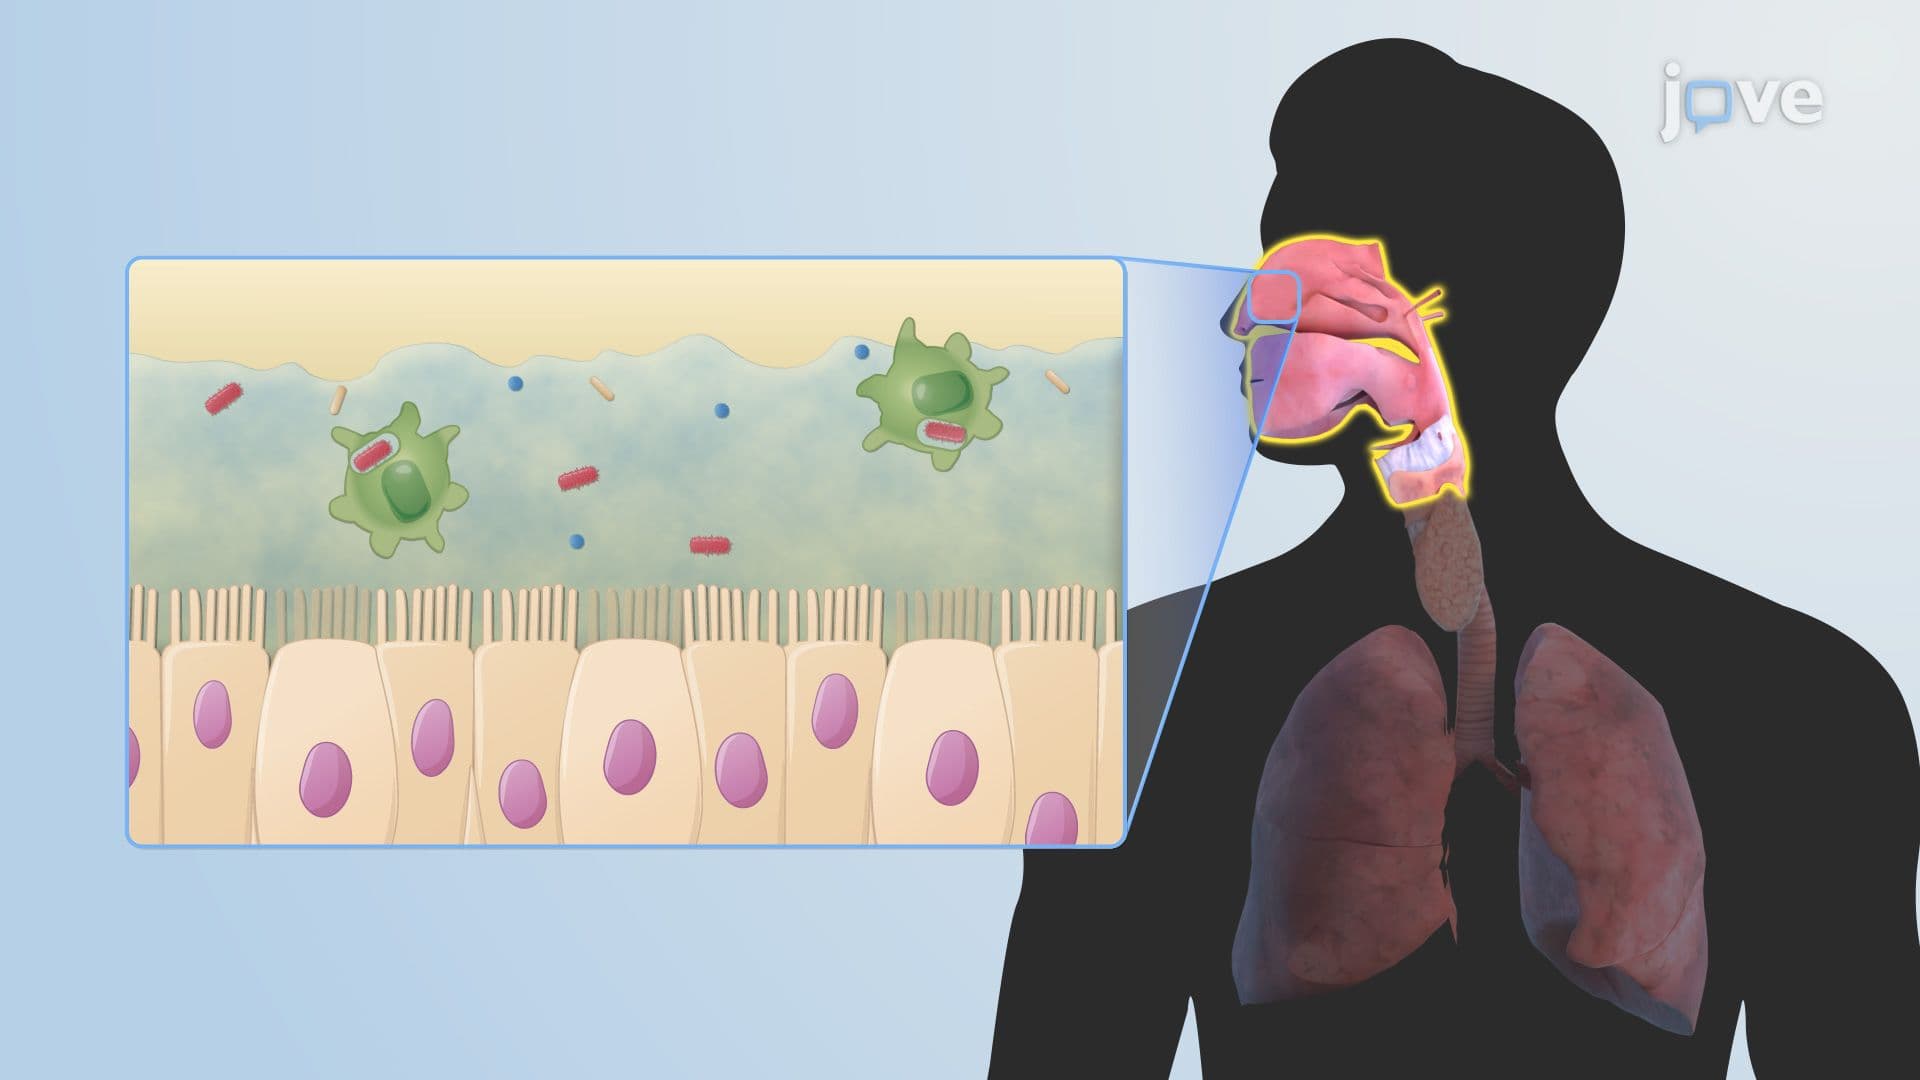
Microbiota of the Respiratory Tract
 01:30
Development of Human Microbiota
 01:28
Development of the Oral Microbiota
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:27
Microbiota of the Stomach and Small Intestine