Related Experiment Video
Updated: Mar 23, 2026
Modification and Functionalization of the Guanidine Group by Tailor-made Precursors
Published on: April 27, 2017
Conformational Restriction and Enantioseparation Increase Potency and Selectivity of Cyanoguanidine-Type Histamine H4
Roland Geyer1, Uwe Nordemann1, Andrea Strasser1
1Institute of Pharmacy, Pharmaceutical/Medicinal Chemistry, Faculty of Chemistry and Pharmacy, University of Regensburg , Universitätsstraße 31, D-93053 Regensburg, Germany.

Abstract:
2-Cyano-1-[4-(1H-imidazol-4-yl)butyl]-3-[2-(phenylsulfanyl)ethyl]guanidine (UR-PI376, 1) is a potent and selective agonist of the human histamine H4 receptor (hH4R). To gain information on the active conformation, we synthesized analogues of 1 with a cyclopentane-1,3-diyl linker. Affinities and functional activities were determined at recombinant hHxR (x: 1-4) subtypes on Sf9 cell membranes (radioligand binding, [(35)S]GTPγS, or GTPase assays) and in part in luciferase assays on human or mouse H4R (HEK-293 cells). The most potent H4R agonists among 14 racemates were separated by chiral HPLC, yielding eight enantiomerically pure compounds. Configurations were assigned based on X-ray structures of intermediates and a stereocontrolled synthetic pathway. (+)-2-Cyano-1-{[trans-(1S,3S)-3-(1H-imidazol-4-yl)cyclopentyl]methyl}-3-[2-(phenylsulfanyl)ethyl]guanidine ((1S,3S)-UR-RG98, 39a) was the most potent H4R agonist in this series (EC50 11 nM; H4R vs H3R, >100-fold selectivity; H1R, H2R, negligible activities), whereas the optical antipode proved to be an H4R antagonist ([(35)S]GTPγS assay). MD simulations confirmed differential stabilization of the active and inactive H4R state by the enantiomers.
More Related Videos
 06:13
Stepwise Dosing Protocol for Increased Throughput in Label-Free Impedance-Based GPCR Assays
Published on: February 21, 2020
 09:39
Drug-induced Sensitization of Adenylyl Cyclase: Assay Streamlining and Miniaturization for Small Molecule and siRNA Screening Applications
Published on: January 27, 2014
Related Concept Videos
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...
 01:28
Acid Suppressive Drugs for Peptic Ulcer Disease: Histamine H2-Receptor Antagonists
 01:30
Adrenergic Agonists: Direct-Acting Agents
These agents can be classified...
 01:17
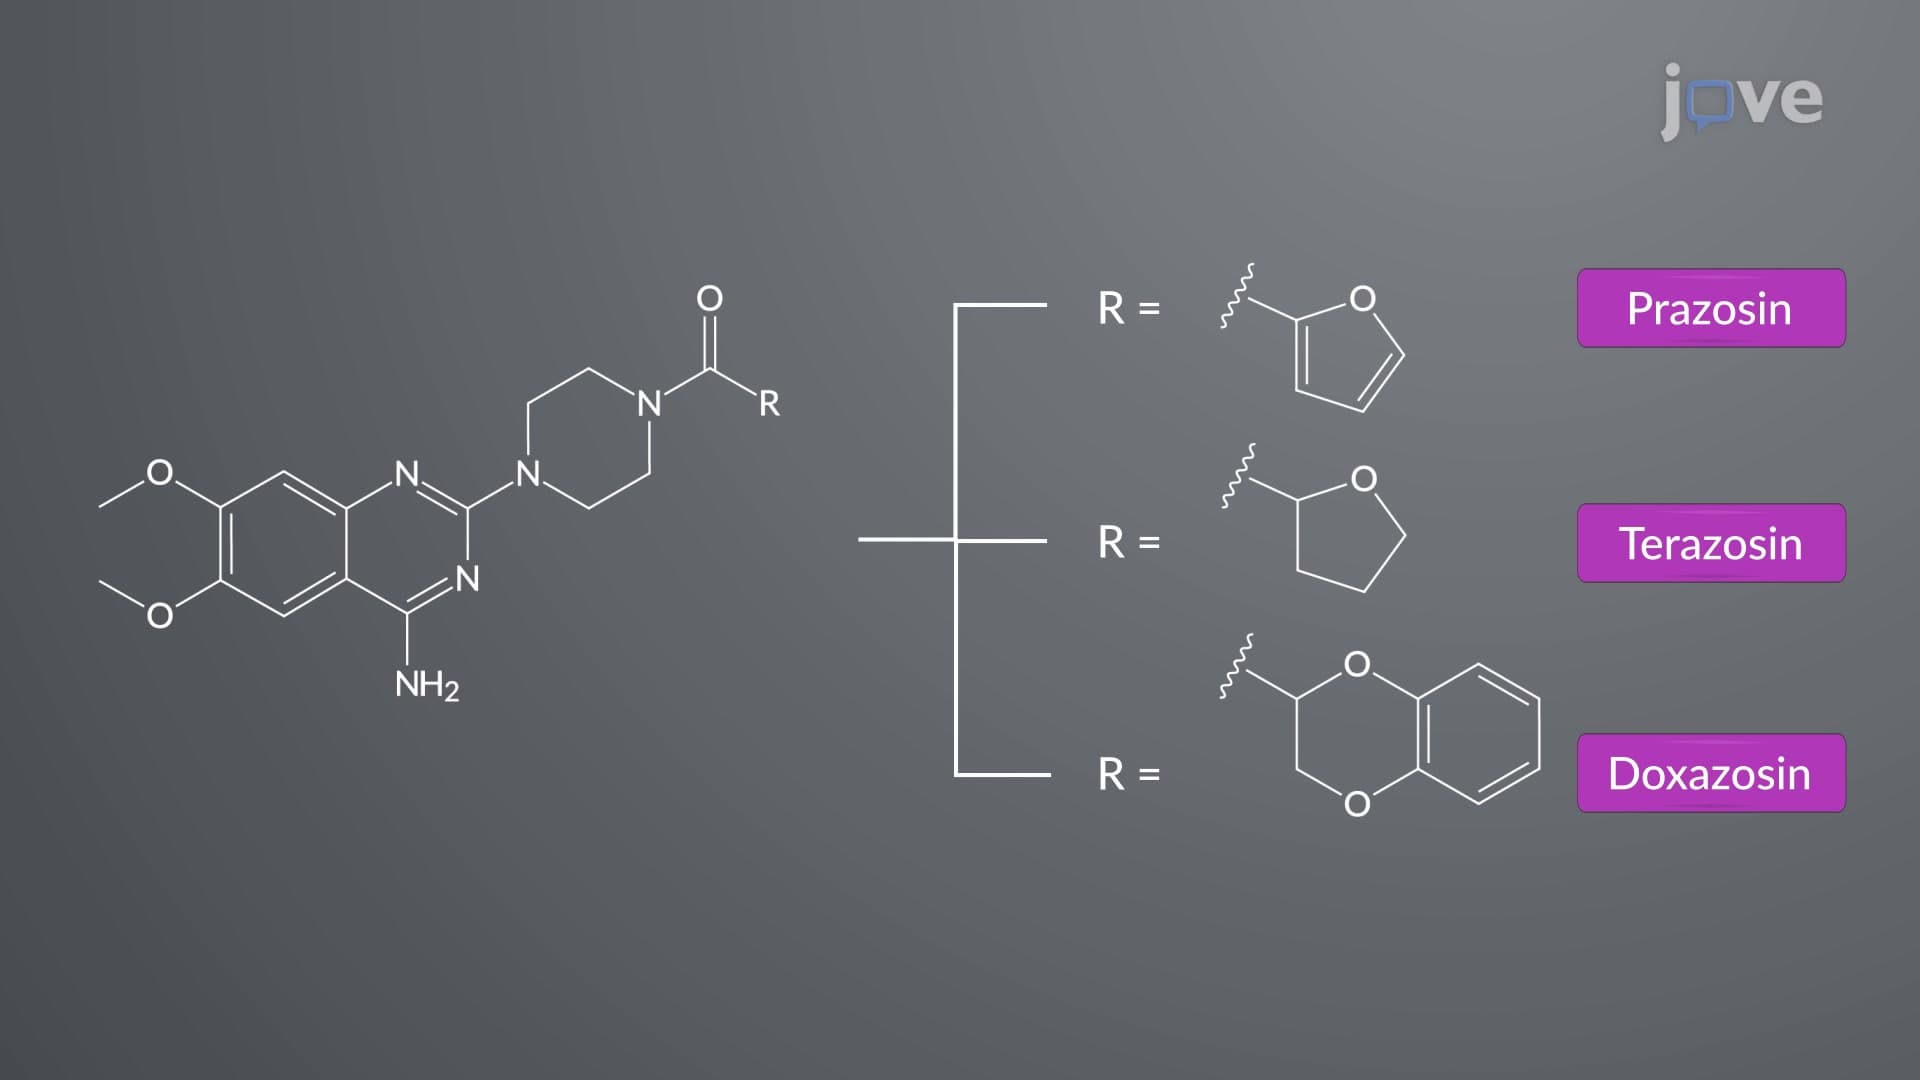
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline...
 01:29
Indirect-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
Reversible inhibitors display short to medium durations of action. Short-acting agents include simple alcohols with...
 01:25
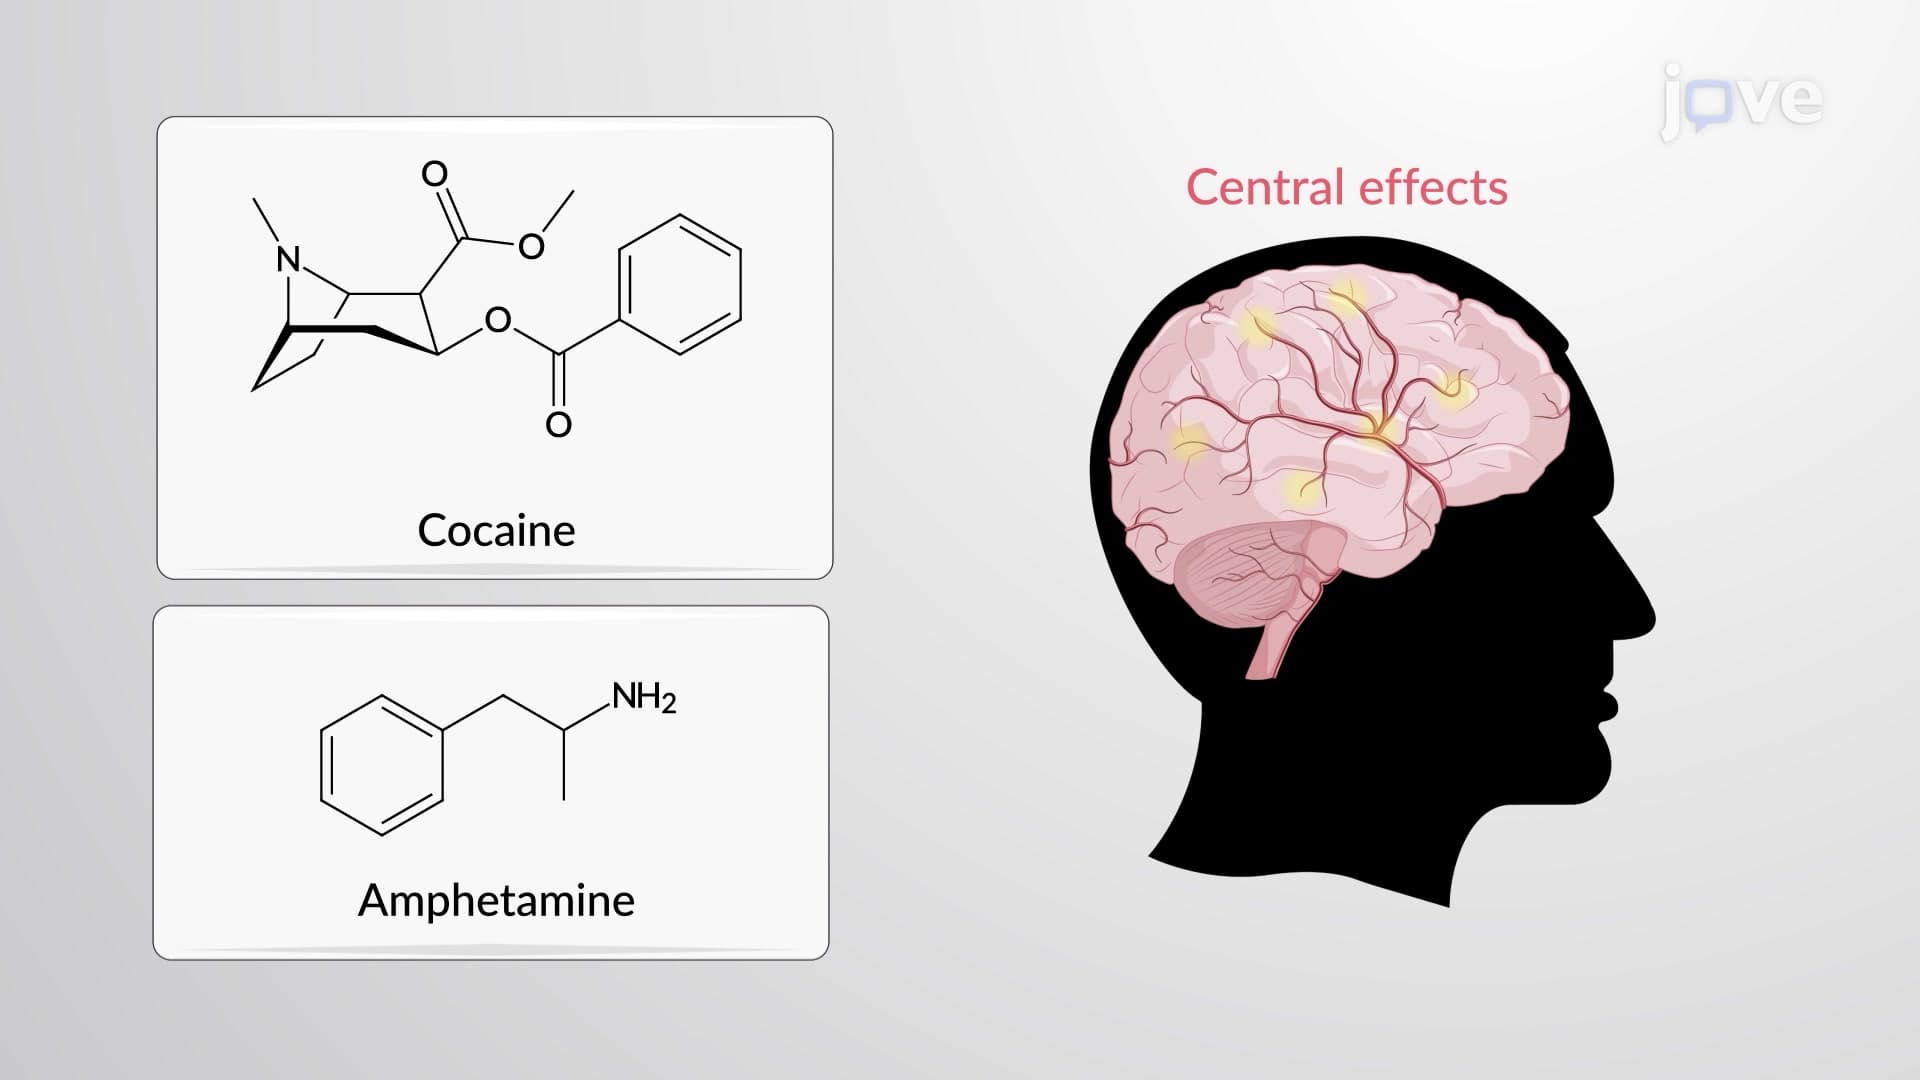
Adrenergic Agonists: Indirect-Acting Agents
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral...