Related Experiment Video
Updated: Mar 23, 2026

Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Highly efficient implementation of pseudospectral time-dependent density-functional theory for the calculation of
Yixiang Cao1, Thomas Hughes1, Dave Giesen1
1Schrödinger Inc, 120 West 45th Street, Tower 45, 17th Floor, New York, New York, 10036.

Abstract:
We have developed and implemented pseudospectral time-dependent density-functional theory (TDDFT) in the quantum mechanics package Jaguar to calculate restricted singlet and restricted triplet, as well as unrestricted excitation energies with either full linear response (FLR) or the Tamm-Dancoff approximation (TDA) with the pseudospectral length scales, pseudospectral atomic corrections, and pseudospectral multigrid strategy included in the implementations to improve the chemical accuracy and to speed the pseudospectral calculations. The calculations based on pseudospectral time-dependent density-functional theory with full linear response (PS-FLR-TDDFT) and within the Tamm-Dancoff approximation (PS-TDA-TDDFT) for G2 set molecules using B3LYP/6-31G*(*) show mean and maximum absolute deviations of 0.0015 eV and 0.0081 eV, 0.0007 eV and 0.0064 eV, 0.0004 eV and 0.0022 eV for restricted singlet excitation energies, restricted triplet excitation energies, and unrestricted excitation energies, respectively; compared with the results calculated from the conventional spectral method. The application of PS-FLR-TDDFT to OLED molecules and organic dyes, as well as the comparisons for results calculated from PS-FLR-TDDFT and best estimations demonstrate that the accuracy of both PS-FLR-TDDFT and PS-TDA-TDDFT. Calculations for a set of medium-sized molecules, including Cn fullerenes and nanotubes, using the B3LYP functional and 6-31G(**) basis set show PS-TDA-TDDFT provides 19- to 34-fold speedups for Cn fullerenes with 450-1470 basis functions, 11- to 32-fold speedups for nanotubes with 660-3180 basis functions, and 9- to 16-fold speedups for organic molecules with 540-1340 basis functions compared to fully analytic calculations without sacrificing chemical accuracy. The calculations on a set of larger molecules, including the antibiotic drug Ramoplanin, the 46-residue crambin protein, fullerenes up to C540 and nanotubes up to 14×(6,6), using the B3LYP functional and 6-31G(**) basis set with up to 8100 basis functions show that PS-FLR-TDDFT CPU time scales as N(2.05) with the number of basis functions. © 2016 Wiley Periodicals, Inc.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:14
Molecular Spectroscopy: Absorption and Emission
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:02
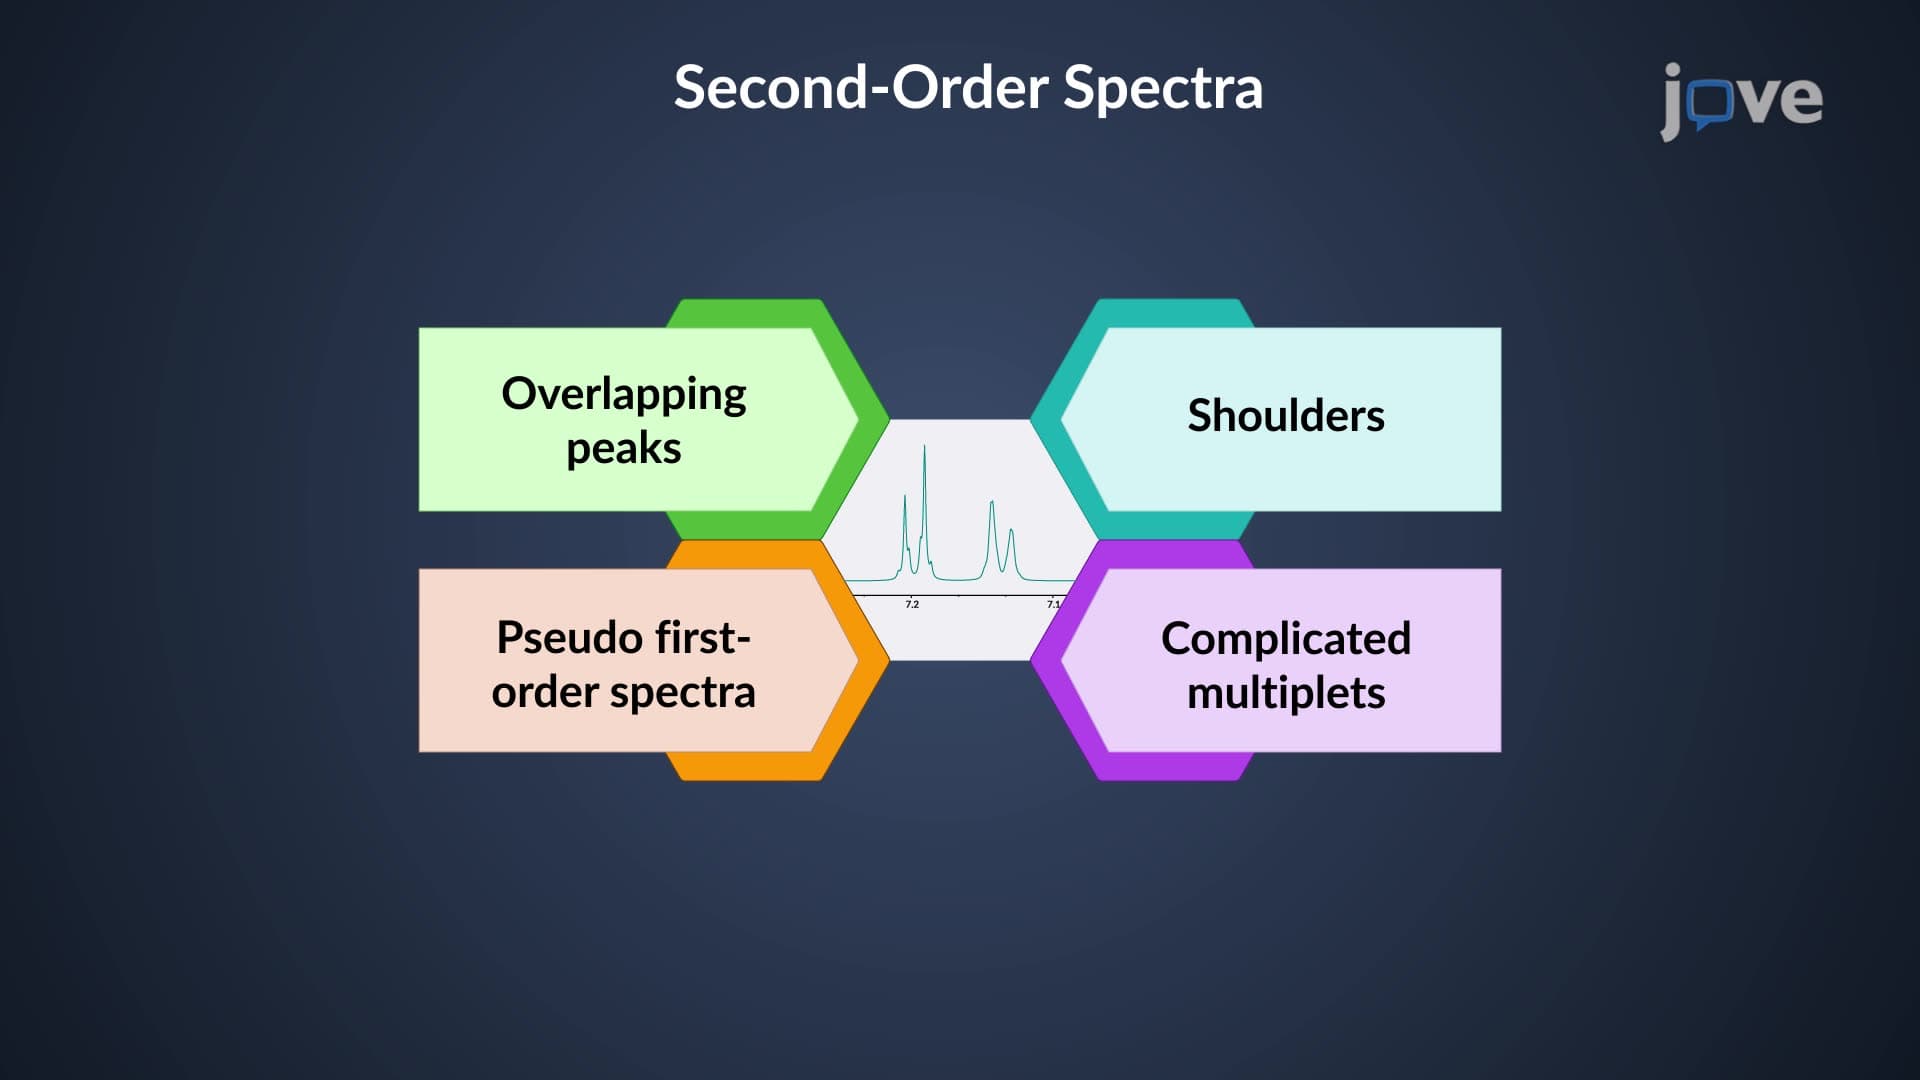
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 03:51
Molecular Orbital Theory II