Related Experiment Video
Updated: Mar 23, 2026
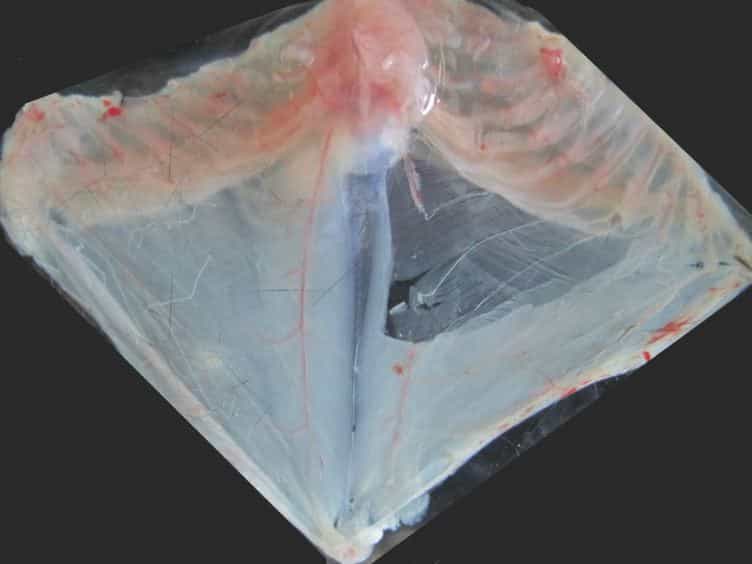

Dissection of the Transversus Abdominis Muscle for Whole-mount Neuromuscular Junction Analysis
Published on: January 11, 2014
Motor neuron degeneration in spastic paraplegia 11 mimics amyotrophic lateral sclerosis lesions
Paola S Denora1, Katrien Smets2, Federica Zolfanelli3
11 Ecole Pratique des Hautes Etudes, EPHE, PSL université, laboratoire de neurogénétique, F-75013, Paris, France 2 Inserm, U1127, F-75013, Paris, France 3 CNRS, UMR7225, F-75013, Paris, France 4 Sorbonne Universités, UPMC Univ Paris 06, UMR_S1127, Institut du Cerveau et de la Moelle épinière - ICM, Pitié-Salpêtrière Hospital, F-75013, Paris, France 5 Department of Genetics and Rare Diseases, IRCCS Bambino Gesu' Children Hospital, Rome, Italy.

Abstract:
The most common form of autosomal recessive hereditary spastic paraplegia is caused by mutations in the SPG11/KIAA1840 gene on chromosome 15q. The nature of the vast majority of SPG11 mutations found to date suggests a loss-of-function mechanism of the encoded protein, spatacsin. The SPG11 phenotype is, in most cases, characterized by a progressive spasticity with neuropathy, cognitive impairment and a thin corpus callosum on brain MRI. Full neuropathological characterization has not been reported to date despite the description of >100 SPG11 mutations. We describe here the clinical and pathological features observed in two unrelated females, members of genetically ascertained SPG11 families originating from Belgium and Italy, respectively. We confirm the presence of lesions of motor tracts in medulla oblongata and spinal cord associated with other lesions of the central nervous system. Interestingly, we report for the first time pathological hallmarks of SPG11 in neurons that include intracytoplasmic granular lysosome-like structures mainly in supratentorial areas, and others in subtentorial areas that are partially reminiscent of those observed in amyotrophic lateral sclerosis, such as ubiquitin and p62 aggregates, except that they are never labelled with anti-TDP-43 or anti-cystatin C. The neuropathological overlap with amyotrophic lateral sclerosis, associated with some shared clinical manifestations, opens up new fields of investigation in the physiopathological continuum of motor neuron degeneration.
More Related Videos
 06:35
In Vivo Electrophysiological Measurement of Compound Muscle Action Potential from the Forelimbs in Mouse Models of Motor Neuron Degeneration
Published on: June 15, 2018
 07:43
Modeling Charcot-Marie-Tooth Disease In Vitro by Transfecting Mouse Primary Motoneurons
Published on: January 7, 2019
Related Concept Videos
 01:26
Cross-bridge Cycle
 01:21
Satellite Stem Cells and Muscular Dystrophy
 01:37
Neural Regulation
 01:15
Parkinson's Disease: Overview