Related Experiment Video
Updated: Mar 22, 2026

Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
Molecular phylogeny and genome size evolution of the genus Betula (Betulaceae)
Nian Wang1, Hugh A McAllister2, Paul R Bartlett3
1School of Biological and Chemical Sciences, Queen Mary University of London, London E1 4NS, UK.

Background And Aims:
Betula L. (birch) is a genus of approx. 60 species, subspecies or varieties with a wide distribution in the northern hemisphere, of ecological and economic importance. A new classification of Betula has recently been proposed based on morphological characters. This classification differs somewhat from previously published molecular phylogenies, which may be due to factors such as convergent evolution, hybridization, incomplete taxon sampling or misidentification of samples. While chromosome counts have been made for many species, few have had their genome size measured. The aim of this study is to produce a new phylogenetic and genome size analysis of the genus.
Methods:
Internal transcribed spacer (ITS) regions of nuclear ribosomal DNA were sequenced for 76 Betula samples verified by taxonomic experts, representing approx. 60 taxa, of which approx. 24 taxa have not been included in previous phylogenetic analyses. A further 49 samples from other collections were also sequenced, and 108 ITS sequences were downloaded from GenBank. Phylogenetic trees were built for these sequences. The genome sizes of 103 accessions representing nearly all described species were estimated using flow cytometry.
Key Results:
As expected for a gene tree of a genus where hybridization and allopolyploidy occur, the ITS tree shows clustering, but not resolved monophyly, for the morphological subgenera recently proposed. Most sections show some clustering, but species of the dwarf section Apterocaryon are unusually scattered. Betula corylifolia (subgenus Nipponobetula) unexpectedly clusters with species of subgenus Aspera Unexpected placements are also found for B. maximowicziana, B. bomiensis, B. nigra and B. grossa Biogeographical disjunctions were found within Betula between Europe and North America, and also disjunctions between North-east and South-west Asia. The 2C-values for Betula ranged from 0·88 to 5·33 pg, and polyploids are scattered widely throughout the ITS phylogeny. Species with large genomes tend to have narrow ranges.
Conclusions:
Betula grossa may have formed via allopolyploidization between parents in subgenus Betula and subgenus Aspera. Betula bomiensis may also be a wide allopolyploid. Betula corylifolia may be a parental species of allopolyploids in the subsection Chinenses Placements of B. maximowicziana, B. michauxii and B. nigra need further investigation. This analysis, in line with previous studies, suggests that section Apterocaryon is not monophyletic and thus dwarfism has evolved repeatedly in different lineages of Betula Polyploidization has occurred many times independently in the evolution of Betula.
More Related Videos
 10:23
A Concoction Pipeline for Generating Molecular Operational Taxonomic Units (MOTUs) Among Riparian and Aquatic Beetles
Published on: July 11, 2025
 12:05
A Workflow for the Quantitative Assessment of the Endophytic and Epiphytic Bacterial Microbiomes of the Bark of Populus trichocarpa
Published on: June 27, 2025
Related Concept Videos
 02:54
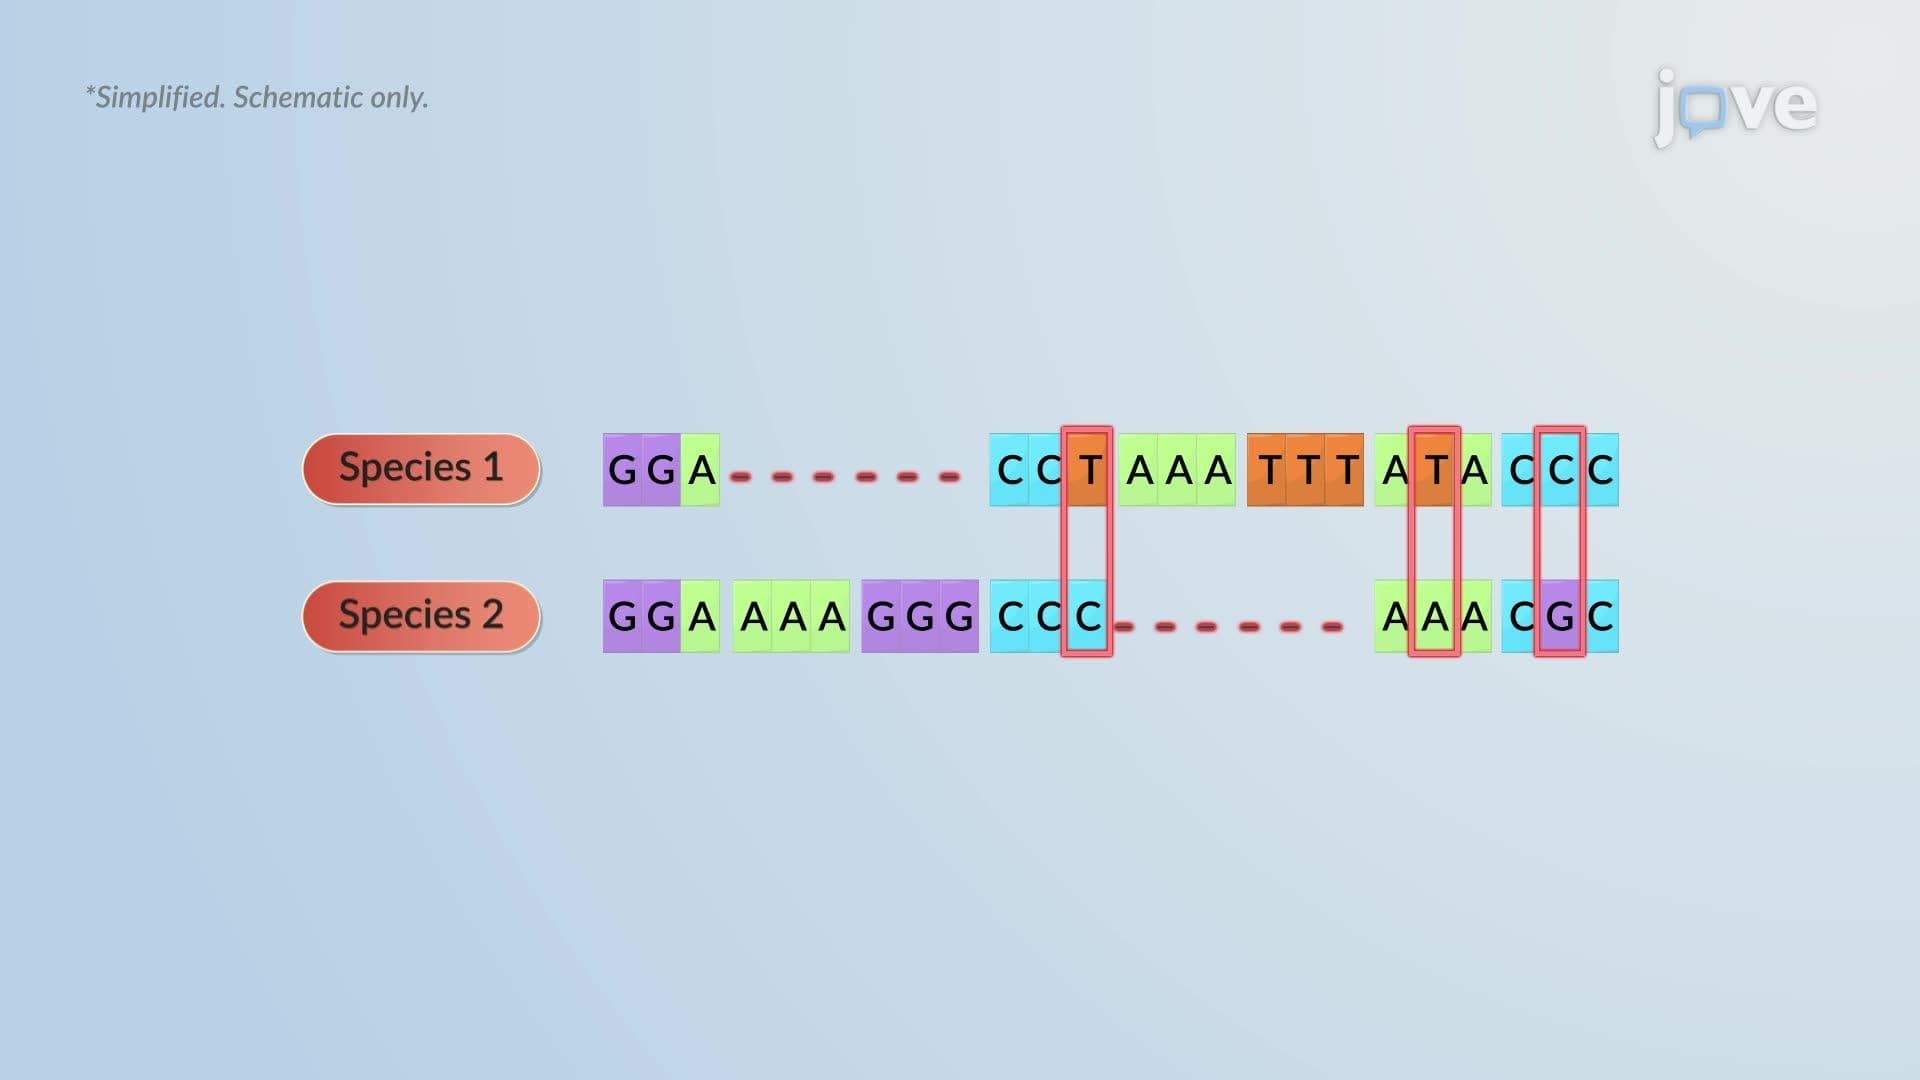
Evolutionary Relationships through Genome Comparisons
 01:28
Microbial Phylogeny
 03:21
Genome Size and the Evolution of New Genes
03:21Genome Size and the Evolution of New Genes
 02:16
Comparing Mitochondrial, Chloroplast, and Prokaryotic Genomes
 01:29
Modern Molecular Taxonomy