Related Experiment Video
Updated: Mar 22, 2026

Novel Techniques for Observing Structural Dynamics of Photoresponsive Liquid Crystals
Published on: May 29, 2018
New insights into photodissociation dynamics of cyclobutanone from the AIMS dynamic simulation
1Key Laboratory of Theoretical and Computational Photochemistry Ministry of Education College of Chemistry, Beijing Normal University, Beijing 100875, China.

Abstract:
In this work, the combined electronic structure calculations and non-adiabatic dynamics simulations were performed for understanding mechanistic photodissociation of cyclobutanone at ∼248 nm. Besides the stationary and intersection structures reported before, two new conical intersections between the ground (S0) and the first excited singlet (S1) states were determined in the present study, which were confirmed to be the new S1 → S0 funnels by the ab initio multiple spawning dynamic simulation, giving rise to products in the S0 state selectively. The time evolution of the S1 electronic population was fitted with the pure exponential formulae, from which the S1 lifetime was estimated to be 484.0 fs. The time constant for the S1 α-cleavage is calculated to be 176.6 fs, which is based on the present dynamics simulation. As a result of the ultrafast S1 processes, the statistical distribution of the excess energies is prevented in the S1 state. The S1 dynamic effect (the nonergodic behavior) was predicted to be an important factor that is responsible for the wavelength dependence of the branching ratio of photodissociation products, which will be discussed in detail.
More Related Videos
 08:51
Coulomb Explosion Imaging as a Tool to Distinguish Between Stereoisomers
Published on: August 18, 2017
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:05
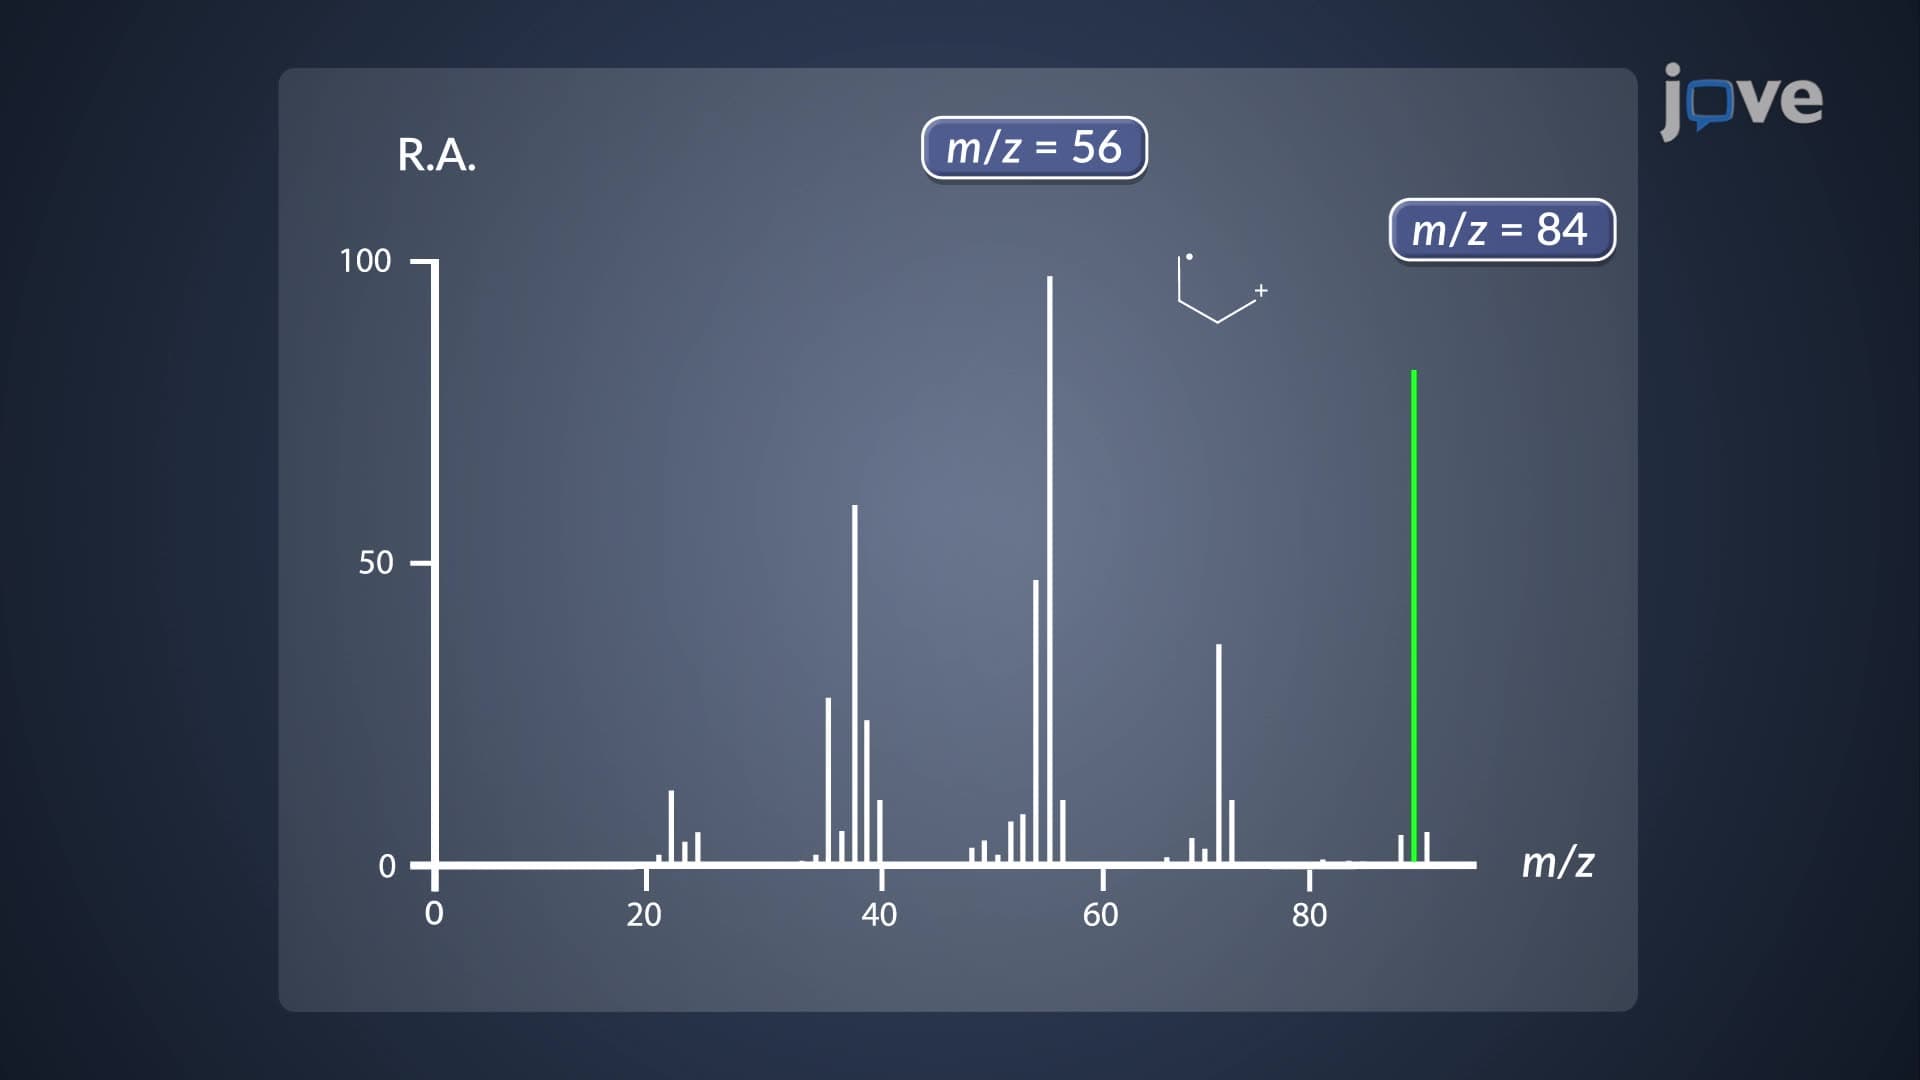
Mass Spectrometry: Cycloalkane Fragmentation
For example, cyclohexane molecular ions have a mass-to-charge ratio (m/z) of 84, which tends to produce a stronger signal than linear alkanes like hexane. This stability comes from...
 00:54
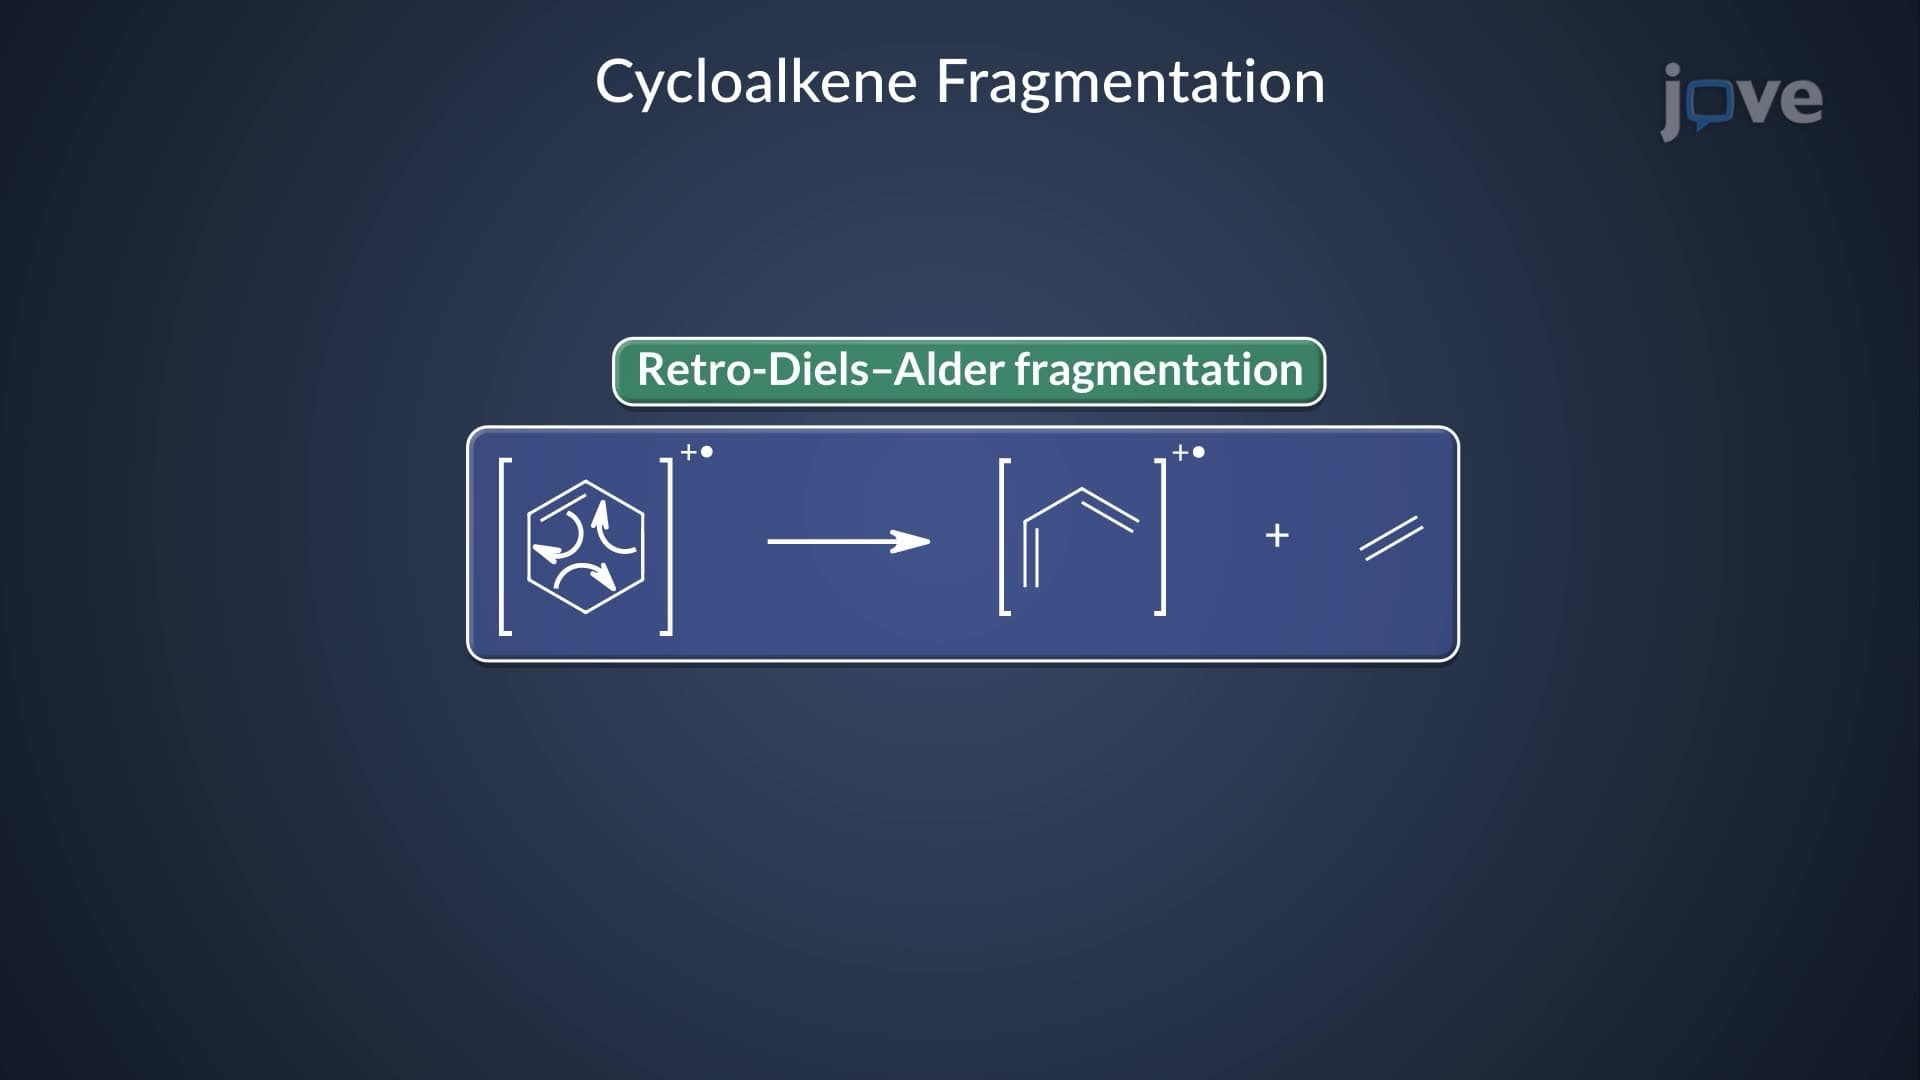
Mass Spectrometry: Cycloalkene Fragmentation
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 02:02
Chair Conformation of Cyclohexane
The hydrogen atoms linked to carbons are arranged in two different axial and equatorial orientations to achieve this...