Related Experiment Video
Updated: Mar 22, 2026

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Is It Reliable to Use Common Molecular Docking Methods for Comparing the Binding Affinities of Enantiomer Pairs for
David Ramírez1, Julio Caballero2
1Centro de Bioinformática y Simulación Molecular (CBSM), Universidad de Talca. 2 Norte 685, Casilla 721, Talca, Chile. damach.david@gmail.com.

Abstract:
Molecular docking is a computational chemistry method which has become essential for the rational drug design process. In this context, it has had great impact as a successful tool for the study of ligand-receptor interaction modes, and for the exploration of large chemical datasets through virtual screening experiments. Despite their unquestionable merits, docking methods are not reliable for predicting binding energies due to the simple scoring functions they use. However, comparisons between two or three complexes using the predicted binding energies as a criterion are commonly found in the literature. In the present work we tested how wise is it to trust the docking energies when two complexes between a target protein and enantiomer pairs are compared. For this purpose, a ligand library composed by 141 enantiomeric pairs was used, including compounds with biological activities reported against seven protein targets. Docking results using the software Glide (considering extra precision (XP), standard precision (SP), and high-throughput virtual screening (HTVS) modes) and AutoDock Vina were compared with the reported biological activities using a classification scheme. Our test failed for all modes and targets, demonstrating that an accurate prediction when binding energies of enantiomers are compared using docking may be due to chance. We also compared pairs of compounds with different molecular weights and found the same results.
More Related Videos
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 02:18
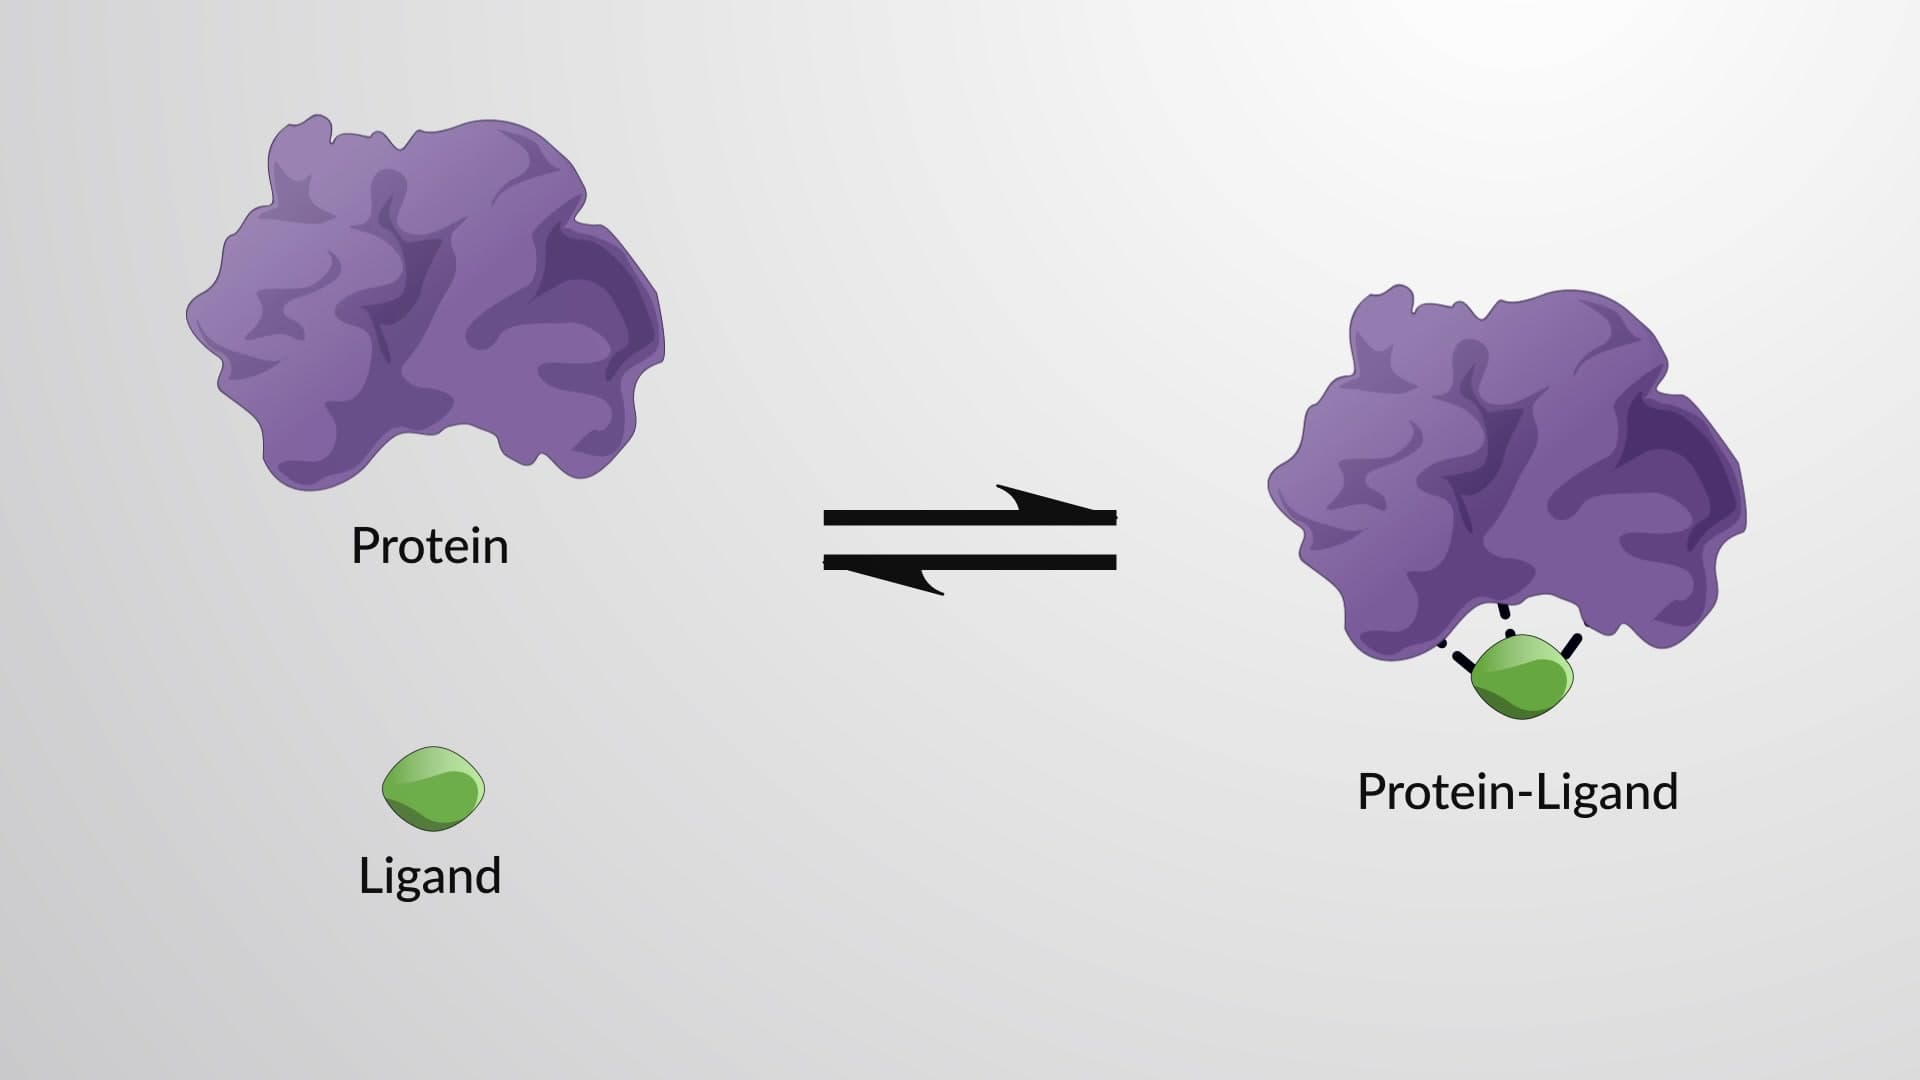
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
 01:22
Protein-Drug Binding: Determination Methods
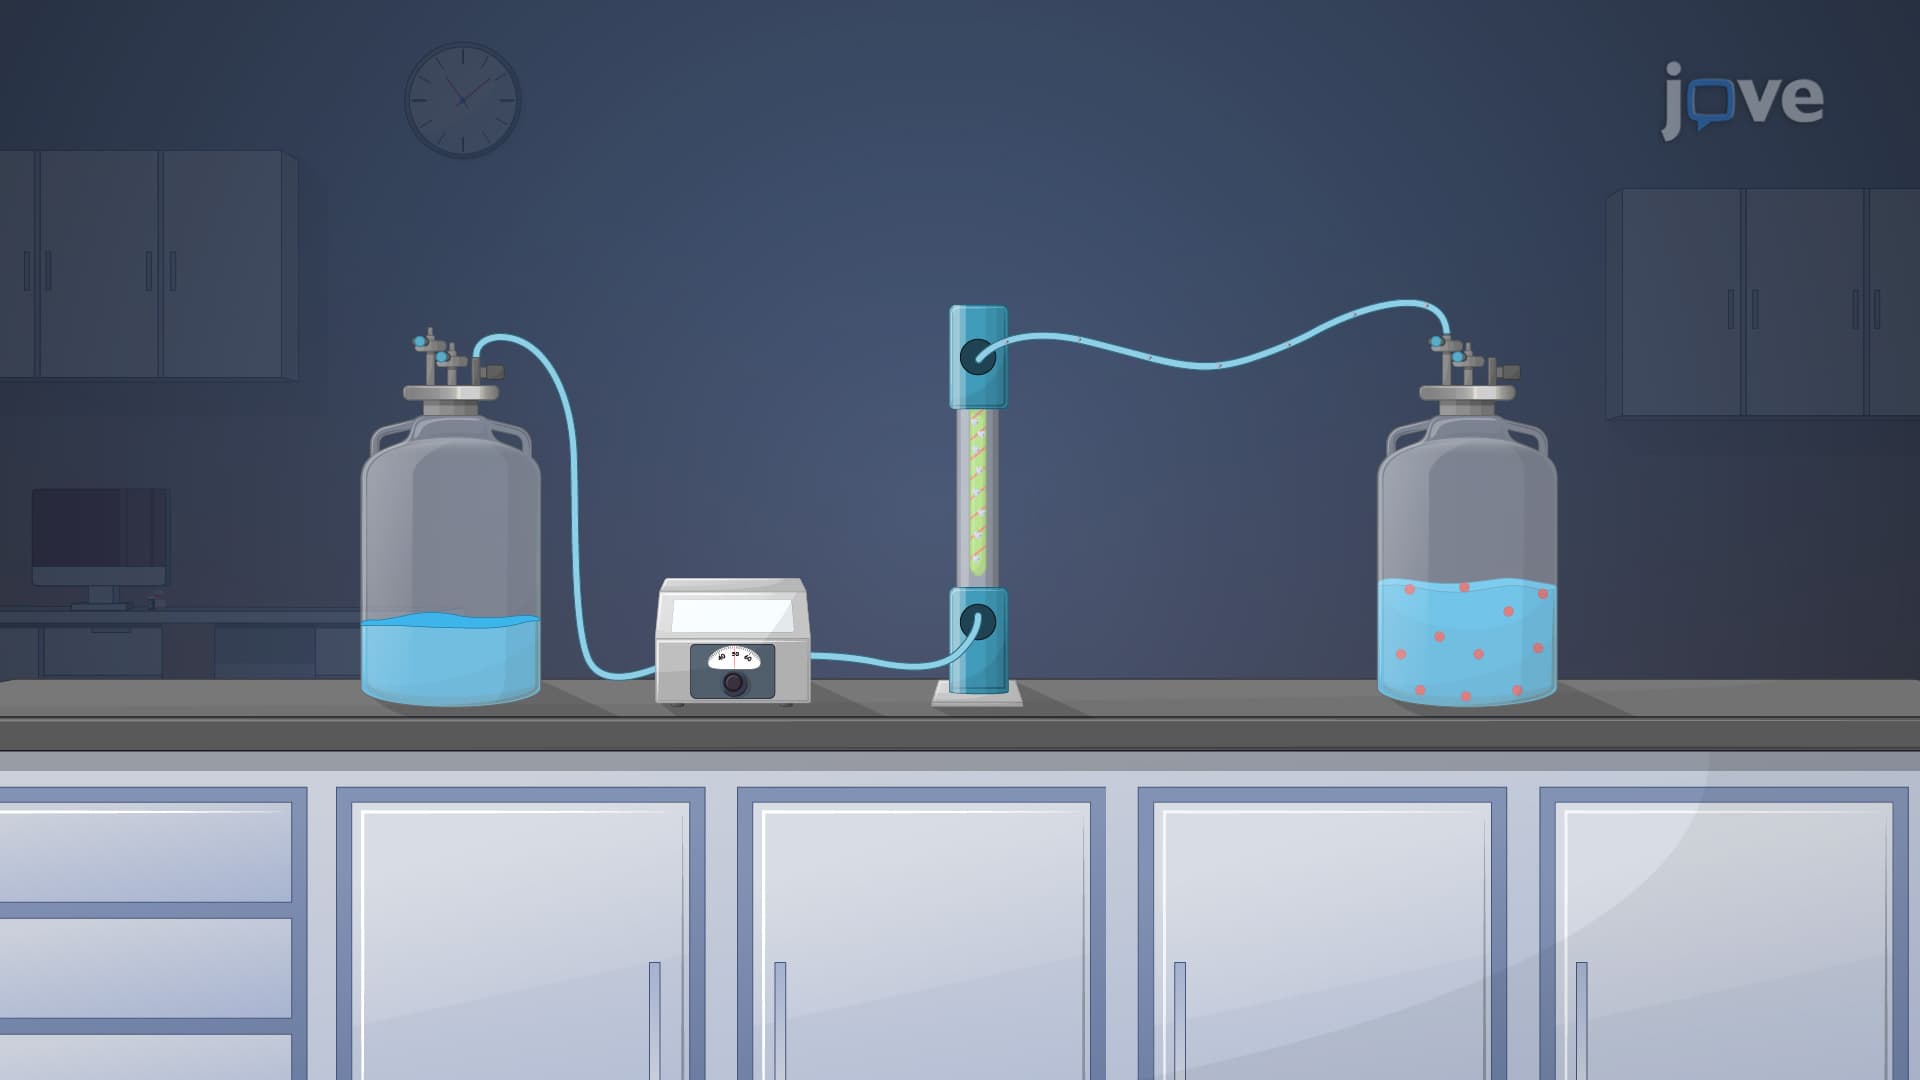
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:24
Properties of Enantiomers and Optical Activity