Related Experiment Video
Updated: Mar 21, 2026


Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
A Simple and Accurate Method To Calculate Free Energy Profiles and Reaction Rates from Restrained Molecular
Victor Ovchinnikov1, Kwangho Nam2, Martin Karplus1,3
1Department of Chemistry and Chemical Biology, Harvard University , Cambridge, Massachusetts 02138, United States.

Abstract:
A method is developed to obtain simultaneously free energy profiles and diffusion constants from restrained molecular simulations in diffusive systems. The method is based on low-order expansions of the free energy and diffusivity as functions of the reaction coordinate. These expansions lead to simple analytical relationships between simulation statistics and model parameters. The method is tested on 1D and 2D model systems; its accuracy is found to be comparable to or better than that of the existing alternatives, which are briefly discussed. An important aspect of the method is that the free energy is constructed by integrating its derivatives, which can be computed without need for overlapping sampling windows. The implementation of the method in any molecular simulation program that supports external umbrella potentials (e.g., CHARMM) requires modification of only a few lines of code. As a demonstration of its applicability to realistic biomolecular systems, the method is applied to model the α-helix ↔ β-sheet transition in a 16-residue peptide in implicit solvent, with the reaction coordinate provided by the string method. Possible modifications of the method are briefly discussed; they include generalization to multidimensional reaction coordinates [in the spirit of the model of Ermak and McCammon (Ermak, D. L.; McCammon, J. A. J. Chem. Phys. 1978, 69, 1352-1360)], a higher-order expansion of the free energy surface, applicability in nonequilibrium systems, and a simple test for Markovianity. In view of the small overhead of the method relative to standard umbrella sampling, we suggest its routine application in the cases where umbrella potential simulations are appropriate.
Related Concept Videos
 02:23
Gibbs Free Energy and Thermodynamic Favorability
 02:55
Temperature Dependence on Reaction Rate
Atoms, molecules, or ions must collide before they can react with each other. Atoms must be close together to form chemical bonds. This premise is the basis for a theory that explains many observations regarding chemical kinetics, including factors affecting reaction rates.
The collision theory is based on the postulates that (i) the reaction rate is proportional to the rate of reactant collisions, (ii) the reacting species collide in an orientation allowing contact between...
 03:48
Behavior of Gas Molecules: Molecular Diffusion, Mean Free Path, and Effusion
 02:56
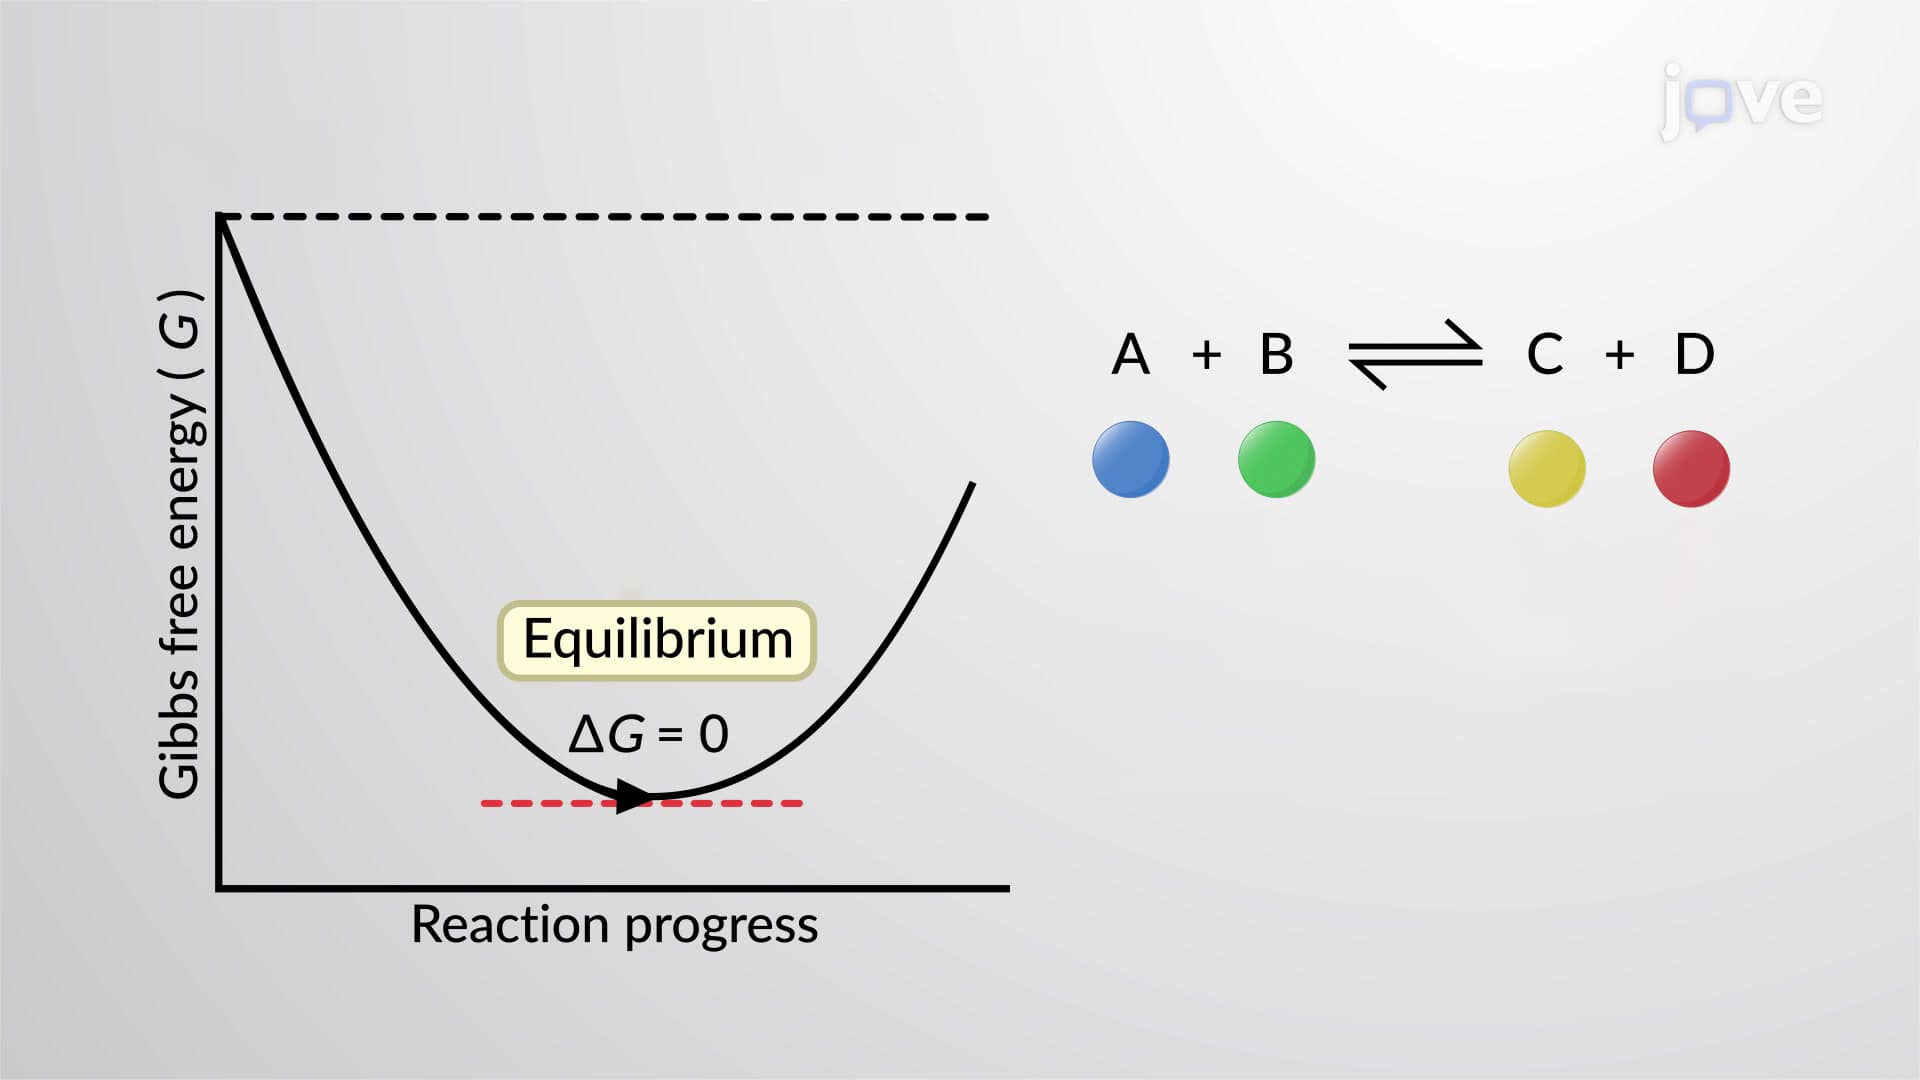
Free Energy and Equilibrium
Recall that Q is the numerical value of the mass action...
 00:55
Free Energy and Equilibrium
The reaction quotient, Q, is a convenient measure of the...
 03:25
Free Energy Changes for Nonstandard States