Related Experiment Video
Updated: Mar 20, 2026

Interactions with and Membrane Permeabilization of Brain Mitochondria by Amyloid Fibrils
Published on: September 28, 2019
Mitofusin-2 knockdown increases ER-mitochondria contact and decreases amyloid β-peptide production
Nuno Santos Leal1, Bernadette Schreiner1, Catarina Moreira Pinho1
1Center for Alzheimer Research, Division of Neurogeriatrics, Department of Neurobiology, Care Sciences and Society, Karolinska Institutet, Stockholm, Sweden.

Abstract:
Mitochondria are physically and biochemically in contact with other organelles including the endoplasmic reticulum (ER). Such contacts are formed between mitochondria-associated ER membranes (MAM), specialized subregions of ER, and the outer mitochondrial membrane (OMM). We have previously shown increased expression of MAM-associated proteins and enhanced ER to mitochondria Ca(2+) transfer from ER to mitochondria in Alzheimer's disease (AD) and amyloid β-peptide (Aβ)-related neuronal models. Here, we report that siRNA knockdown of mitofusin-2 (Mfn2), a protein that is involved in the tethering of ER and mitochondria, leads to increased contact between the two organelles. Cells depleted in Mfn2 showed increased Ca(2+) transfer from ER to mitchondria and longer stretches of ER forming contacts with OMM. Interestingly, increased contact resulted in decreased concentrations of intra- and extracellular Aβ40 and Aβ42 . Analysis of γ-secretase protein expression, maturation and activity revealed that the low Aβ concentrations were a result of impaired γ-secretase complex function. Amyloid-β precursor protein (APP), β-site APP-cleaving enzyme 1 and neprilysin expression as well as neprilysin activity were not affected by Mfn2 siRNA treatment. In summary, our data shows that modulation of ER-mitochondria contact affects γ-secretase activity and Aβ generation. Increased ER-mitochondria contact results in lower γ-secretase activity suggesting a new mechanism by which Aβ generation can be controlled.
Insights
Altering contact between the endoplasmic reticulum (ER) and mitochondria impacts Alzheimer's disease (AD) pathology. Increased ER-mitochondria contact reduces amyloid-beta (Aβ) generation by impairing γ-secretase activity.
Area of Science:
- Cell Biology
- Neuroscience
- Biochemistry
Background:
- Mitochondria and endoplasmic reticulum (ER) interact at mitochondria-associated ER membranes (MAM).
- Previous studies indicated increased MAM proteins and ER-mitochondria calcium (Ca2+) transfer in Alzheimer's disease (AD) models.
- Amyloid-beta (Aβ) peptide is implicated in AD pathogenesis.
Purpose of the Study:
- To investigate the role of mitofusin-2 (Mfn2) in ER-mitochondria contact and its effect on Aβ production.
- To determine if modulating ER-mitochondria contact influences γ-secretase activity.
Main Methods:
- Utilized siRNA to knockdown Mfn2, a protein mediating ER-mitochondria tethering.
- Quantified ER-mitochondria contact and Ca2+ transfer.
- Measured intra- and extracellular Aβ40 and Aβ42 concentrations.
- Analyzed γ-secretase complex components, maturation, and activity.
Main Results:
- Mfn2 knockdown increased ER-mitochondria contact and Ca2+ transfer.
- Reduced intra- and extracellular Aβ40 and Aβ42 levels were observed with Mfn2 depletion.
- Impaired γ-secretase complex function, not altered expression of APP or other proteases, accounted for decreased Aβ.
- Increased ER-mitochondria contact led to decreased γ-secretase activity.
Conclusions:
- Modulating ER-mitochondria contact influences γ-secretase activity and Aβ generation.
- Enhanced ER-mitochondria physical interaction presents a novel mechanism for controlling Aβ production in AD.
- Targeting Mfn2 or ER-mitochondria contact could be a therapeutic strategy for AD.
More Related Videos
 06:41
Quantitative Analysis of Mitochondria-Associated Endoplasmic Reticulum Membrane (MAM) Stabilization in a Neural Model of Alzheimer's Disease (AD)
Published on: January 10, 2025
 08:40
In Vitro and In Vivo Detection of Mitophagy in Human Cells, C. Elegans, and Mice
Published on: November 22, 2017
Related Concept Videos
 01:19
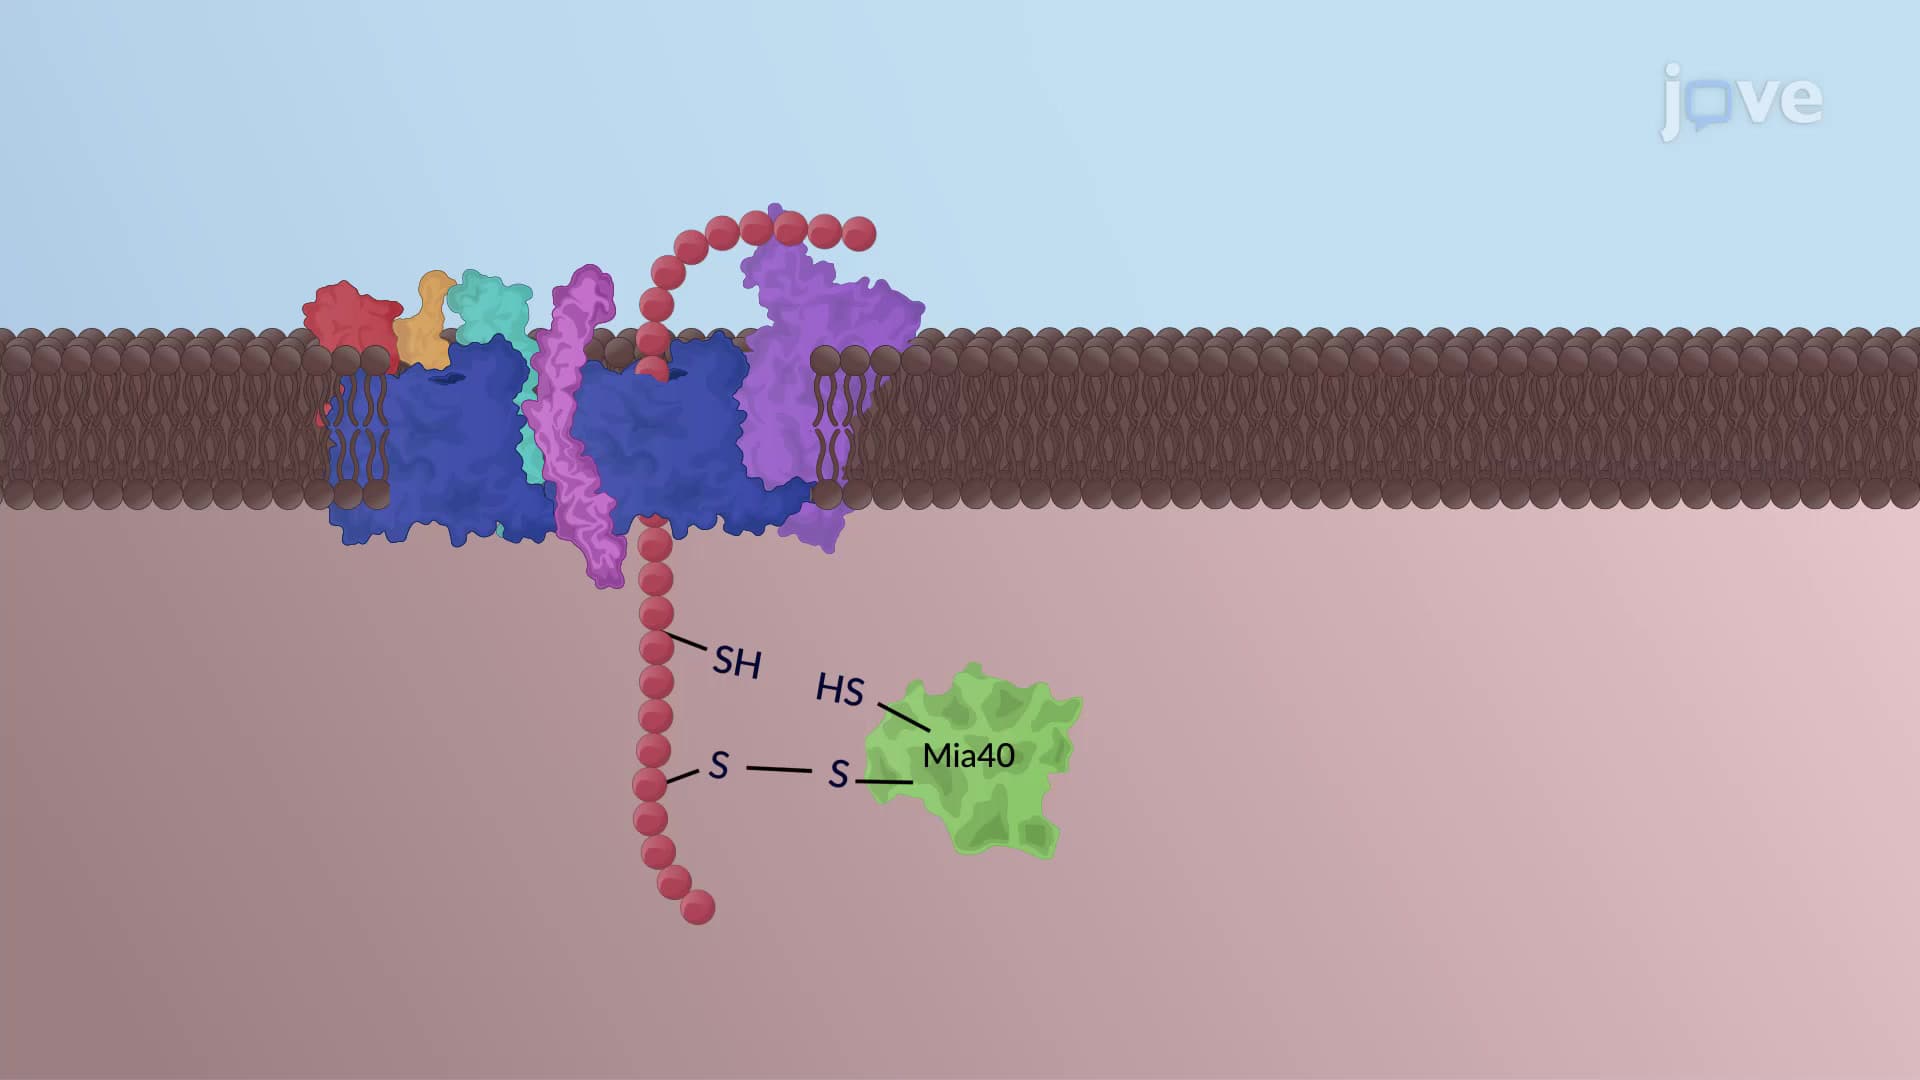
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining,...