Related Experiment Video
Updated: Mar 19, 2026
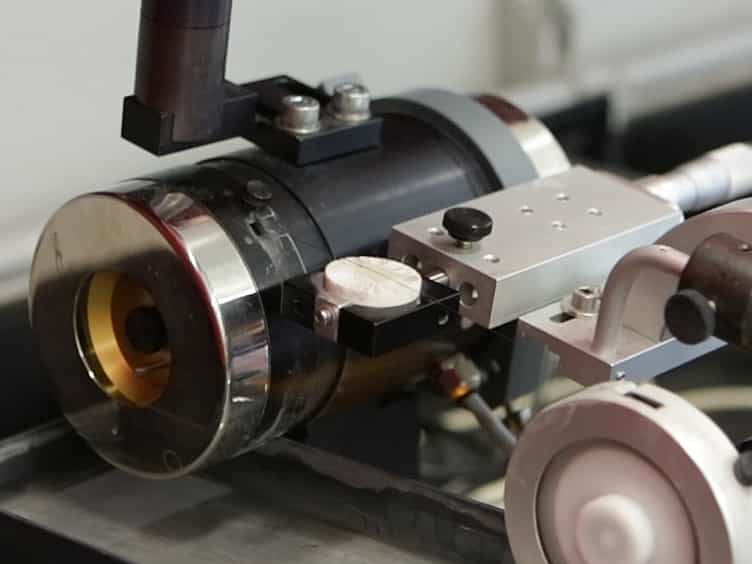

Spark Plasma Sintering Apparatus Used for the Formation of Strontium Titanate Bicrystals
Published on: February 9, 2017
Effect of the Local Atomic Ordering on the Stability of β-Spodumene
Radhika L Moore1, Brian S Haynes1, Alejandro Montoya1
1The University of Sydney , Faculty of Engineering and Information Technologies, School of Chemical and Biomolecular Engineering, Sydney, New South Wales 2006, Australia.

Abstract:
This study focuses on the relative energetic stability of β-spodumene configurations with different atomic ordering, evaluated using electronic structure methods based on static periodic density functional theory. We found that β-spodumene configurations with a framework containing exclusively Al-O-Si linkages are energetically the most stable, consistent with the aluminum avoidance principle. A correlation between the interstitial sites occupied by lithium and the stability of the configuration was established: highly stable configurations contain greater proportions of lithium associated with the edges of AlO4 tetrahedrons. The identified low-energy configurations have a band gap of ∼4.8 eV, and similar electronic band structures and densities of states. Both the PBE and PBEsol functionals predict small differences in the relative stabilities of the different configurations of β-spodumene. However, only PBEsol is able to reproduce the experimentally observed stability differences between α-spodumene and β-spodumene. β-Spodumene is the preferred polymorph at high temperatures, with the PBEsol inversion temperature from α- to β-spodumene predicted to occur at 1070 K.
More Related Videos
 07:24
Hyperspectral Imaging as a Tool to Study Optical Anisotropy in Lanthanide-Based Molecular Single Crystals
Published on: April 14, 2020
 07:24
Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
Related Concept Videos
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:26
Imperfections in Crystal Structure: Stoichiometric Point Defects
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:54
Trends in Lattice Energy: Ion Size and Charge
 02:40
MO Theory and Covalent Bonding