Related Experiment Video
Updated: Mar 19, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Common SAR Derived from Multiple QSAR Models on Vorinostat Derivatives Targeting HDACs in Tumor Treatment
Sugathan Praseetha, Srinivas Bandaru, Mukesh Yadav
1Research and Development Centre, Bharathiyar University, Coimbatore, 641046, India. suresh@kufos.ac.in.

Background:
Dysregulation of HDACs has been associated with tumour development and therefore inhibiting HDAC's have surfaced as promising therapeutic strategy in malignancy.
Methods:
Vorinostat analogues with different biological activities were investigated for underlying structure-activity relationship.
Results:
Out of six activities and their multiple QSAR models, HDAC1 and HDAC8 produced statistically fit, stable and predictive linear (MLR) and non-linear (SVM) QSAR models. In case of HDAC1 activity as end point, linear (R2=0.8089, R2 CV=0.7343) and non-linear (R2=0.9801, R2 CV=0.8952) QSAR models turned reliable to investigate SAR. Similarly, HDAC8 activity based linear (R2=0.9454, R2 CV=0.9049) and non-linear (R2=0.9899, R2 CV=0.9232) QSAR models produced statistically improved and stable models.
Conclusion:
Molecular descriptors derived from 3-D Morse and Radial Distribution Function indices were found to be selective in all the models. These molecular descriptors which encode common SAR among Vorinostat derivatives were evaluated for their potent HDAC inhibition activity.
Insights
Histone deacetylase (HDAC) inhibitors are promising cancer therapeutics. This study developed quantitative structure-activity relationship (QSAR) models for Vorinostat analogues, identifying key molecular descriptors for potent HDAC inhibition.
Area of Science:
- Medicinal Chemistry
- Computational Chemistry
- Pharmacology
Background:
- Dysregulation of histone deacetylases (HDACs) is linked to cancer development.
- HDAC inhibition presents a promising therapeutic strategy for malignancies.
Purpose of the Study:
- To investigate the structure-activity relationship (SAR) of Vorinostat analogues.
- To develop predictive quantitative structure-activity relationship (QSAR) models for HDAC inhibition.
Main Methods:
- Synthesis and evaluation of Vorinostat analogues for biological activity.
- Development of multiple linear regression (MLR) and support vector machine (SVM) QSAR models.
- Utilized 3-D Morse and Radial Distribution Function indices as molecular descriptors.
Main Results:
- Statistically significant and predictive QSAR models were developed for HDAC1 and HDAC8 inhibition.
- High statistical fit and stability were achieved for both linear and non-linear models.
- R-squared values for HDAC1 models: linear (0.8089), non-linear (0.9801).
- R-squared values for HDAC8 models: linear (0.9454), non-linear (0.9899).
Conclusions:
- Molecular descriptors from 3-D Morse and Radial Distribution Function indices were selective and crucial for modeling HDAC inhibition.
- These descriptors capture common SAR among Vorinostat derivatives, indicating their potential for potent HDAC inhibition.
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:15
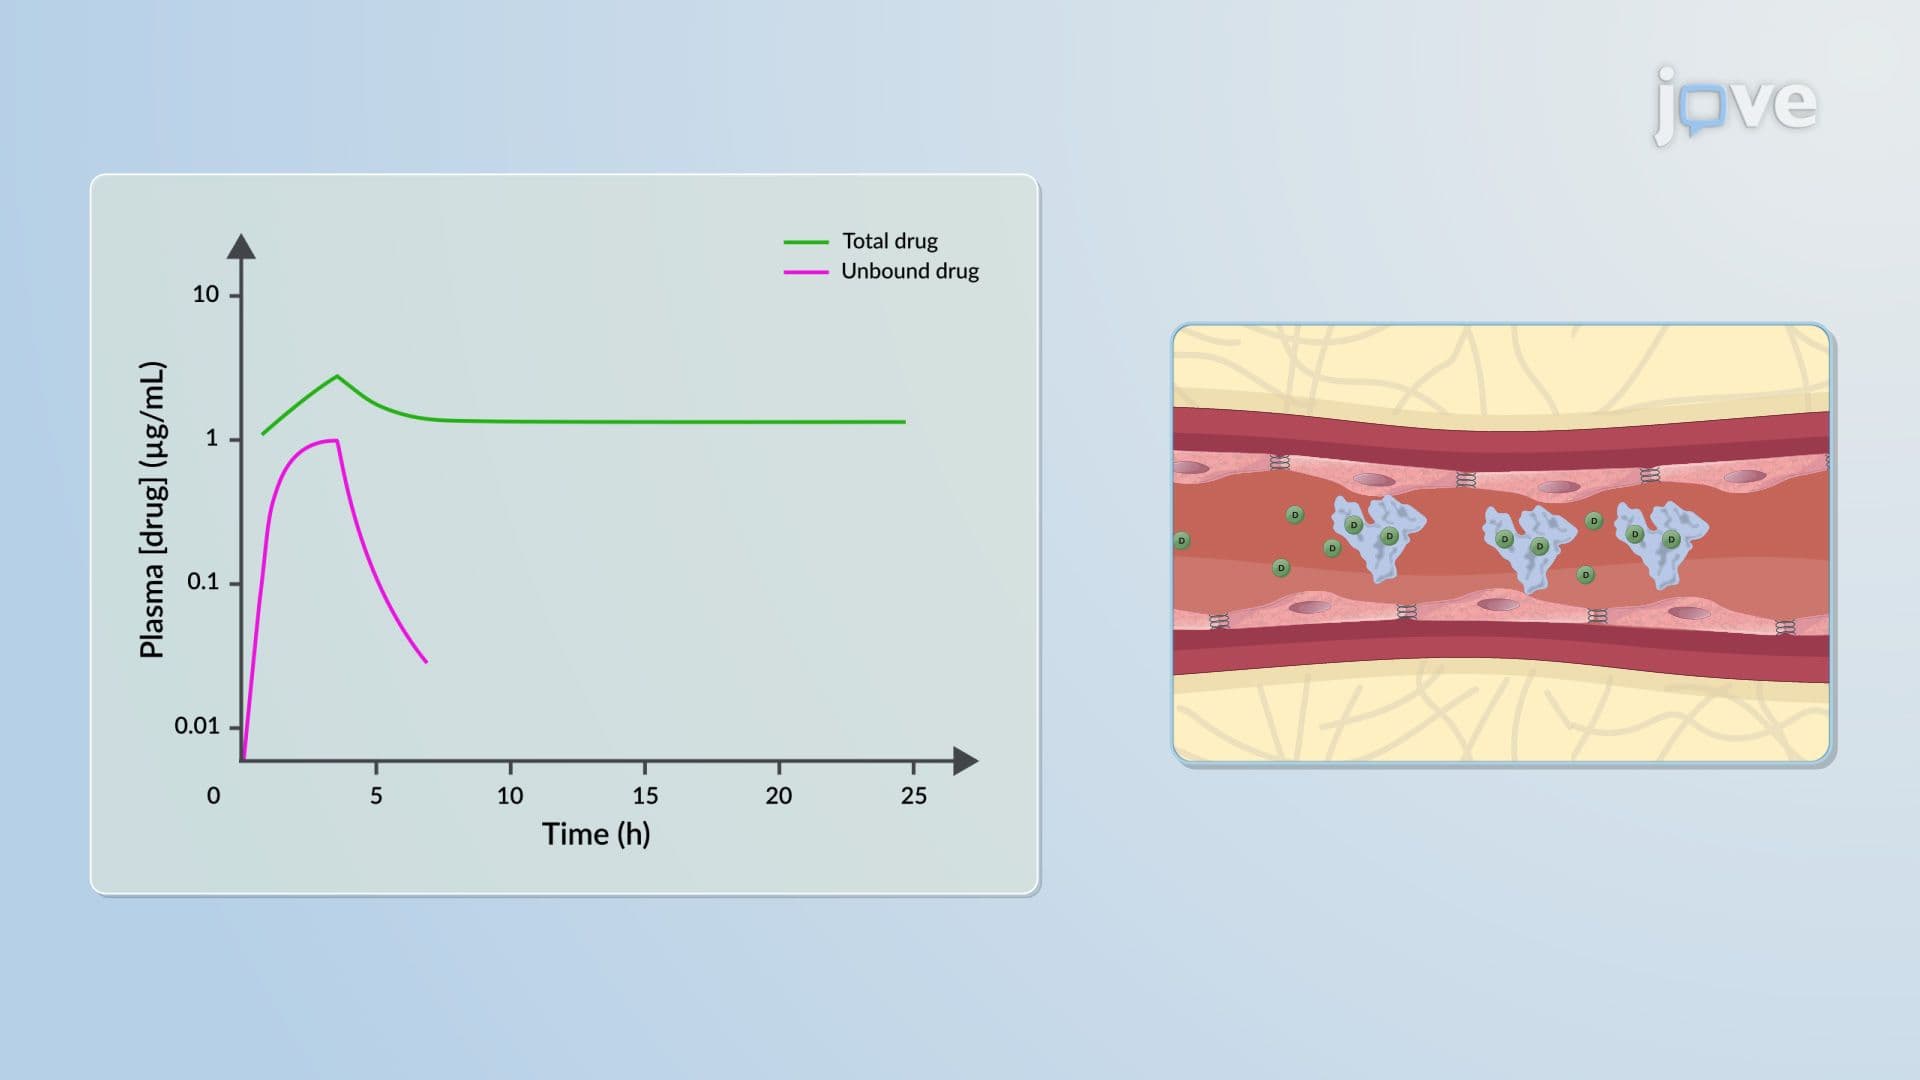
Impact of Pharmacokinetic–Pharmacodynamic Models: Regulatory Decisions
 01:24
Pharmacokinetic–Pharmacodynamic Relationship: Problems
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:20
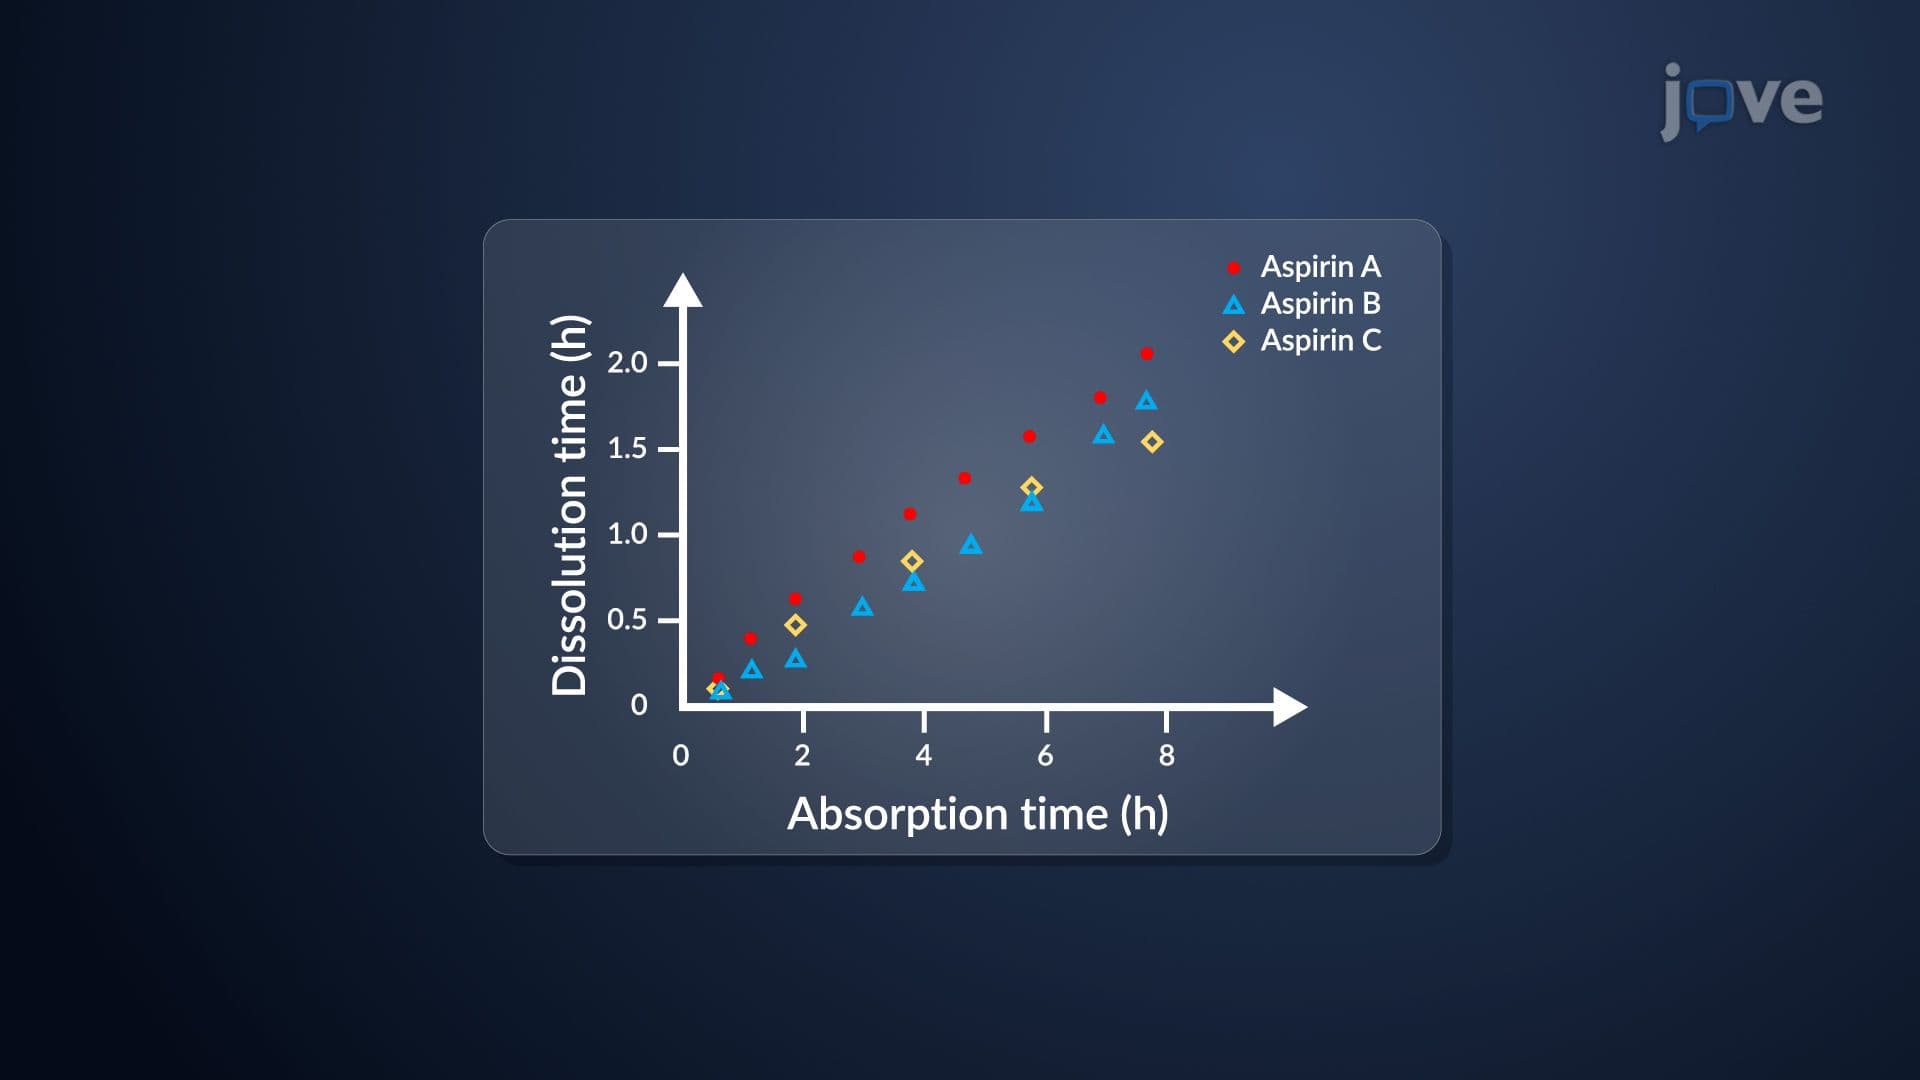
Drug Product Performance: In Vitro–In Vivo Correlation
 01:14
Pharmacokinetic–Pharmacodynamic Relationship: Model Components