Related Experiment Video
Updated: Mar 18, 2026


In Vitro Enzyme Measurement to Test Pharmacological Chaperone Responsiveness in Fabry and Pompe Disease
Published on: December 20, 2017
Expanding the molecular diversity and phenotypic spectrum of glycerol 3-phosphate dehydrogenase 1 deficiency
Carlo Dionisi-Vici1, Eyal Shteyer2, Marcello Niceta3
1Division of Metabolism, Bambino Gesù Children's Hospital, Piazza S. Onofrio 4, 00165, Rome, Italy. carlo.dionisivici@opbg.net.

Abstract:
Transient infantile hypertriglyceridemia (HTGT1; OMIM #614480) is a rare autosomal recessive disorder, which manifests in early infancy with transient hypertriglyceridemia, hepatomegaly, elevated liver enzymes, persistent fatty liver and hepatic fibrosis. This rare clinical entity is caused by inactivating mutations in the GPD1 gene, which encodes the cytosolic isoform of glycerol-3-phosphate dehydrogenase. Here we report on four patients from three unrelated families of diverse ethnic origins, who presented with hepatomegaly, liver steatosis, hypertriglyceridemia, with or without fasting ketotic hypoglycemia. Whole exome sequencing revealed the affected individuals to harbor deleterious biallelic mutations in the GPD1 gene, including the previously undescribed c.806G > A (p.Arg269Gln) and c.640T > C (p.Cys214Arg) mutations. The clinical features in three of our patients showed several differences compared to the original reports. One subject presented with recurrent episodes of fasting hypoglycemia along with hepatomegaly, hypetriglyceridemia, and elevated liver enzymes; the second showed a severe liver disease, with intrahepatic cholestasis associated with kidney involvement; finally, the third presented persistent hypertriglyceridemia at the age of 30 years. These findings expand the current knowledge of this rare disorder, both with regard to the phenotype and molecular basis. The enlarged phenotypic spectrum of glycerol-3-phosphate dehydrogenase 1 deficiency can mimic other inborn errors of metabolism with liver involvement and should alert clinicians to recognize this entity by considering GPD1 mutations in appropriate clinical settings.
More Related Videos
Related Concept Videos
 01:24
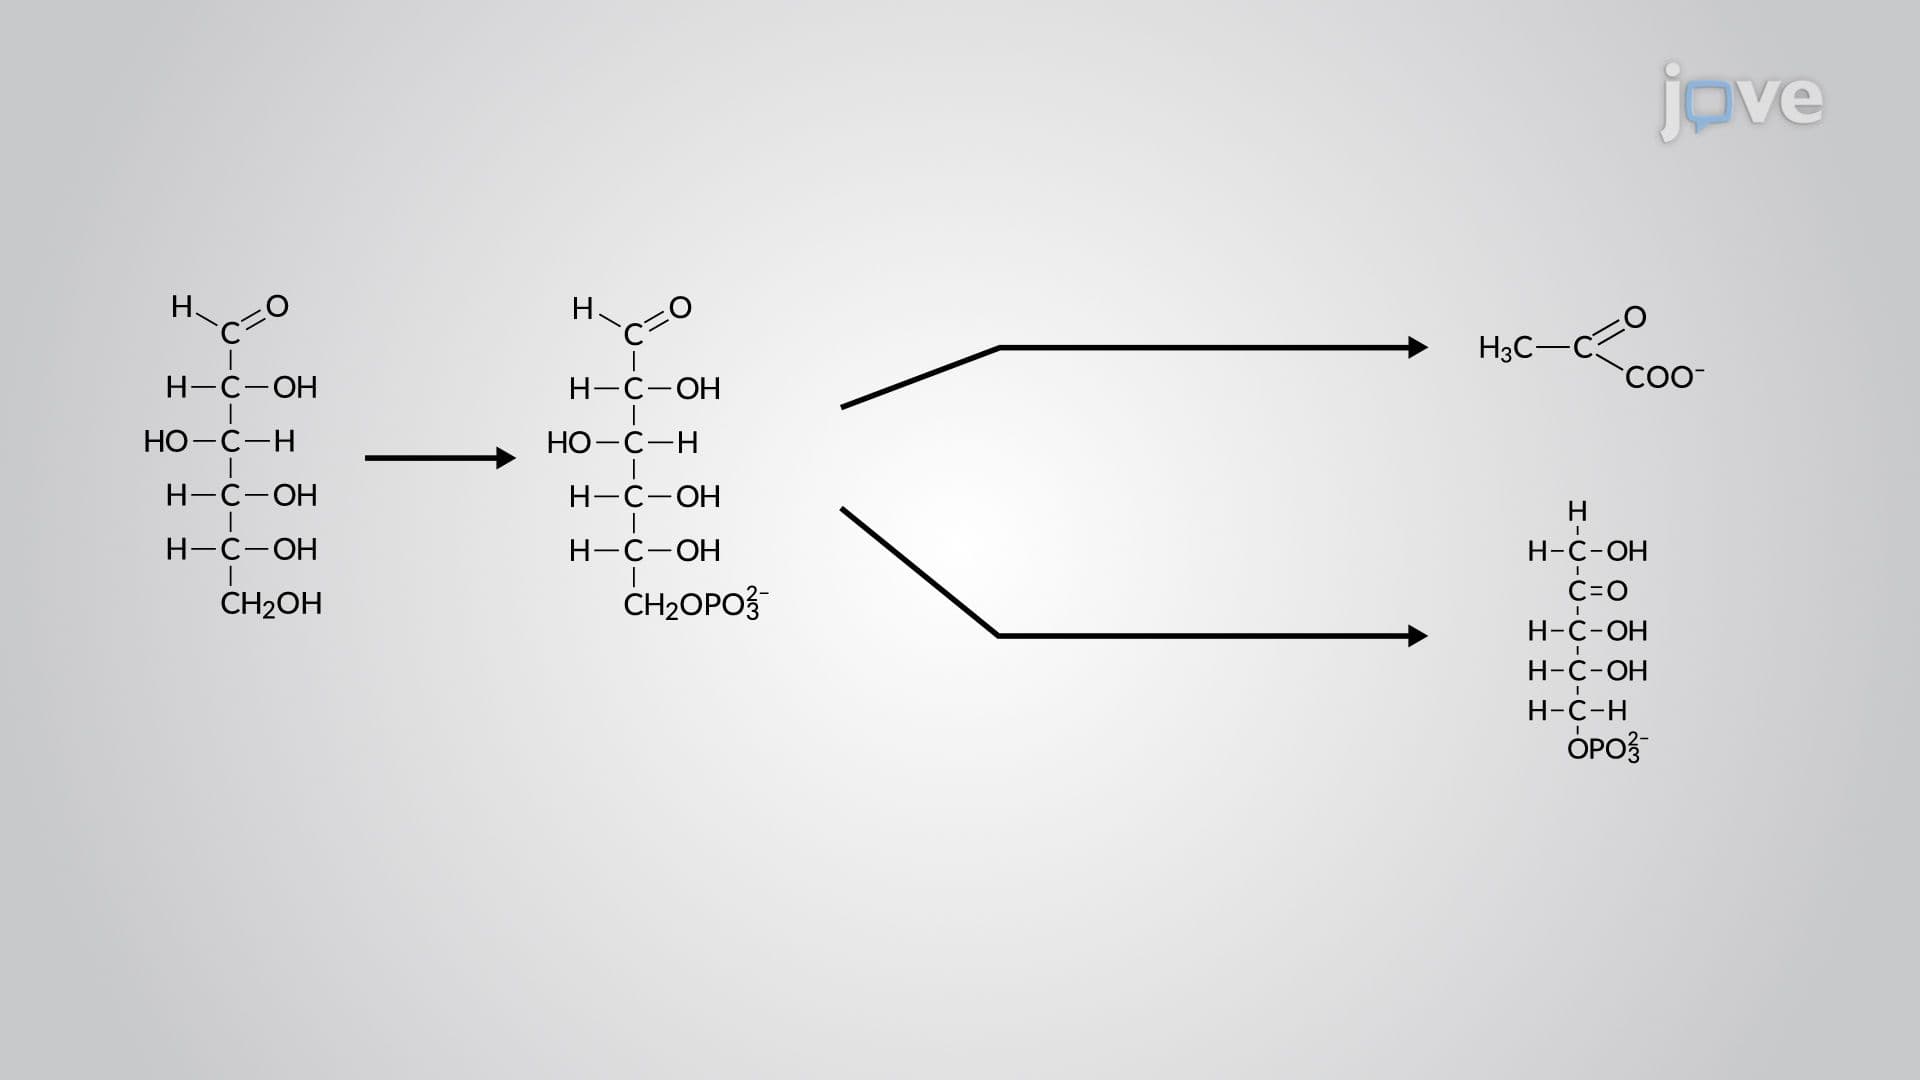
Other Glycolytic Pathways
 01:20
Inborn Errors of Metabolism
 01:22
Lysosomal Hydrolases
 01:27
Glucose Transporters
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes:
 01:29
Pharmacogenomics: Identification of New Drug Targets
 02:32
Exon Recombination
Exon shuffling follows “splice frame rules.” Each exon...