Related Experiment Video For Arrhythmia
Updated: Mar 18, 2026
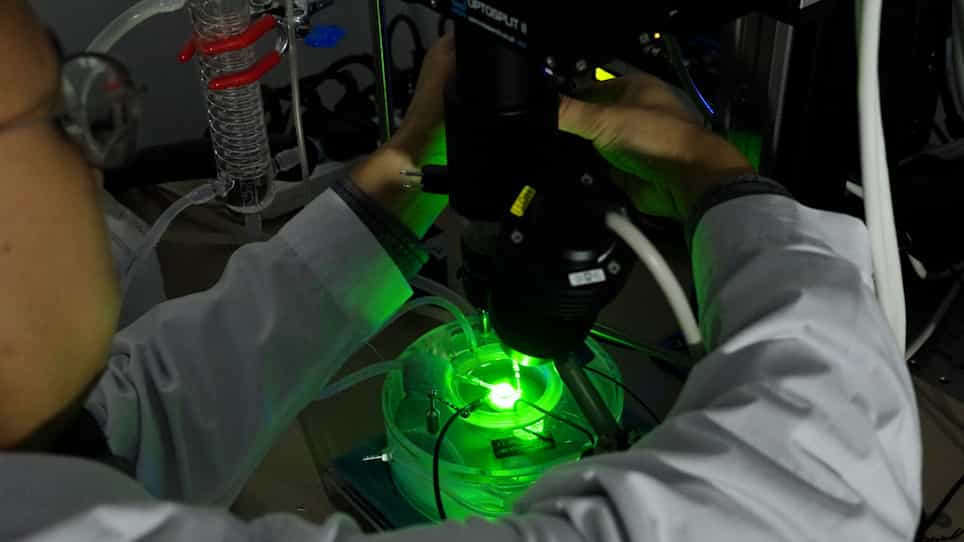

Dual-Dye Optical Mapping of Hearts from RyR2R2474S Knock-In Mice of Catecholaminergic Polymorphic Ventricular Tachycardia
Published on: December 22, 2023
Novel calmodulin mutations associated with congenital long QT syndrome affect calcium current in human cardiomyocytes
Daniel C Pipilas1, Christopher N Johnson2, Gregory Webster3
1Department of Pharmacology, Northwestern University Feinberg School of Medicine, Chicago, Illinois.

Background:
Calmodulin (CaM) mutations are associated with cardiac arrhythmia susceptibility including congenital long QT syndrome (LQTS).
Objective:
The purpose of this study was to determine the clinical, genetic, and functional features of 2 novel CaM mutations in children with life-threatening ventricular arrhythmias.
Methods:
The clinical and genetic features of 2 congenital arrhythmia cases associated with 2 novel CaM gene mutations were ascertained. Biochemical and functional investigations were conducted on the 2 mutations.
Results:
A novel de novo CALM2 mutation (D132H) was discovered by candidate gene screening in a male infant with prenatal bradycardia born to healthy parents. Postnatal course was complicated by profound bradycardia, prolonged corrected QT interval (651 ms), 2:1 atrioventricular block, and cardiogenic shock. He was resuscitated and was treated with a cardiac device. A second novel de novo mutation in CALM1 (D132V) was discovered by clinical exome sequencing in a 3-year-old boy who suffered a witnessed cardiac arrest secondary to ventricular fibrillation. Electrocardiographic recording after successful resuscitation revealed a prolonged corrected QT interval of 574 ms. The Ca(2+) affinity of CaM-D132H and CaM-D132V revealed extremely weak binding to the C-terminal domain, with significant structural perturbations noted for D132H. Voltage-clamp recordings of human induced pluripotent stem cell-derived cardiomyocytes transiently expressing wild-type or mutant CaM demonstrated that both mutations caused impaired Ca(2+)-dependent inactivation of voltage-gated Ca(2+) current. Neither mutant affected voltage-dependent inactivation.
Conclusion:
Our findings implicate impaired Ca(2+)-dependent inactivation in human cardiomyocytes as the plausible mechanism for long QT syndrome associated with 2 novel CaM mutations. The data further expand the spectrum of genotype and phenotype associated with calmodulinopathy.
Related Concept Videos
 01:16
Calmodulin-dependent Signaling
The Ca2+-CaM complex does not have enzymatic activity by itself. Instead, the complex binds downstream target proteins, including membrane proteins or enzymes,...
 01:28
Mechanism of Cardiac Arrhythmias
 01:29
Cardiomyopathy III: Hypertrophic Cardiomyopathy
 01:18
Antihypertensive Drugs: Action of Calcium Channel Blockers
 01:26
Voltage-gated Ion Channels
Generally, all voltage-gated ion channels have a 'voltage-sensing domain' that spans the lipid bilayer. The charged residues in the sensor move in response to the membrane potential changes that open the channel allowing ions movement. There are several types of...