Related Experiment Video
Updated: Mar 17, 2026

Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
Benchmark fragment-based (1)H, (13)C, (15)N and (17)O chemical shift predictions in molecular crystals
Joshua D Hartman1, Ryan A Kudla, Graeme M Day
1Department of Chemistry, University of California, Riverside, California 92521, USA. gregory.beran@ucr.edu.

Abstract:
The performance of fragment-based ab initio(1)H, (13)C, (15)N and (17)O chemical shift predictions is assessed against experimental NMR chemical shift data in four benchmark sets of molecular crystals. Employing a variety of commonly used density functionals (PBE0, B3LYP, TPSSh, OPBE, PBE, TPSS), we explore the relative performance of cluster, two-body fragment, and combined cluster/fragment models. The hybrid density functionals (PBE0, B3LYP and TPSSh) generally out-perform their generalized gradient approximation (GGA)-based counterparts. (1)H, (13)C, (15)N, and (17)O isotropic chemical shifts can be predicted with root-mean-square errors of 0.3, 1.5, 4.2, and 9.8 ppm, respectively, using a computationally inexpensive electrostatically embedded two-body PBE0 fragment model. Oxygen chemical shieldings prove particularly sensitive to local many-body effects, and using a combined cluster/fragment model instead of the simple two-body fragment model decreases the root-mean-square errors to 7.6 ppm. These fragment-based model errors compare favorably with GIPAW PBE ones of 0.4, 2.2, 5.4, and 7.2 ppm for the same (1)H, (13)C, (15)N, and (17)O test sets. Using these benchmark calculations, a set of recommended linear regression parameters for mapping between calculated chemical shieldings and observed chemical shifts are provided and their robustness assessed using statistical cross-validation. We demonstrate the utility of these approaches and the reported scaling parameters on applications to 9-tert-butyl anthracene, several histidine co-crystals, benzoic acid and the C-nitrosoarene SnCl2(CH3)2(NODMA)2.
More Related Videos
Related Concept Videos
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:15
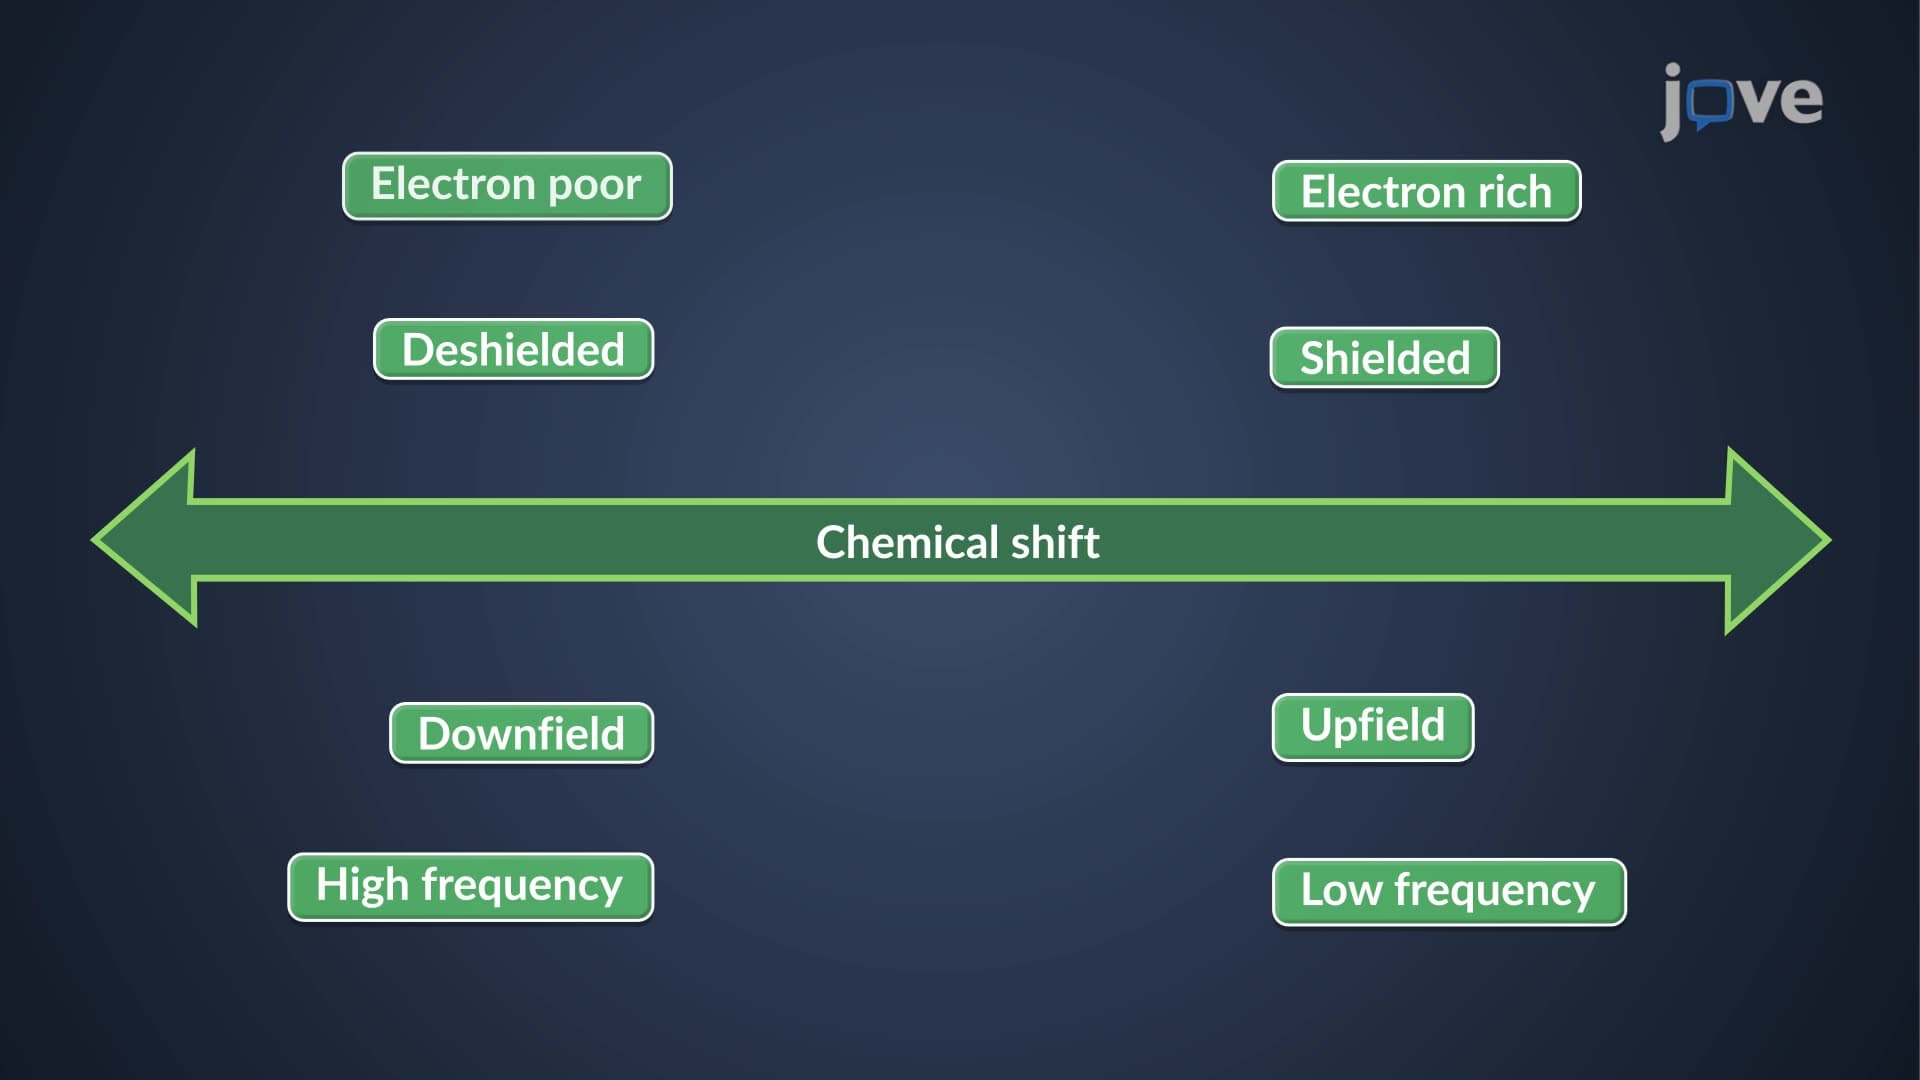
NMR Spectroscopy: Chemical Shift Overview
For instance, the proton...
 01:27
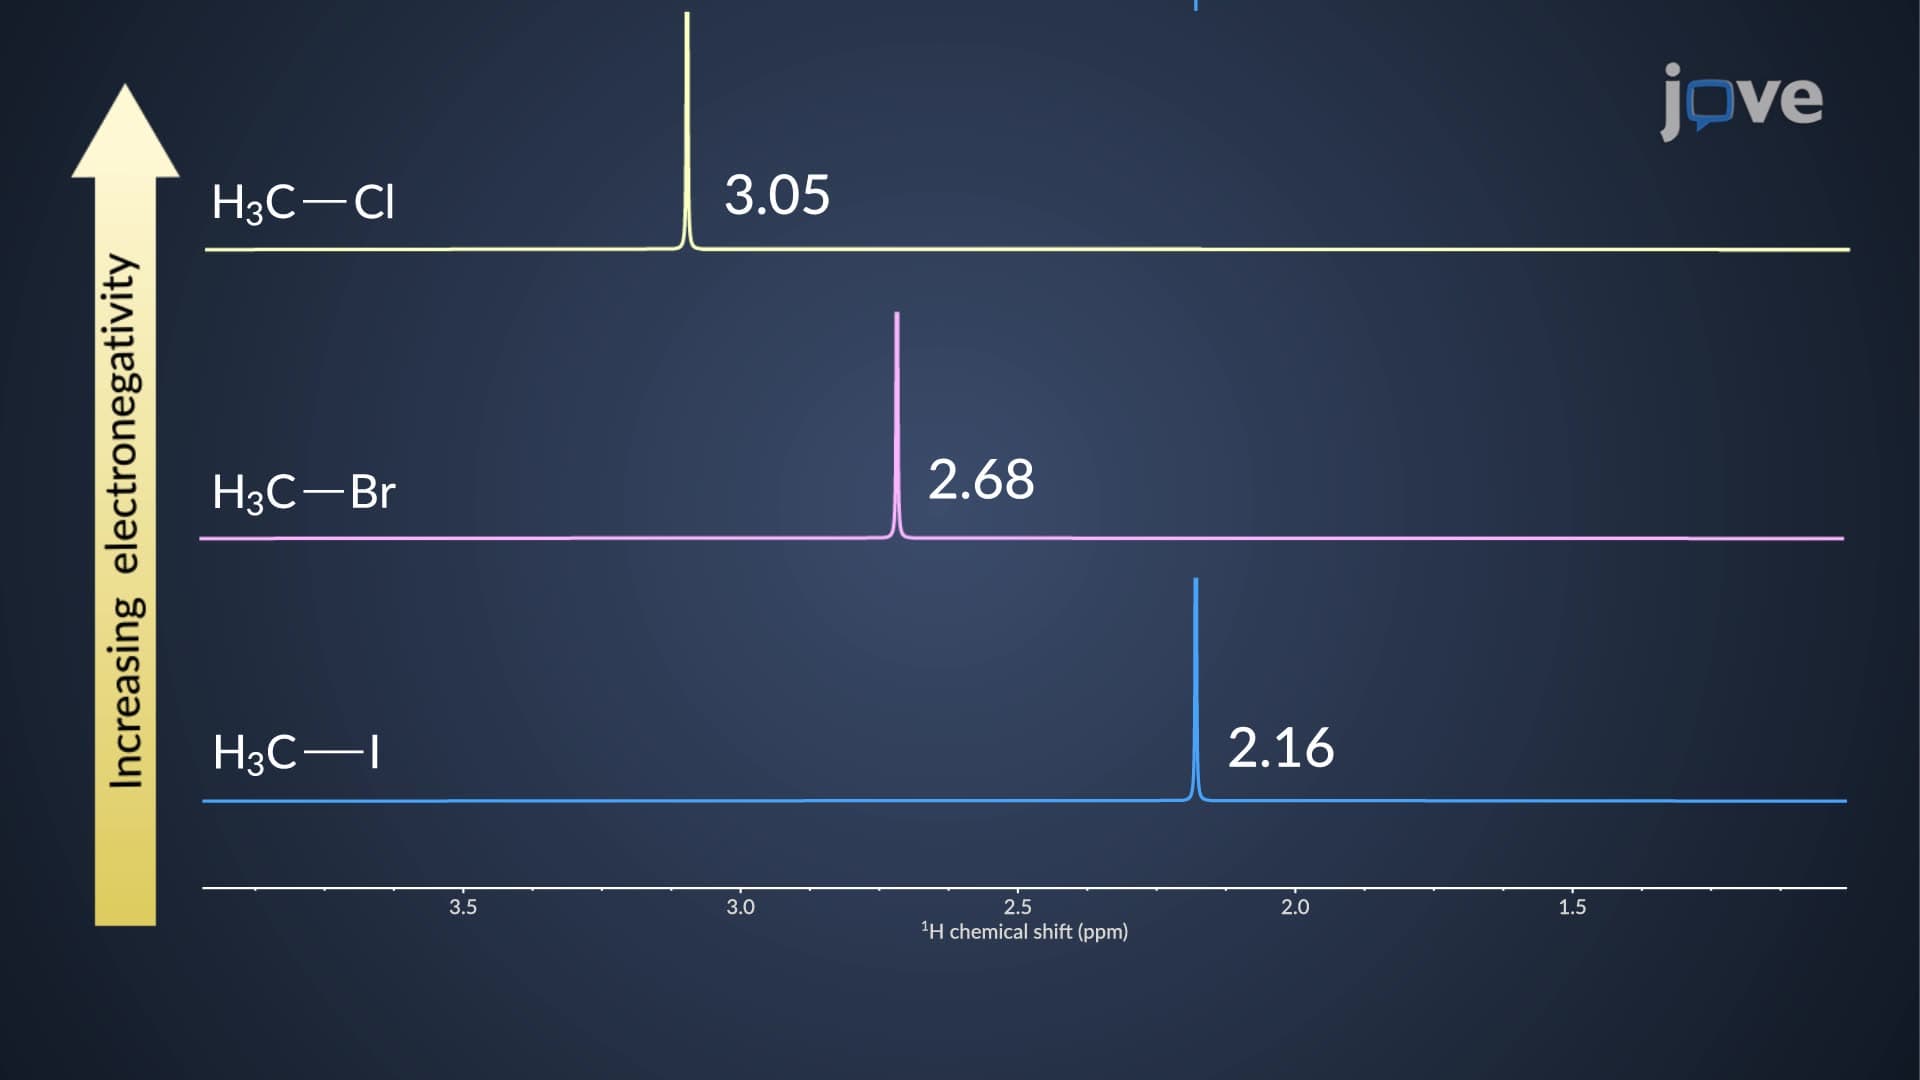
Inductive Effects on Chemical Shift: Overview
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 01:29
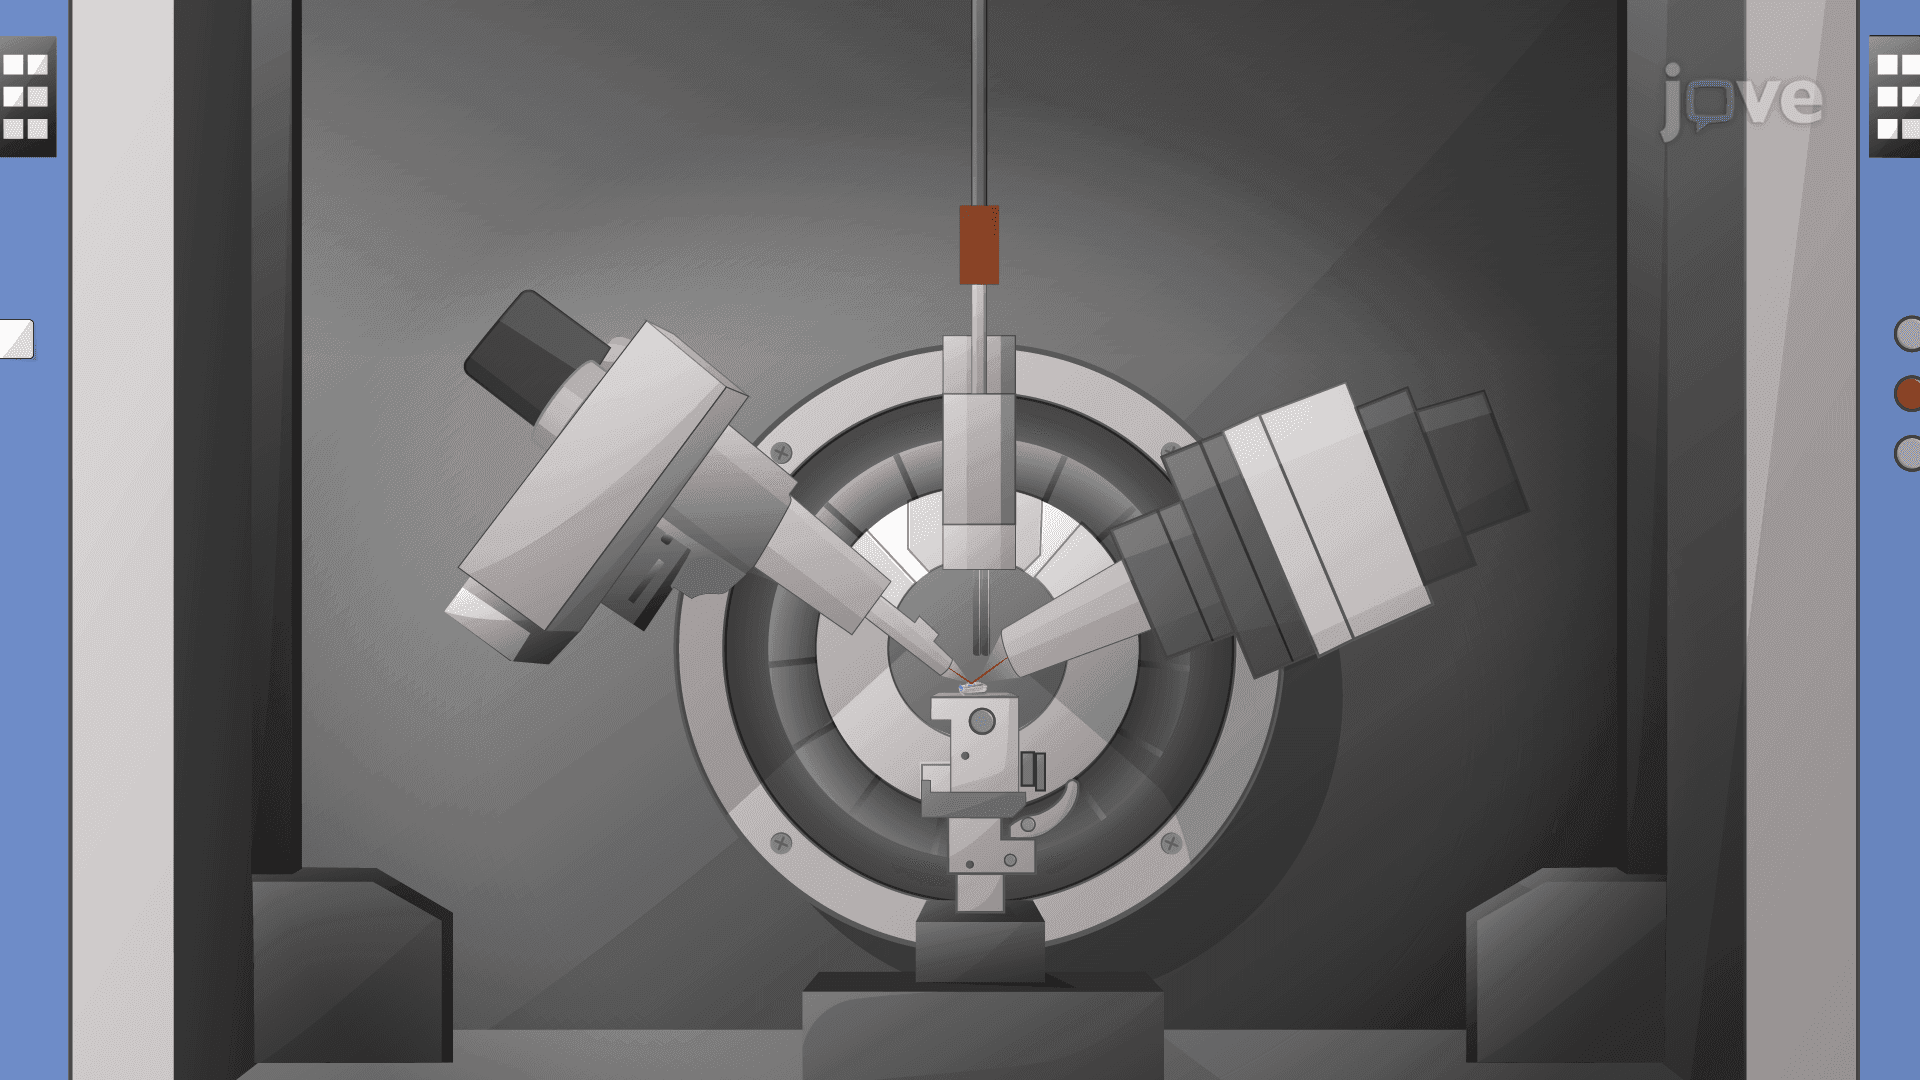
Determination of Crystal Structures
 01:03
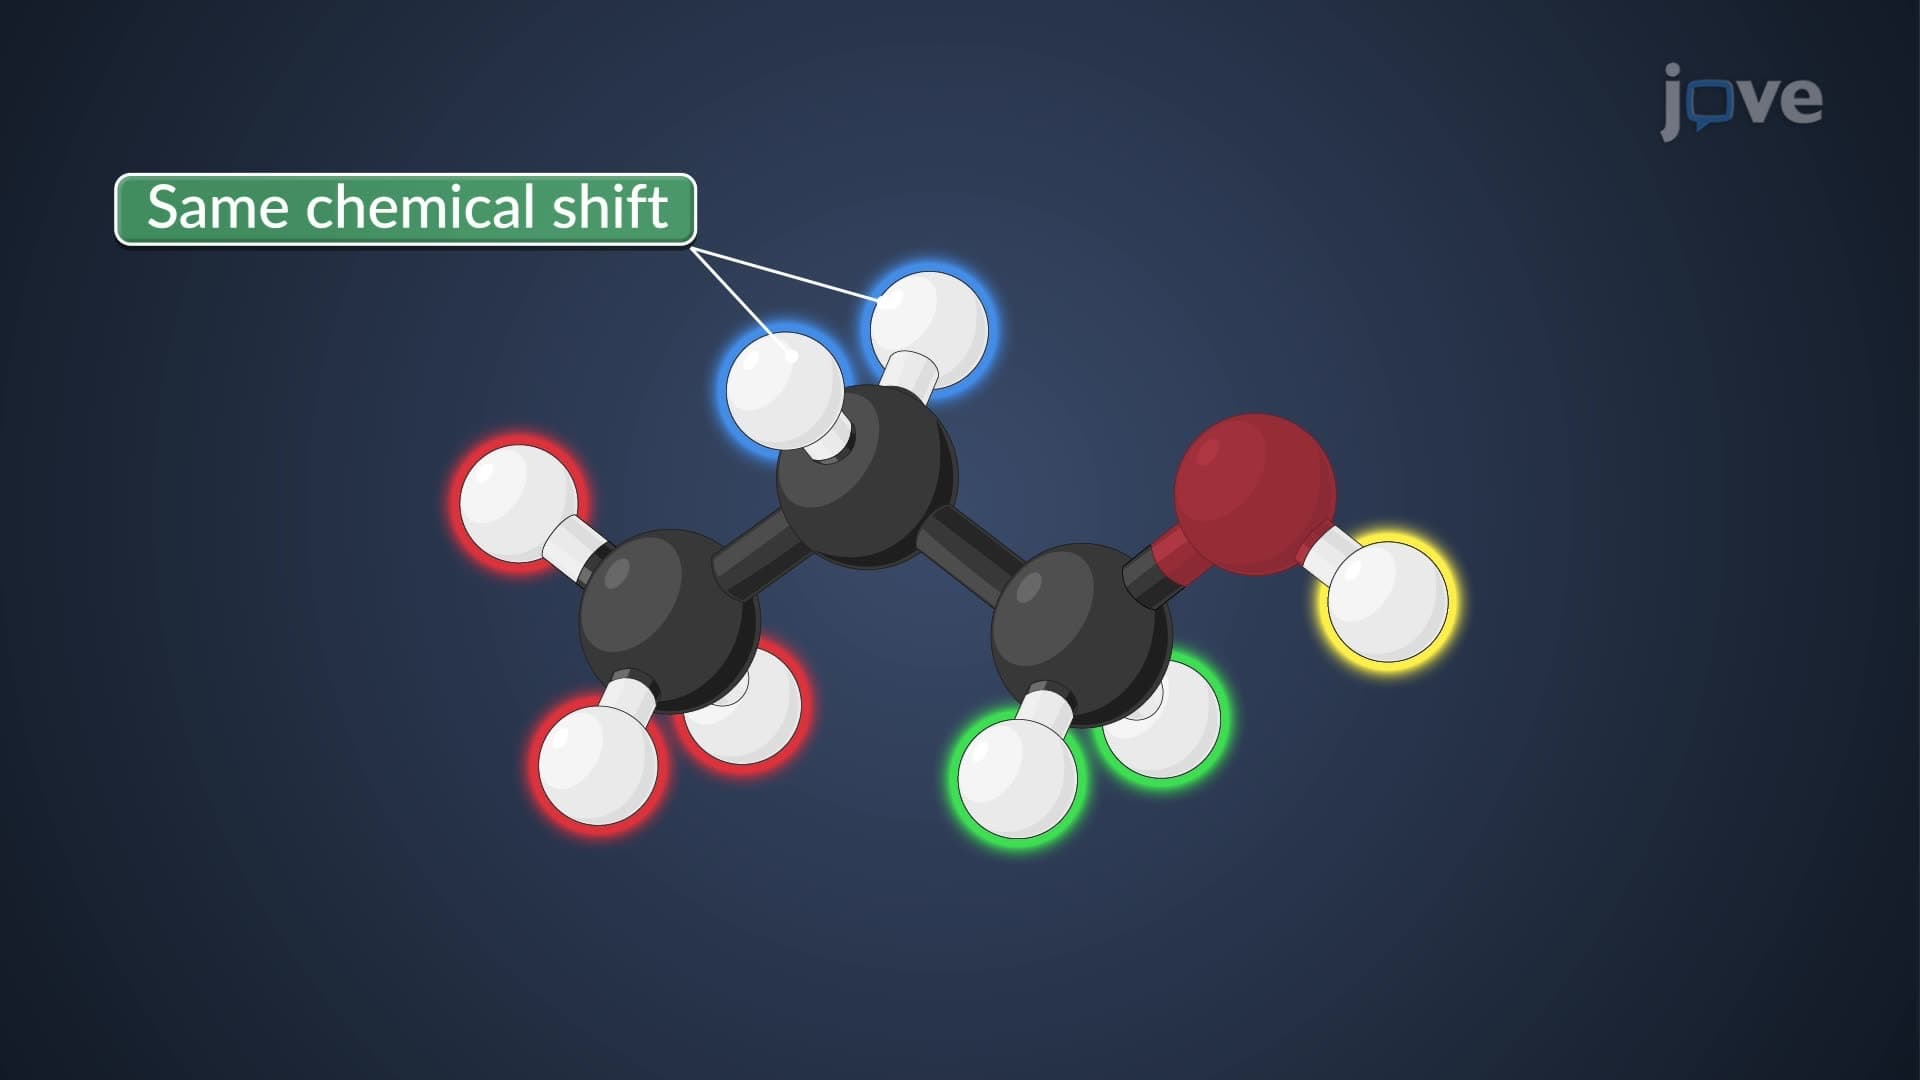
¹H NMR Chemical Shift Equivalence: Homotopic and Heterotopic Protons