Calculating protein-ligand binding affinities with MMPBSA: Method and error analysis
Related Experiment Video
Updated: Mar 16, 2026
Protein WISDOM: A Workbench for In silico De novo Design of BioMolecules
Published on: July 25, 2013
Changhao Wang1,2,3, Peter H Nguyen2, Kevin Pham2
1Chemical and Materials Physics Graduate Program, Irvine, California, 92697.

Abstract:
Molecular Mechanics Poisson-Boltzmann Surface Area (MMPBSA) methods have become widely adopted in estimating protein-ligand binding affinities due to their efficiency and high correlation with experiment. Here different computational alternatives were investigated to assess their impact to the agreement of MMPBSA calculations with experiment. Seven receptor families with both high-quality crystal structures and binding affinities were selected. First the performance of nonpolar solvation models was studied and it was found that the modern approach that separately models hydrophobic and dispersion interactions dramatically reduces RMSD's of computed relative binding affinities. The numerical setup of the Poisson-Boltzmann methods was analyzed next. The data shows that the impact of grid spacing to the quality of MMPBSA calculations is small: the numerical error at the grid spacing of 0.5 Å is already small enough to be negligible. The impact of different atomic radius sets and different molecular surface definitions was further analyzed and weak influences were found on the agreement with experiment. The influence of solute dielectric constant was also analyzed: a higher dielectric constant generally improves the overall agreement with experiment, especially for highly charged binding pockets. The data also showed that the converged simulations caused slight reduction in the agreement with experiment. Finally the direction of estimating absolute binding free energies was briefly explored. Upon correction of the binding-induced rearrangement free energy and the binding entropy lost, the errors in absolute binding affinities were also reduced dramatically when the modern nonpolar solvent model was used, although further developments were apparently necessary to further improve the MMPBSA methods. © 2016 Wiley Periodicals, Inc.
More Related Videos
 06:50
Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 02:18
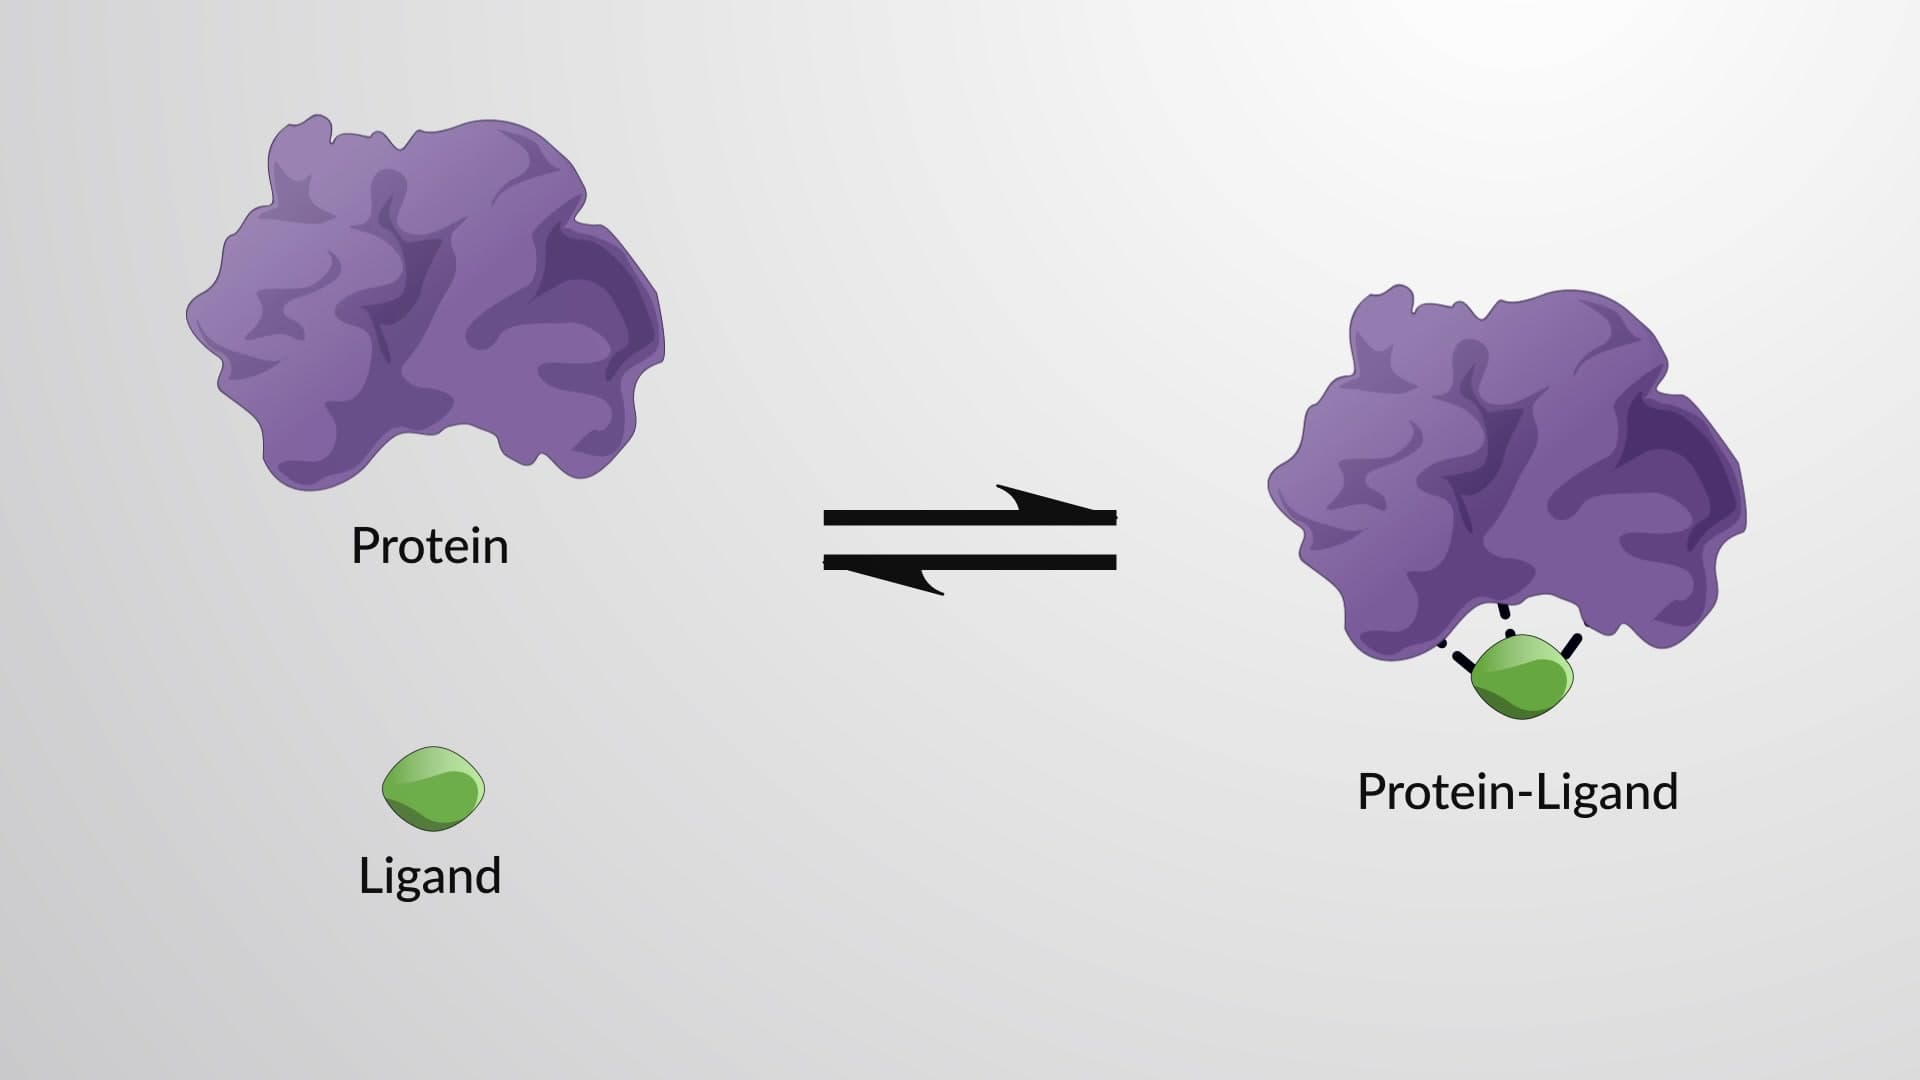
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:04
Protein-protein Interfaces