Related Experiment Video
Updated: Mar 16, 2026
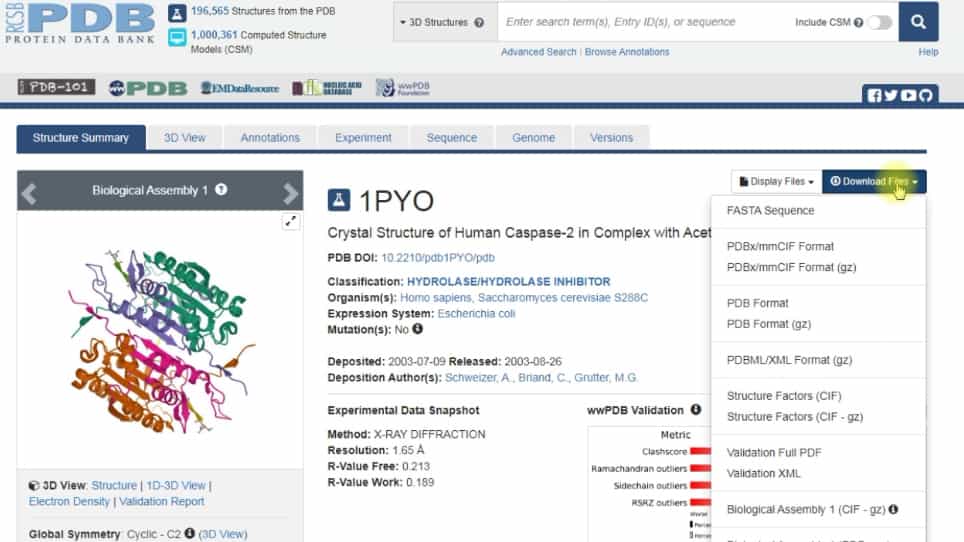
Exploring Caspase Mutations and Post-Translational Modification by Molecular Modeling Approaches
Published on: October 13, 2022
Efficient Atomistic Simulation of Pathways and Calculation of Rate Constants for a Protein-Peptide Binding Process:
Matthew C Zwier1, Adam J Pratt2, Joshua L Adelman3
1Department of Chemistry, Drake University , Des Moines, Iowa 50311, United States.

Abstract:
The characterization of protein binding processes - with all of the key conformational changes - has been a grand challenge in the field of biophysics. Here, we have used the weighted ensemble path sampling strategy to orchestrate molecular dynamics simulations, yielding atomistic views of protein-peptide binding pathways involving the MDM2 oncoprotein and an intrinsically disordered p53 peptide. A total of 182 independent, continuous binding pathways were generated, yielding a kon that is in good agreement with experiment. These pathways were generated in 15 days using 3500 cores of a supercomputer, substantially faster than would be possible with "brute force" simulations. Many of these pathways involve the anchoring of p53 residue F19 into the MDM2 binding cleft when forming the metastable encounter complex, indicating that F19 may be a kinetically important residue. Our study demonstrates that it is now practical to generate pathways and calculate rate constants for protein binding processes using atomistic simulation on typical computing resources.
More Related Videos
Related Concept Videos
 02:18
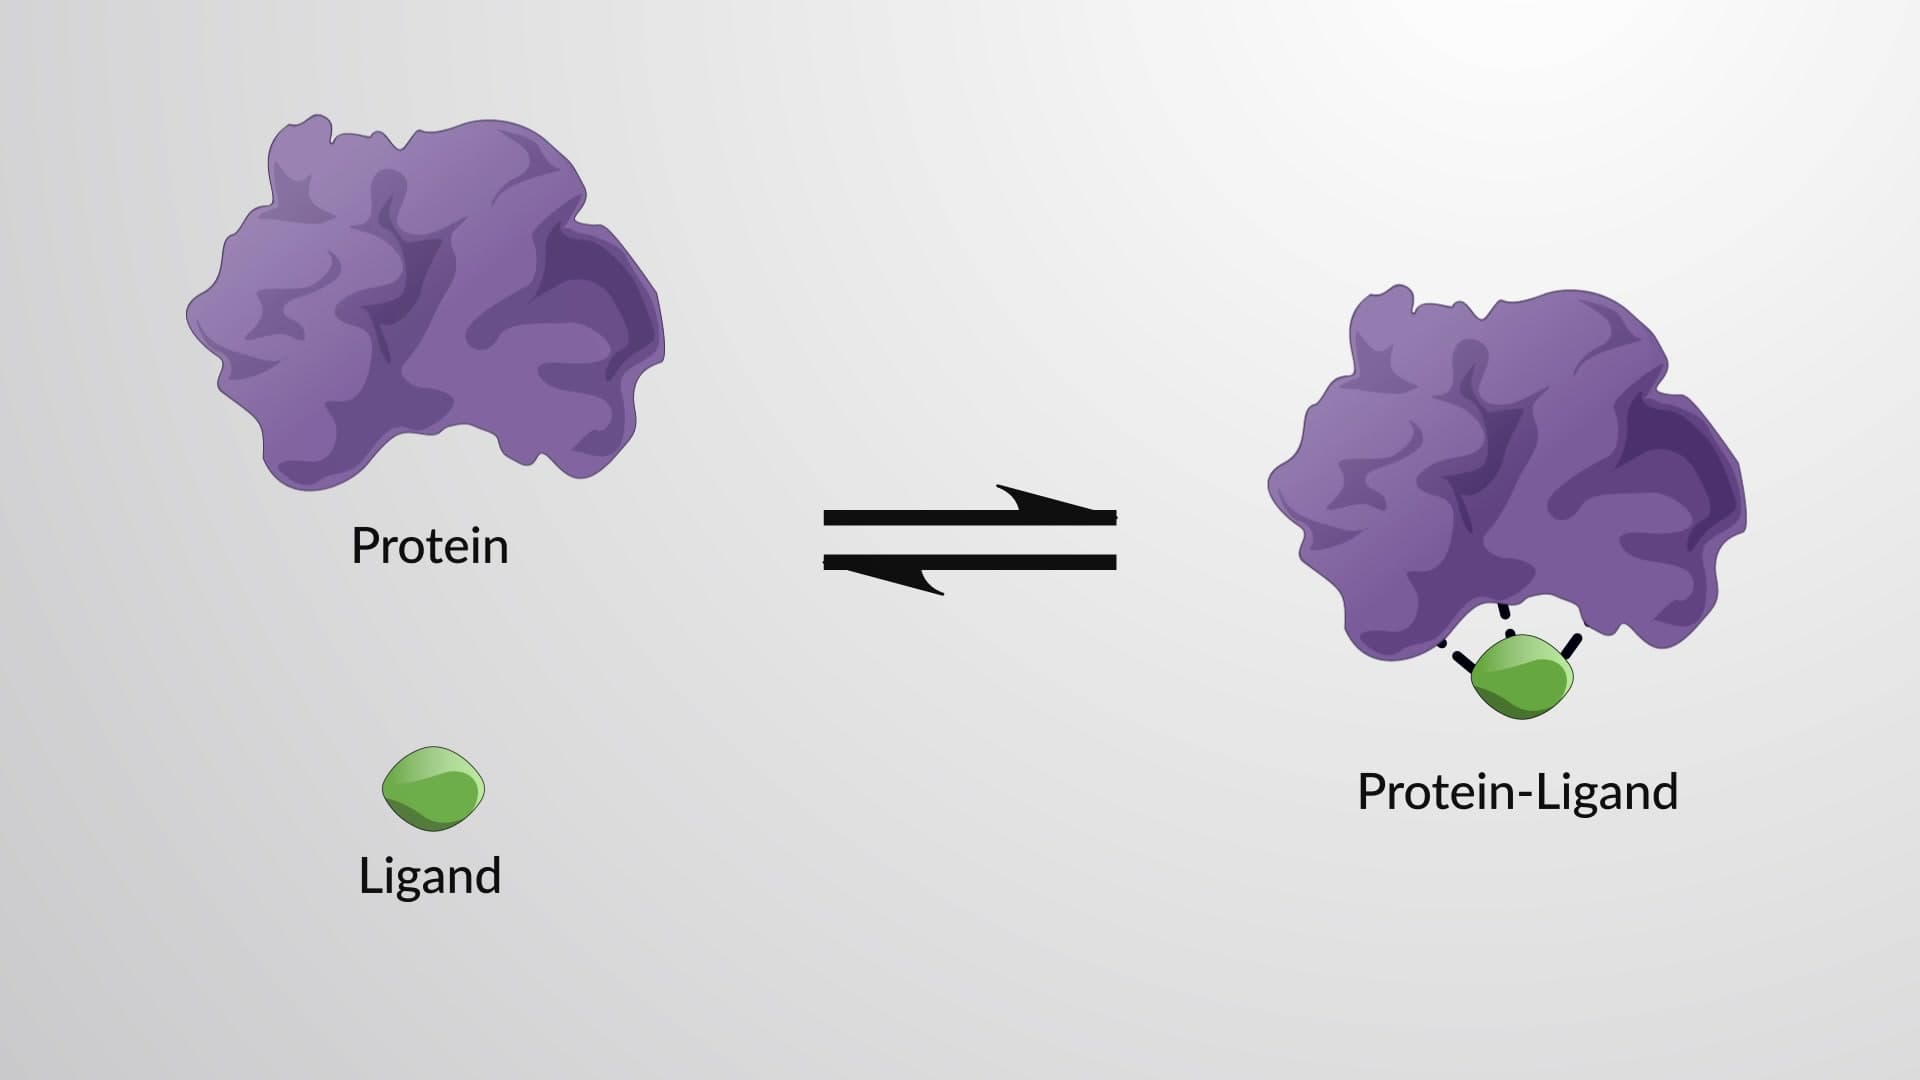
The Equilibrium Binding Constant and Binding Strength
 02:04
Covalently Linked Protein Regulators
These groups modify specific amino acids in a protein....
 02:04
Protein-protein Interfaces