
Development of Xe and Kr empirical potentials for CeO2, ThO2, UO2 and PuO2, combining DFT with high temperature MD
M W D Cooper1, N Kuganathan2, P A Burr3
1Materials Science and Technology Division, Los Alamos National Laboratory, PO Box 1663, Los Alamos, NM 87545, USA.

Abstract:
The development of embedded atom method (EAM) many-body potentials for actinide oxides and associated mixed oxide (MOX) systems has motivated the development of a complementary parameter set for gas-actinide and gas-oxygen interactions. A comprehensive set of density functional theory (DFT) calculations were used to study Xe and Kr incorporation at a number of sites in CeO2, ThO2, UO2 and PuO2. These structures were used to fit a potential, which was used to generate molecular dynamics (MD) configurations incorporating Xe and Kr at 300 K, 1500 K, 3000 K and 5000 K. Subsequent matching to the forces predicted by DFT for these MD configurations was used to refine the potential set. This fitting approach ensured weighted fitting to configurations that are thermodynamically significant over a broad temperature range, while avoiding computationally expensive DFT-MD calculations. The resultant gas potentials were validated against DFT trapping energies and are suitable for simulating combinations of Xe and Kr in solid solutions of CeO2, ThO2, UO2 and PuO2, providing a powerful tool for the atomistic simulation of conventional nuclear reactor fuel UO2 as well as advanced MOX fuels.
More Related Videos
 10:42
Combining Solid-state and Solution-based Techniques: Synthesis and Reactivity of ChalcogenidoplumbatesII or IV
Published on: December 29, 2016
 07:24
Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Electron Configurations
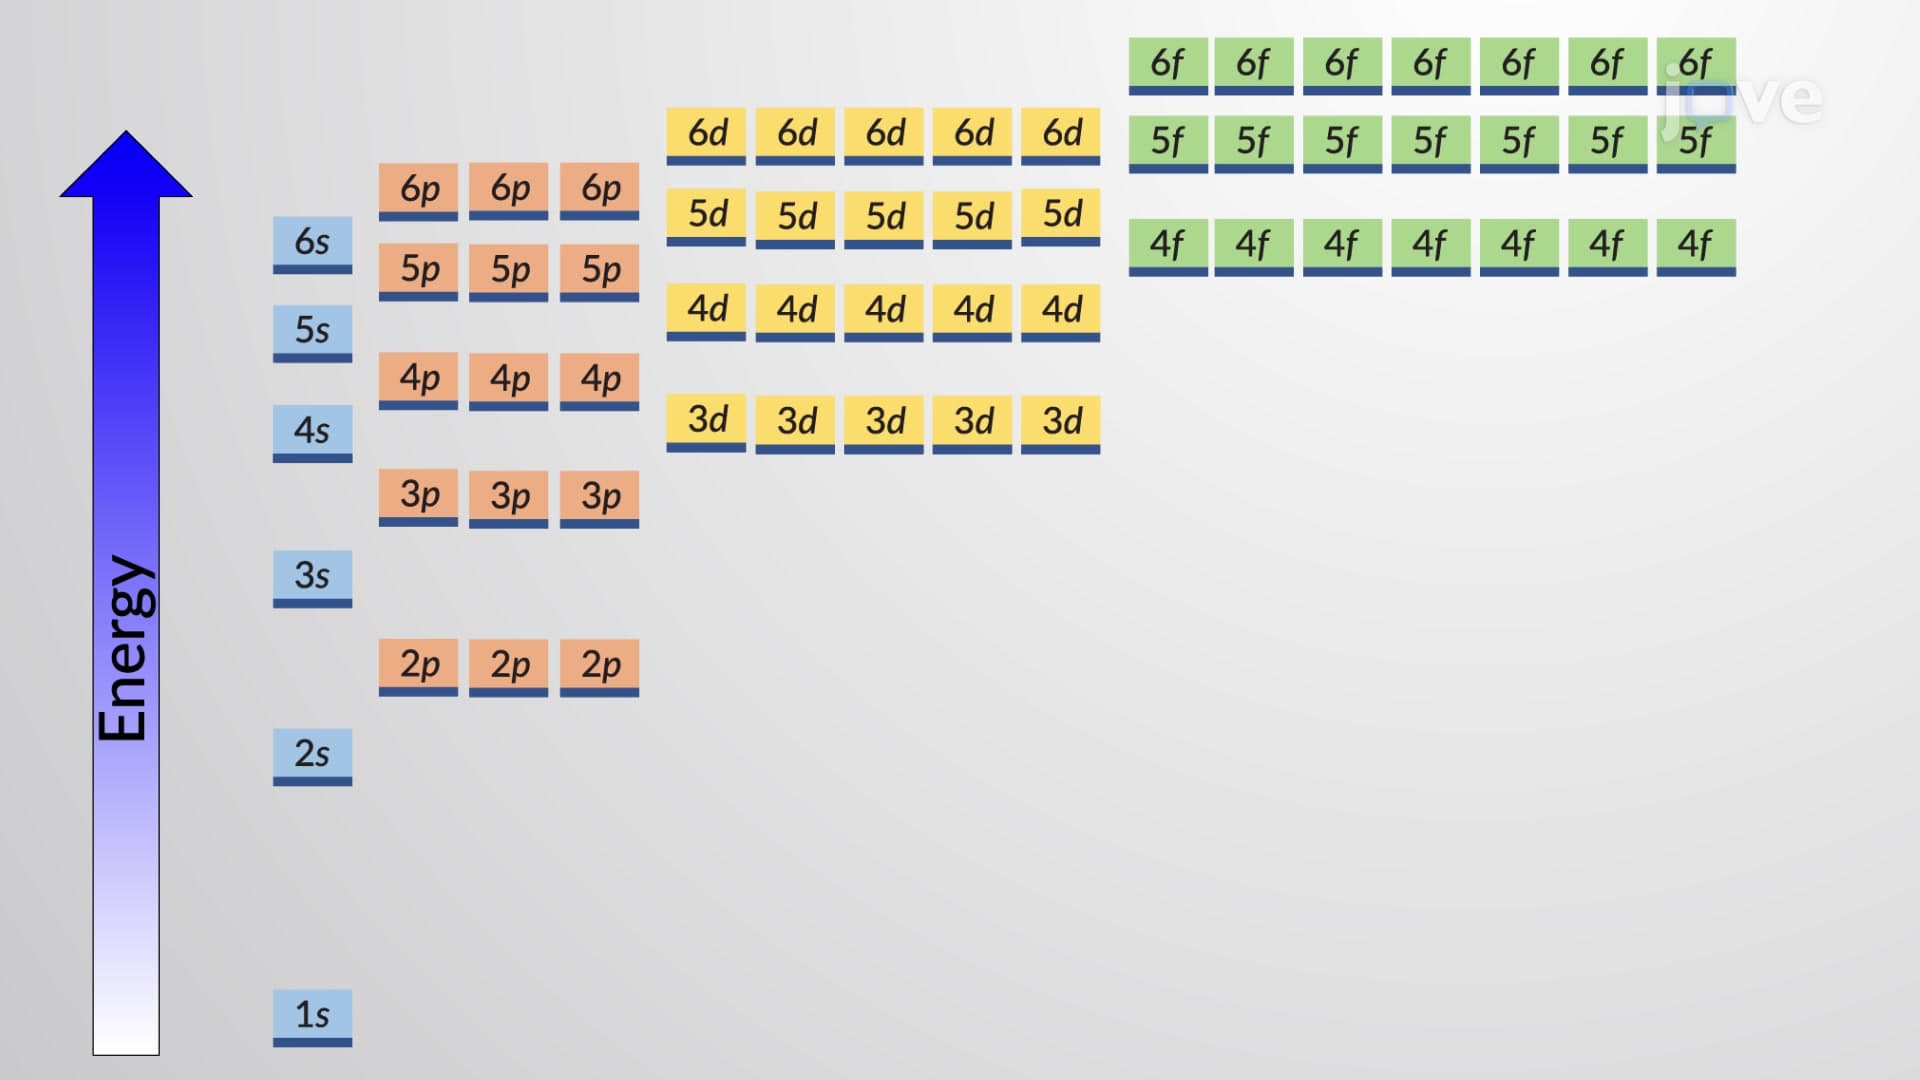
The relative energies of the subshells determine the order in which atomic orbitals are filled (1s, 2s, 2p, 3s, 3p,...
 03:51
Molecular Orbital Theory II
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:26
Electron Configuration of Multielectron Atoms