Related Experiment Video
Updated: Mar 16, 2026
Author Spotlight: Impact of Intergenic Interactions on Disease-Identifying Dark Biomarkers
Published on: March 1, 2024
Interpreting transcriptional changes using causal graphs: new methods and their practical utility on public networks
Carl Tony Fakhry1, Parul Choudhary2, Alex Gutteridge2
1Department of Computer Science, University of Massachusetts Boston, 100 Morrissey Boulevard, Boston, 02125, USA.

Background:
Inference of active regulatory cascades under specific molecular and environmental perturbations is a recurring task in transcriptional data analysis. Commercial tools based on large, manually curated networks of causal relationships offering such functionality have been used in thousands of articles in the biomedical literature. The adoption and extension of such methods in the academic community has been hampered by the lack of freely available, efficient algorithms and an accompanying demonstration of their applicability using current public networks.
Results:
In this article, we propose a new statistical method that will infer likely upstream regulators based on observed patterns of up- and down-regulated transcripts. The method is suitable for use with public interaction networks with a mix of signed and unsigned causal edges. It subsumes and extends two previously published approaches and we provide a novel algorithmic method for efficient statistical inference. Notably, we demonstrate the feasibility of using the approach to generate biological insights given current public networks in the context of controlled in-vitro overexpression experiments, stem-cell differentiation data and animal disease models. We also provide an efficient implementation of our method in the R package QuaternaryProd available to download from Bioconductor.
Conclusions:
In this work, we have closed an important gap in utilizing causal networks to analyze differentially expressed genes. Our proposed Quaternary test statistic incorporates all available evidence on the potential relevance of an upstream regulator. The new approach broadens the use of these types of statistics for highly curated signed networks in which ambiguities arise but also enables the use of networks with unsigned edges. We design and implement a novel computational method that can efficiently estimate p-values for upstream regulators in current biological settings. We demonstrate the ready applicability of the implemented method to analyze differentially expressed genes using the publicly available networks.
More Related Videos
 09:58
Mapping the Structure-Function Relationships of Disordered Oncogenic Transcription Factors Using Transcriptomic Analysis
Published on: June 27, 2020
 12:54
Real-time Analysis of Transcription Factor Binding, Transcription, Translation, and Turnover to Display Global Events During Cellular Activation
Published on: March 7, 2018
Related Concept Videos
 01:30
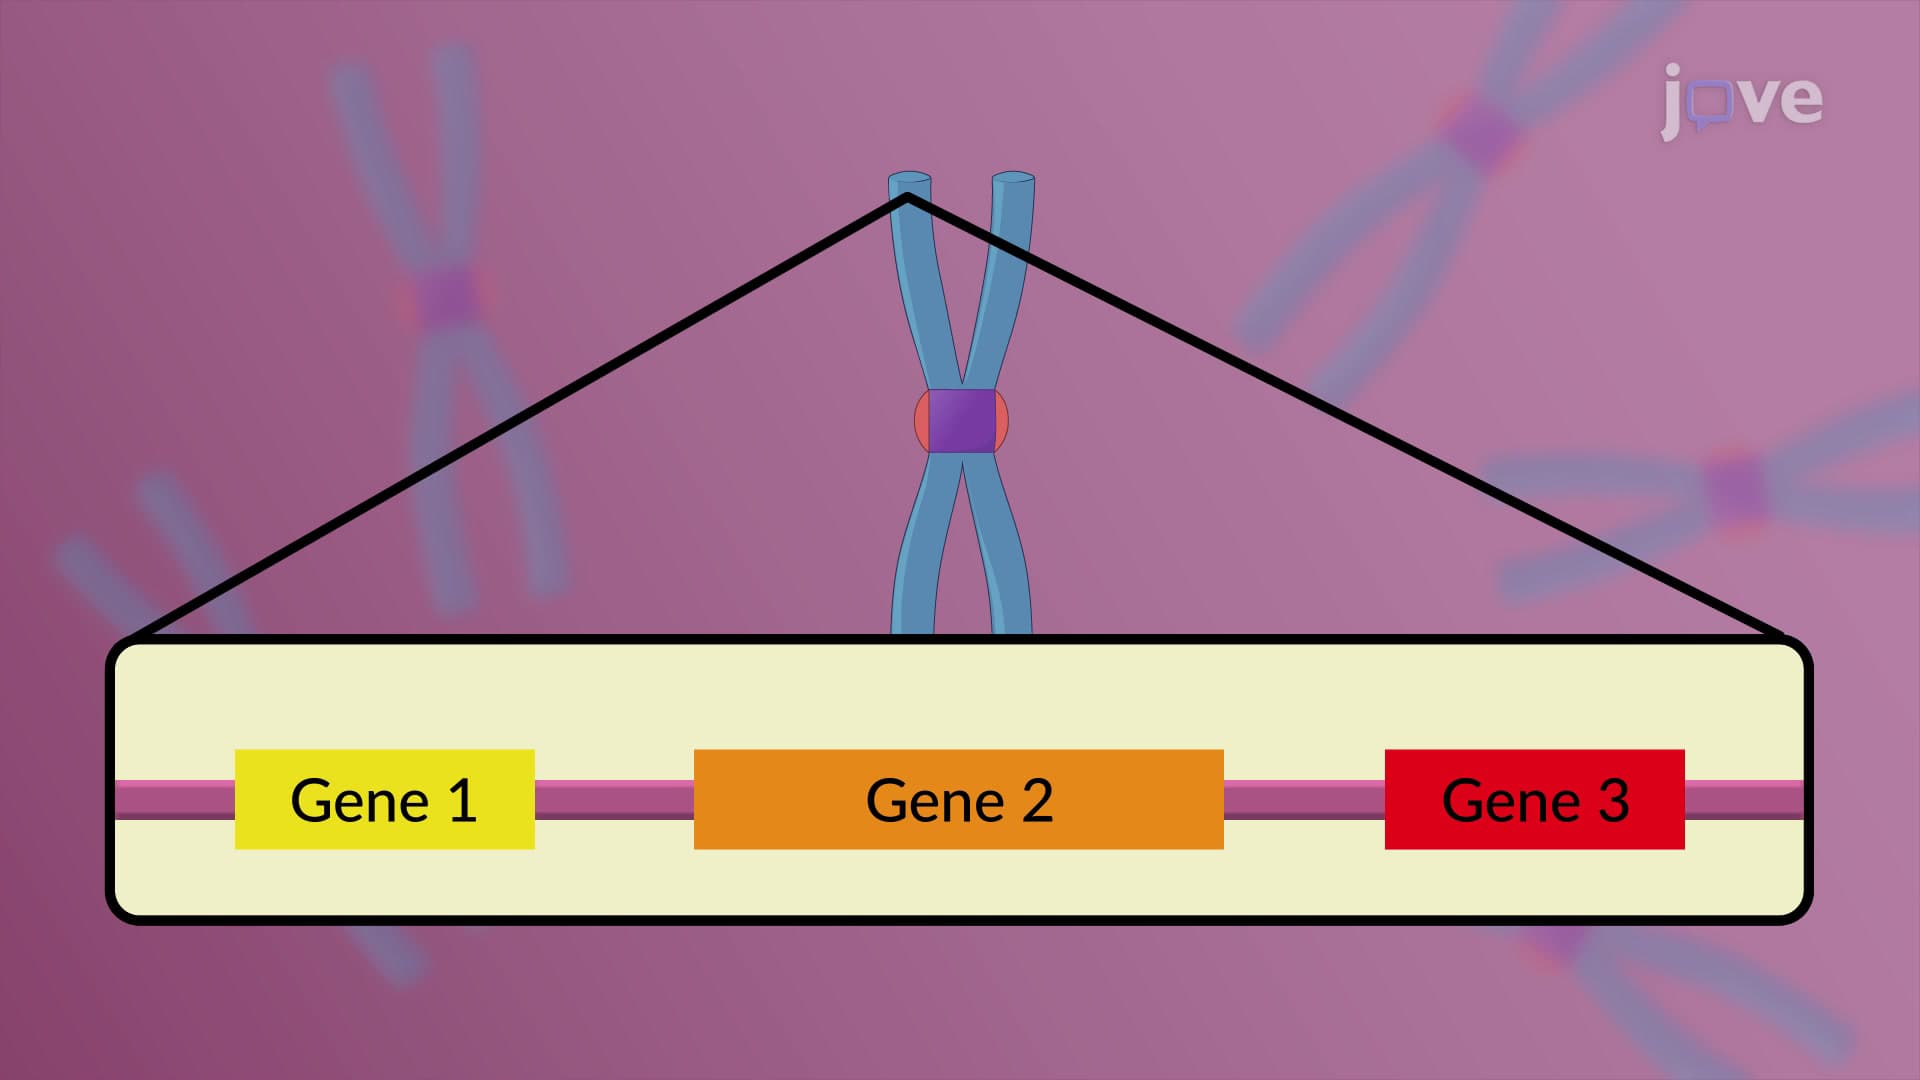
Structure of a Gene
However, only 1% of the DNA is composed of genes that encode proteins; the rest, 99% is non-coding DNA. This non-coding DNA performs...
 02:23
Master Transcription Regulators
 02:02
Cis-regulatory Sequences
 01:09
Epistasis Analysis
 02:13
Cooperative Binding of Transcription Regulators
 01:36
What is Gene Expression?