Related Experiment Video
Updated: Mar 16, 2026

Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
Intrinsically disordered regions stabilize the helical form of the C-terminal domain of RfaH: A molecular dynamics
Sangni Xun1, Fan Jiang1, Yun-Dong Wu2
1Laboratory of Computational Chemistry and Drug Design, Laboratory of Chemical Genomics, Peking University Shenzhen Graduate School, Shenzhen 518055, China.

Abstract:
RfaH protein functions as both transcription anti-terminator and translation enhancer in bacteria. Recent studies have shown that the C-terminal domain (CTD) is an α-helical hairpin (two-helix bundle) in full-length RfaH, despite the intrinsically favored β-barrel structure. Here, we carried out μs-timescale molecular dynamics (MD) simulations for the wild-type (WT) RfaH, its E48S mutant and an established model without the intrinsically disordered region (IDR1) linking the CTD and the N-terminal domain (NTD). Our simulations showed that the WT can be well stabilized by our RSFF1 force field, while the E48S mutant and the model without IDR1 undergo considerable structural change, which is in good agreement with experimental observations. The IDR1 plays important roles in stabilizing the hydrophobic environment near the crucial E48-R138 salt-bridge as well as in tethering α4 helix in CTD to α3 helix in NTD. In the absence of the IDR1, destabilization of key interdomain contacts and unfolding of the CTD α5 helix were observed in the simulation. In addition, the intrinsically disordered tail of the CTD (IDR2) is also of great significance to stabilize the bound conformation of CTD. These findings provide important implications for consideration of simulations in revealing the functions of residues invisible in a crystal structure.
More Related Videos
 09:49
Sedimentation Equilibrium of a Small Oligomer-forming Membrane Protein: Effect of Histidine Protonation on Pentameric Stability
Published on: April 2, 2015
 06:50
Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Related Concept Videos
 01:22
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 02:18
Intrinsically Disordered Proteins
02:18Intrinsically Disordered Proteins
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:15
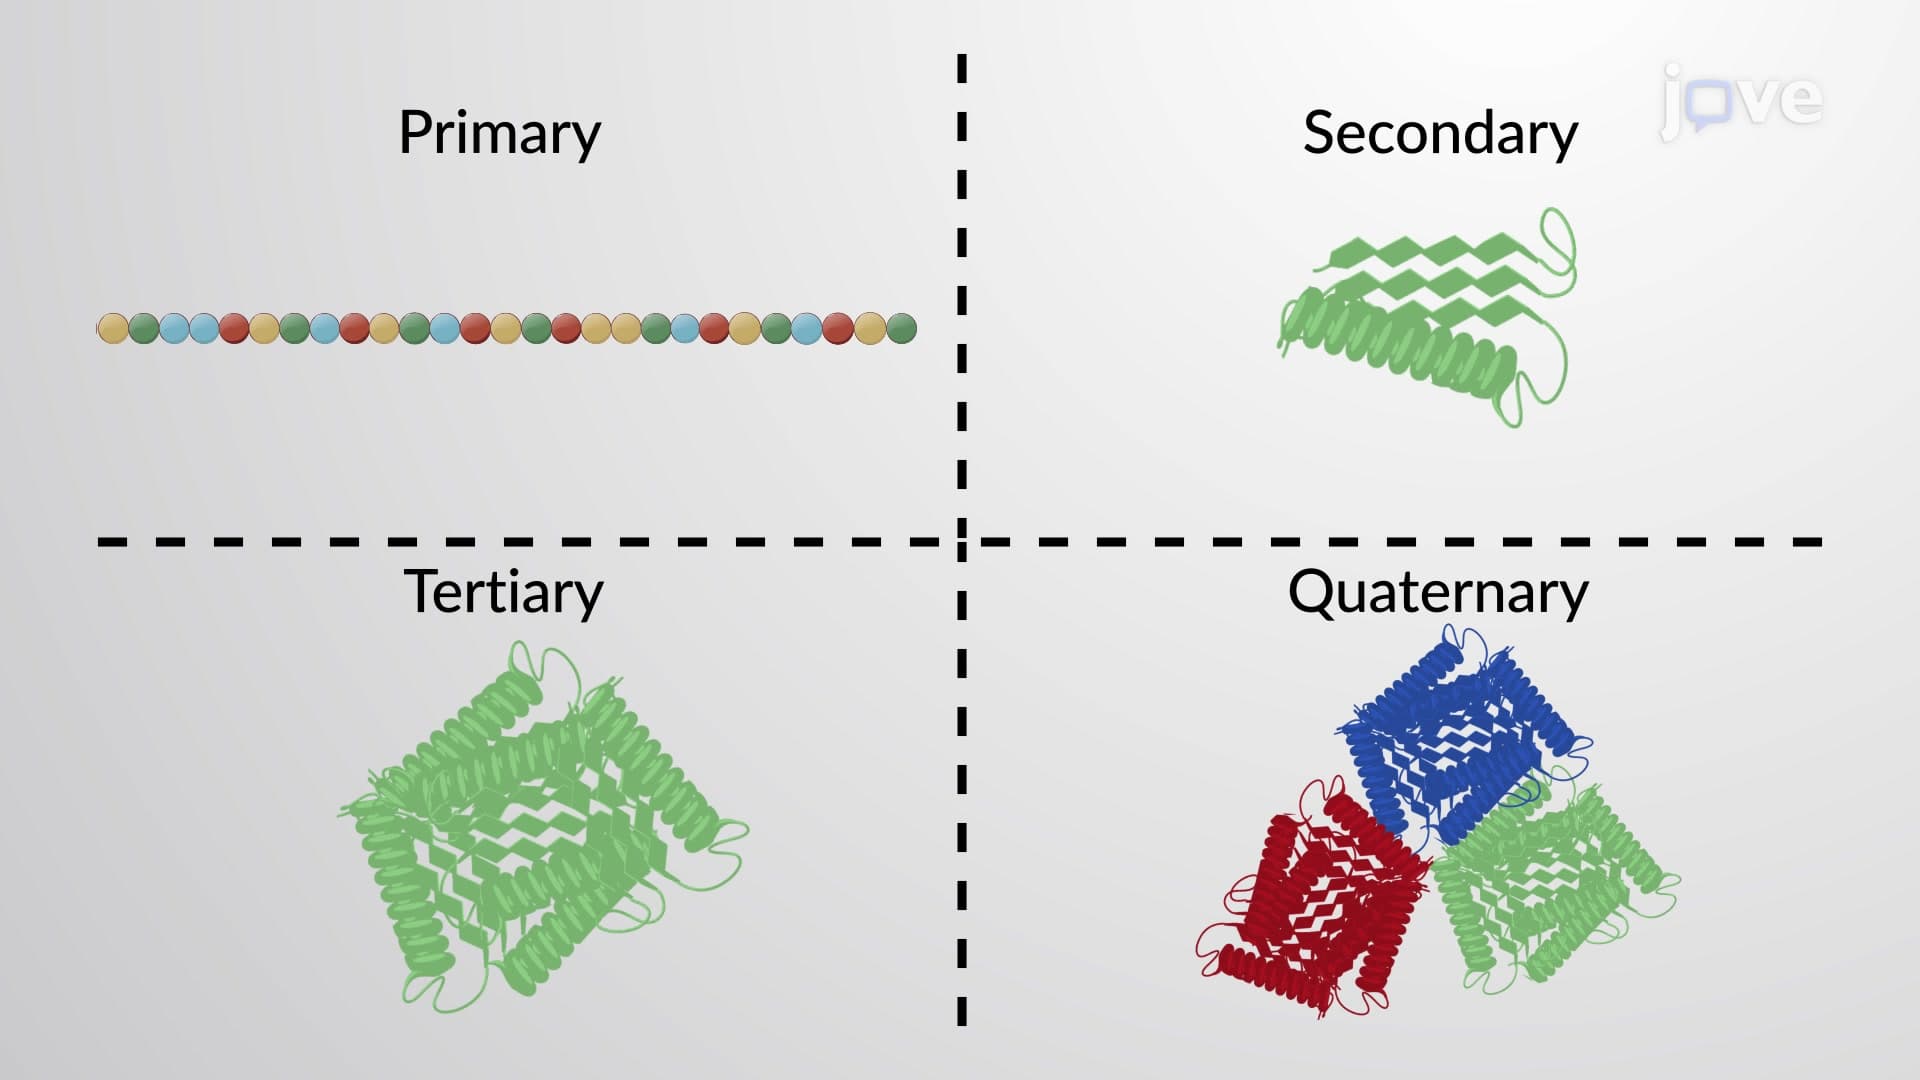
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...