Related Experiment Video
Updated: Mar 15, 2026
Reverse Yeast Two-hybrid System to Identify Mammalian Nuclear Receptor Residues that Interact with Ligands and/or Antagonists
Published on: November 15, 2013
Sintokamide A Is a Novel Antagonist of Androgen Receptor That Uniquely Binds Activation Function-1 in Its
Carmen A Banuelos1, Iran Tavakoli1, Amy H Tien1
1From the Department of Genome Sciences Centre, British Columbia Cancer Agency, 675 West 10th Avenue, Vancouver, British Columbia V5Z 1L3, Canada.

Abstract:
Androgen receptor (AR) is a validated drug target for all stages of prostate cancer including metastatic castration-resistant prostate cancer (CRPC). All current hormone therapies for CRPC target the C-terminal ligand-binding domain of AR and ultimately all fail with resumed AR transcriptional activity. Within the AR N-terminal domain (NTD) is activation function-1 (AF-1) that is essential for AR transcriptional activity. Inhibitors of AR AF-1 would potentially block most AR mechanisms of resistance including constitutively active AR splice variants that lack the ligand-binding domain. Here we provide evidence that sintokamide A (SINT1) binds AR AF-1 region to specifically inhibit transactivation of AR NTD. Consistent with SINT1 targeting AR AF-1, it attenuated transcriptional activities of both full-length AR and constitutively active AR splice variants, which correlated with inhibition of growth of enzalutamide-resistant prostate cancer cells expressing AR splice variants. In vivo, SINT1 caused regression of CRPC xenografts and reduced expression of prostate-specific antigen, a gene transcriptionally regulated by AR. Inhibition of AR activity by SINT1 was additive to EPI-002, a known AR AF-1 inhibitor that is in clinical trials (NCT02606123). This implies that SINT1 binds to a site on AF-1 that is unique from EPI. Consistent with this suggestion, these two compounds showed differences in blocking AR interaction with STAT3. This work provides evidence that the intrinsically disordered NTD of AR is druggable and that SINT1 analogs may provide a novel scaffold for drug development for the treatment of prostate cancer or other diseases of the AR axis.
Insights
Sintokamide A (SINT1) inhibits the androgen receptor (AR) N-terminal domain (NTD) activation function-1 (AF-1), blocking prostate cancer cell growth and CRPC xenografts. This novel AR AF-1 inhibitor offers a potential new drug scaffold.
Area of Science:
- Oncology
- Molecular Biology
- Drug Discovery
Background:
- Androgen receptor (AR) is a key target in prostate cancer, but therapies targeting its ligand-binding domain eventually fail due to resistance.
- Resistance mechanisms include constitutively active AR splice variants lacking the ligand-binding domain.
- The AR N-terminal domain (NTD), specifically activation function-1 (AF-1), is crucial for AR transcriptional activity and a potential target for overcoming resistance.
Purpose of the Study:
- To investigate sintokamide A (SINT1) as a novel inhibitor targeting the AR AF-1 region.
- To evaluate SINT1's efficacy against both full-length AR and AR splice variants in prostate cancer models.
- To explore SINT1's potential as a therapeutic scaffold for prostate cancer treatment.
Main Methods:
- Biochemical assays to confirm SINT1 binding to the AR AF-1 region.
- Cell-based assays measuring AR transcriptional activity of full-length AR and splice variants.
- In vitro studies assessing the effect of SINT1 on enzalutamide-resistant prostate cancer cell growth.
- In vivo studies using CRPC xenografts to evaluate SINT1's anti-tumor efficacy and impact on prostate-specific antigen (PSA) expression.
- Comparative studies of SINT1 and EPI-002, another AR AF-1 inhibitor, to assess binding site and mechanism.
Main Results:
- Sintokamide A (SINT1) specifically binds to the AR AF-1 region, inhibiting AR NTD transactivation.
- SINT1 attenuated the transcriptional activity of both full-length AR and AR splice variants.
- SINT1 inhibited the growth of enzalutamide-resistant prostate cancer cells expressing AR splice variants and caused regression of CRPC xenografts in vivo.
- SINT1 demonstrated additive effects with EPI-002, suggesting a unique binding site on AF-1, and showed differential effects on AR-STAT3 interaction.
Conclusions:
- The intrinsically disordered AR NTD is druggable, with SINT1 representing a novel inhibitor targeting AR AF-1.
- SINT1 effectively inhibits AR activity, including resistance mechanisms mediated by splice variants.
- SINT1 analogs hold promise as a new class of drugs for treating prostate cancer and other AR-axis-related diseases.
More Related Videos
Related Concept Videos
 01:28
Drug-Receptor Interaction: Antagonist
Antagonists can be classified as competitive or noncompetitive based on their...
 01:17
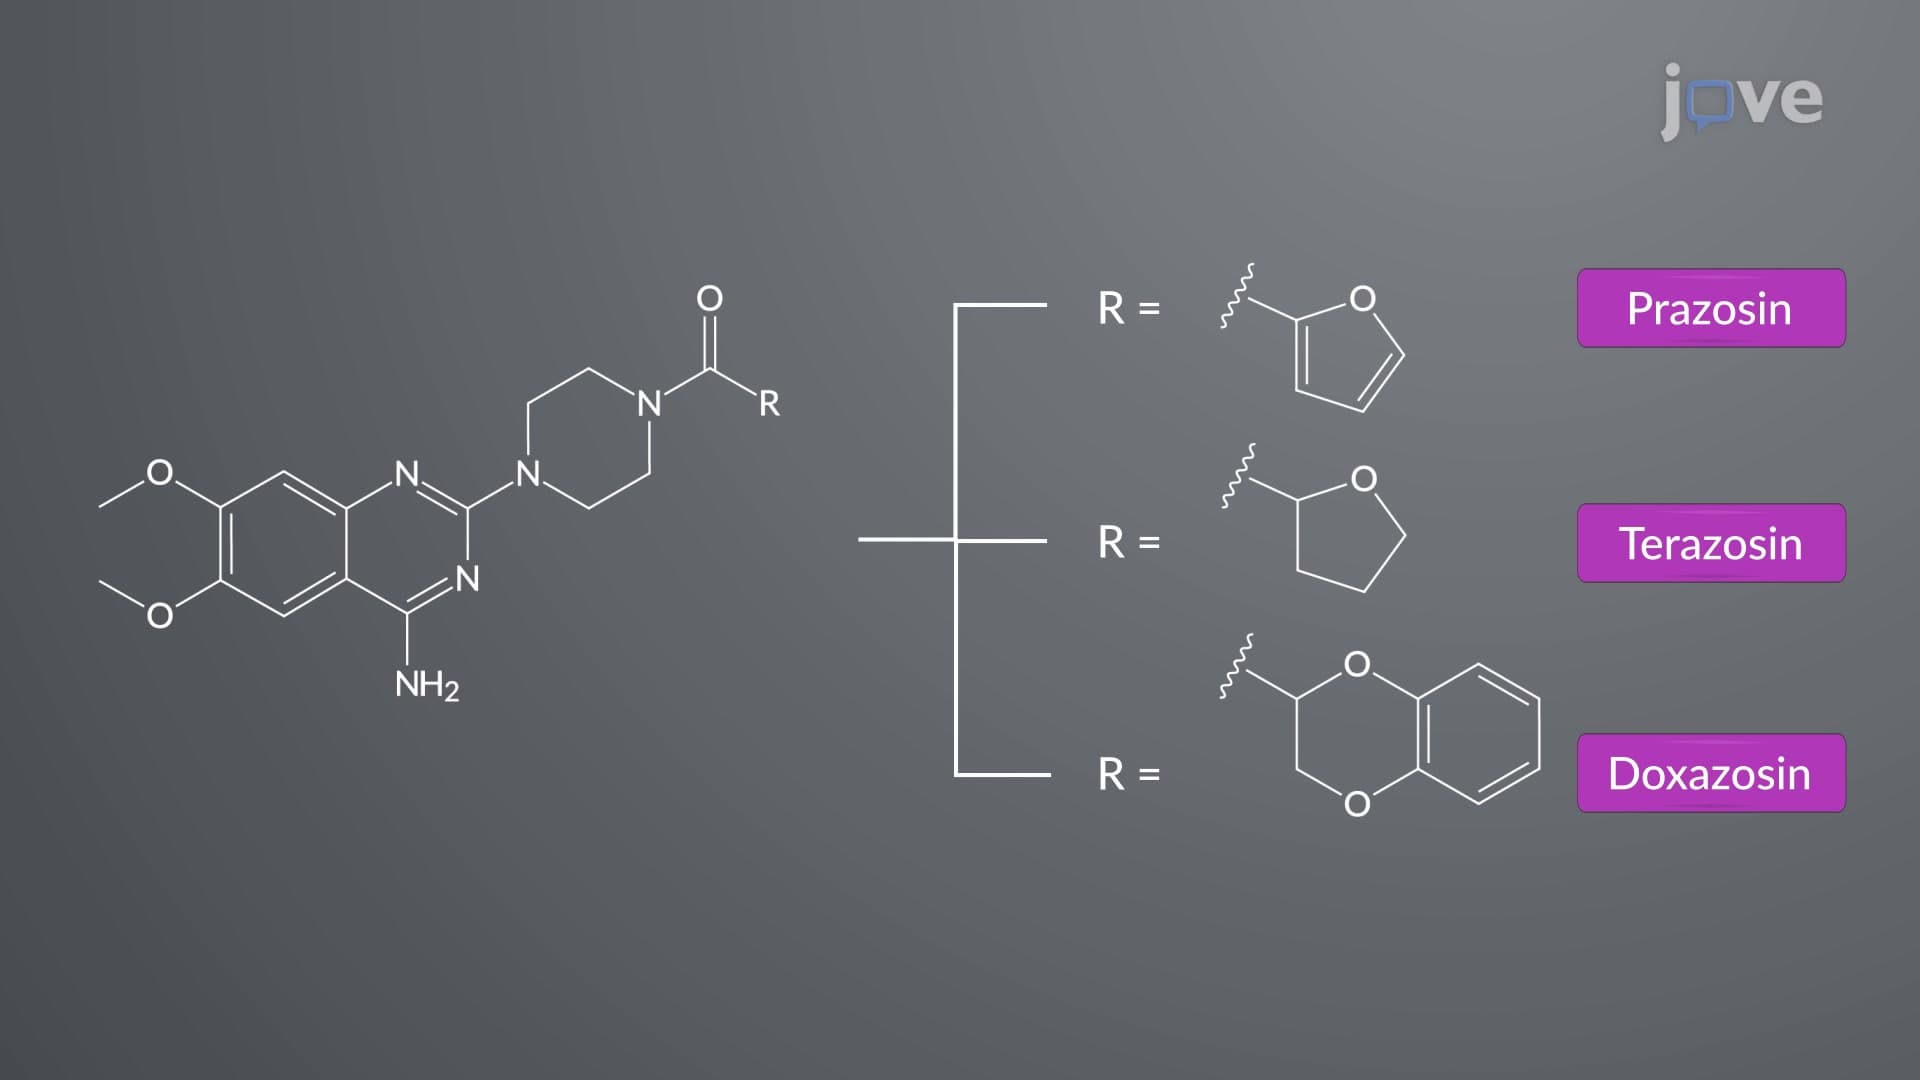
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline...
 01:17
Sedatives and Hypnotics Drugs: Miscellaneous Agents
Melatonin congeners like ramelteon (Rozerem) and tasimelteon (Hetlioz) selectively bind to melatonin receptors (MT1 and MT2) and thus mimic the actions of melatonin, a hormone that regulates sleep-wake cycles. Tasimelteon is primarily used for non-24-hour sleep-wake disorder, common in blind patients. They are also used to treat conditions like insomnia...
 01:22
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally,...
 01:30
Adrenergic Agonists: Direct-Acting Agents
These agents can be classified...
 01:25
Adrenergic Agonists: Indirect-Acting Agents
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral...