In Silico Prediction of Volume of Distribution in Humans. Extensive Data Set and the Exploration of Linear and Nonlinear Methods Coupled with Molecular Interaction Fields Descriptors
Related Experiment Video
Updated: Mar 15, 2026

Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Franco Lombardo1, Yankang Jing1
1Modelling, Computation and Molecular Properties Group, Biogen, 225 Binney Street, Cambridge, Massachusetts 02142, United States.

Abstract:
We present three in silico volume of distribution at steady state (VDss) models generated on a training set comprising 1096 compounds, which goes well beyond the conventional drug space delineated by the Rule of 5 or similar approaches. We have performed a careful selection of descriptors and kept a homogeneous Molecular Interaction Field-based descriptor set and linear (Partial Least Squares, PLS) and nonlinear (Random Forest, RF) models. We have tested the models, which we deem orthogonal in nature due to different descriptors and statistical approaches, with good results. In particular we tested the RF model, via a leave-class-out approach and by using a set of 34 additional compounds not used for training. We report comparable results against in vivo scaling approaches with geometric mean-fold error at or below 2 (for a set of 60 compounds with animal data available) and discuss the predictive performance based on the ionization states of the compounds. Lastly, we report the findings using a two-tier approach (classification followed by regression) based on VDss ranges, in an attempt to improve the prediction of compounds with very high VDss. We would recommend, overall, the RF model, with 33 descriptors, as the primary choice for VDss prediction in humans.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 06:50
Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Related Concept Videos
 01:20
Volume of Distribution
 01:25
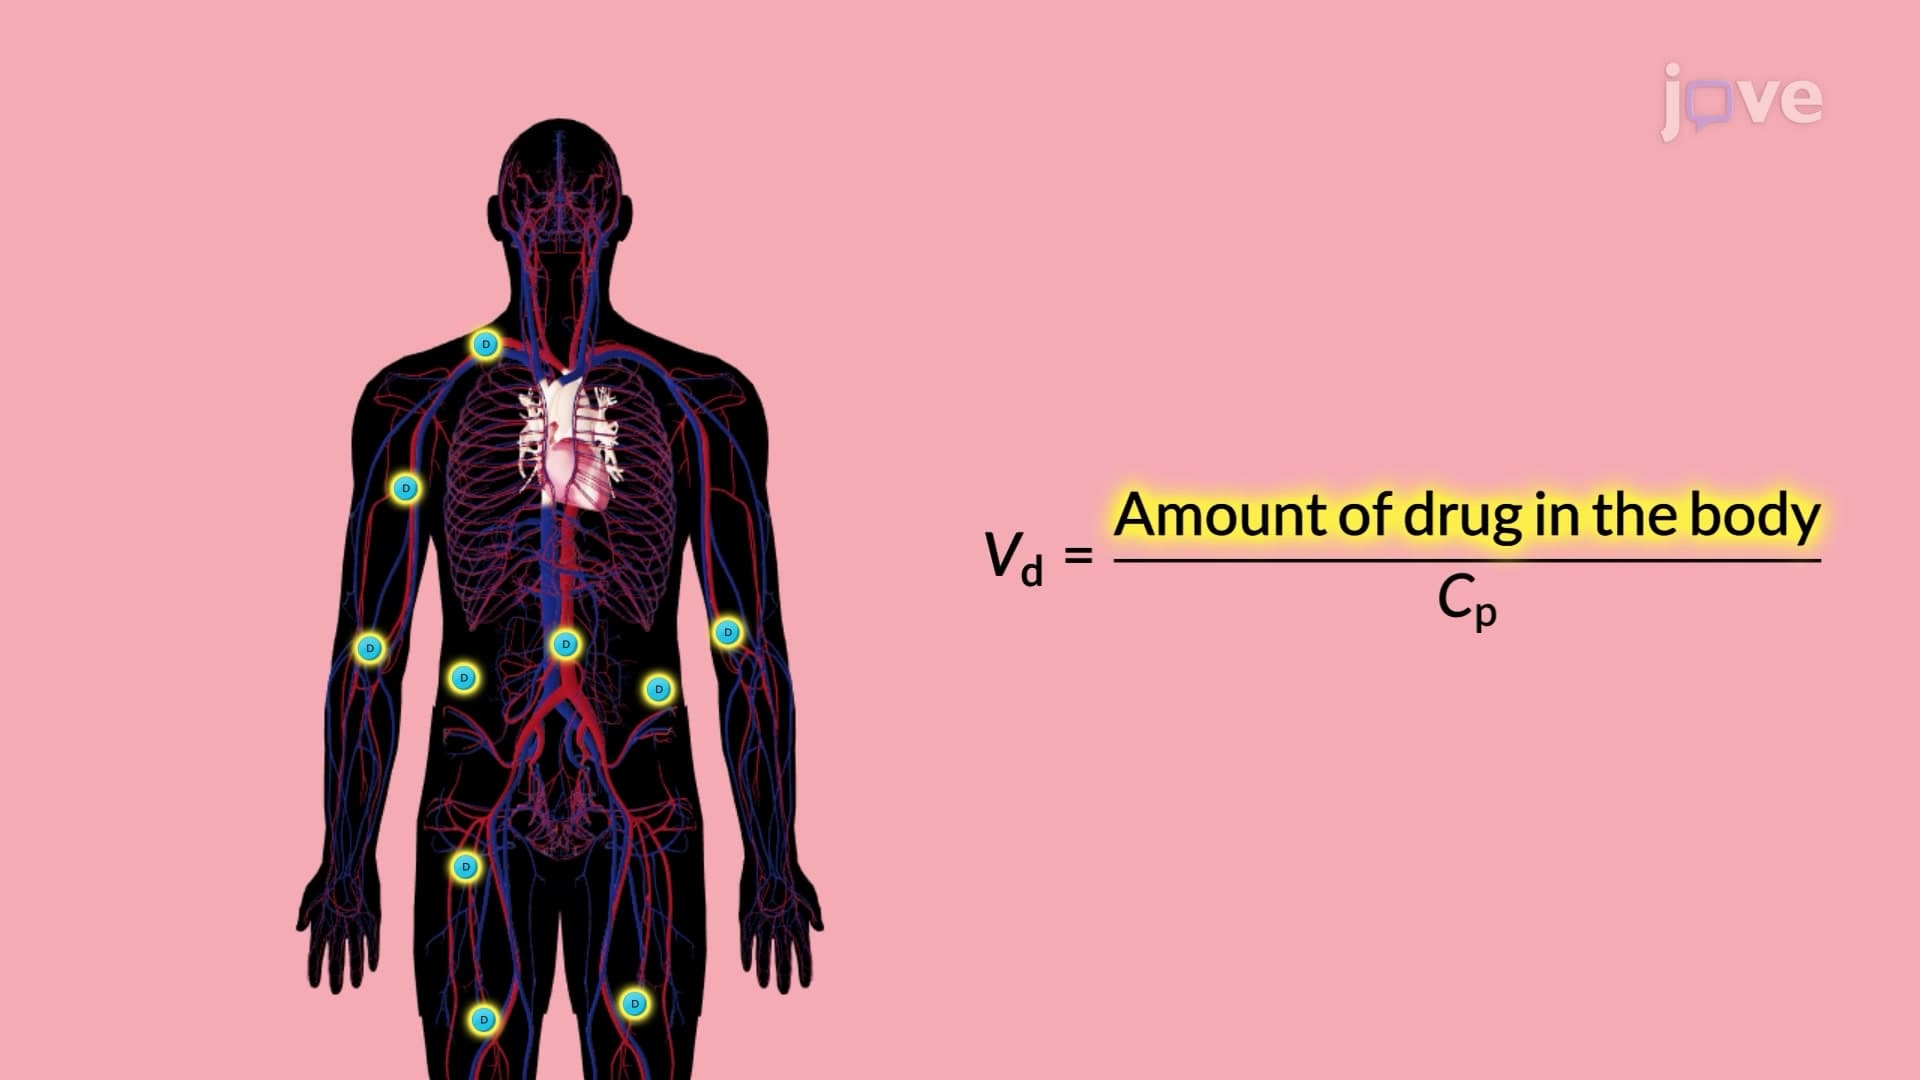
Drug Distribution: Volume of Distribution
 02:04
Protein-protein Interfaces
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...