
Impact of metal ion's charge on the interatomic Coulombic decay widths in microsolvated clusters
V Stumpf1, C Brunken1, K Gokhberg1
1Theoretische Chemie, Physikalisch-Chemisches Institut, Universität Heidelberg, Im Neuenheimer Feld 229, D-69120 Heidelberg, Germany.

Abstract:
Interatomic Coulombic decay (ICD) is an efficient electronic decay mechanism of electronically excited atoms and molecules embedded in an environment. For the series of isoelectronic Na(+), Mg(2+), and Al(3+) ions in aqueous solution, ultrashort ICD lifetimes of 3.1 fs, 1.5 fs, and 0.9 fs, respectively, were observed experimentally. The magnitude of the ICD lifetimes and their variation within the series were qualitatively explained by shortening metal-oxygen equilibrium distances and the increasing polarization of the water molecules as the metal's charge grows. We carried out an extensive ab initio investigation of the variation of the ICD widths with the metal-oxygen distances and the number of water neighbors in Na(+)-(H2O)m (m = 1-4) and Mg(2+)-(H2O)n (n = 1-6) clusters including and excluding polarization effects in the decaying state. We demonstrated that the effect of the induced polarization of the water ligand and the equilibrium cation-oxygen distance are equally important in determining the ordering and ratios of the ICD lifetimes in the series. Moreover, we showed that the induced polarization of the water molecules leads to a slower than linear growth of ICD width with the number of equivalent water neighbors; the non-linearity is stronger for Mg(2+). The ab initio ICD widths in microsolvated Na(+)-(H2O)4 and Mg(2+)-(H2O)6 clusters are found to be in good agreement with the experimental values.
More Related Videos
 16:11
Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
 06:53
Author Spotlight: Magnetometric Characterization of Intermediates in the Solid-State Electrochemistry of Redox-Active Metal-Organic Frameworks
Published on: June 9, 2023
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:09
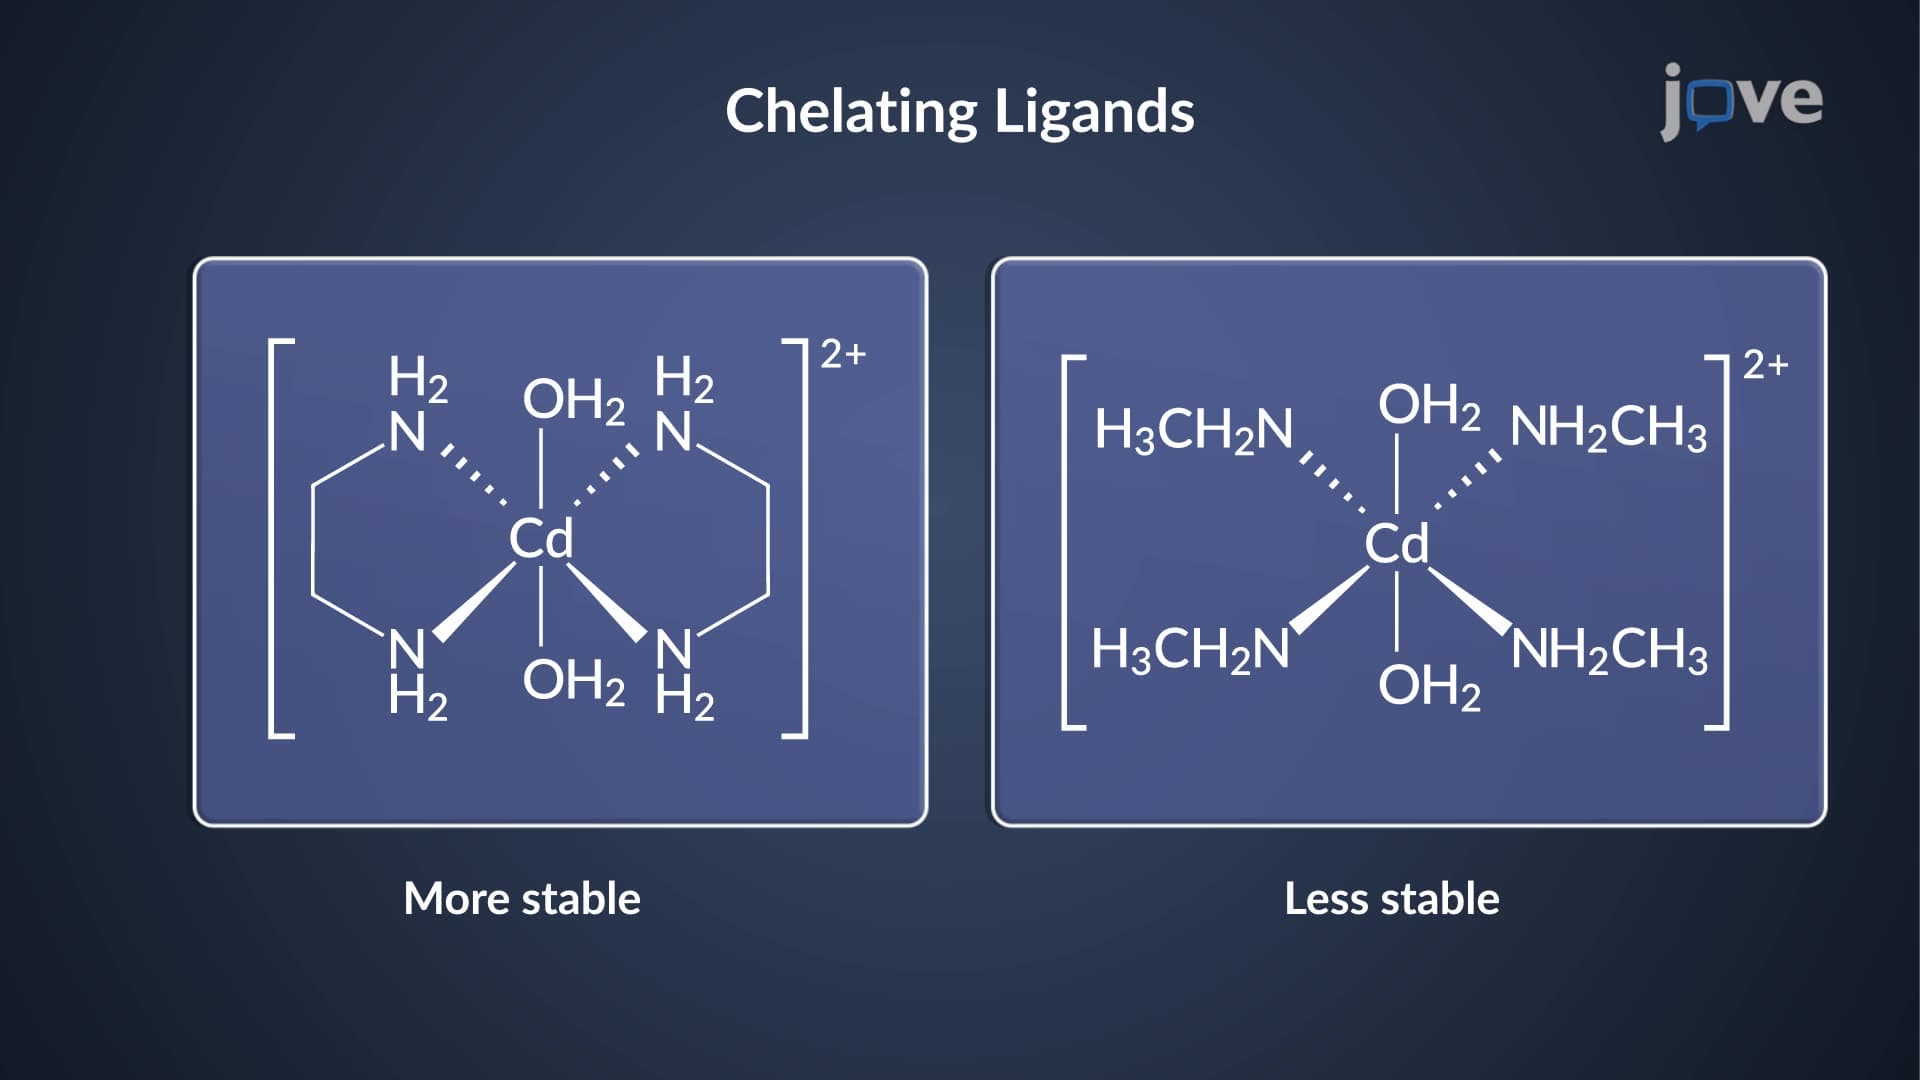
Complexation Equilibria: Factors Influencing Stability of Complexes
 03:45
Formation of Complex Ions
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:54
Trends in Lattice Energy: Ion Size and Charge
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...