Related Experiment Video
Updated: Mar 14, 2026

Interactive Molecular Model Assembly with 3D Printing
Published on: August 13, 2020
Potential Energy Surface-Based Automatic Deduction of Conformational Transition Networks and Its Application on
Hiroko Satoh1,2,3,4, Tomohiro Oda5, Kumiyo Nakakoji6
1Research Organization of Information and Systems (ROIS), Tokyo 105-0001, Japan.

Abstract:
This paper describes our approach that is built upon the potential energy surface (PES)-based conformational analysis. This approach automatically deduces a conformational transition network, called a conformational reaction route map (r-map), by using the Scaled Hypersphere Search of the Anharmonic Downward Distortion Following method (SHS-ADDF). The PES-based conformational search has been achieved by using large ADDF, which makes it possible to trace only low transition state (TS) barriers while restraining bond lengths and structures with high free energy. It automatically performs sampling the minima and TS structures by simply taking into account the mathematical feature of PES without requiring any a priori specification of variable internal coordinates. An obtained r-map is composed of equilibrium (EQ) conformers connected by reaction routes via TS conformers, where all of the reaction routes are already confirmed during the process of the deduction using the intrinsic reaction coordinate (IRC) method. The postcalculation analysis of the deduced r-map is interactively carried out using the RMapViewer software we have developed. This paper presents computational details of the PES-based conformational analysis and its application to d-glucose. The calculations have been performed for an isolated glucose molecule in the gas phase at the RHF/6-31G level. The obtained conformational r-map for α-d-glucose is composed of 201 EQ and 435 TS conformers and that for β-d-glucose is composed of 202 EQ and 371 TS conformers. For the postcalculation analysis of the conformational r-maps by using the RMapViewer software program we have found multiple minimum energy paths (MEPs) between global minima of 1C4 and 4C1 chair conformations. The analysis using RMapViewer allows us to confirm the thermodynamic and kinetic predominance of 4C1 conformations; that is, the potential energy of the global minimum of 4C1 is lower than that of 1C4 (thermodynamic predominance) and that the highest energy of those of all the TS structures along a route from 4C1 to 1C4 is lower than that of 1C4 to 4C1 (kinetic predominance).
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 10:27
Contrast-Matching Detergent in Small-Angle Neutron Scattering Experiments for Membrane Protein Structural Analysis and Ab Initio Modeling
Published on: October 21, 2018
Related Concept Videos
 02:18
Fischer Projections
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 02:06
Newman Projections
The organic molecules rotate across the single bonds leading to numerous temporary three-dimensional structures of varying energy known as...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 02:20
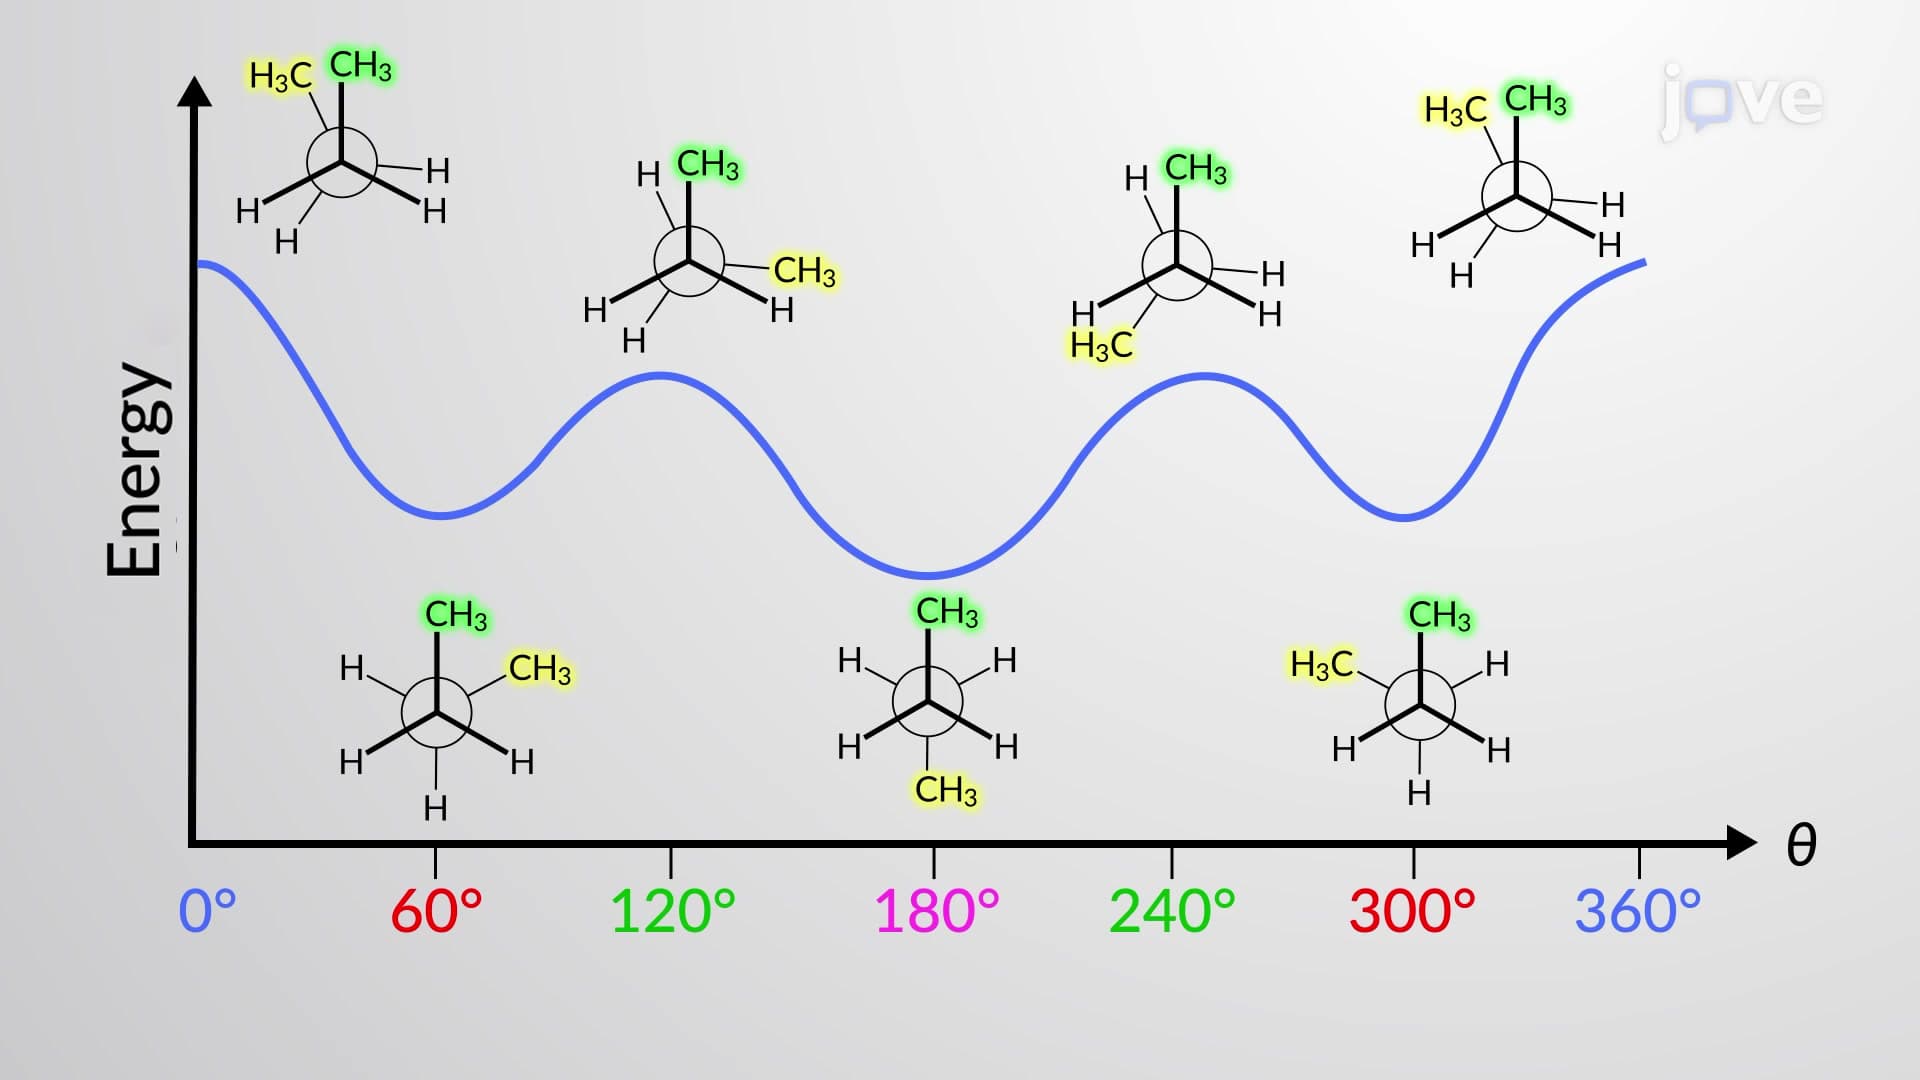
Conformations of Butane
 02:13
Energy Diagrams, Transition States, and Intermediates