Related Experiment Video
Updated: Mar 14, 2026

Setting-up an In Vitro Model of Rat Blood-brain Barrier BBB: A Focus on BBB Impermeability and Receptor-mediated Transport
Published on: June 28, 2014
QSAR Model of Unbound Brain-to-Plasma Partition Coefficient, Kp,uu,brain: Incorporating P-glycoprotein Efflux as a
Elena Dolgikh1, Ian A Watson1, Prashant V Desai1
1Global Scientific Informatics, ‡Advanced Analytics, §Computational ADME, ∥IT Informatics and ⊥Drug Disposition, Lilly Research Laboratories, Eli Lilly and Company , Indianapolis, Indiana 46285, United States.

Abstract:
We report development and prospective validation of a QSAR model of the unbound brain-to-plasma partition coefficient, Kp,uu,brain, based on the in-house data set of ∼1000 compounds. We discuss effects of experimental variability, explore the applicability of both regression and classification approaches, and evaluate a novel, model-within-a-model approach of including P-glycoprotein efflux prediction as an additional variable. When tested on an independent test set of 91 internal compounds, incorporation of P-glycoprotein efflux information significantly improves the model performance resulting in an R2 of 0.53, RMSE of 0.57, Spearman's Rho correlation coefficient of 0.73, and qualitative prediction accuracy of 0.8 (kappa = 0.6). In addition to improving the performance, one of the key advantages of this approach is the larger chemical space coverage provided indirectly through incorporation of the in vitro, higher throughput data set that is 4 times larger than the in vivo data set.
Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:16
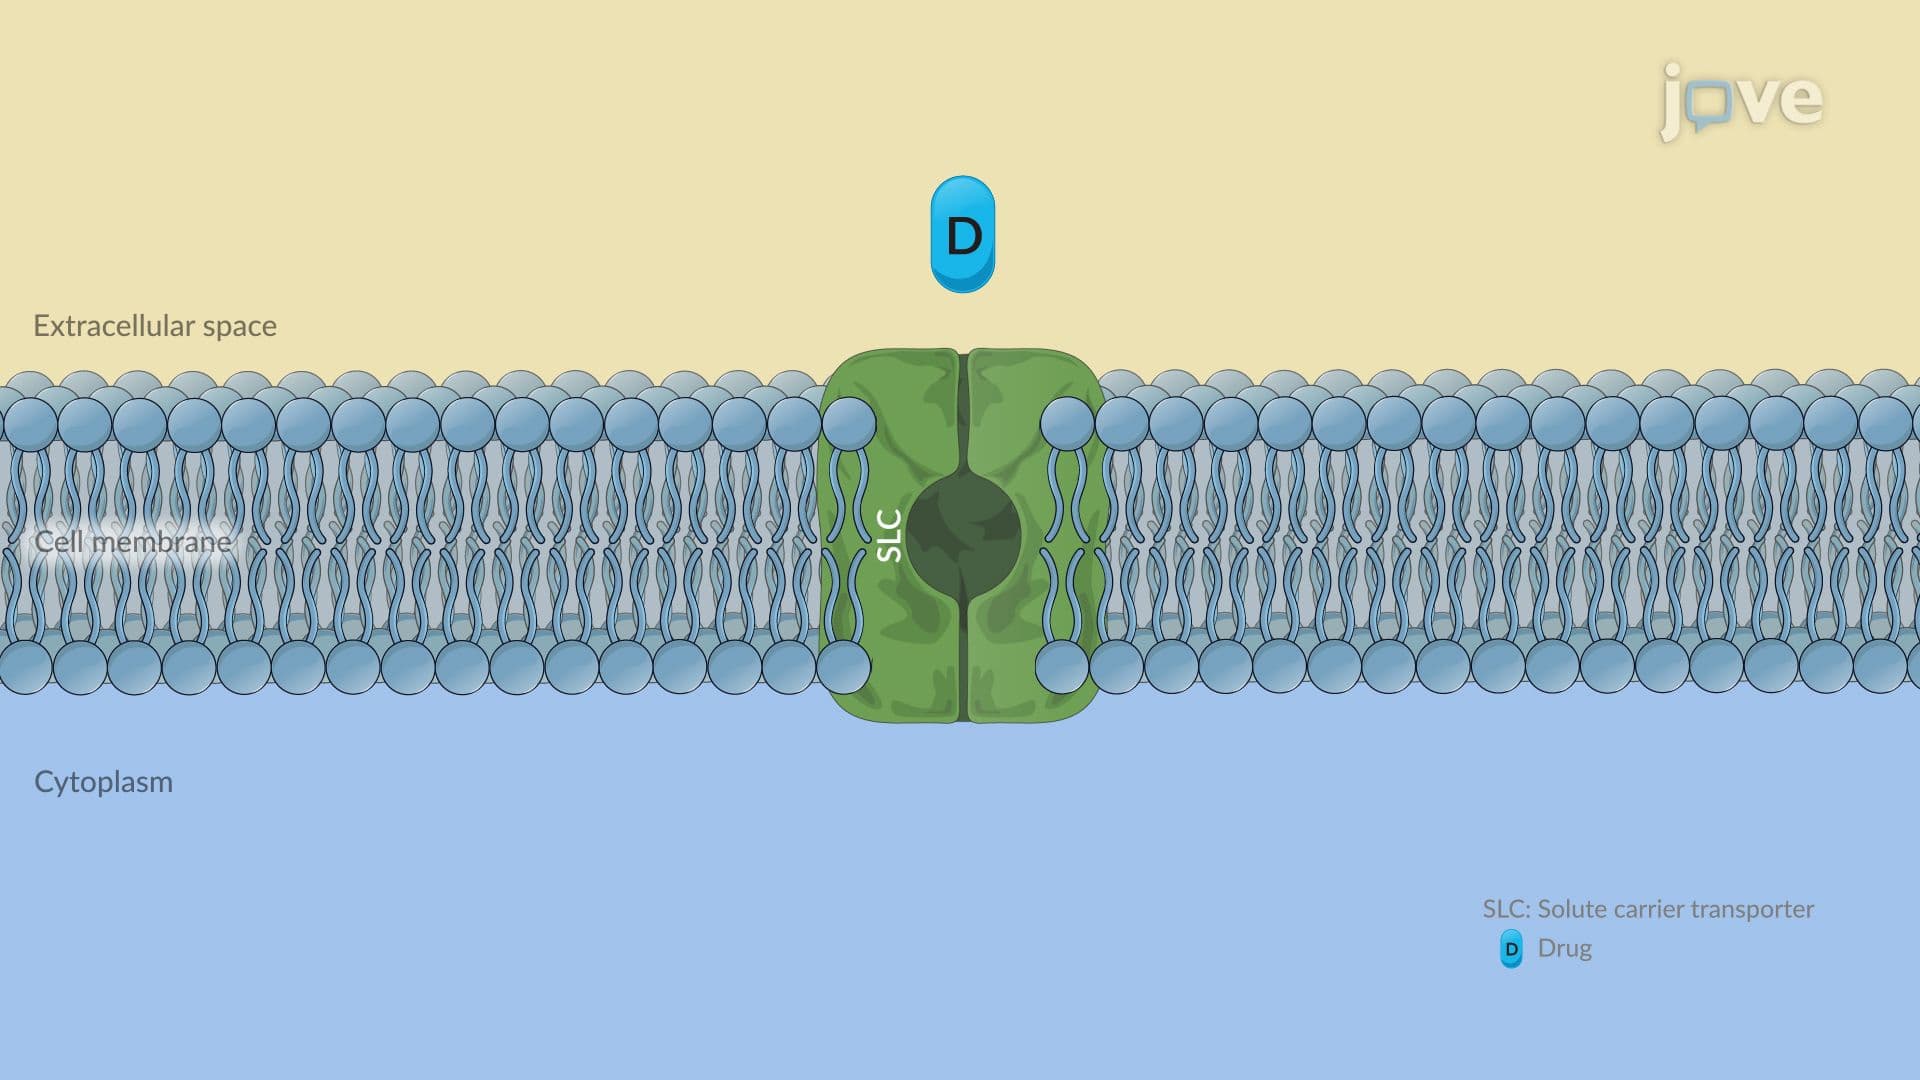
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters
 01:29
Drug Distribution: Plasma Protein Binding
 01:14
Hepatic Drug Clearance: Role of Transporters
 01:22
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
To quantify the extent of bioavailability, pharmacologists often use a parameter called .