Related Experiment Video
Updated: Mar 14, 2026
Modeling Mitochondrial Disease Using Brain Organoids: A Focus on Mitochondrial Encephalomyopathy, Lactic Acidosis, and Stroke-like Episodes
Published on: October 10, 2025
Pathogenic variants in HTRA2 cause an early-onset mitochondrial syndrome associated with 3-methylglutaconic aciduria
Monika Oláhová1, Kyle Thompson1, Steven A Hardy1
1Wellcome Trust Centre for Mitochondrial Research, Institute of Neuroscience, The Medical School, Newcastle University, Framlington Place, Newcastle upon Tyne, NE2 4HH, UK.

Abstract:
Mitochondrial diseases collectively represent one of the most heterogeneous group of metabolic disorders. Symptoms can manifest at any age, presenting with isolated or multiple-organ involvement. Advances in next-generation sequencing strategies have greatly enhanced the diagnosis of patients with mitochondrial disease, particularly where a mitochondrial aetiology is strongly suspected yet OXPHOS activities in biopsied tissue samples appear normal. We used whole exome sequencing (WES) to identify the molecular basis of an early-onset mitochondrial syndrome-pathogenic biallelic variants in the HTRA2 gene, encoding a mitochondria-localised serine protease-in five subjects from two unrelated families characterised by seizures, neutropenia, hypotonia and cardio-respiratory problems. A unifying feature in all affected children was 3-methylglutaconic aciduria (3-MGA-uria), a common biochemical marker observed in some patients with mitochondrial dysfunction. Although functional studies of HTRA2 subjects' fibroblasts and skeletal muscle homogenates showed severely decreased levels of mutant HTRA2 protein, the structural subunits and complexes of the mitochondrial respiratory chain appeared normal. We did detect a profound defect in OPA1 processing in HTRA2-deficient fibroblasts, suggesting a role for HTRA2 in the regulation of mitochondrial dynamics and OPA1 proteolysis. In addition, investigated subject fibroblasts were more susceptible to apoptotic insults. Our data support recent studies that described important functions for HTRA2 in programmed cell death and confirm that patients with genetically-unresolved 3-MGA-uria should be screened by WES with pathogenic variants in the HTRA2 gene prioritised for further analysis.
Related Concept Videos
 01:27
Pharmacogenetics of Phase II Enzymes: N-acetyltransferase, Thiopurine S-methyltransferase, UDP-glucuronosyltransferase
 01:19
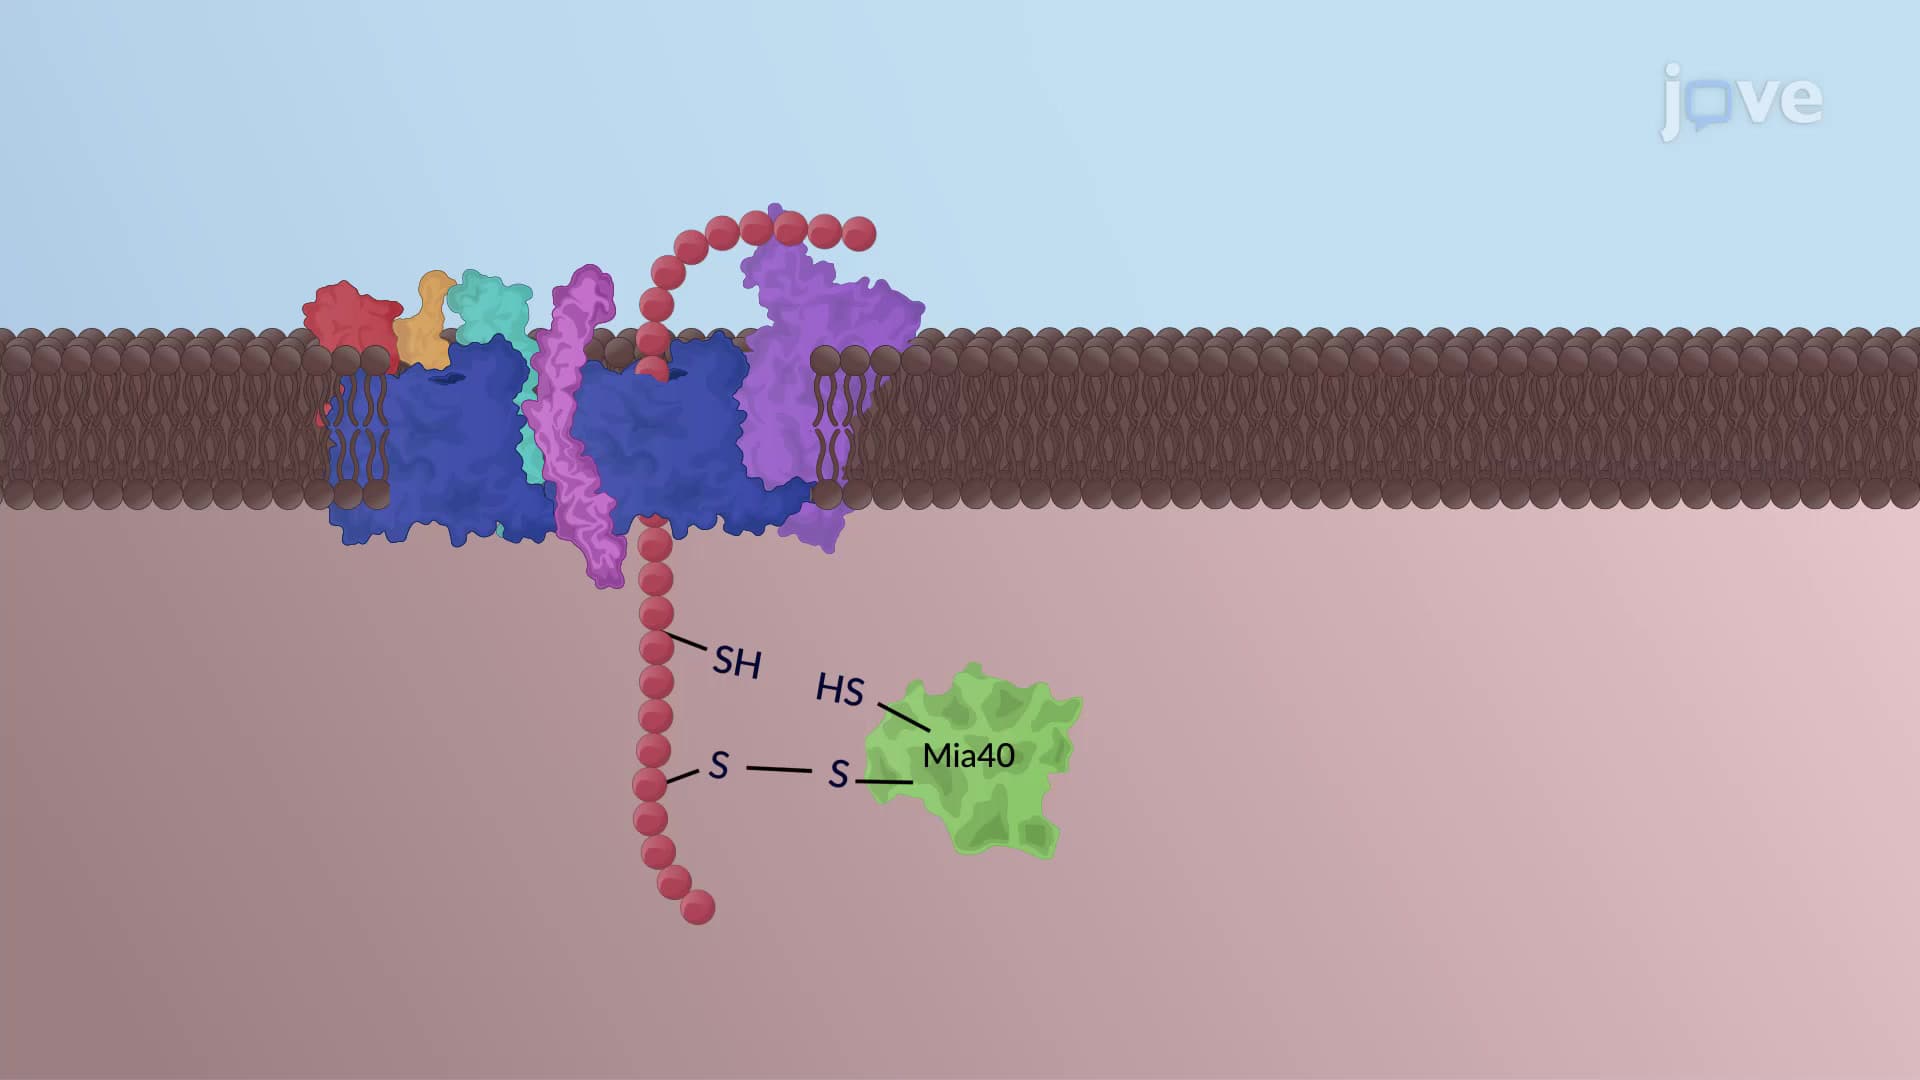
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:48
ATP Synthase: Mechanism
 01:37
Energy to Drive Translocation
Generally, polypeptides are unfolded by two distinct...
 02:59
Animal Mitochondrial Genetics
 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial...