Related Experiment Video
Updated: Mar 14, 2026
An Electrochemiluminescence-Based Assay for MeCP2 Protein Variants
Published on: May 22, 2020
Methylmalonyl-coA epimerase deficiency: A new case, with an acute metabolic presentation and an intronic splicing
Paula J Waters1, Fanny Thuriot1, Joe T R Clarke1
1Medical Genetics Service, Department of Pediatrics, Centre hospitalier universitaire de Sherbrooke (CHUS) and University of Sherbrooke, Sherbrooke, Quebec, Canada.

Abstract:
Methylmalonyl-coA epimerase (MCE) follows propionyl-coA carboxylase and precedes methylmalonyl-coA mutase in the pathway converting propionyl-coA to succinyl-coA. MCE deficiency has previously been described in six patients, one presenting with metabolic acidosis, the others with nonspecific neurological symptoms or asymptomatic. The clinical significance and biochemical characteristics of this rare condition have been incompletely defined. We now describe a patient who presented acutely at 5 years of age with vomiting, dehydration, confusion, severe metabolic acidosis and mild hyperammonemia. At presentation, organic acid profiles were dominated by increased ketones and 3-hydroxypropionate, with moderately elevated methylcitrate and propionylglycine, and acylcarnitine profiles showed marked C3 (propionylcarnitine) elevation with normal C4DC (methylmalonylcarnitine + succinylcarnitine). Propionic acidemia was initially suspected, but it was subsequently noted that methylmalonic acid was mildly but persistently elevated in urine, and clearly elevated in plasma and cerebrospinal fluid. The overall biochemical profile prompted consideration of MCE deficiency. Studies on cultured fibroblasts showed moderately decreased propionate incorporation. Complementation analysis permitted assignment to the MCEE group. A heterozygous p.Arg47Ter (p.R47*) mutation in the MCEE gene was identified by sequencing of exons, and RNA studies identified a novel intronic splicing mutation, c.379-644A > G, confirming the diagnosis of MCE deficiency. Following the initial severe presentation, development has been normal and the clinical course over the subsequent six years has remained relatively uneventful on an essentially normal diet. This report contributes to the clinical and biochemical characterisation of this rare disorder, while highlighting potential causes of under-diagnosis or of diagnostic confusion.
More Related Videos
 08:56
Modeling Mitochondrial Disease Using Brain Organoids: A Focus on Mitochondrial Encephalomyopathy, Lactic Acidosis, and Stroke-like Episodes
Published on: October 10, 2025
 03:45
Investigating the Pathogenesis of MYH7 Mutation Gly823Glu in Familial Hypertrophic Cardiomyopathy using a Mouse Model
Published on: August 8, 2022
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 02:18
Alternative RNA Splicing
There are five types of alternative RNA splicing that vary in the ways the pre-mRNA segments are removed or retained in the mature mRNA. The first...
 02:59
Animal Mitochondrial Genetics
 01:29
Insertion of Multi-pass Transmembrane Proteins in the RER
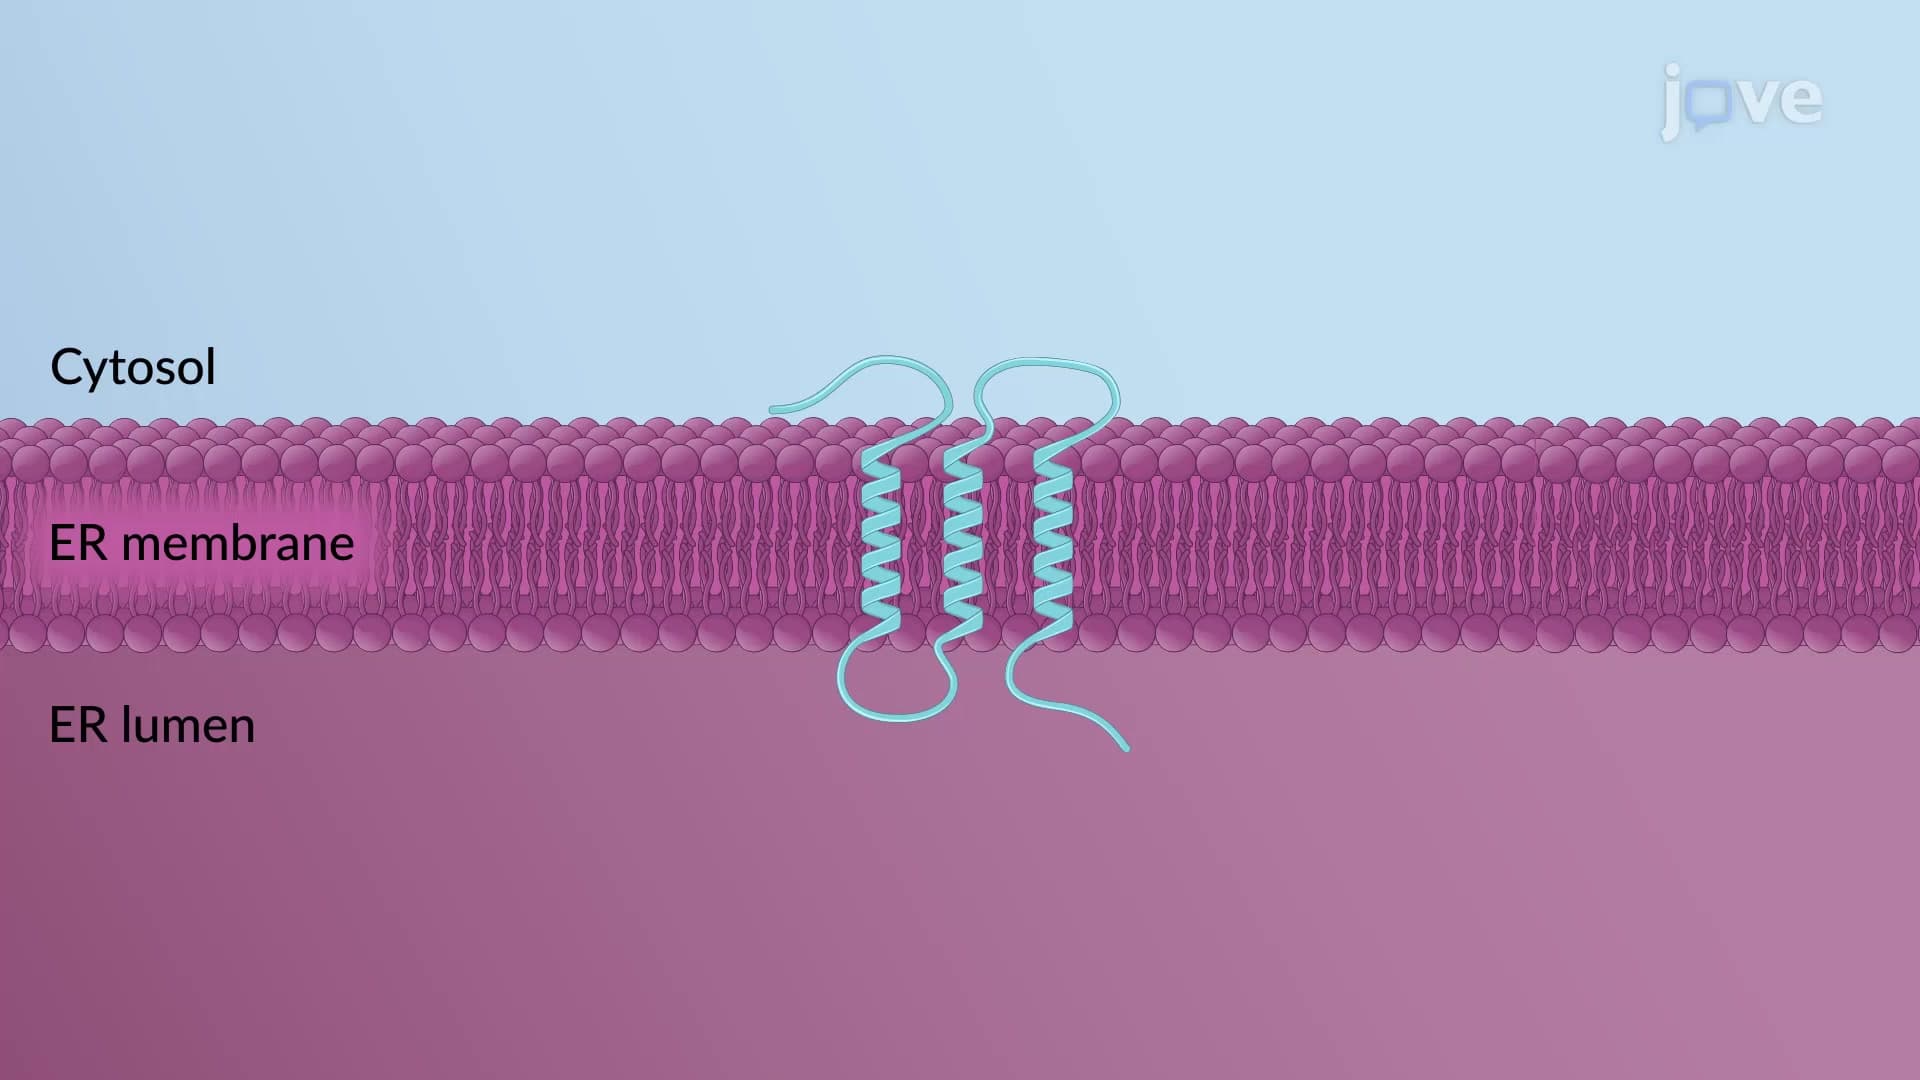
The multipass transmembrane proteins are the type IV integral membrane proteins with multiple topogenic sequences determining their spatial arrangement in the ER membrane. Nearly all multipass proteins lack a cleavable signal sequence and use...
 01:32
RNA Splicing