The evolution of a stereoselective synthesis of the C1-C16 fragment of bryostatins
Related Experiment Video
Updated: Mar 14, 2026
Synthesis of a Borylated Ibuprofen Derivative Through Suzuki Cross-Coupling and Alkene Boracarboxylation Reactions
Published on: November 30, 2022
Matthew Ball1, Anne Baron1, Ben Bradshaw1
1The School of Chemistry, The University of Manchester, Manchester M13 9PL, UK. e.j.thomas@manchester.ac.uk.

Abstract:
The development of a synthesis of the C1-C16 fragment of bryostatins in which the key step is a stereoselective oxy-Michael reaction used to assemble the cis-2,6-disubstituted tetrahydropyran with the exocyclic alkene already installed, is described. Following early work using Stille reactions to prepare precursors for oxy-Michael reactions, a more efficient route was devised based on a ketophosphonate-aldehyde condensation to prepare the key dienone. Synthesis of the aldehyde required for the ketophosphonate condensation involved the highly stereoselective addition of a diorganocuprate derived from allylmagnesium bromide and copper(i) iodide to the methyl 5-hydroxyhex-2-ynoate prepared by ring-opening of a protected glycidol using lithiated methyl propiolate. Ester reduction, selective alcohol protection and oxidative cleavage of the terminal double bond gave the required aldehyde. The ketophosphonate was prepared in 13 steps from (R)-pantolactone using a synthesis based on a chelation controlled aldol condensation and an anti-selective reduction of a 3-hydroxyketone. Following the ketophosphonate-aldehyde reaction, selective deprotection followed by treatment with base effected the oxy-Michael reaction that gave the cis-2,6-disubstituted tetrahydropyran via thermodynamic control. Subsequent functional group manipulation led to the synthesis of a hydroxy ester that corresponded to the C1-C16 fragment of the bryostatins in 23 steps from (R)-pantolactone. The synthesis was repeated using slightly different protecting groups for a study of a ring-closing metathesis approach to the bryostatins.
More Related Videos
 10:17
Efficient Construction of Drug-like Bispirocyclic Scaffolds Via Organocatalytic Cycloadditions of α-Imino γ-Lactones and Alkylidene Pyrazolones
Published on: February 7, 2019
 08:46
Regioselective O-Glycosylation of Nucleosides via the Temporary 2',3'-Diol Protection by a Boronic Ester for the Synthesis of Disaccharide Nucleosides
Published on: July 26, 2018
Related Concept Videos
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 02:05
Prochirality
 01:14
Alkylation of β-Diester Enolates: Malonic Ester Synthesis
 01:29
Diels–Alder Reaction Forming Bridged Bicyclic Products: Stereochemistry
 01:07
Alkylation of β-Ketoester Enolates: Acetoacetic Ester Synthesis
 02:33
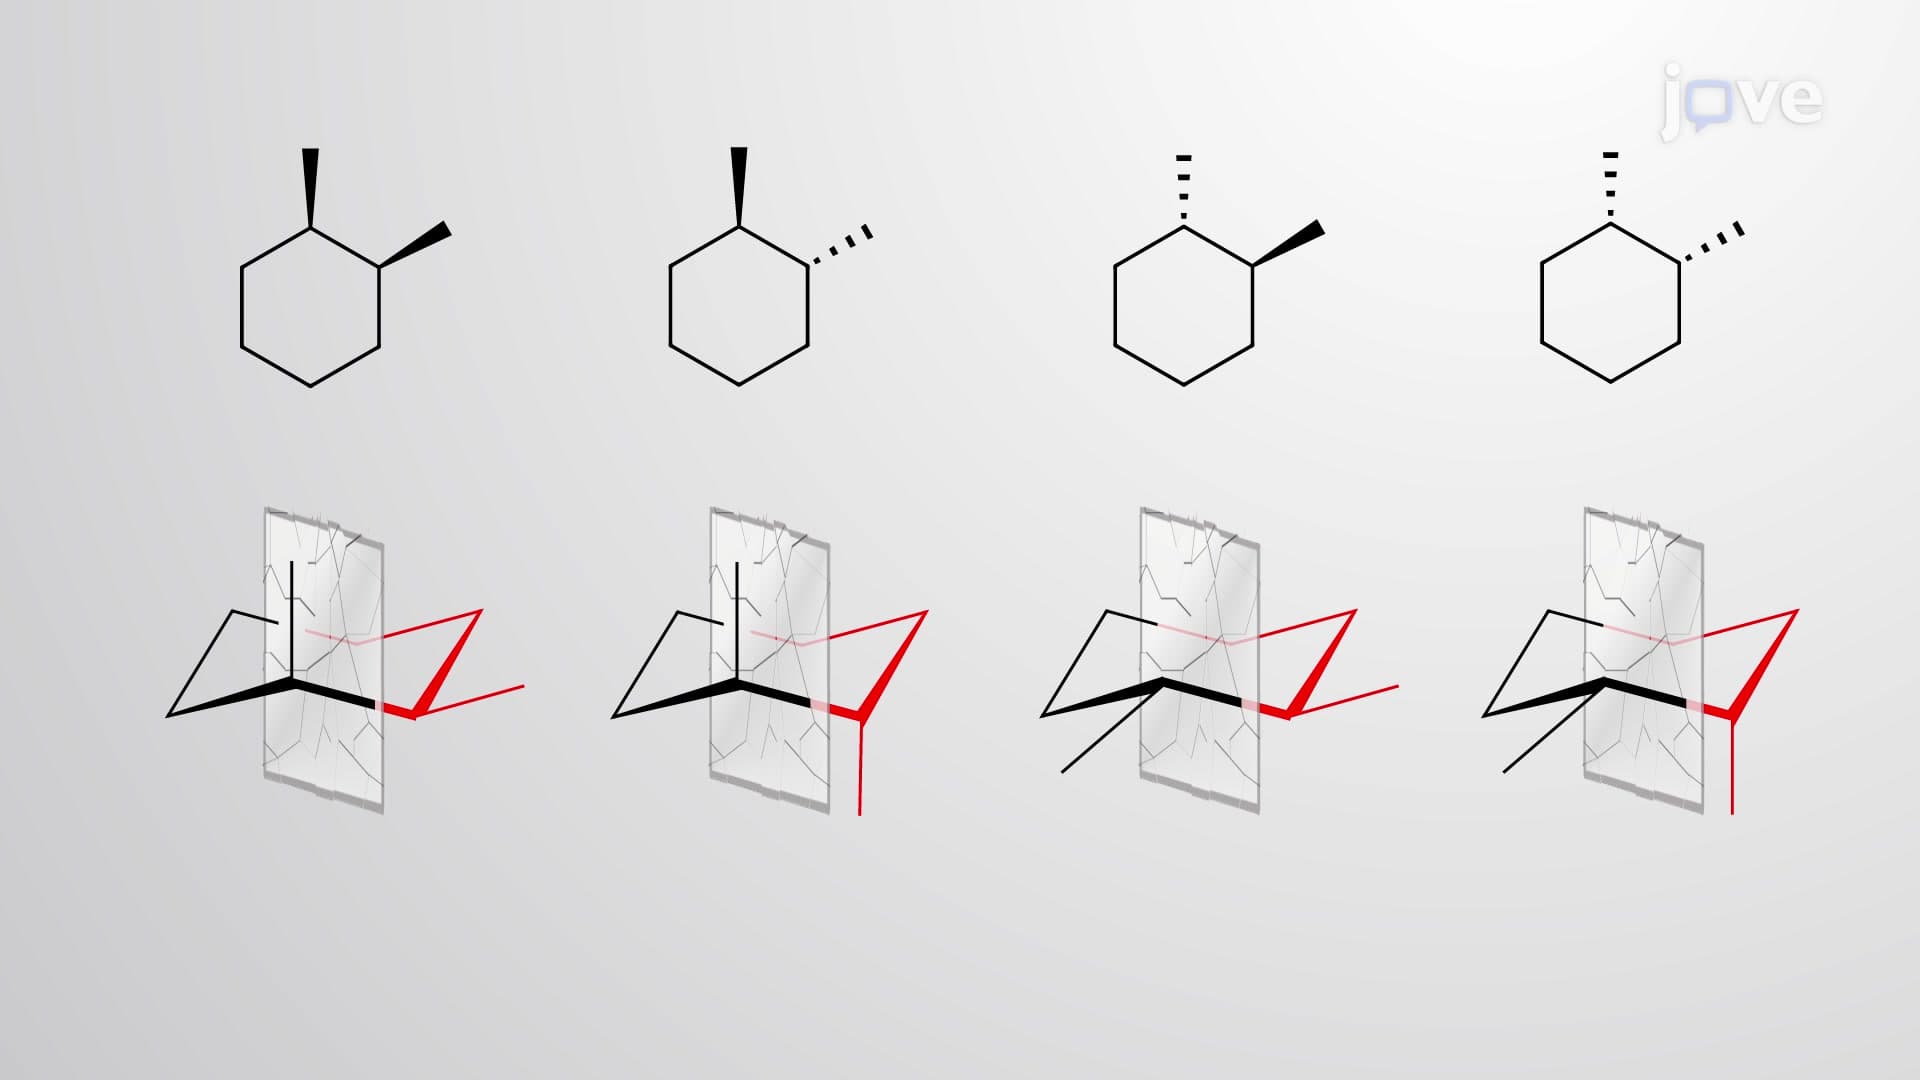
Stereoisomerism of Cyclic Compounds