Related Experiment Video
Updated: Mar 14, 2026


From Molecules to Materials: Engineering New Ionic Liquid Crystals Through Halogen Bonding
Published on: March 24, 2018
Assessment of DFT Functionals for QTAIM Topological Analysis of Halogen Bonds with Benzene
Alessandra Forni1, Stefano Pieraccini1,2, Davide Franchini2
1Istituto di Scienze e Tecnologie Molecolari, CNR (CNR-ISTM) and INSTM UdR , Via Golgi 19, 20133 Milano, Italy.

Abstract:
Halogen bonding, a noncovalent interaction between a halogen atom and a nucleophilic site, is receiving a growing attention in the chemical community stimulating a large number of theoretical investigations. The density functional theory (DFT) approach revealed to be one of the most suitable methods owing to its accuracy and low computational cost. We report here a detailed analysis of the performance of an extensive set of DFT functionals in reproducing accurate binding energies and topological properties for the halogen-bonding interaction of either NCX or PhX molecules (X = F, Cl, Br, I) with the aromatic system of benzene in the T-shaped configuration. It was found that the better performance for both sets of properties is provided by a small subset of functionals able to take into account, implicitly or explicitly (by inclusion of an additive pairwise potential), the dispersion contribution, that is, ωB97X, M06-2X, M11, mPW2PLYP-D, and B2PLYP-D3.
More Related Videos


Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding
 01:07
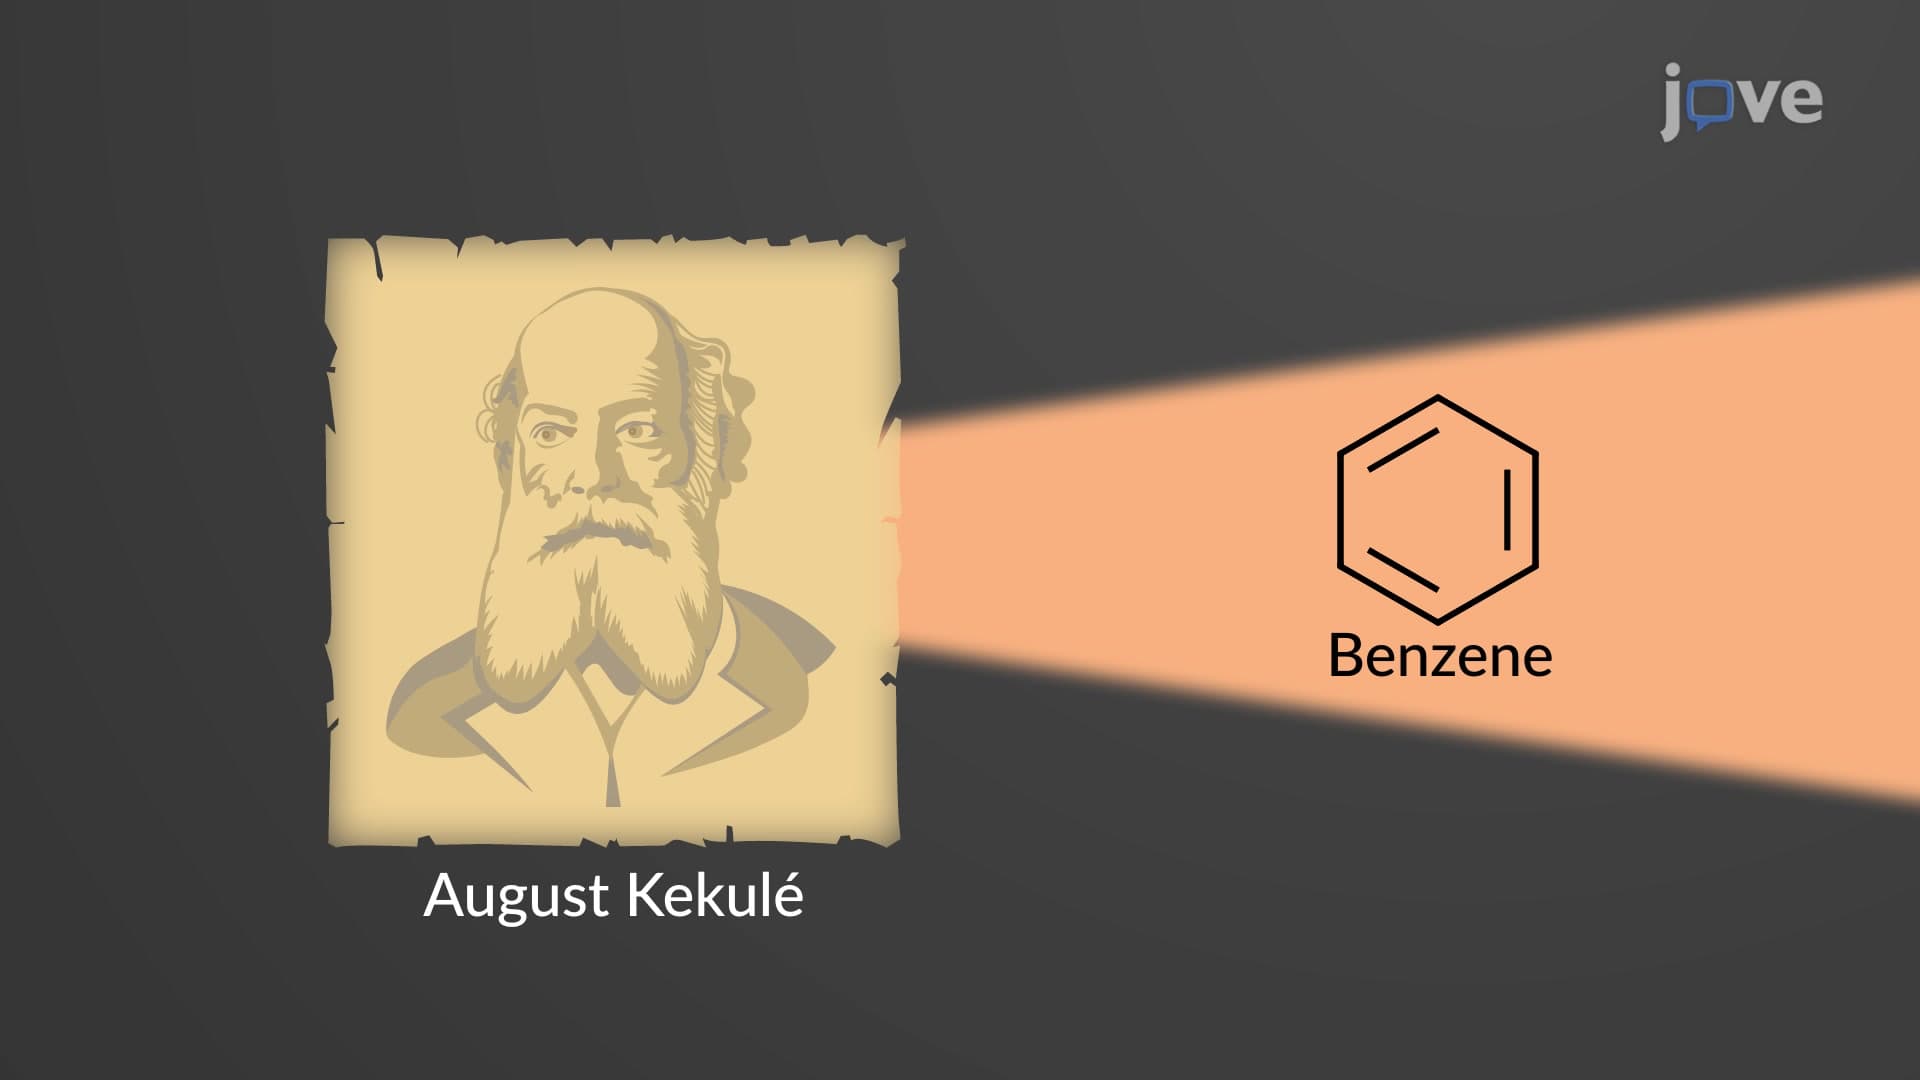
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
 01:11
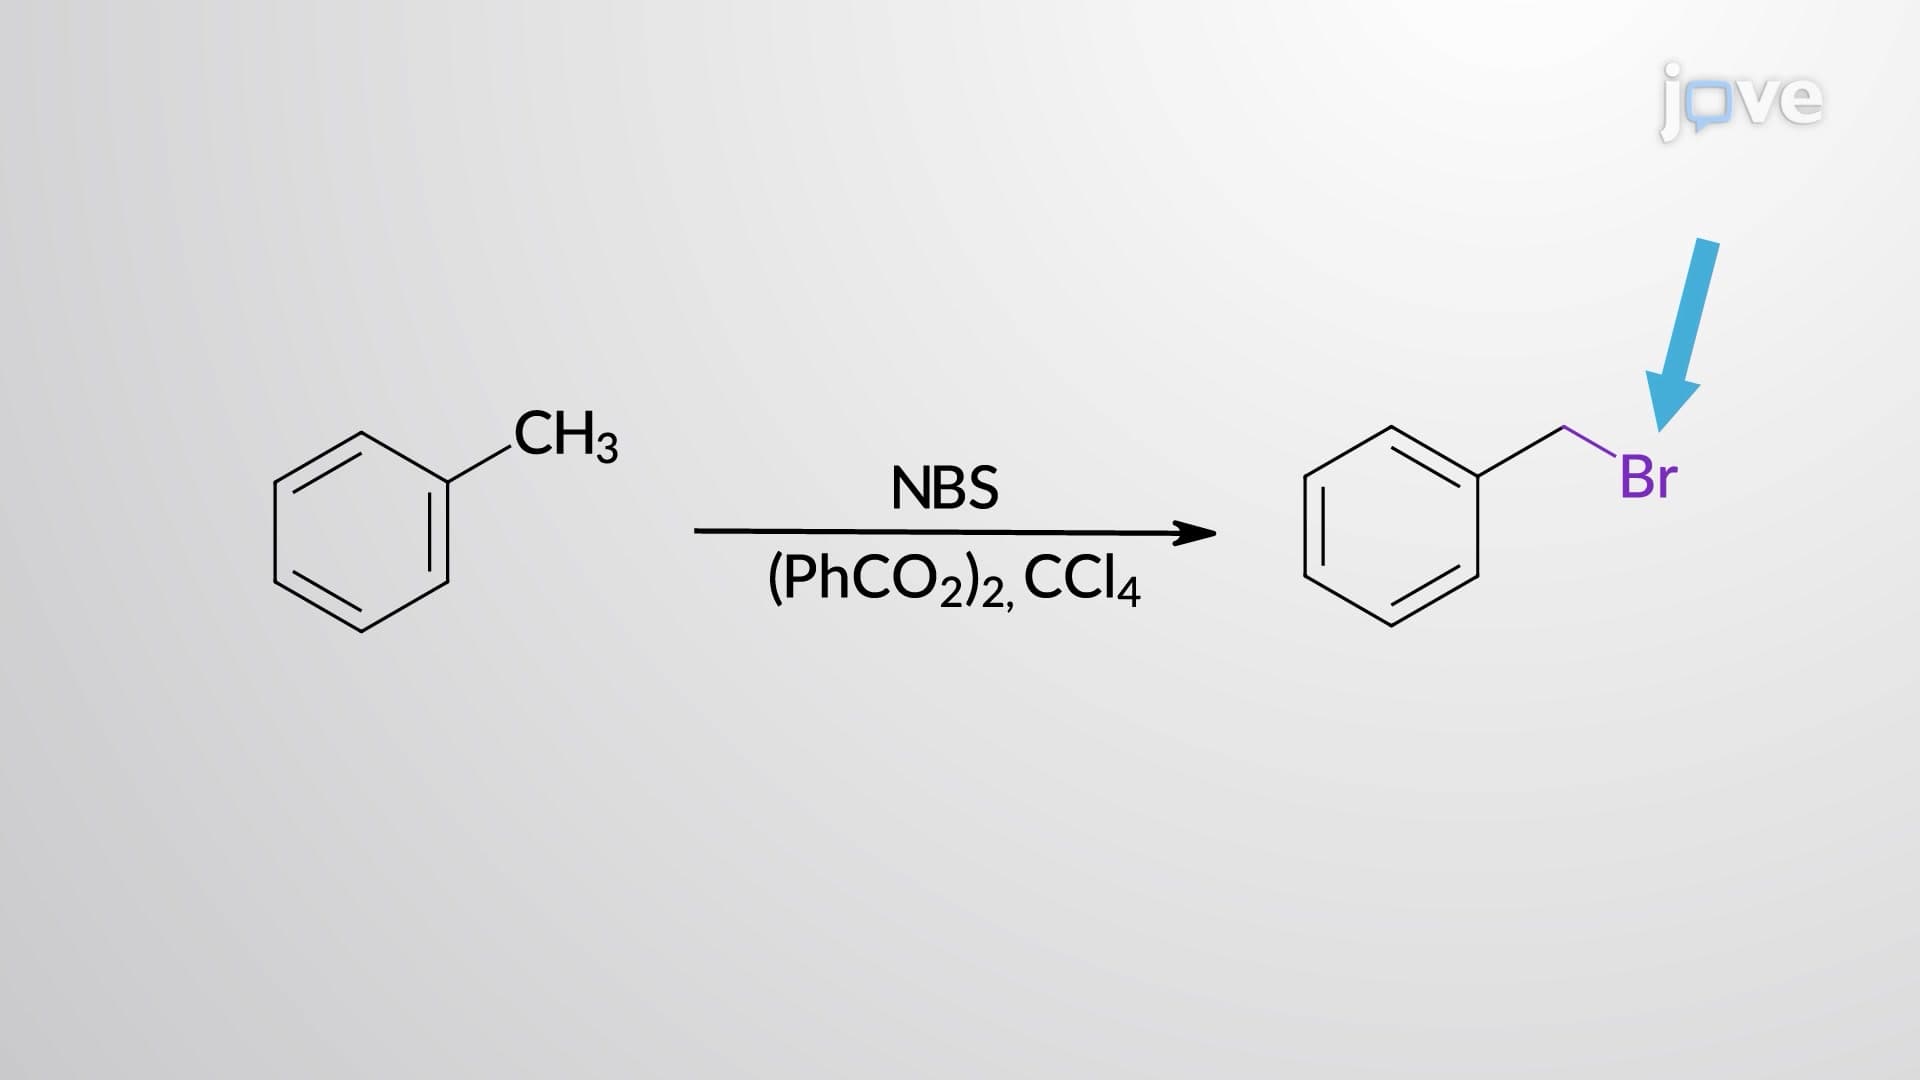
Reactions at the Benzylic Position: Halogenation
 01:15
Electrophilic Aromatic Substitution: Chlorination and Bromination of Benzene