Related Experiment Video
Updated: Mar 13, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Implementation of Constrained DFT for Computing Charge Transfer Rates within the Projector Augmented Wave Method
Marko Melander1, Elvar Ö Jónsson2, Jens J Mortensen3
1Department of Energy Conversion and Storage, Technical University of Denmark , DK-4000 Roskilde, Denmark.

Abstract:
Combining constrained density function theory (cDFT) with Marcus theory is an efficient and promising way to address charge transfer reactions. Here, we present a general and robust implementation of cDFT within the projector augmented wave (PAW) framework. PAW pseudopotentials offer a reliable frozen-core electron description across the whole periodic table, with good transferability, as well as facilitate the extraction of all-electron quantities. The present implementation is applicable to two different wave function representations, atomic-centered basis sets (LCAO) and the finite-difference (FD) approximation utilizing real-space grids. LCAO can be used for large systems, molecular dynamics, or quick initialization, while more accurate calculations are achieved with the FD basis. Furthermore, the calculations can be performed with flexible boundary conditions, ranging from isolated molecules to periodic systems in one-, two-, or three-dimensions. As such, this implementation is relevant for a wide variety of applications. We also present how to extract the electronic coupling element and reorganization energy from the resulting diabatic cDFT-PAW wave functions for the parametrization of Marcus theory. Here, the combined method is applied to important test cases where practical implementations of DFT fail due to the self-interaction error, such as the dissociation of the helium dimer cation, and it is compared to other established cDFT codes. Moreover, for charge localization in a diamine cation, where it was recently shown that the commonly used generalized gradient and hybrid functionals of DFT failed to produce the localized state, cDFT produces qualitatively and quantitatively accurate results when benchmarked against self-interaction corrected DFT and high-level CCSD(T) calculations at a fraction of the computational cost.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 01:28
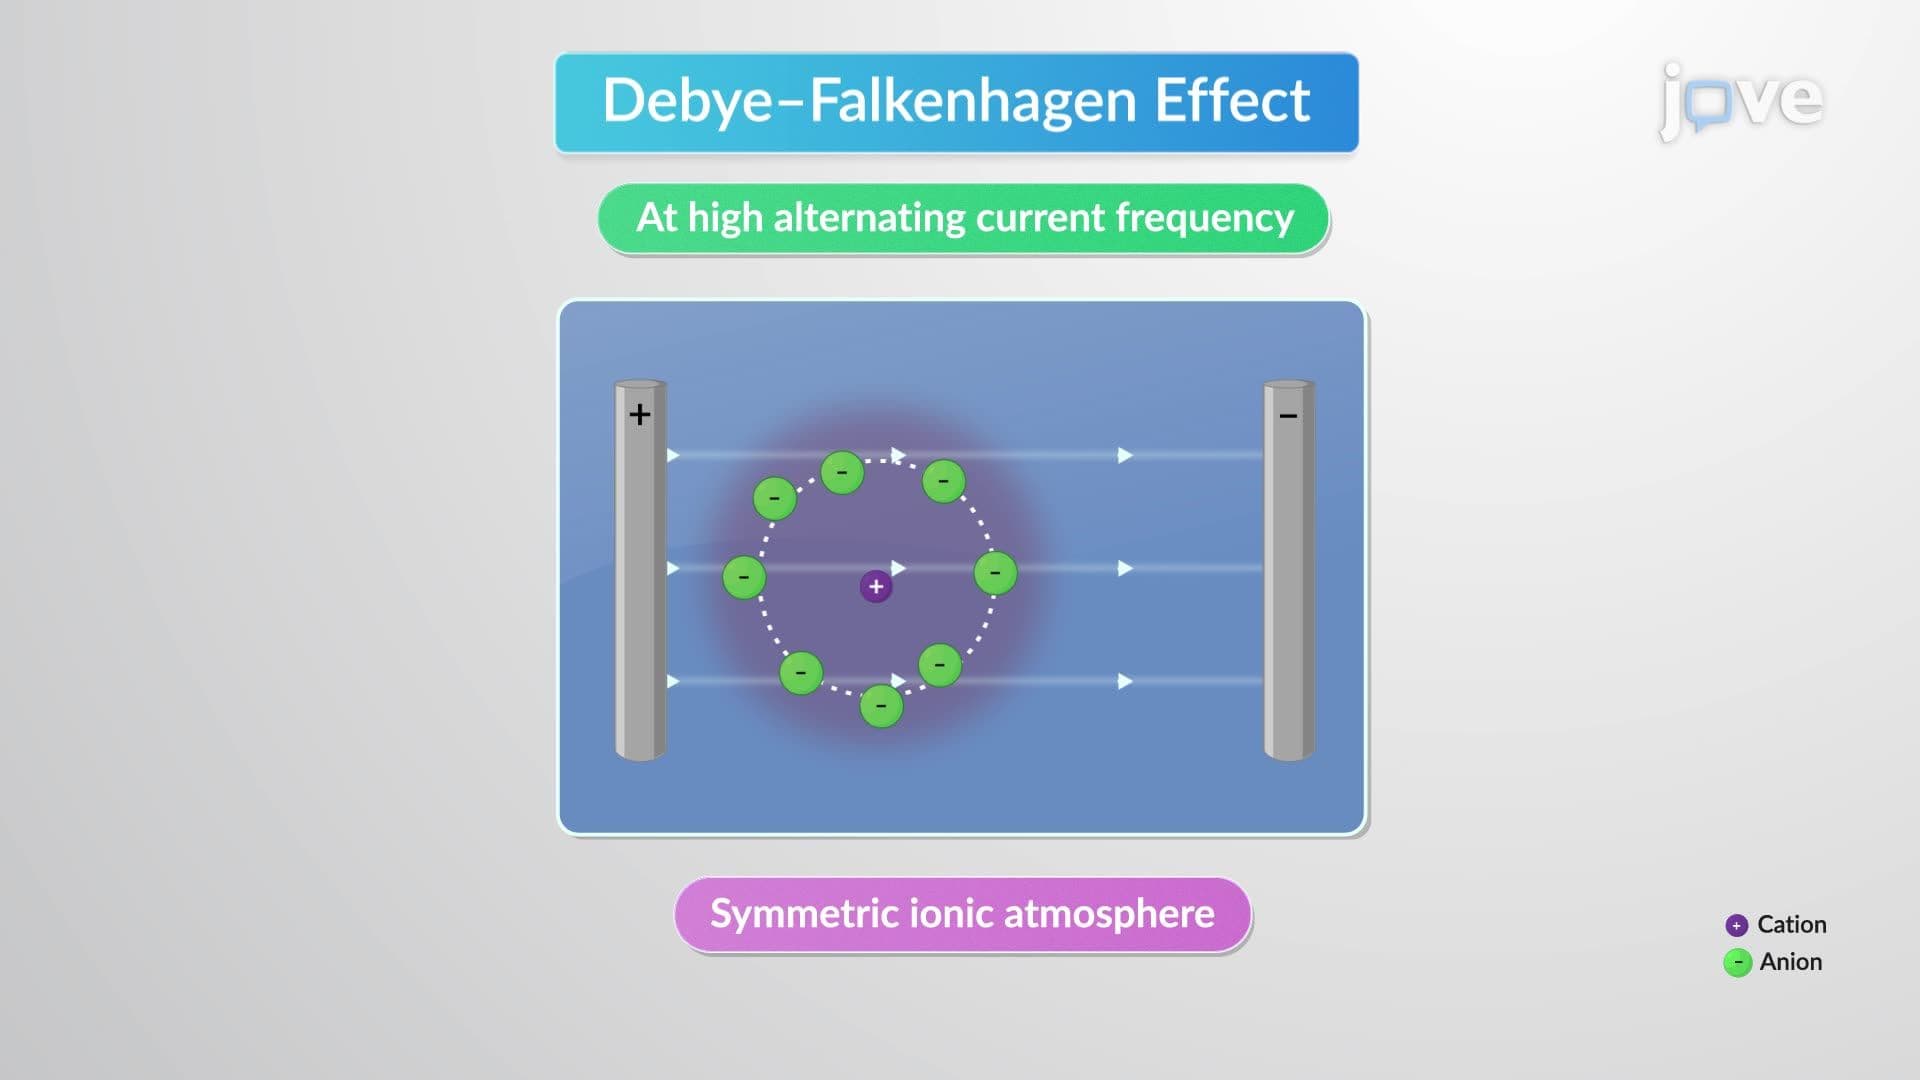
Debye–Huckel–Onsager Conductance Equation
 01:17
Continuous Charge Distributions
The electric charge can also be subjected to an analogical...
 01:30
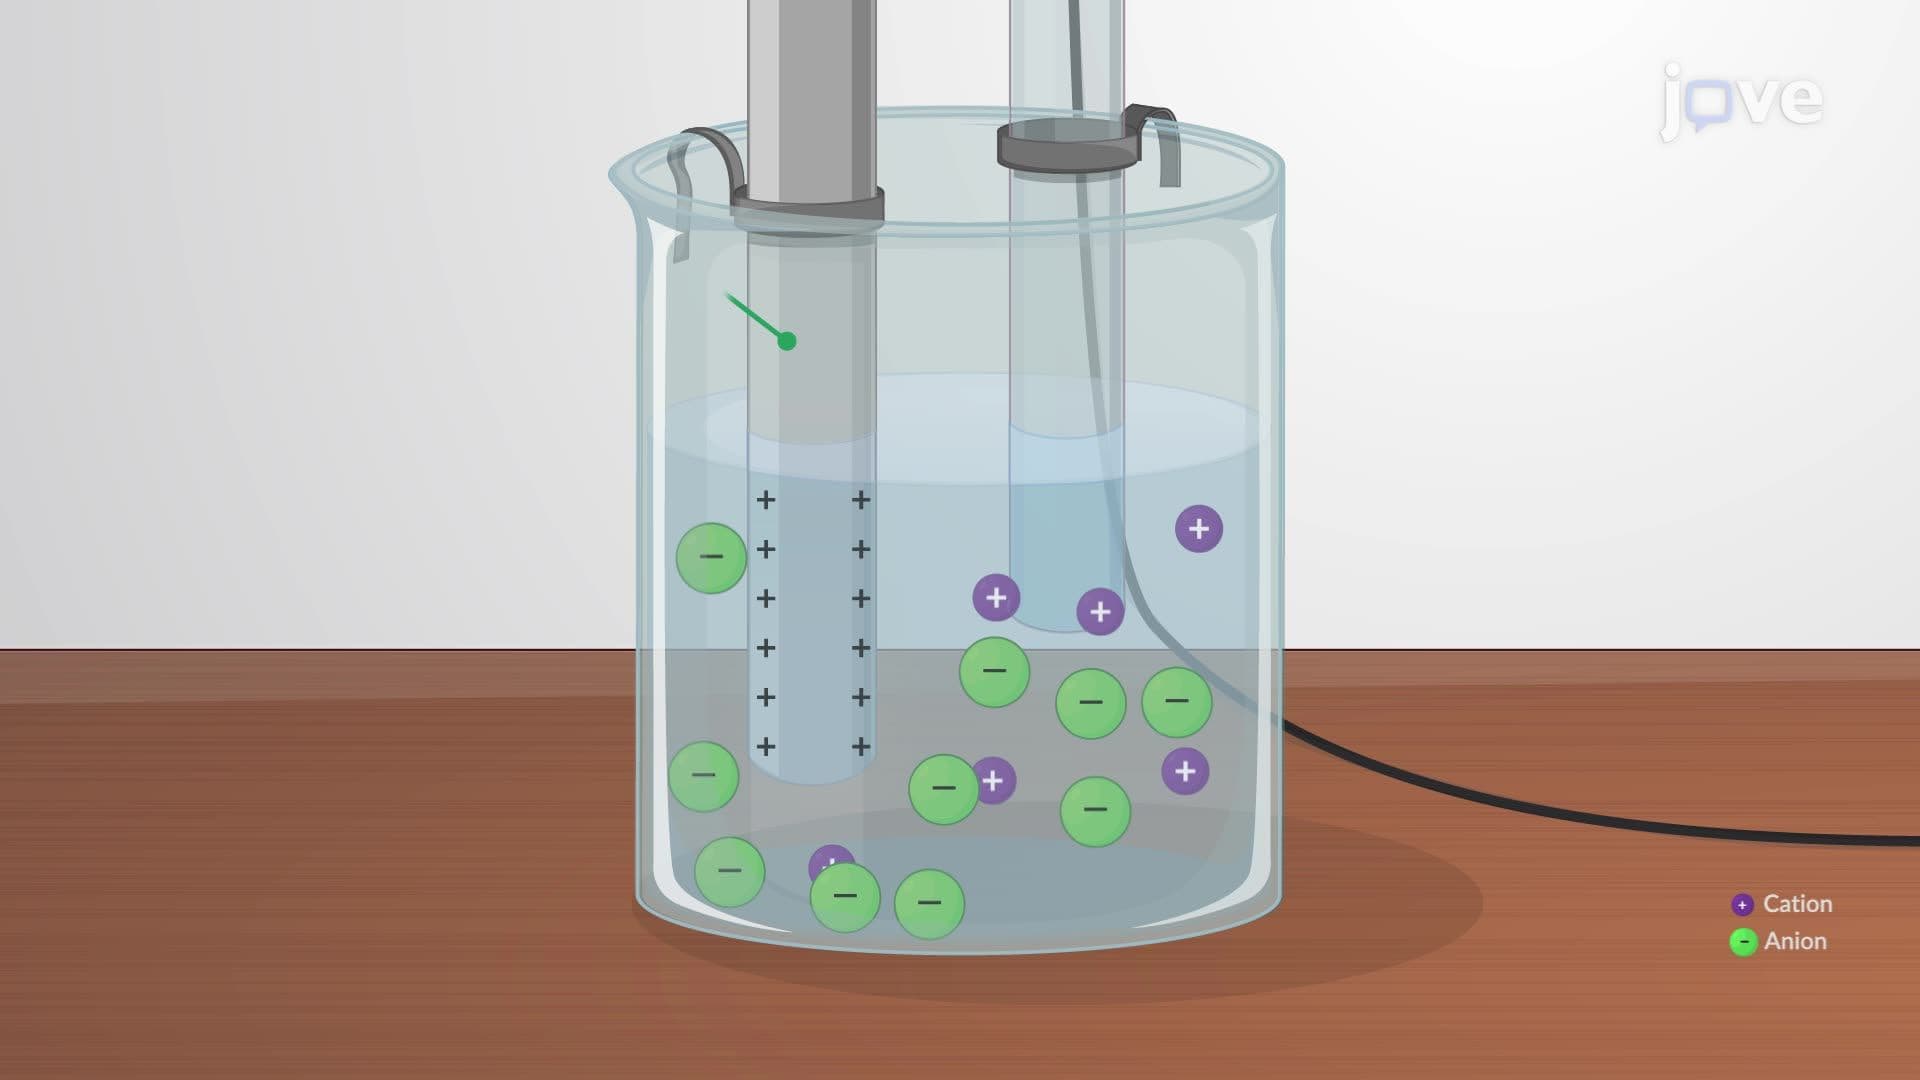
The Electrical Double Layer
 01:30
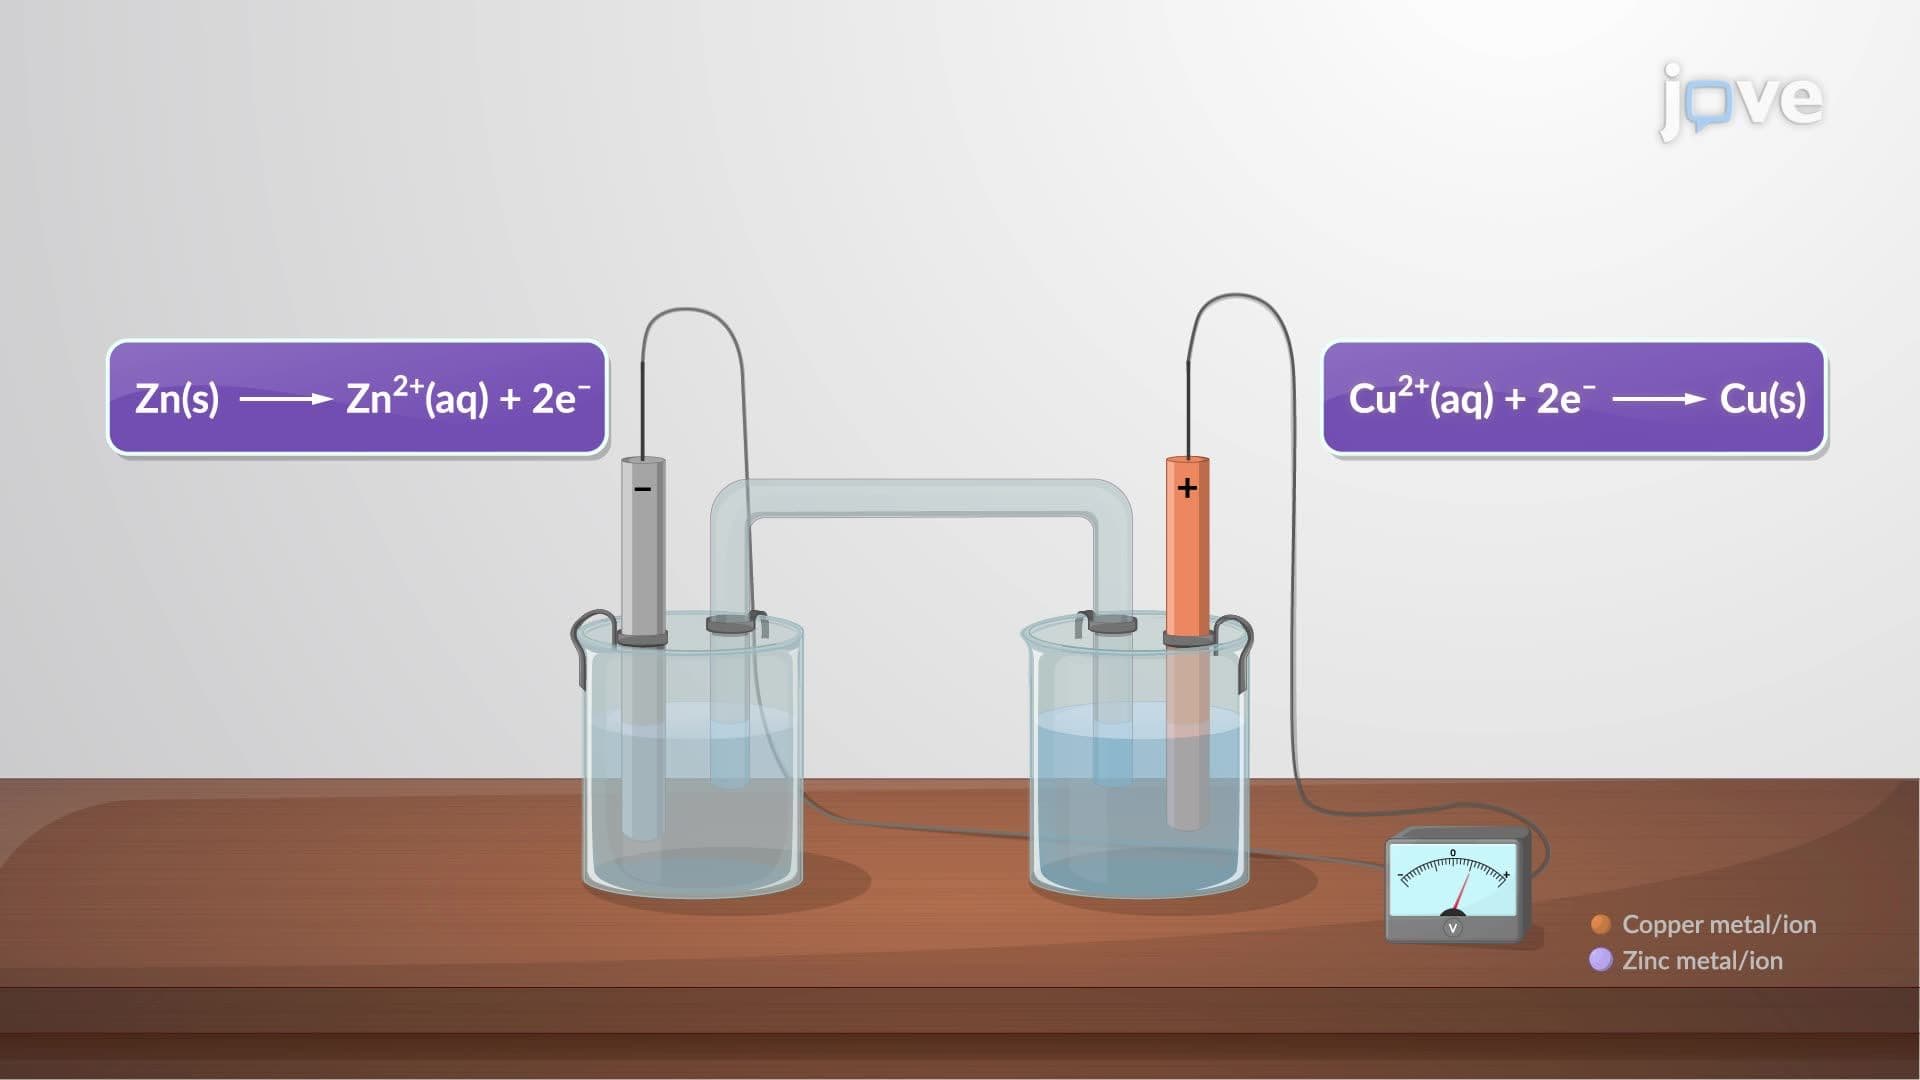
Processes at Electrodes
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:06
Interfacial Electrochemical Methods: Overview