Related Experiment Video
Updated: Mar 13, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Deprotonation of formic acid in collisions with a liquid water surface studied by molecular dynamics and metadynamics
Garold Murdachaew1, Gilbert M Nathanson, R Benny Gerber
1Laboratory of Physical Chemistry, Department of Chemistry, P. O. Box 55 (A.I. Virtasen aukio 1), FI-00014 University of Helsinki, Finland. garold.murdachaew@helsinki.fi.

Abstract:
Deprotonation of organic acids at aqueous surfaces has important implications in atmospheric chemistry and other disciplines, yet it is not well-characterized or understood. This article explores the interactions of formic acid (FA), including ionization, in collisions at the air-water interface. Ab initio molecular dynamics simulations with dispersion-corrected density functional theory were used. The 8-50 picosecond duration trajectories all resulted in the adsorption of FA within the interfacial region, with no scattering, absorption into the bulk or desorption into the vapor. Despite the known weak acidity of FA, spontaneous deprotonation of the acid was observed at the interface on a broad picosecond timescale, ranging from a few picoseconds typical for stronger acids to tens of picoseconds. Deprotonation occurred in 4% of the trajectories, and was followed by Grotthuss proton transfer through adjacent water molecules. Both sequential and ultrafast concerted proton transfer were observed. The formation of contact ion pairs and solvent-separated ion pairs, and finally the reformation of neutral FA, both trans and cis conformers, occurred in different stages of the dynamics. To better understand the deprotonation mechanisms at the interface compared with the process in bulk water, we used well-tempered metadynamics to obtain deprotonation free energy profiles. While in bulk water FA deprotonation has a free energy barrier of 14.8 kJ mol-1, in fair agreement with the earlier work, the barrier at the interface is only 7.5 kJ mol-1. Thus, at the air-water interface, FA may dissociate more rapidly than in the bulk. This finding can be rationalized with reference to the dissimilar aqueous solvation and hydrogen-bonding environments in the interface compared to those in bulk liquid water.

Related Concept Videos
 02:30
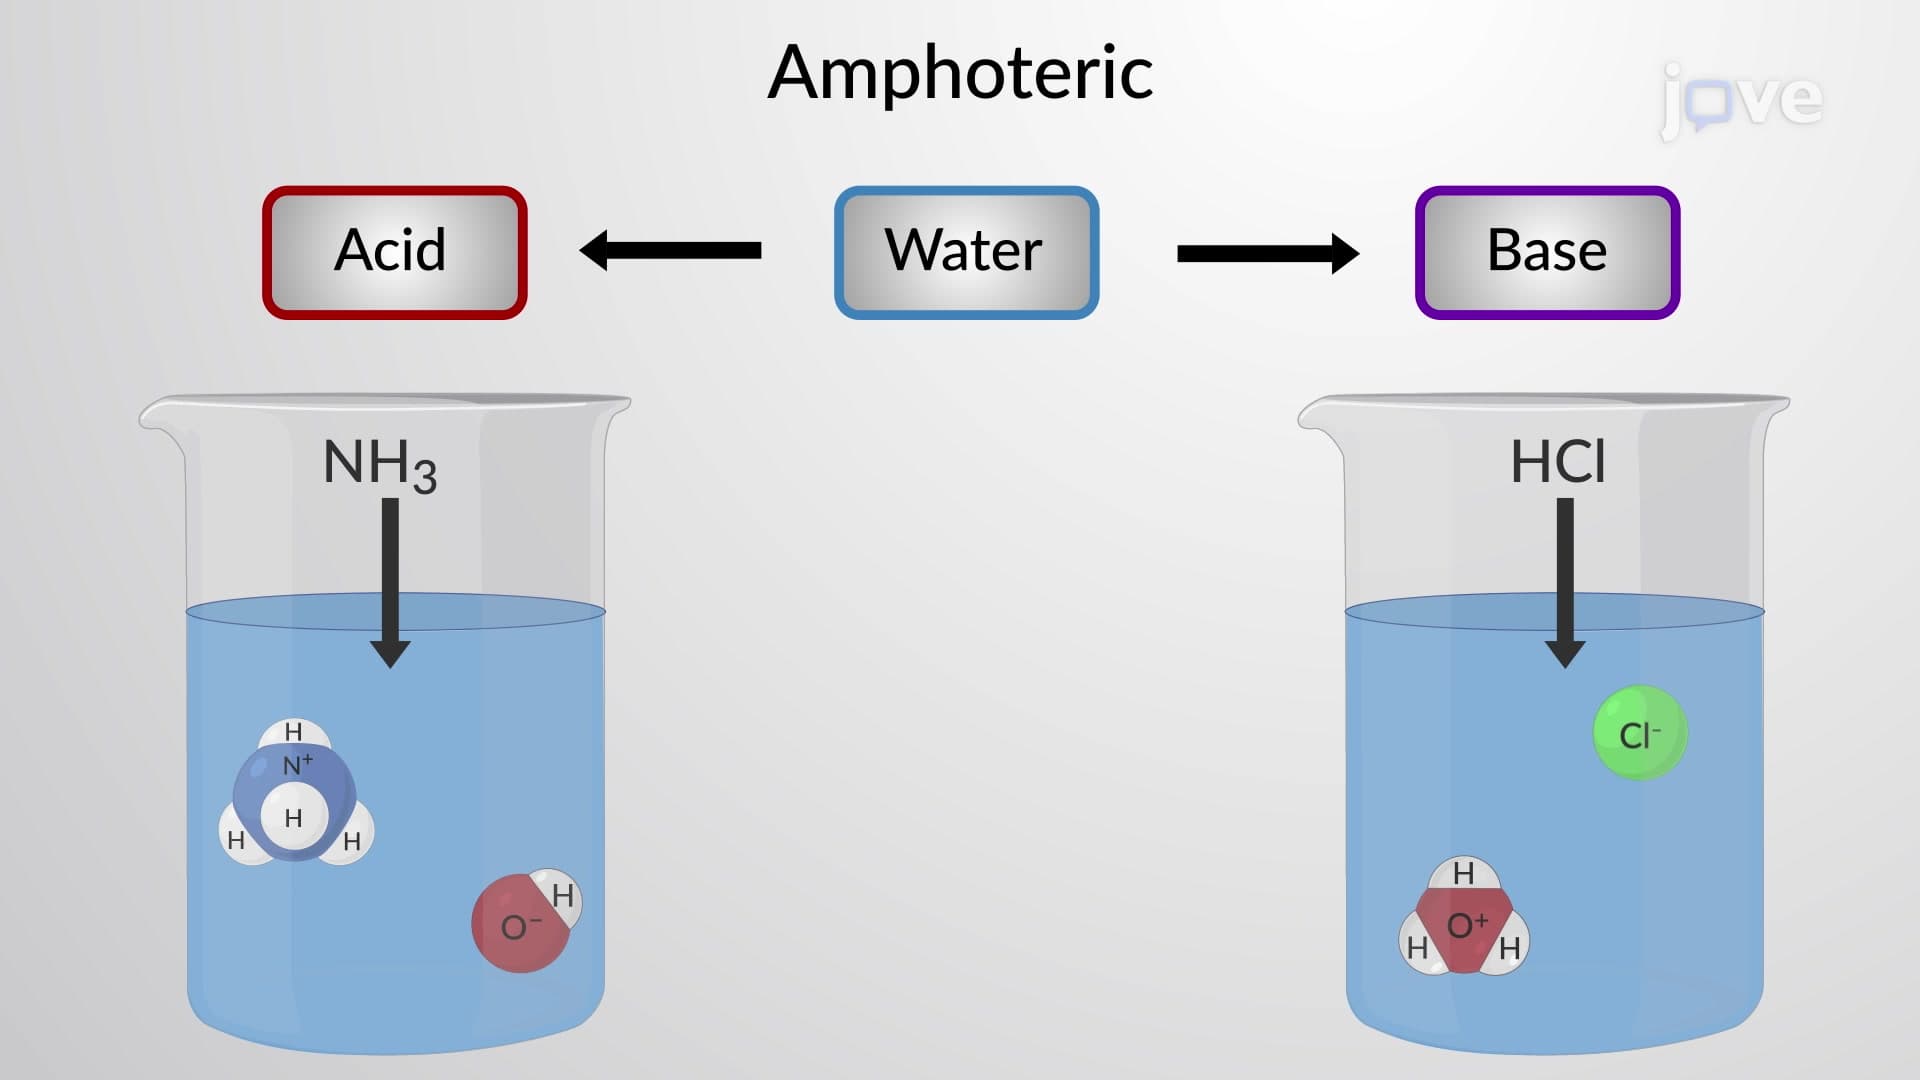
Water: A Bronsted-Lowry Acid and Base
 01:01
Acid Halides to Carboxylic Acids: Hydrolysis
As shown below, the mechanism involves a nucleophilic attack by water at the carbonyl carbon to form a tetrahedral intermediate. This is followed by the reformation of the carbon–oxygen π bond along with the departure of a halide ion. A final proton transfer step yields carboxylic...
 02:45
Acid-Catalyzed Hydration of Alkenes
 01:16
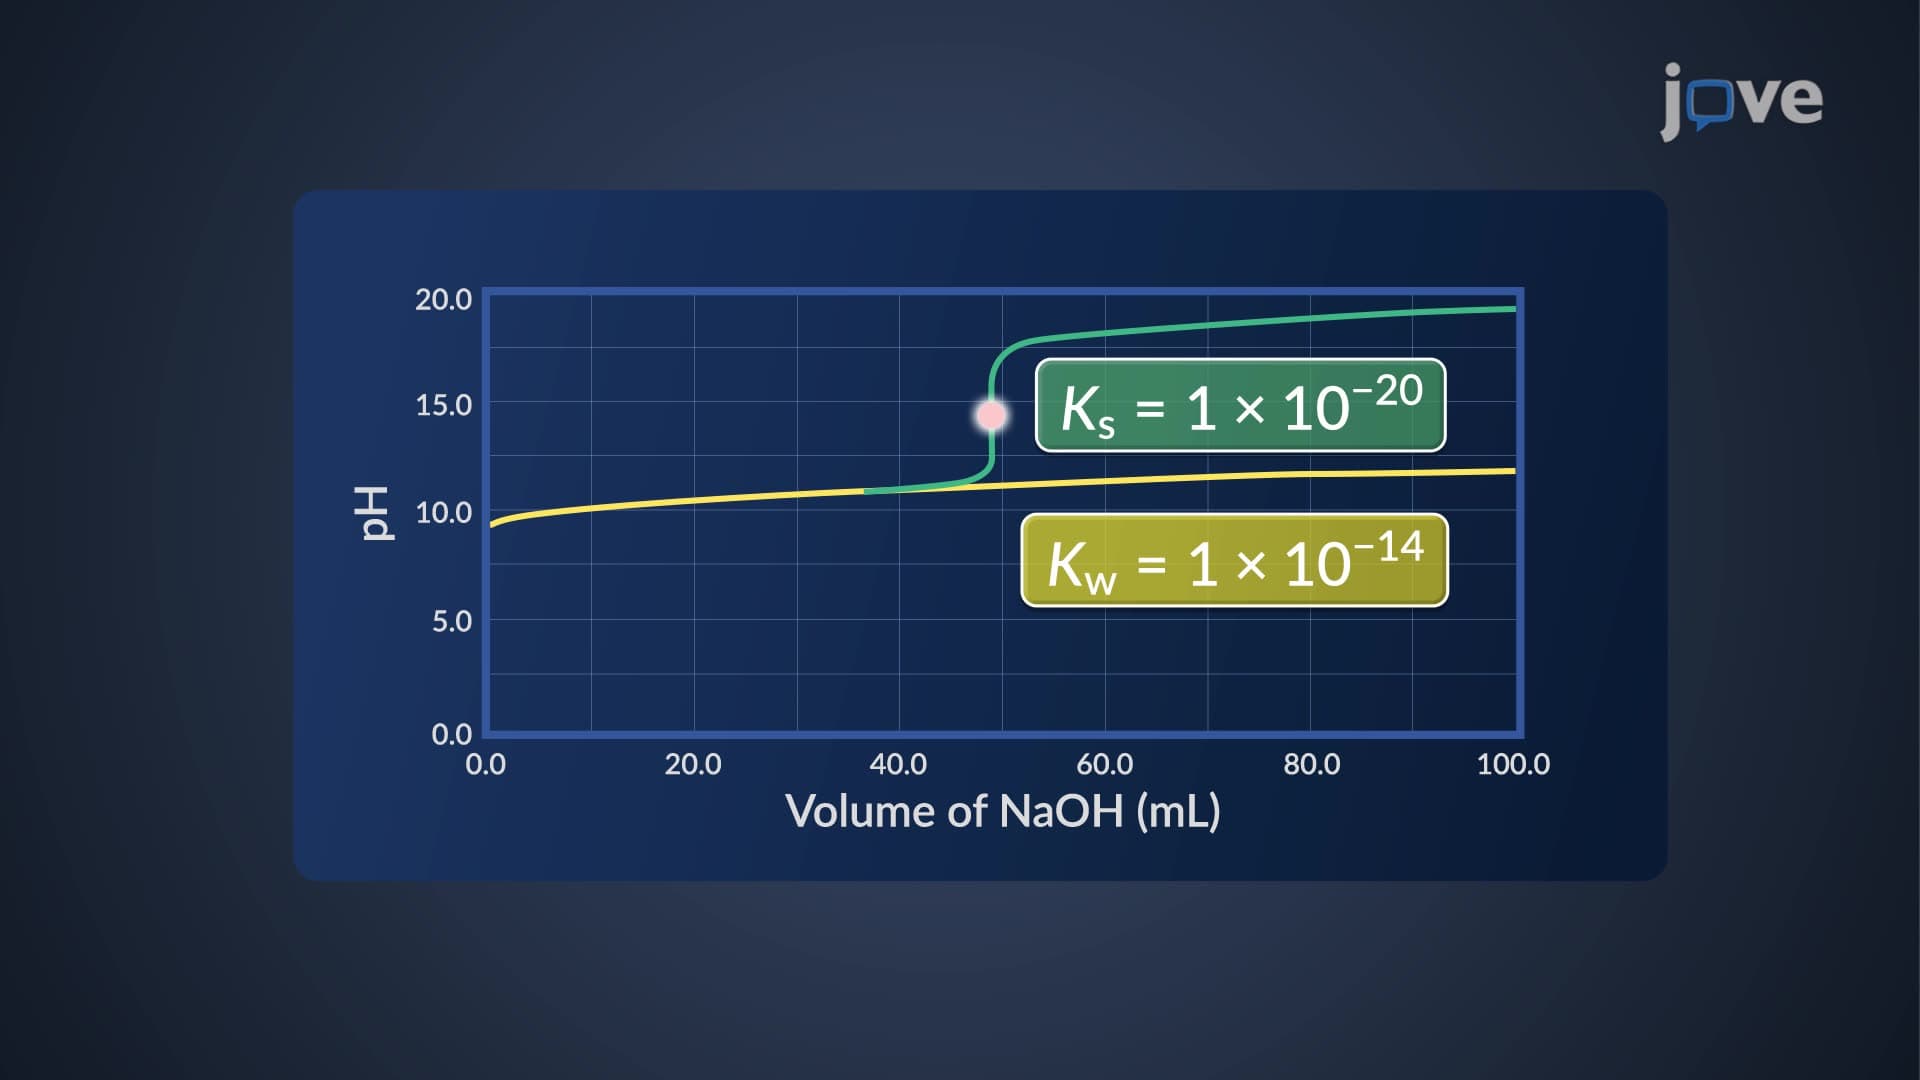
Titration in Nonaqueous Solvents
 04:02
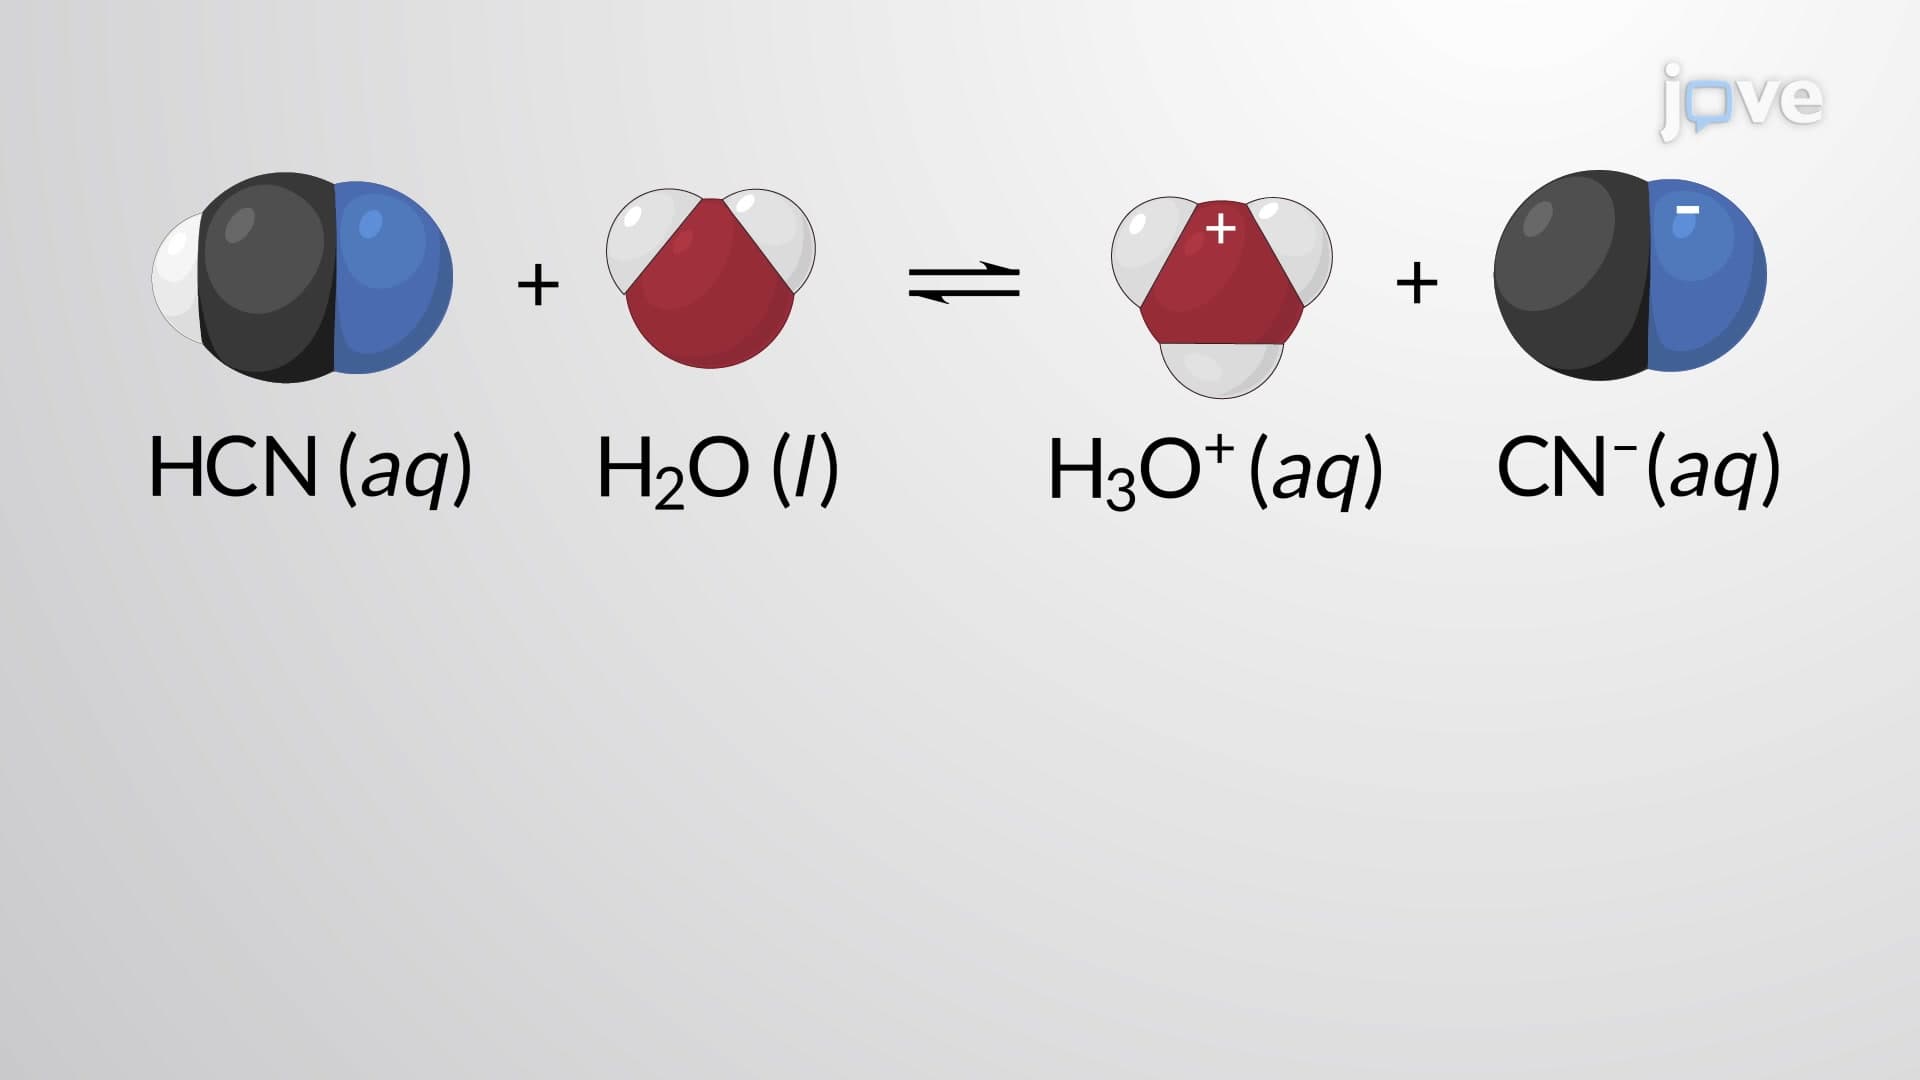
Weak Acid Solutions
 01:29
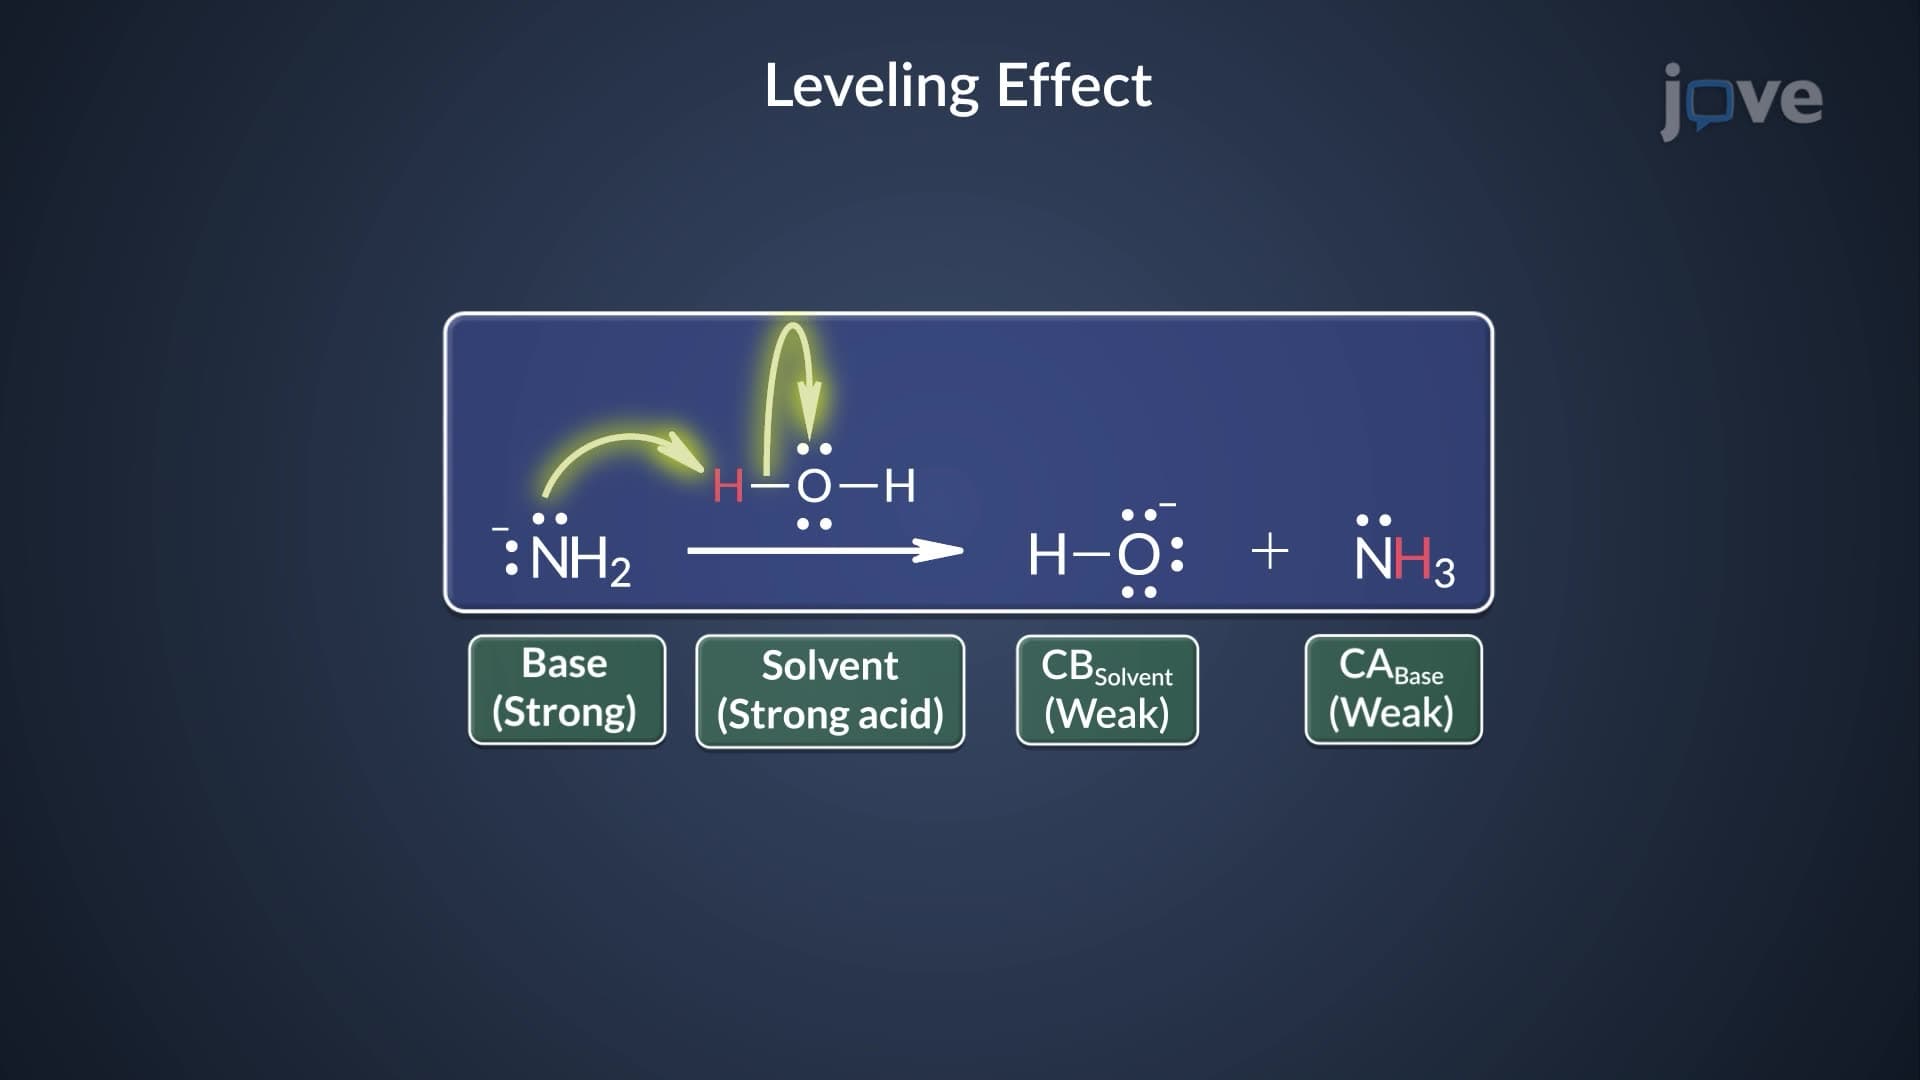
Leveling Effect