Classical molecular dynamics simulation of electronically non-adiabatic processes
William H Miller1, Stephen J Cotton1
1Department of Chemistry and Kenneth S. Pitzer Center for Theoretical Chemistry, University of California, and Chemical Sciences Division, Lawrence Berkeley National Laboratory, Berkeley, California 94720, USA. MillerWH@berkeley.edu StephenJCotton47@gmail.com.

Abstract:
Both classical and quantum mechanics (as well as hybrids thereof, i.e., semiclassical approaches) find widespread use in simulating dynamical processes in molecular systems. For large chemical systems, however, which involve potential energy surfaces (PES) of general/arbitrary form, it is usually the case that only classical molecular dynamics (MD) approaches are feasible, and their use is thus ubiquitous nowadays, at least for chemical processes involving dynamics on a single PES (i.e., within a single Born-Oppenheimer electronic state). This paper reviews recent developments in an approach which extends standard classical MD methods to the treatment of electronically non-adiabatic processes, i.e., those that involve transitions between different electronic states. The approach treats nuclear and electronic degrees of freedom (DOF) equivalently (i.e., by classical mechanics, thereby retaining the simplicity of standard MD), and provides "quantization" of the electronic states through a symmetrical quasi-classical (SQC) windowing model. The approach is seen to be capable of treating extreme regimes of strong and weak coupling between the electronic states, as well as accurately describing coherence effects in the electronic DOF (including the de-coherence of such effects caused by coupling to the nuclear DOF). A survey of recent applications is presented to illustrate the performance of the approach. Also described is a newly developed variation on the original SQC model (found universally superior to the original) and a general extension of the SQC model to obtain the full electronic density matrix (at no additional cost/complexity).
More Related Videos
 06:37
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:18
Adiabatic Processes for an Ideal Gas
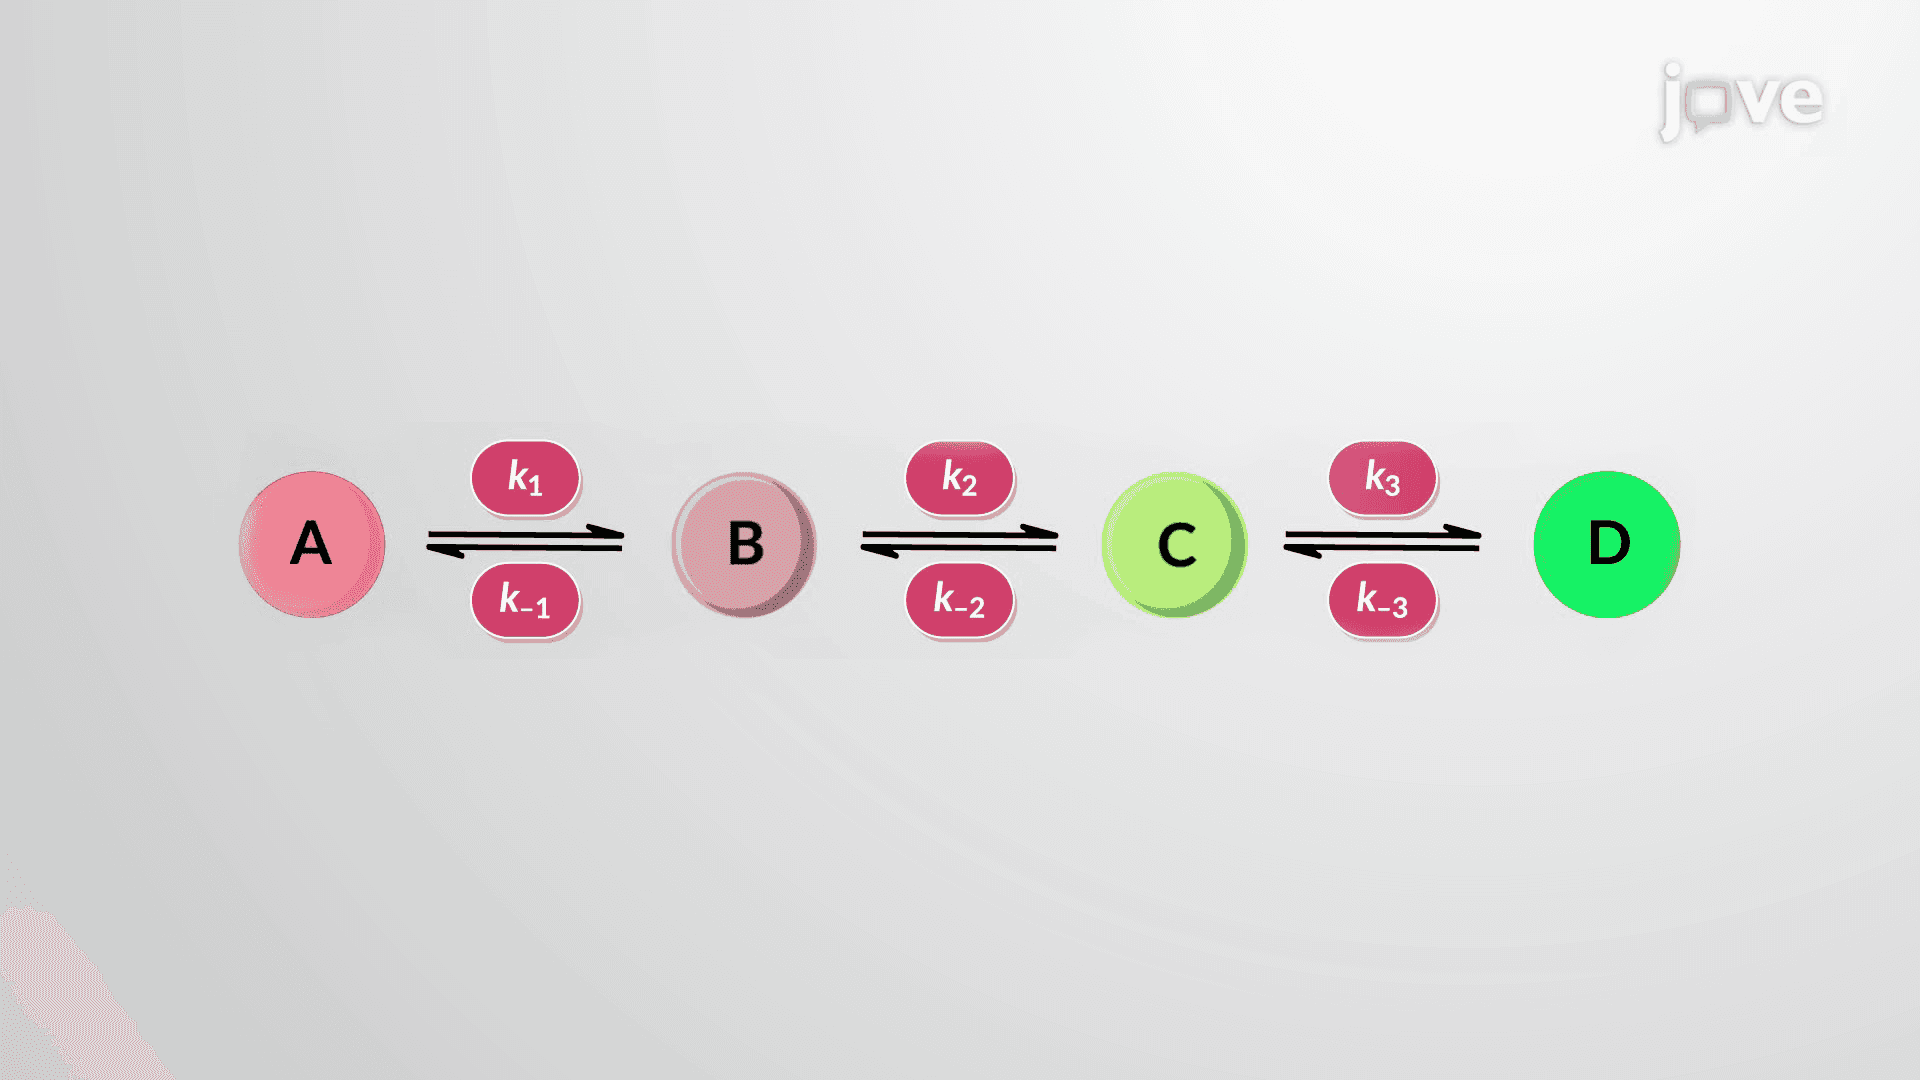 01:29
Reaction Mechanisms: Rate-limiting Step Approximation
 01:16
Thermal Sigmatropic Reactions: Overview
Sigmatropic shifts are classified based on an order term [i, j ], where i and j indicate the number of atoms across which each end of the σ bond migrates. Below are examples of a [3,3] sigmatropic shift in 1,5-hexadiene, referred...
 00:54
Electron Behavior
Electrons are negatively charged subatomic particles that are attracted to an orbit around the positively-charged nucleus of an atom. They reside in locations that are associated with energy levels called shells and are further organized into sub-shells and orbitals within each shell.
Electrons Orbit the Nucleus
Electrons are found in specific locations outside of the nucleus. The shell in which an electron resides indicates the general energy level of the electron: those closer to the...
 01:20
Work Done in an Adiabatic Process
 01:12
Classical Mechanics