Related Experiment Video
Updated: Mar 12, 2026
Functional Reconstitution and Channel Activity Measurements of Purified Wildtype and Mutant CFTR Protein
Published on: March 9, 2015
Emerging relationship between CFTR, actin and tight junction organization in cystic fibrosis airway epithelium
Stefano Castellani1, Maria Favia2, Lorenzo Guerra2
1Laboratory of Experimental and Regenerative Medicine, Department of Medical and Surgical Sciences, University of Foggia, Foggia, Italy.

Abstract:
Cystic fibrosis (CF), one of the most common genetic disorders affecting primarily Caucasians, is due to mutations in the CF Transmembrane Conductance Regulator (CFTR) gene, encoding for a chloride channel also acting as regulator of other transmembrane proteins. In healthy subjects, CFTR is maintained in its correct apical plasma membrane location via the formation of a multiprotein complex in which scaffold proteins (such as NHERF1) and signaling molecules (such as cAMP and protein kinases) guarantee its correct functioning. In CF, a disorganized and dysfunctional airway epithelium brings an altered flux of ions and water into the lumen of bronchioles, consequent bacterial infections and an enormous influx of inflammatory cells (mainly polymorphonuclear neutrophils) into the airway lumen. Recent evidence in healthy airway cells supports the notion that CFTR protein/function is strictly correlated with the actin cytoskeleton and tight junctions status. In CF cells, the most frequent CFTR gene mutation, F508del, has been shown to be associated with a disorganized actin cytoskeleton and altered tight junction permeability. Thus, the correct localization of CFTR on the apical plasma membrane domain through the formation of the scaffolding and signaling complex is likely fundamental to determine a physiological airway epithelium. The correction of CFTR mutations by either gene or drug therapies, as well as by stem cell-based interventions, can determine the resumption of a physiological organization of actin stress fibers and TJ structure and barrier function, further indicating the close interrelationship among these processes.
Related Concept Videos
 01:29
Tight Junctions
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:10
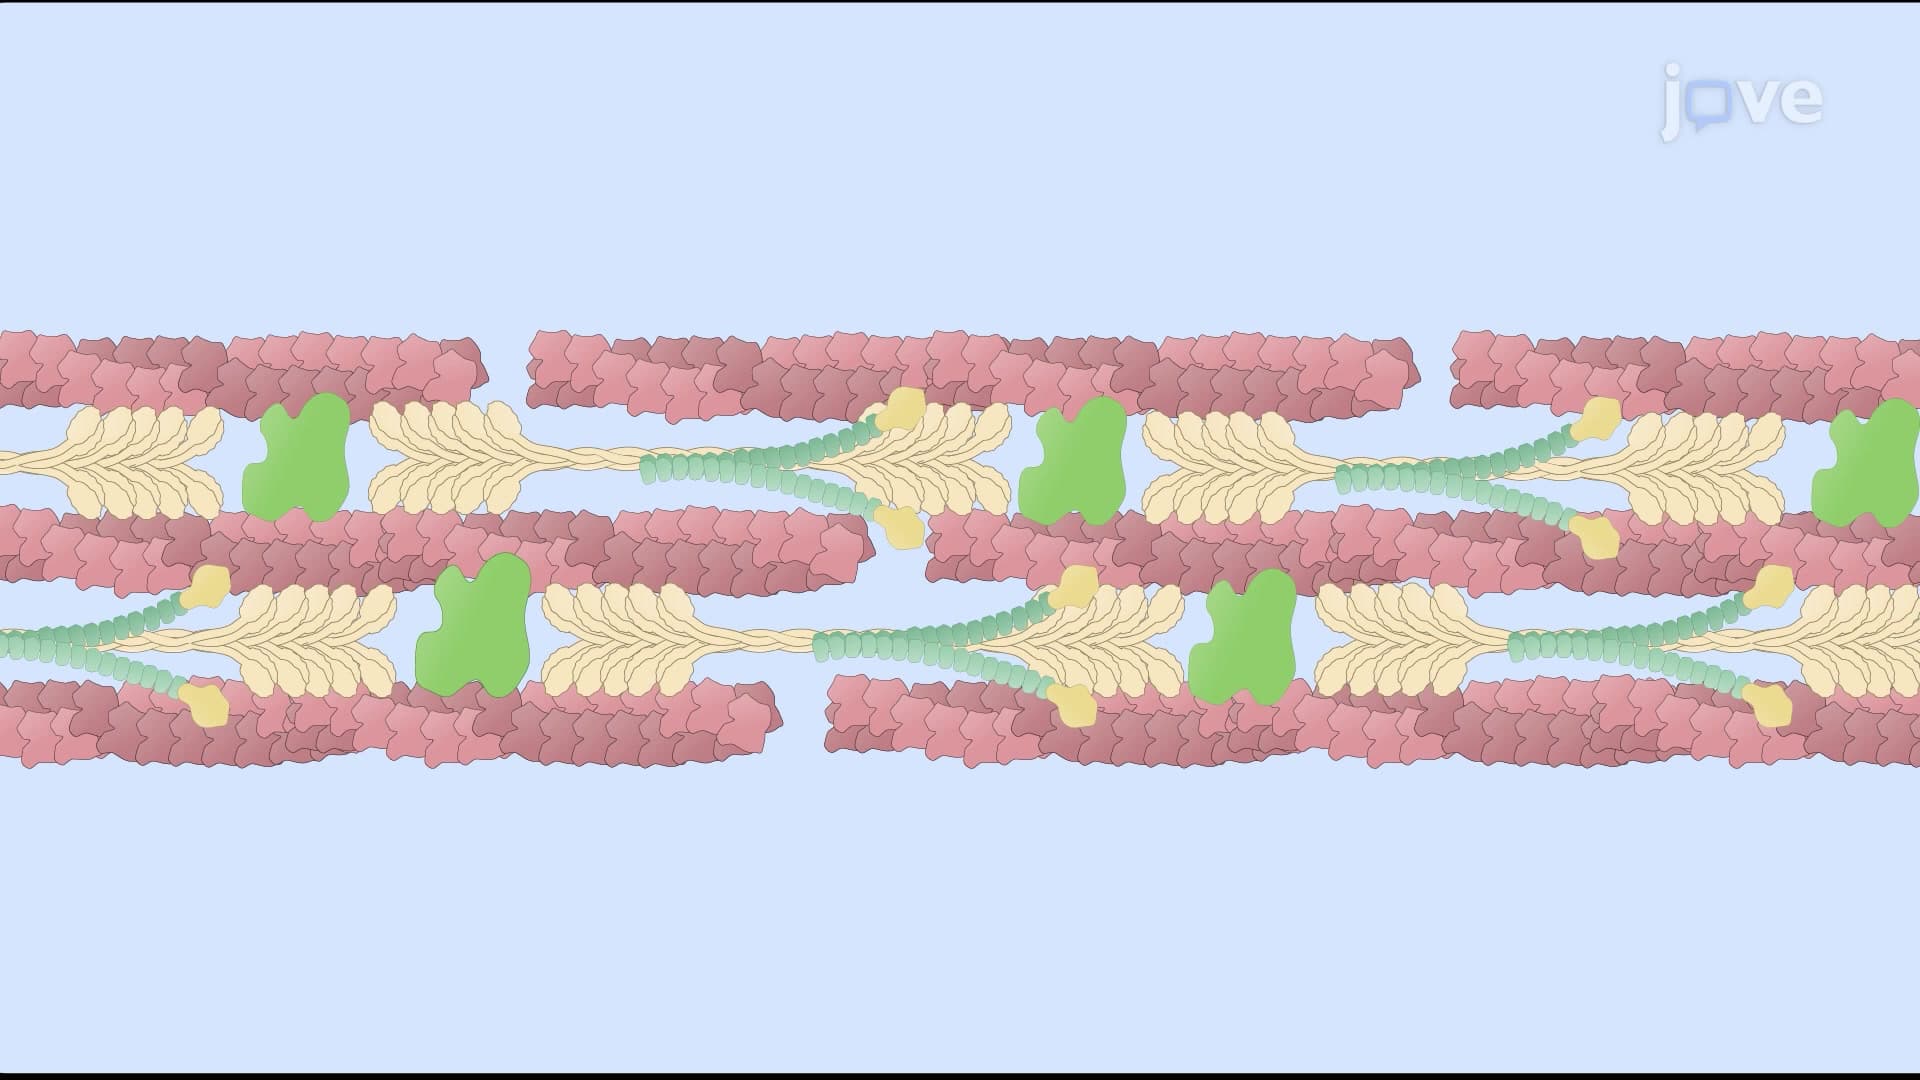
The Role of Actin and Myosin in Non-muscle Cells
 01:26
Tension Response at Adherens Junctions
α-Catenin as a Mechanosensory Protein
The α-catenin of adherens junctions is an allosteric protein with three VH (vinculin...
 01:17
Intracellular Signaling Affects Focal Adhesions
Some...