Related Experiment Video
Updated: Mar 12, 2026

Quantification of Coenzyme A in Cells and Tissues
Published on: September 27, 2019
Screening of MCAD deficiency in Japan: 16years' experience of enzymatic and genetic evaluation
Go Tajima1, Keiichi Hara2, Miyuki Tsumura3
1Department of Pediatrics, Hiroshima University Graduate School of Biomedical & Health Sciences, 1-2-3 Kasumi, Minami-ku, Hiroshima 734-8551, Japan; Division of Neonatal Screening, Research Institute, National Center for Child Health and Development, 2-10-1 Okura, Setagaya-ku, Tokyo 157-8535, Japan.

Background:
Medium-chain acyl-CoA dehydrogenase (MCAD) deficiency is a representative disorder of fatty acid oxidation and is one of the most prevalent inborn errors of metabolism among Caucasian populations. In Japan, however, it was as late as 2000 when the first patient was found, and enzymatic and genetic evaluation of MCAD deficiency began.
Methods:
We measured octanoyl-CoA dehydrogenase activity in lymphocytes of symptomatic children and newborn screening (NBS)-positive subjects who showed elevated levels of C8-acylcarnitine in blood. The results were further confirmed by direct sequencing of the ACADM gene.
Results:
The disease was diagnosed in 9 out of 18 symptomatic children. The affected patients showed residual activities from 0% to 3% of the normal average value, except for one patient with 10% activity. Concerning 50 NBS-positive subjects, 18 with enzymatic activities around 10% or lower and 14 with activities ranging from 13% to 30% were judged to be affected patients, and biallelic variants were detected in most of the cases tested. Newborns with higher enzymatic activities were estimated to be heterozygous carriers or healthy subjects, though biallelic variants were detected in 5 of them. Genetic analysis detected 22 kinds of variant alleles. The most prevalent was c.449_452delCTGA (p.T150Rfs), which was followed by c.50G>A (p.R17H), c.1085G>A (p.G362E), c.157C>T (p.R53C), and c.843A>T (p.R281S); these five variants accounted for approximately 60% of all the alleles examined.
Conclusion:
Our study has revealed the unique genetic backgrounds of MCAD deficiency among Japanese, based on the largest series of non-Caucasian cases. A continuous spectrum of severity was also observed in our series of NBS-positive cases, suggesting that it is essential for every nation and ethnic group to accumulate its own information on gene variants, together with their enzymatic evaluation, in order to establish an efficient NBS system for MCAD deficiency.
More Related Videos
 05:58
Digital Polymerase Chain Reaction Assay for the Genetic Variation in a Sporadic Familial Adenomatous Polyposis Patient Using the Chip-in-a-tube Format
Published on: August 20, 2018
 09:28
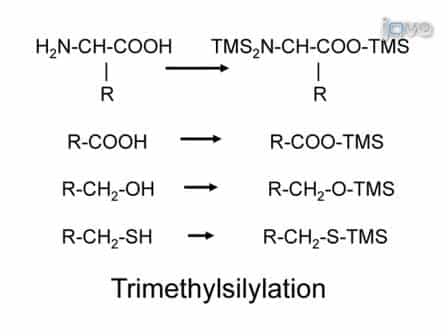
One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 02:59
Animal Mitochondrial Genetics
 01:15
Myasthenia Gravis: Diagnostic Tests
The edrophonium test is a diagnostic tool for myasthenia gravis. It involves...