Related Experiment Video
Updated: Mar 11, 2026
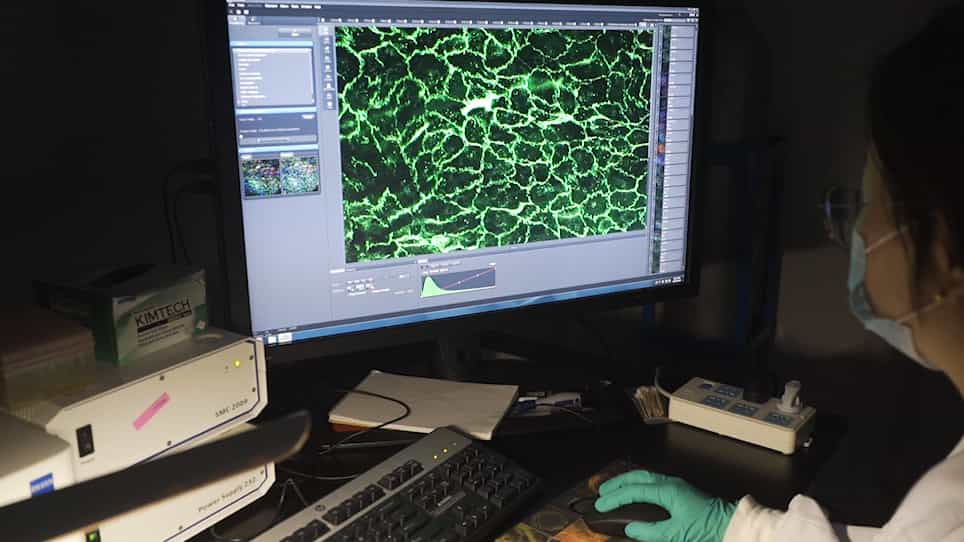

Using En Face Immunofluorescence Staining to Observe Vascular Endothelial Cells Directly
Published on: August 20, 2019
CLIC1 Inhibition Attenuates Vascular Inflammation, Oxidative Stress, and Endothelial Injury
Yingling Xu1, Ji Zhu2, Xiao Hu1
1College of Life Science, Zhejiang Chinese Medical University, Hangzhou, China.

Abstract:
Endothelial dysfunction, which includes endothelial oxidative damage and vascular inflammation, is a key initiating step in the pathogenesis of atherosclerosis (AS) and an independent risk factor for this disorder. Intracellular chloride channel 1 (CLIC1), a novel metamorphic protein, acts as a sensor of cell oxidation and is involved in inflammation. In this study, we hypothesize that CLIC1 plays an important role in AS. Apolipoprotein E-deficient mice were supplied with a normal diet or a high-fat and high-cholesterol diet for 8 weeks. Overexpressed CLIC1 was associated with the accelerated atherosclerotic plaque development, amplified oxidative stress, and in vivo release of inflammatory cytokines. We subsequently examined the underlying molecular mechanisms through in vitro experiments. Treatment of cultured human umbilical vein endothelial cells (HUVECs) with H2O2 induced endothelial oxidative damage and enhanced CLIC1 expression. Suppressing CLIC1 expression through gene knocked-out (CLIC1-/-) or using the specific inhibitor indanyloxyacetic acid-94 (IAA94) reduced ROS production, increased SOD enzyme activity, and significantly decreased MDA level. CLIC1-/- HUVECs exhibited significantly reduced expression of TNF-α and IL-1β as well as ICAM-1 and VCAM-1 at the protein levels. In addition, H2O2 promoted CLIC1 translocation to the cell membrane and insertion into lipid membranes, whereas IAA94 inhibited CLIC1 membrane translocation induced by H2O2. By contrast, the majority of CLIC1 did not aggregate on the cell membrane in normal HUVECs, and this finding is consistent with the changes in cytoplasmic chloride ion concentration. This study demonstrates for the first time that CLIC1 is overexpressed during AS development both in vitro and in vivo and can regulate the accumulation of inflammatory cytokines and production of oxidative stress. Our results also highlight that deregulation of endothelial functions may be associated with the membrane translocation of CLIC1 and active chloride-selective ion channels in endothelial cells.
Insights
Intracellular chloride channel 1 (CLIC1) is overexpressed in atherosclerosis, promoting oxidative stress and inflammation. Inhibiting CLIC1 reduces these effects, suggesting it
Area of Science:
- Cardiovascular Biology
- Molecular Medicine
- Cellular Physiology
Background:
- Endothelial dysfunction, characterized by oxidative damage and inflammation, initiates atherosclerosis (AS).
- Intracellular chloride channel 1 (CLIC1) is a novel protein involved in cellular oxidation and inflammation.
- The role of CLIC1 in AS pathogenesis remains largely unexplored.
Purpose of the Study:
- To investigate the role of CLIC1 in the development of atherosclerosis.
- To elucidate the molecular mechanisms by which CLIC1 influences endothelial dysfunction and AS.
Main Methods:
- Apolipoprotein E-deficient mice fed high-fat diets to induce AS.
- In vitro studies using human umbilical vein endothelial cells (HUVECs) treated with H2O2.
- Gene knockout (CLIC1-/-) and pharmacological inhibition (IAA94) of CLIC1.
- Assessment of oxidative stress markers (ROS, SOD, MDA), inflammatory cytokines (TNF-α, IL-1β), and adhesion molecules (ICAM-1, VCAM-1).
Main Results:
- Overexpressed CLIC1 correlated with accelerated AS plaque development, increased oxidative stress, and elevated inflammatory cytokines in vivo.
- H2O2-induced endothelial oxidative damage enhanced CLIC1 expression and membrane translocation in HUVECs.
- CLIC1 suppression (CLIC1-/- or IAA94) reduced oxidative stress, inflammatory markers, and adhesion molecules, while inhibiting H2O2-induced CLIC1 membrane translocation.
Conclusions:
- CLIC1 is significantly overexpressed in atherosclerosis and contributes to its pathogenesis by regulating oxidative stress and inflammation.
- CLIC1 membrane translocation and chloride channel activity in endothelial cells are implicated in endothelial dysfunction.
- Targeting CLIC1 may offer a novel therapeutic strategy for atherosclerosis.
Related Concept Videos
 01:24
Regulation of Angiogenesis and Blood Supply
 01:28
Nitric Oxide Signaling Pathway
 01:30
Atherosclerosis I: Introduction
 01:29
Hypertension II: Pathophysiology
 01:30
Inflammatory Response I: Vascular and Cellular
 01:26
Atherosclerosis III: Management