Related Experiment Video
Updated: Mar 11, 2026

One-pot Microwave-assisted Conversion of Anomeric Nitrate-esters to Trichloroacetimidates
Published on: January 15, 2018
Organocatalyzed asymmetric Michael addition by an efficient bifunctional carbohydrate-thiourea hybrid with
Chandra S Azad1, Imran A Khan, Anudeep K Narula
1"Hygeia" Centre of Excellence in Pharmaceutical Sciences (CEPS), GGS Indraprastha University, Sec. 16-C, Dwarka, New Delhi, 110078, India. aknarula58@gmail.com.

Abstract:
A series of thiourea based bifunctional organocatalysts having d-glucose as a core scaffold were synthesized and examined as catalysts for the asymmetric Michael addition reaction of aryl/alkyl trans-β-nitrostyrenes over cyclohexanone and other Michael donors having active methylene. Excellent enantioselectivities (<95%), diastereoselectivities (<99%), and yields (<99%) were attained under solvent free conditions using 10 mol% of 1d0. The obtained results were explained through DFT calculations using the B3LYP/6-311G(d,p)//B3LYP/6-31G(d) basic set. The QM/MM calculations revealed the role of cyclohexanone as a solvent as well as reactant in the rate determining step imparting 31.91 kcal mol-1 of energy towards the product formation.
Related Concept Videos
 01:08
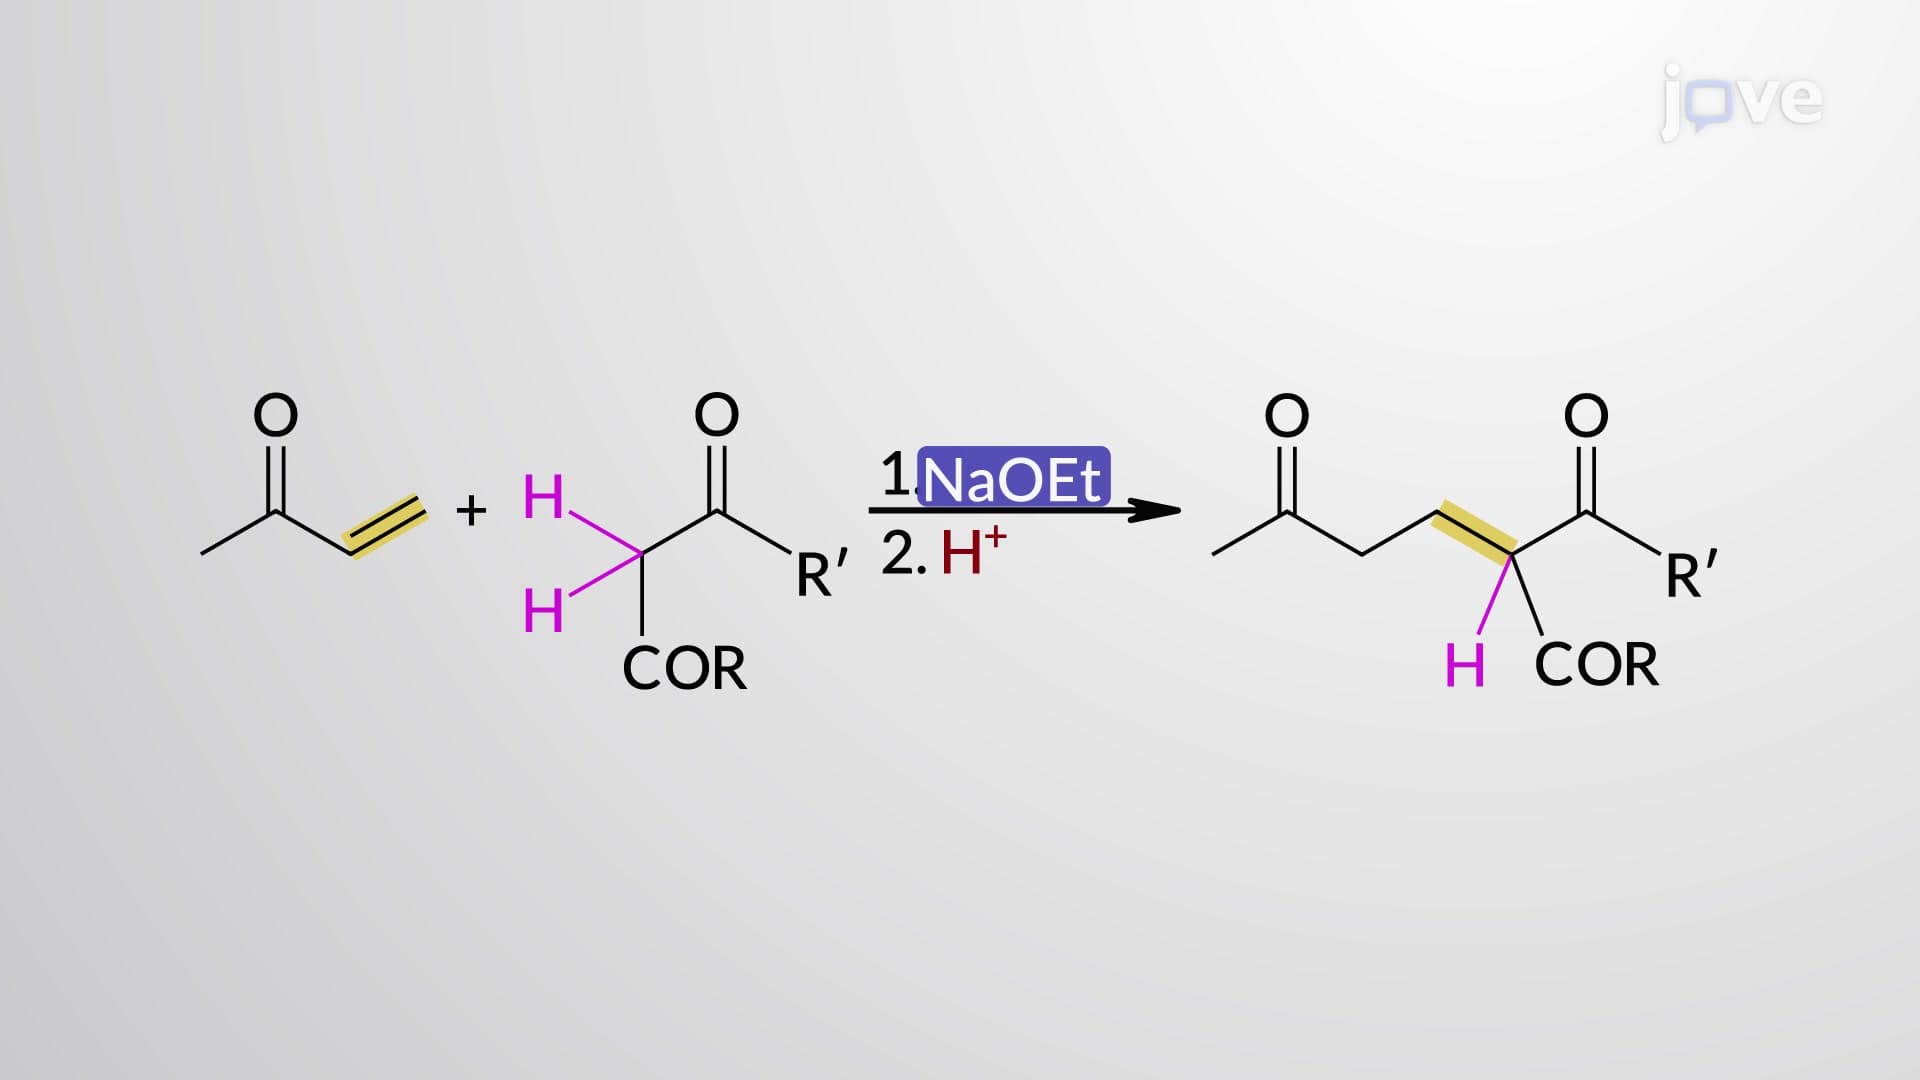
Conjugate Addition of Enolates: Michael Addition
 01:09
Conjugate Addition to α,β-Unsaturated Carbonyl Compounds
 01:27
Cyclohexenones via Michael Addition and Aldol Condensation: The Robinson Annulation
 02:15
Preparation of Alcohols via Addition Reactions
The acid-catalyzed addition of water to the double bond of alkenes is a large-scale industrial method used to synthesize low-molecular-weight alcohols. An acidic atmosphere is required to allow the hydrogen in the water molecule to act as an electrophile and attack the double bond in an alkene. The addition of a proton to the double bond creates a carbocation intermediate. The proton preferentially bonds to the less substituted end of the double bond to create a more stable carbocation...
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration