Related Experiment Video
Updated: Aug 6, 2026

The WinCF Model - An Inexpensive and Tractable Microcosm of a Mucus Plugged Bronchiole to Study the Microbiology of Lung Infections
Published on: May 8, 2017
Ultrastructural morphology of the lung in cystic fibrosis
M Dovey1, C L Wisseman, V L Roggli
1Department of Pathology, Duke University, Medical Center, Durham, North Carolina.

Abstract:
Cystic fibrosis (CF) is the most common lethal genetic disease among Caucasians, with much of the morbidity and most of the mortality related to pulmonary complications. The underlying defect in this disease has yet to be precisely defined, so it is somewhat surprising that a comprehensive study of the ultrastructural morphology of the lung in CF has not heretofore been reported. We used transmission electron microscopy to examine the small airways in 15 patients who had died of CF, and compared the findings with 15 disease controls with non-CF chronic airways disease and 15 patients with normal lung morphology. The lung parenchyma was also examined ultrastructurally in 7 patients with CF, 4 disease controls, and 4 normal lung cases. In addition, the literature regarding the ultrastructural morphology of the large airways in CF was reviewed. Patients with CF showed non-specific ciliary abnormalities, hyperplasia of mucous cells, increased numbers of pulmonary neuroendocrine and indeterminate cells, degeneration and sloughing of epithelial cells, and colonization of bacteria of the mucous layer of the small airways when compared with normal controls. Alveoli showed non-specific injury and regeneration of type II pneumocytes. However, these changes were all similar to those observed in the disease controls. Specifically, no cellular or subcellular ultrastructural abnormality unique to CF was observed. It is probable that the most useful ultrastructural approach to the lung in CF in future studies will involve X-ray microanalytical studies of ionic composition using cryotechniques.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:17
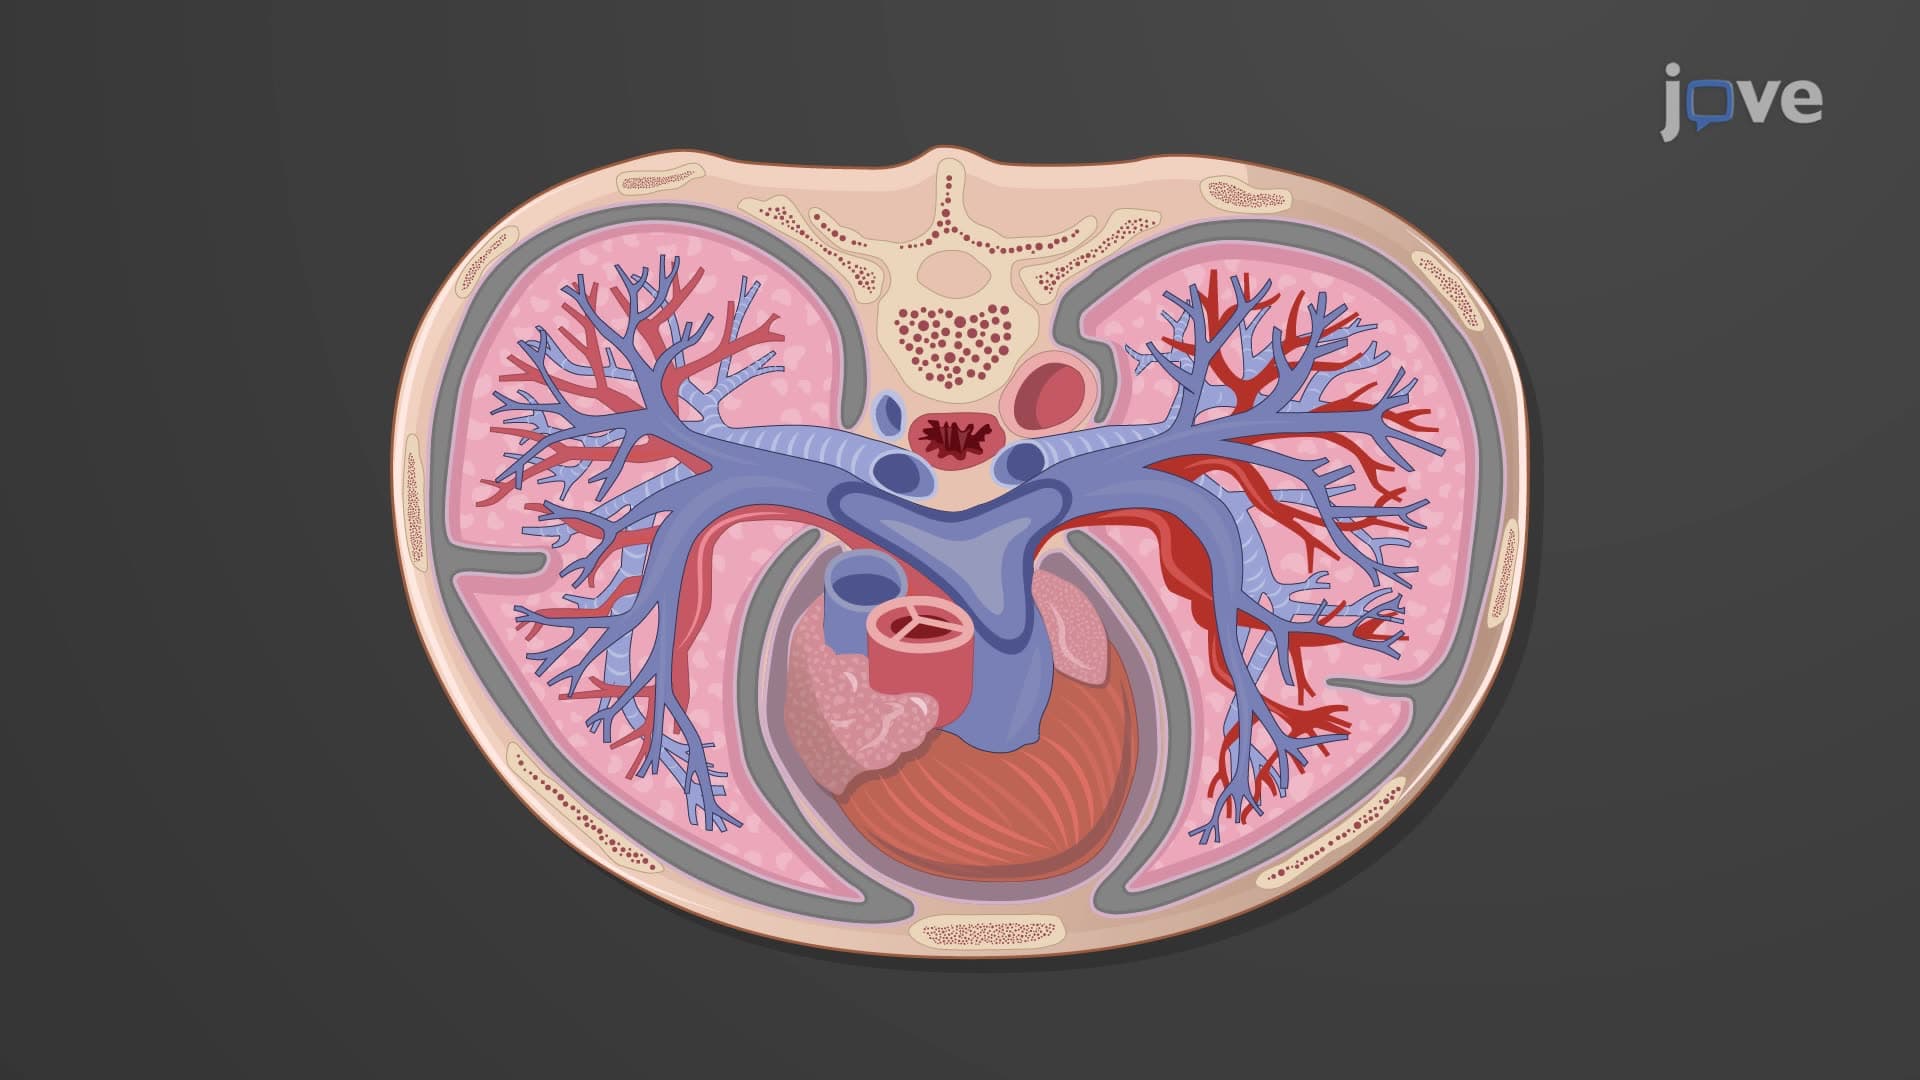
Gross Anatomy of the Lungs
 01:23
Chronic Obstructive Pulmonary Disease II: Emphysema
 01:24
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features