
Accurate Intermolecular Potential for the C60 Dimer: The Performance of Different Levels of Quantum Theory
Dmitry I Sharapa1, Johannes T Margraf2, Andreas Hesselmann3
1Computer-Chemie-Centrum, Department Chemie und Pharmazie, Friedrich-Alexander-Universität Erlangen-Nürnberg , Nägelsbachstraße 25, 91052 Erlangen, Germany.

Abstract:
The self-assembly of molecular building blocks is a promising route to low-cost nanoelectronic devices. It would be very appealing to use computer-aided design to identify suitable molecules. However, molecular self-assembly is guided by weak interactions, such as dispersion, which have long been notoriously difficult to describe with quantum chemical methods. In recent years, several viable techniques have emerged, ranging from empirical dispersion corrections for DFT to fast perturbation and coupled-cluster theories. In this work, we test these methods for the dimer of the prototypical building block for nanoelectronics, C60-fullerene. Benchmark quality data is obtained from DFT-based symmetry-adapted perturbation theory (SAPT), the adiabatic-connection fluctuation dissipation (ACFD) theorem using an adiabatic LDA kernel, and domain-based local pair natural orbital (DLPNO) coupled-pair and coupled-cluster methods. These benchmarks are used to evaluate economical dispersion-corrected DFT methods, double-hybrid DFT functionals, and second-order Møller-Plesset theory. Furthermore, we provide analytical fits to the benchmark interaction curves, which can be used for a coarse-grain description of fullerene self-assembly. These analytical expressions differ significantly from those reported previously based on bulk data.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:40
Preparation and Characterization of C60/Graphene Hybrid Nanostructures
Published on: May 15, 2018
Related Concept Videos
 03:13
Intermolecular Forces
 02:36
Molecular Geometry and Dipole Moments
 03:51
Molecular Orbital Theory II
 01:18
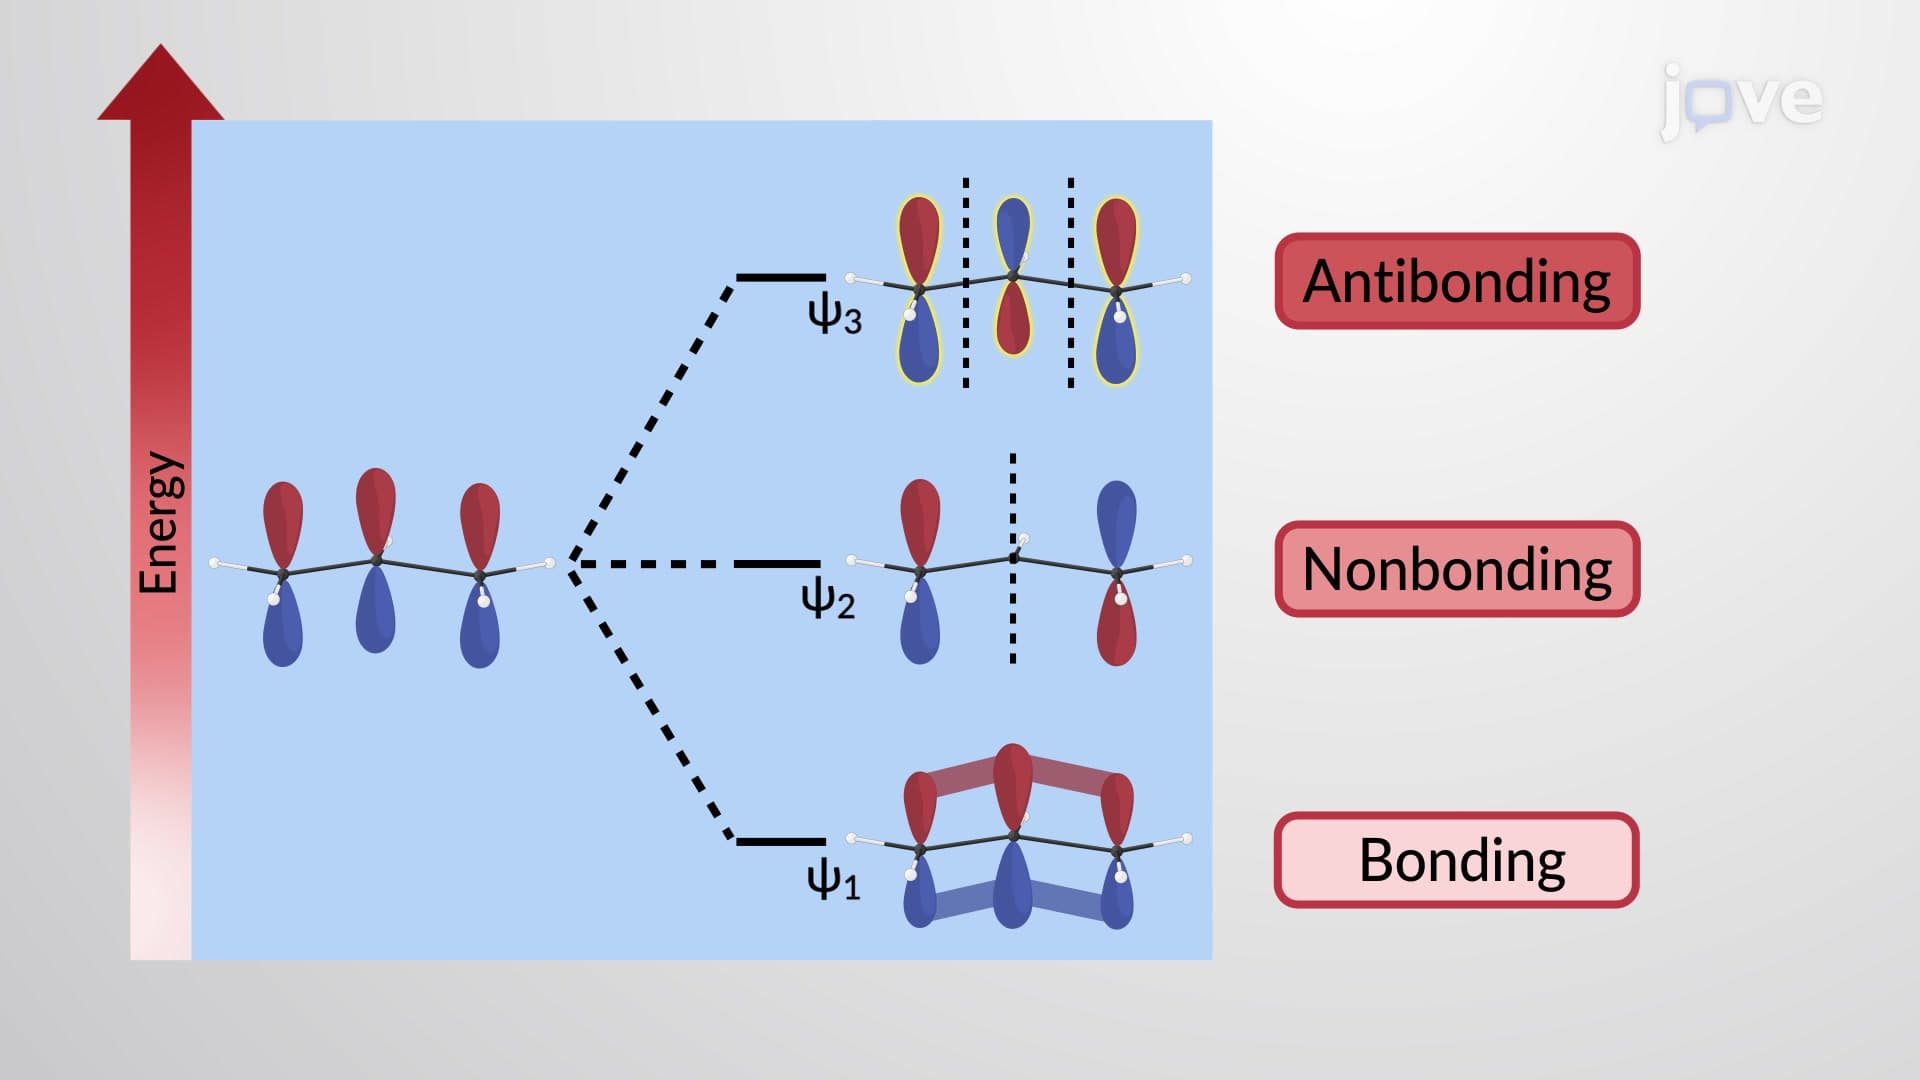
π Molecular Orbitals of the Allyl Cation and Anion
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 02:56
Intermolecular Forces and Physical Properties