Anharmonicity in the Vibrational Spectra of Naphthalene and Naphthalene-d8: Experiment and Theory
Shubhadip Chakraborty1, Subrata Banik2, Puspendu K Das1
1Department of Inorganic and Physical Chemistry, Indian Institute of Science , Bangalore, India.

Abstract:
In this paper, we report the gas phase infrared (IR) spectra of naphthalene and naphthalene-d8 recorded in the mid-infrared region (3200-500 cm-1) using a heated multipass long path gas cell. Several combination bands appear as shoulders and satellite peaks in the 3200-2600 cm-1region along with the C-H stretch fundamental bands. Experimental IR spectra of these molecules were systematically analyzed with vibrational self-consistent field (VSCF) theory, vibrational second order perturbation theory (VPT2) and vibrational couple cluster method (VCCM) with two different potential energy surfaces obtained using B3LYP and MP2 methods. A comparative study between these two PESs was made to match the observed spectra. Final assignment of the IR spectra of naphthalene and naphthalene-d8 was done using the VCCM with MP2 potential, which provided the best match.
More Related Videos
 12:07
Microwave-assisted Intramolecular Dehydrogenative Diels-Alder Reactions for the Synthesis of Functionalized Naphthalenes/Solvatochromic Dyes
Published on: April 1, 2013
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:14
NMR Spectroscopy of Aromatic Compounds
 01:08
IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations
 01:22
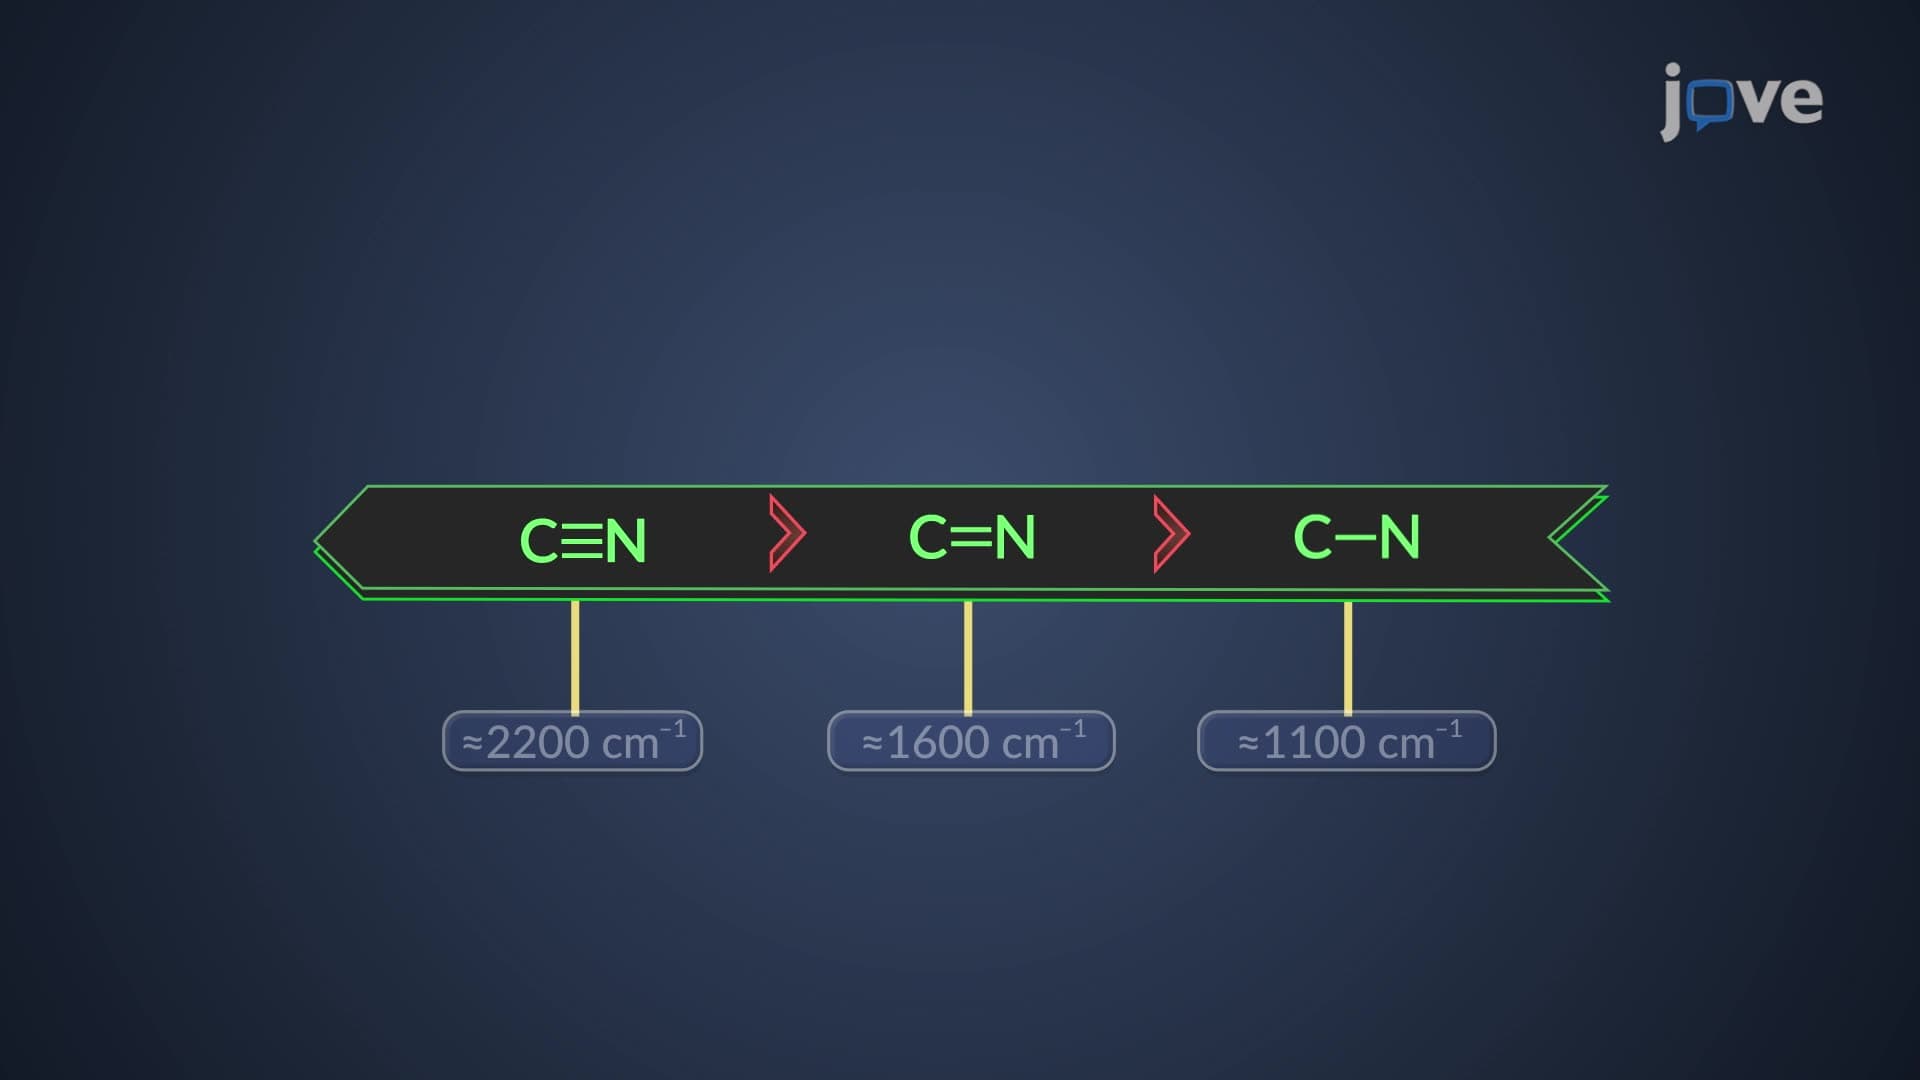
IR Frequency Region: Alkyne and Nitrile Stretching
Comparing the stretching vibrational frequency of C≡C triple bonds with that of double and single bonds, it is evident that C≡C triple bonds exhibit a higher stretching frequency than C=C double and C–C single bonds. Similarly, the C≡N triple bond...
 01:20
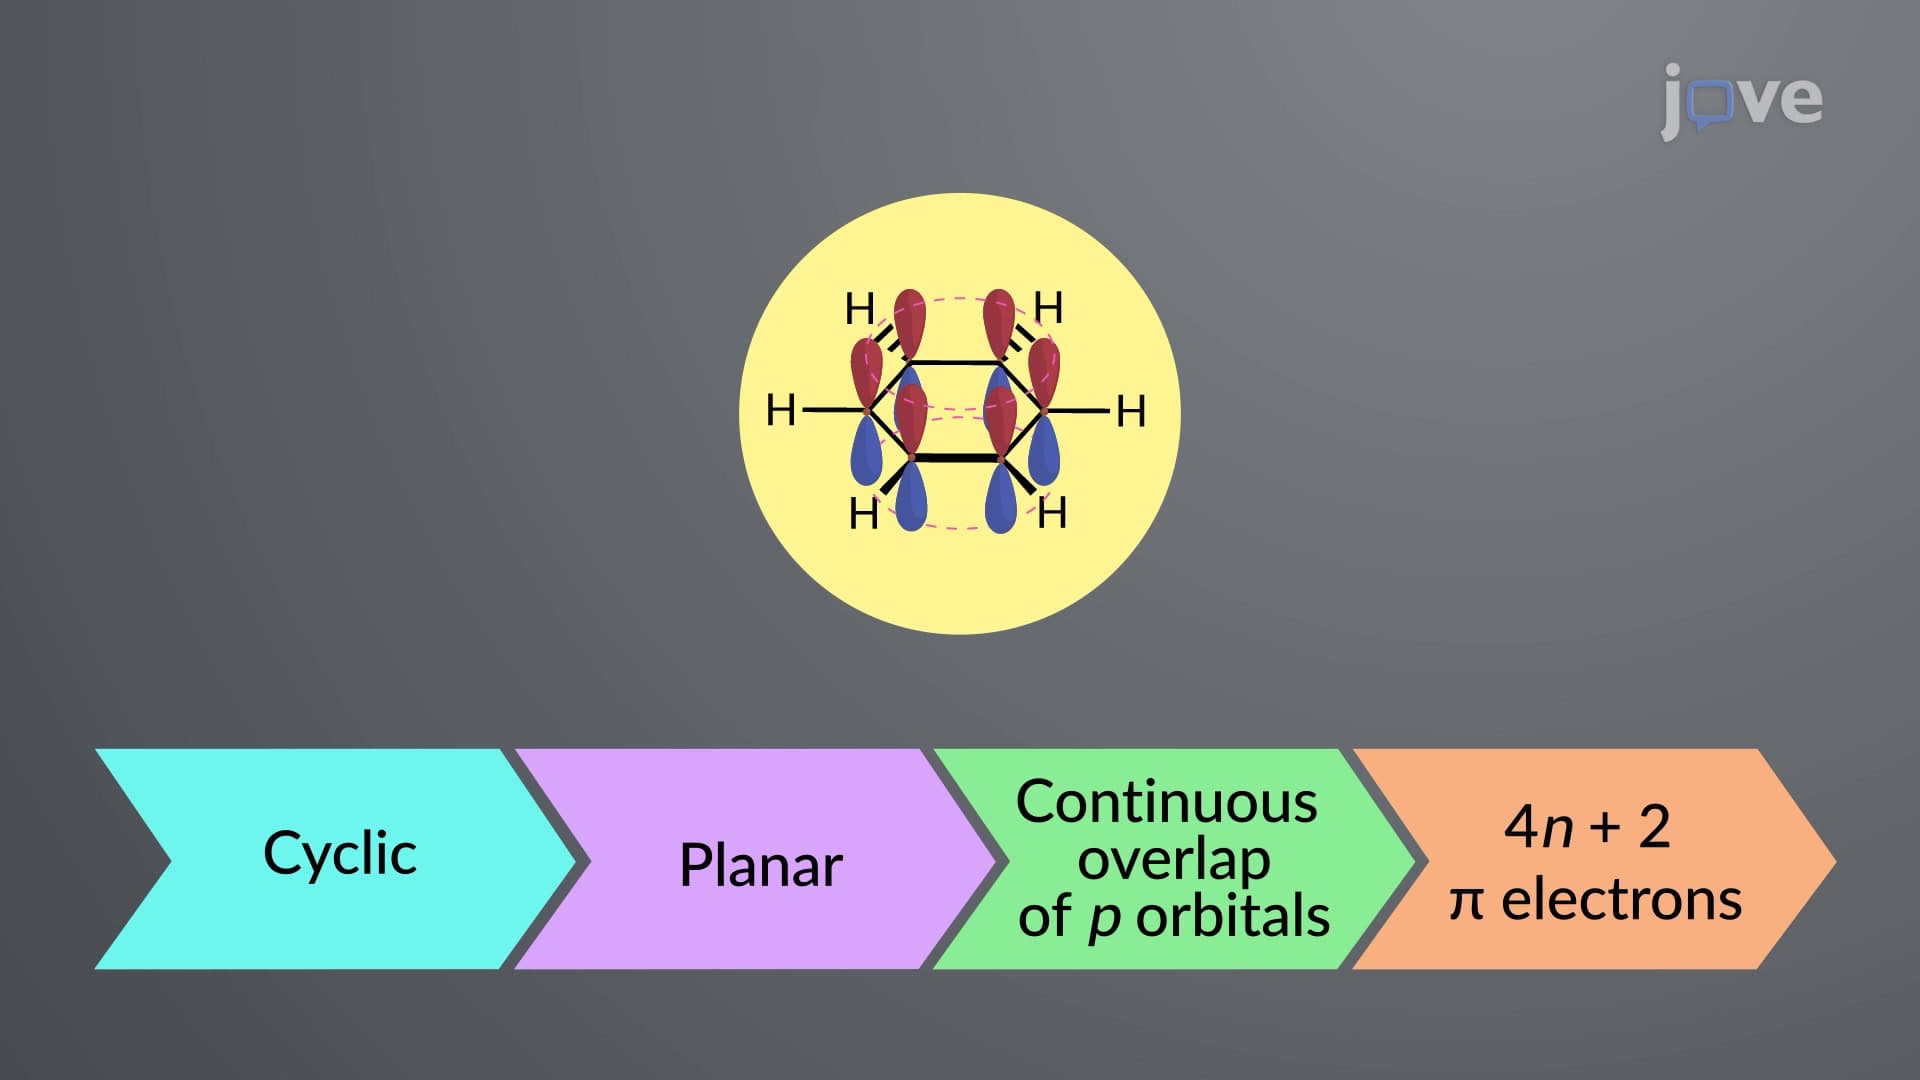
Criteria for Aromaticity and the Hückel 4n + 2 Rule
For the first time, Eric Hückel, a German chemical physicist, derived a set of structural features for a compound to be classified as aromatic. This is now known as Hückel’s rule or the 4n +...
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:37
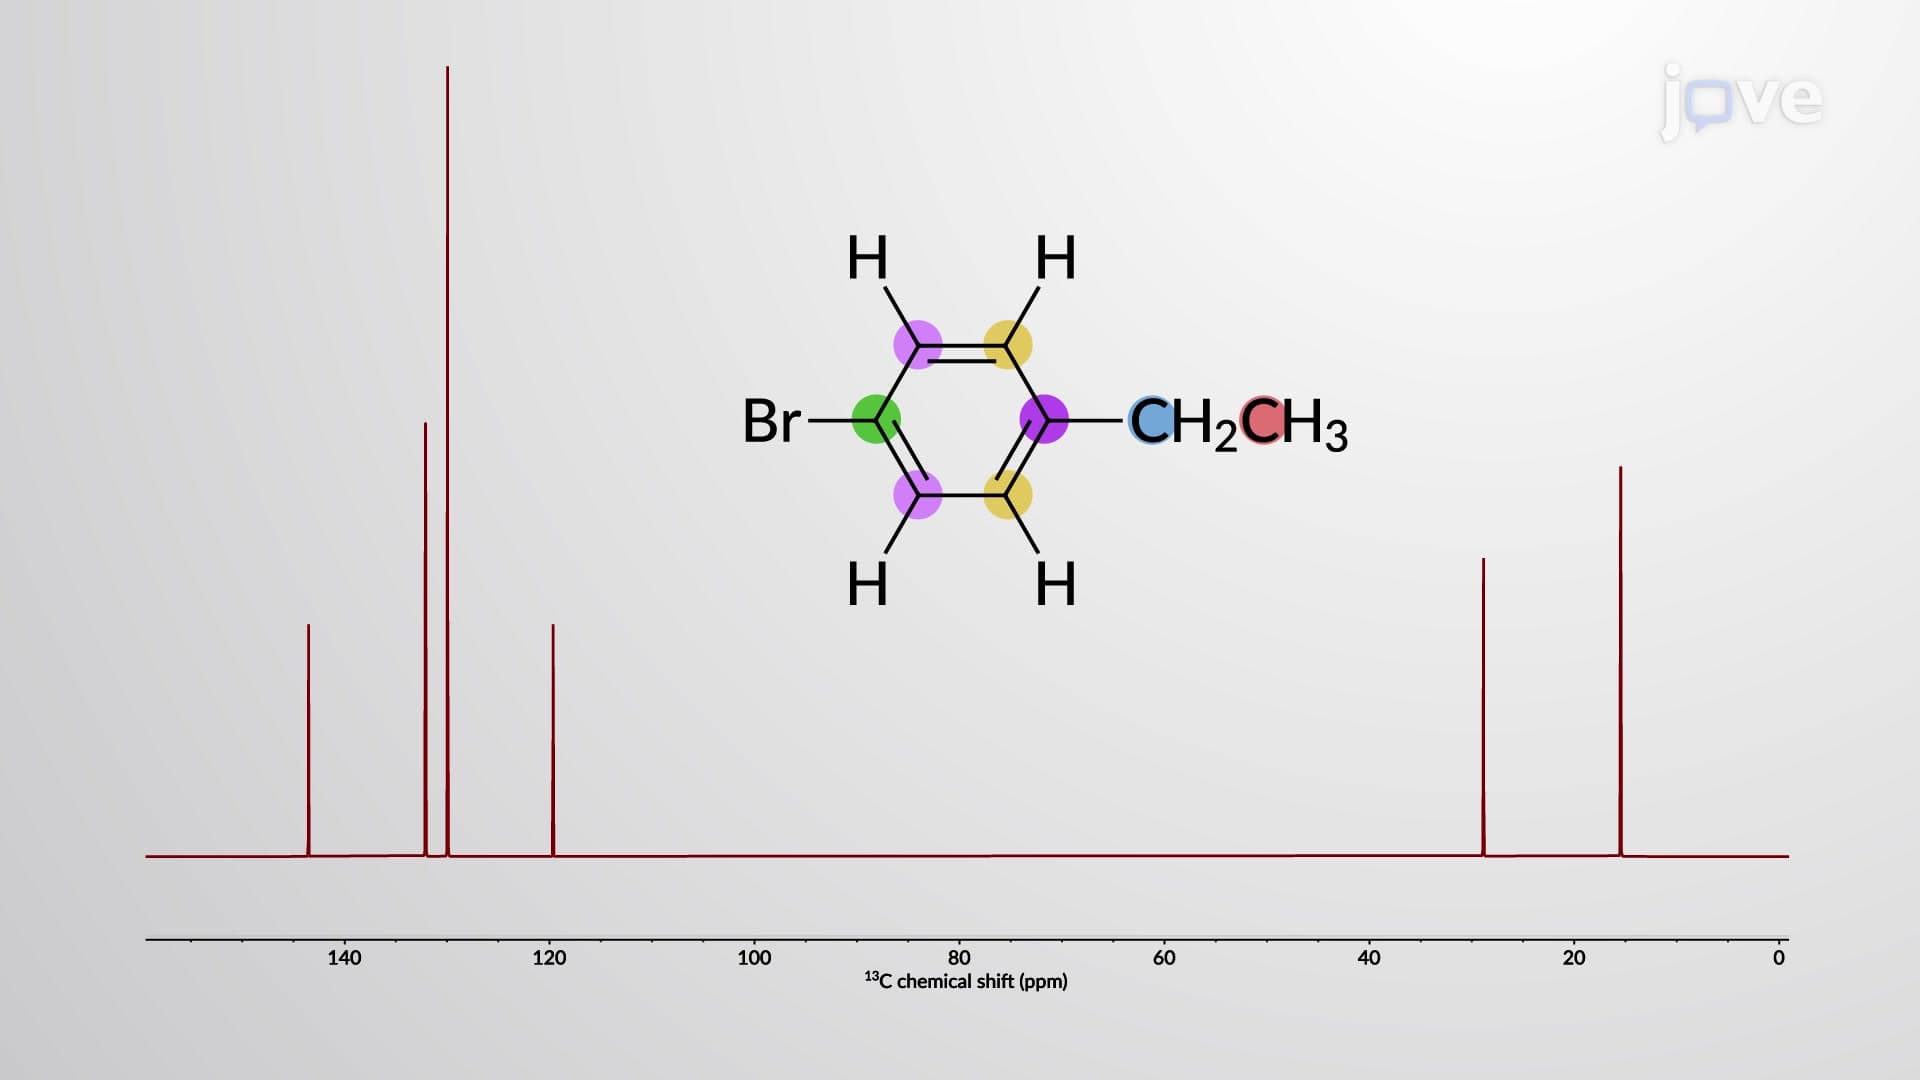
NMR Spectroscopy of Benzene Derivatives