Related Experiment Video
Updated: Mar 9, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Approximate Force Constants from Uncoupled Self-Consistent Field Perturbation Theory Using Nonhybrid Density
Zhigang Ni1, Krzysztof Wolinski2, Peter Pulay1
1Department of Chemistry and Biochemistry, University of Arkansas , Fayetteville, Arkansas 72701, United States.

Abstract:
Second derivatives of the molecular energy with respect to the nuclear coordinates (the nuclear Hessian or force constant matrix) are important for predicting infrared and Raman spectra, for calculating thermodynamic properties, for characterizing stationary states, and for guiding geometry optimization. However, their calculation for larger systems scales with molecular size one power higher than the calculation of the energy and the forces. The step responsible for the steep scaling of the nuclear Hessian is the coupled-perturbed self-consistent field (CP-SCF) iteration. This is omitted in the uncoupled SCF (UC-SCF) approximation. We have found that, though UC-SCF performs rather poorly at the Hartree-Fock and hybrid DFT levels, its performance for "pure" (non-hybrid) DFT is remarkably good. This is valid also for imaginary frequencies that characterize transition states. UC-SCF vibrational frequencies and normal modes are compared with coupled calculations for various exchange-correlation functionals including Hartree-Fock, and with basis sets ranging from simple to large for a variety of organic and some organometallic molecules. Their unexpectedly good performance makes them good candidates for calculating thermodynamic properties and for guiding difficult geometry optimizations, including the determination of transition states.
More Related Videos
 07:31
Author Spotlight: Advancing Cell Membrane Biophysics - Exploring Interactions and Challenges Through Experimental and Computational Approaches
Published on: September 1, 2023
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 01:28
Debye–Huckel–Onsager Conductance Equation
 01:10
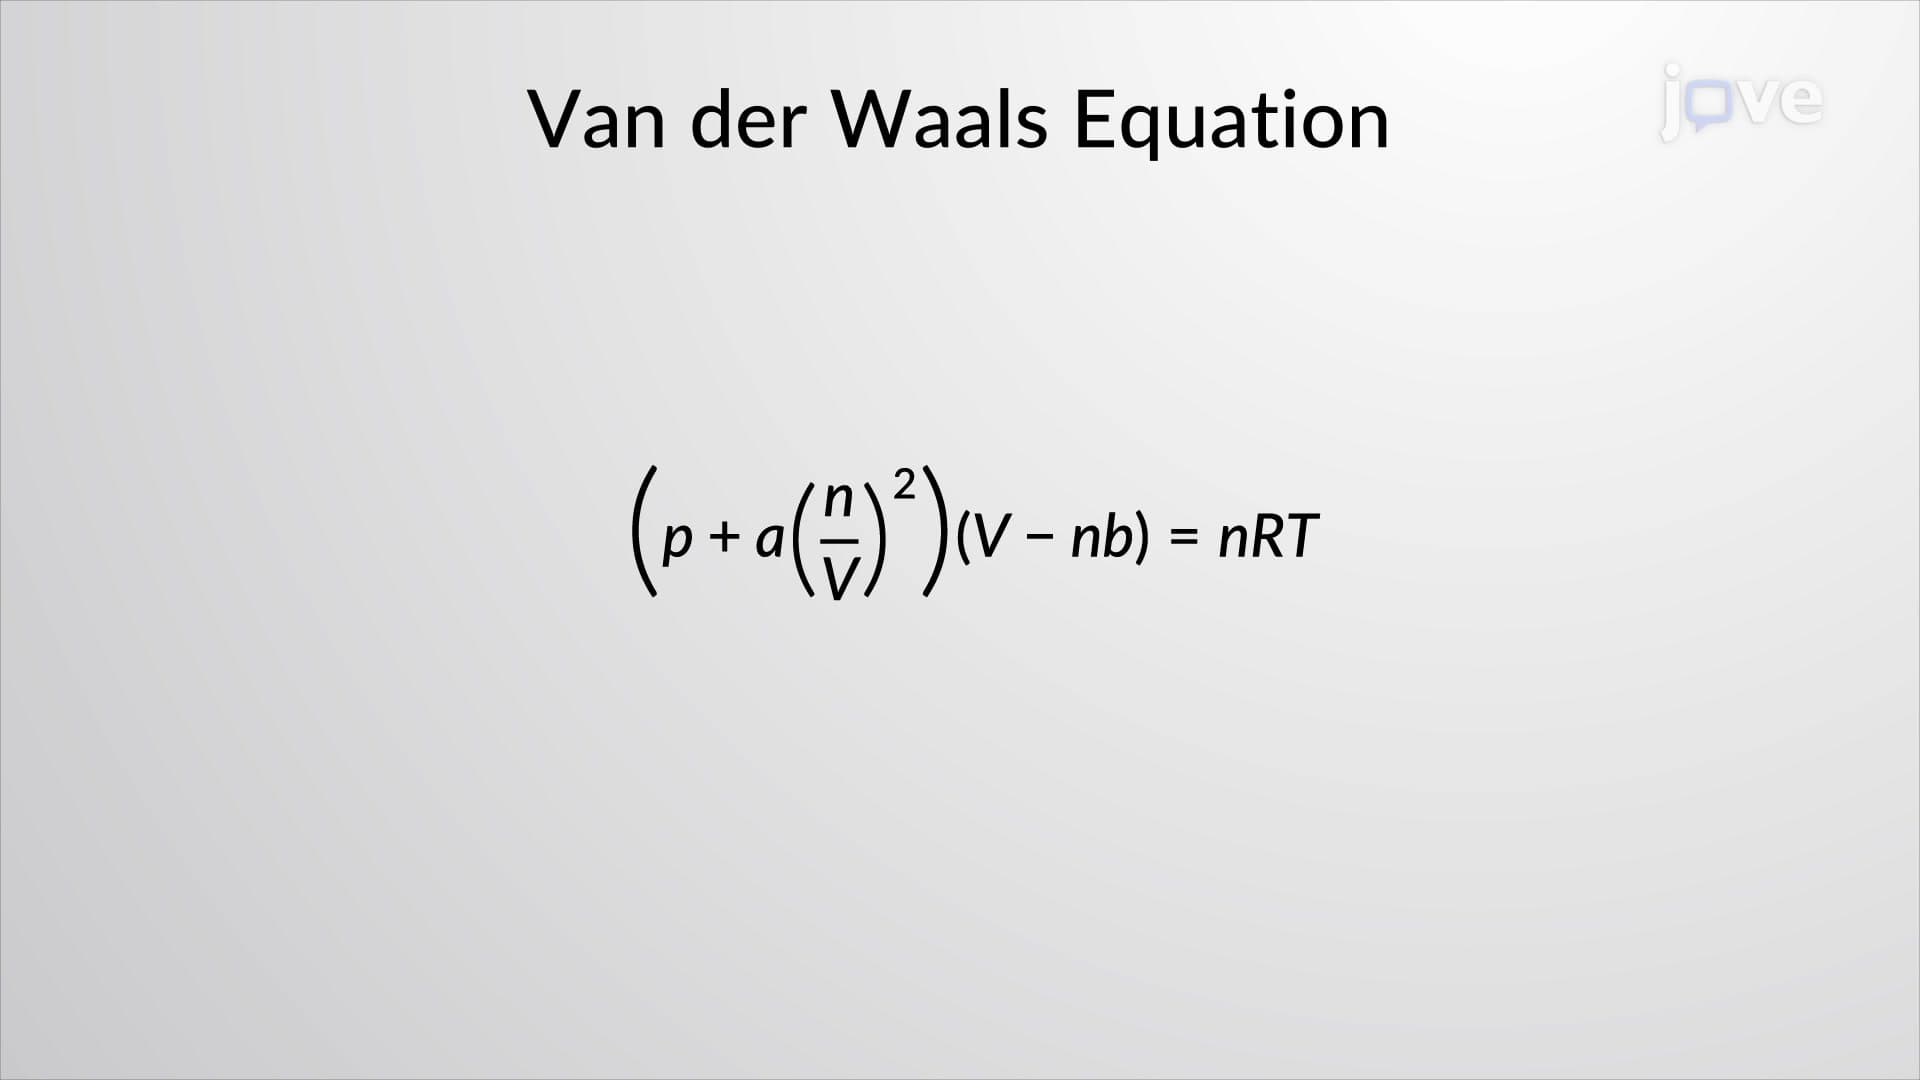
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:17
Non-conservative Forces
Also unlike their conservative counterparts, they are path-dependent; where the object starts and stops does matter. For example, a grinding wheel applies a...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:26
The Van der Waals Equation