Related Experiment Video
Updated: Mar 8, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Reliable prediction of adsorption isotherms via genetic algorithm molecular simulation
L LoftiKatooli1, A Shahsavand2
1Chemical Engineering Department, Ferdowsi university of Mashhad, Mashhad, Iran.

Abstract:
Conventional molecular simulation techniques such as grand canonical Monte Carlo (GCMC) strictly rely on purely random search inside the simulation box for predicting the adsorption isotherms. This blind search is usually extremely time demanding for providing a faithful approximation of the real isotherm and in some cases may lead to non-optimal solutions. A novel approach is presented in this article which does not use any of the classical steps of the standard GCMC method, such as displacement, insertation, and removal. The new approach is based on the well-known genetic algorithm to find the optimal configuration for adsorption of any adsorbate on a structured adsorbent under prevailing pressure and temperature. The proposed approach considers the molecular simulation problem as a global optimization challenge. A detailed flow chart of our so-called genetic algorithm molecular simulation (GAMS) method is presented, which is entirely different from traditions molecular simulation approaches. Three real case studies (for adsorption of CO2 and H2 over various zeolites) are borrowed from literature to clearly illustrate the superior performances of the proposed method over the standard GCMC technique. For the present method, the average absolute values of percentage errors are around 11% (RHO-H2), 5% (CHA-CO2), and 16% (BEA-CO2), while they were about 70%, 15%, and 40% for the standard GCMC technique, respectively.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 15:05
Deciphering the Structural Effects of Activating EGFR Somatic Mutations with Molecular Dynamics Simulation
Published on: May 20, 2020
Related Concept Videos
 01:29
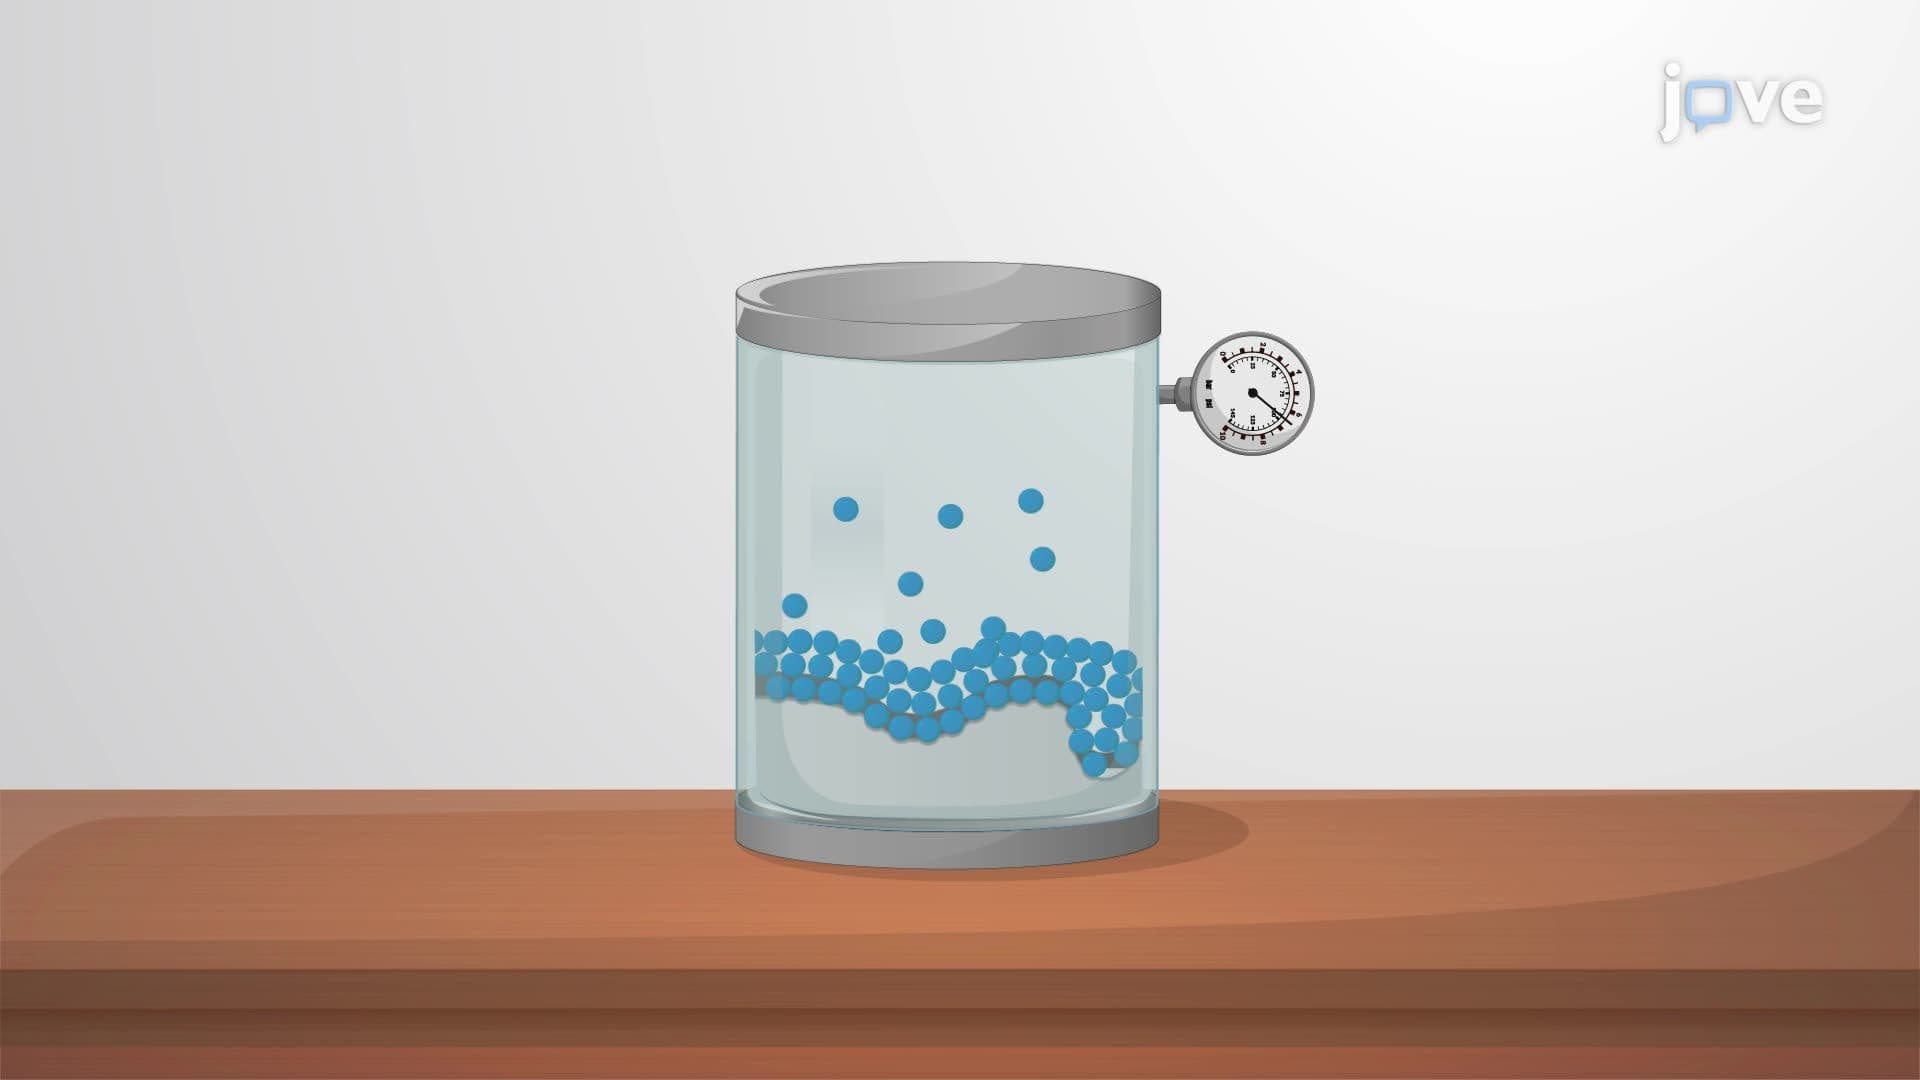
Adsorption Isotherms I
 01:25
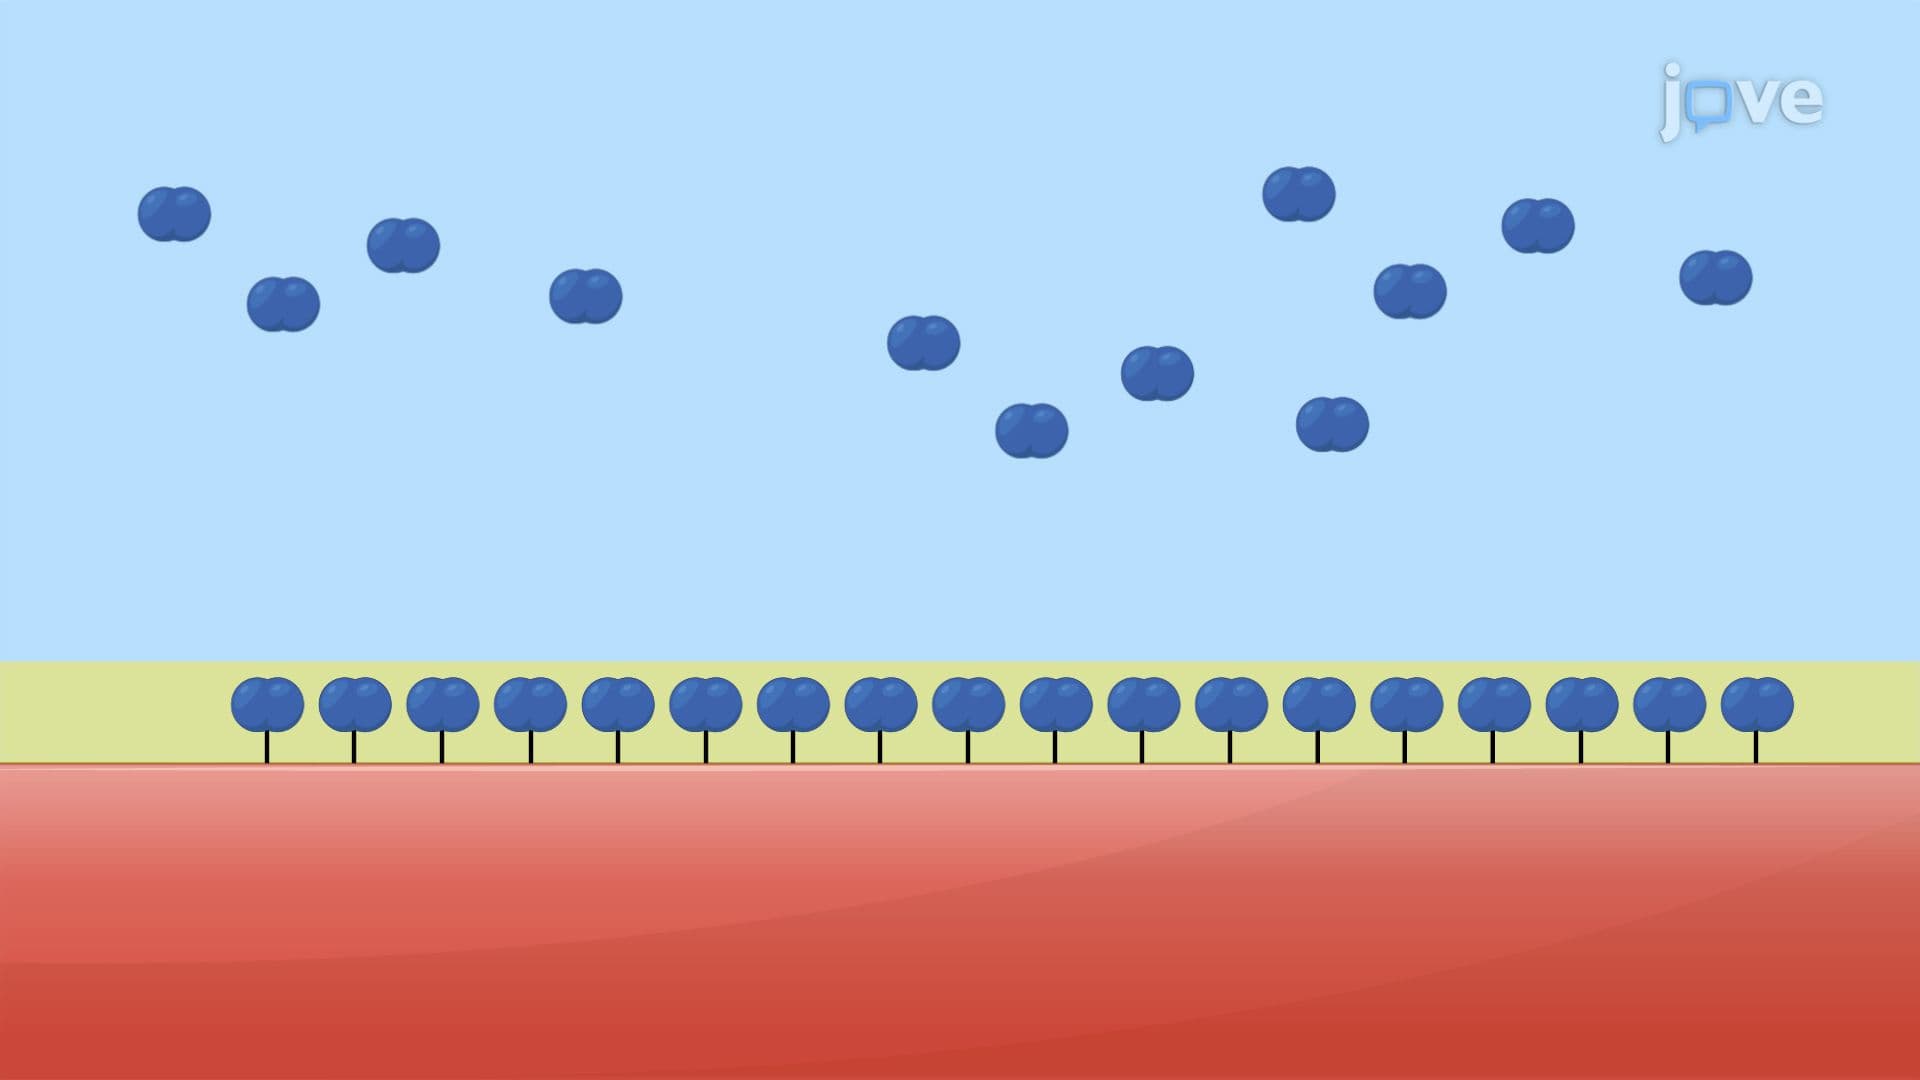
Adsorption Isotherms II
 01:28
Adsorption of Gases on Solids
 02:27
Predicting Molecular Geometry
 01:29
Mechanistic Models: Compartment Models in Algorithms for Numerical Problem Solving
In individual population analyses, different algorithms are employed, such as Cauchy's method, which uses a...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...